Time-dependent density functional theory study on direction-dependent electron and hole transfer processes in molecular systems
Related Experiment Video
Updated: Mar 8, 2026
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Pouya Partovi-Azar1, Payam Kaghazchi1
1Institute of Chemistry and Biochemistry, Physical and Theoretical Chemistry, Freie Universität Berlin, Takustr. 3, Berlin, 14195, Germany.

Abstract:
We report on real-time time-dependent density functional theory calculations on direction-dependent electron and hole transfer processes in molecular systems. As a model system, we focus on α-sulfur. It is shown that time scale of the electron transfer process from a negatively charged S8 molecule to a neighboring neutral monomer is comparable to that of a strong infrared-active molecular vibrations of the dimer with one negatively charged monomer. This results in a strong coupling between the electrons and the nuclei motion which eventually leads to S8 ring opening before the electron transfer process is completed. The open-ring structure is found to be stable. The similar infrared-active peak in the case of hole transfer, however, is shown to be very weak and hence no significant scattering by the nuclei is possible. The presented approach to study the charge transfer processes in sulfur has direct applications in the increasingly growing research field of charge transport in molecular systems. © 2017 Wiley Periodicals, Inc.
More Related Videos
Related Concept Videos
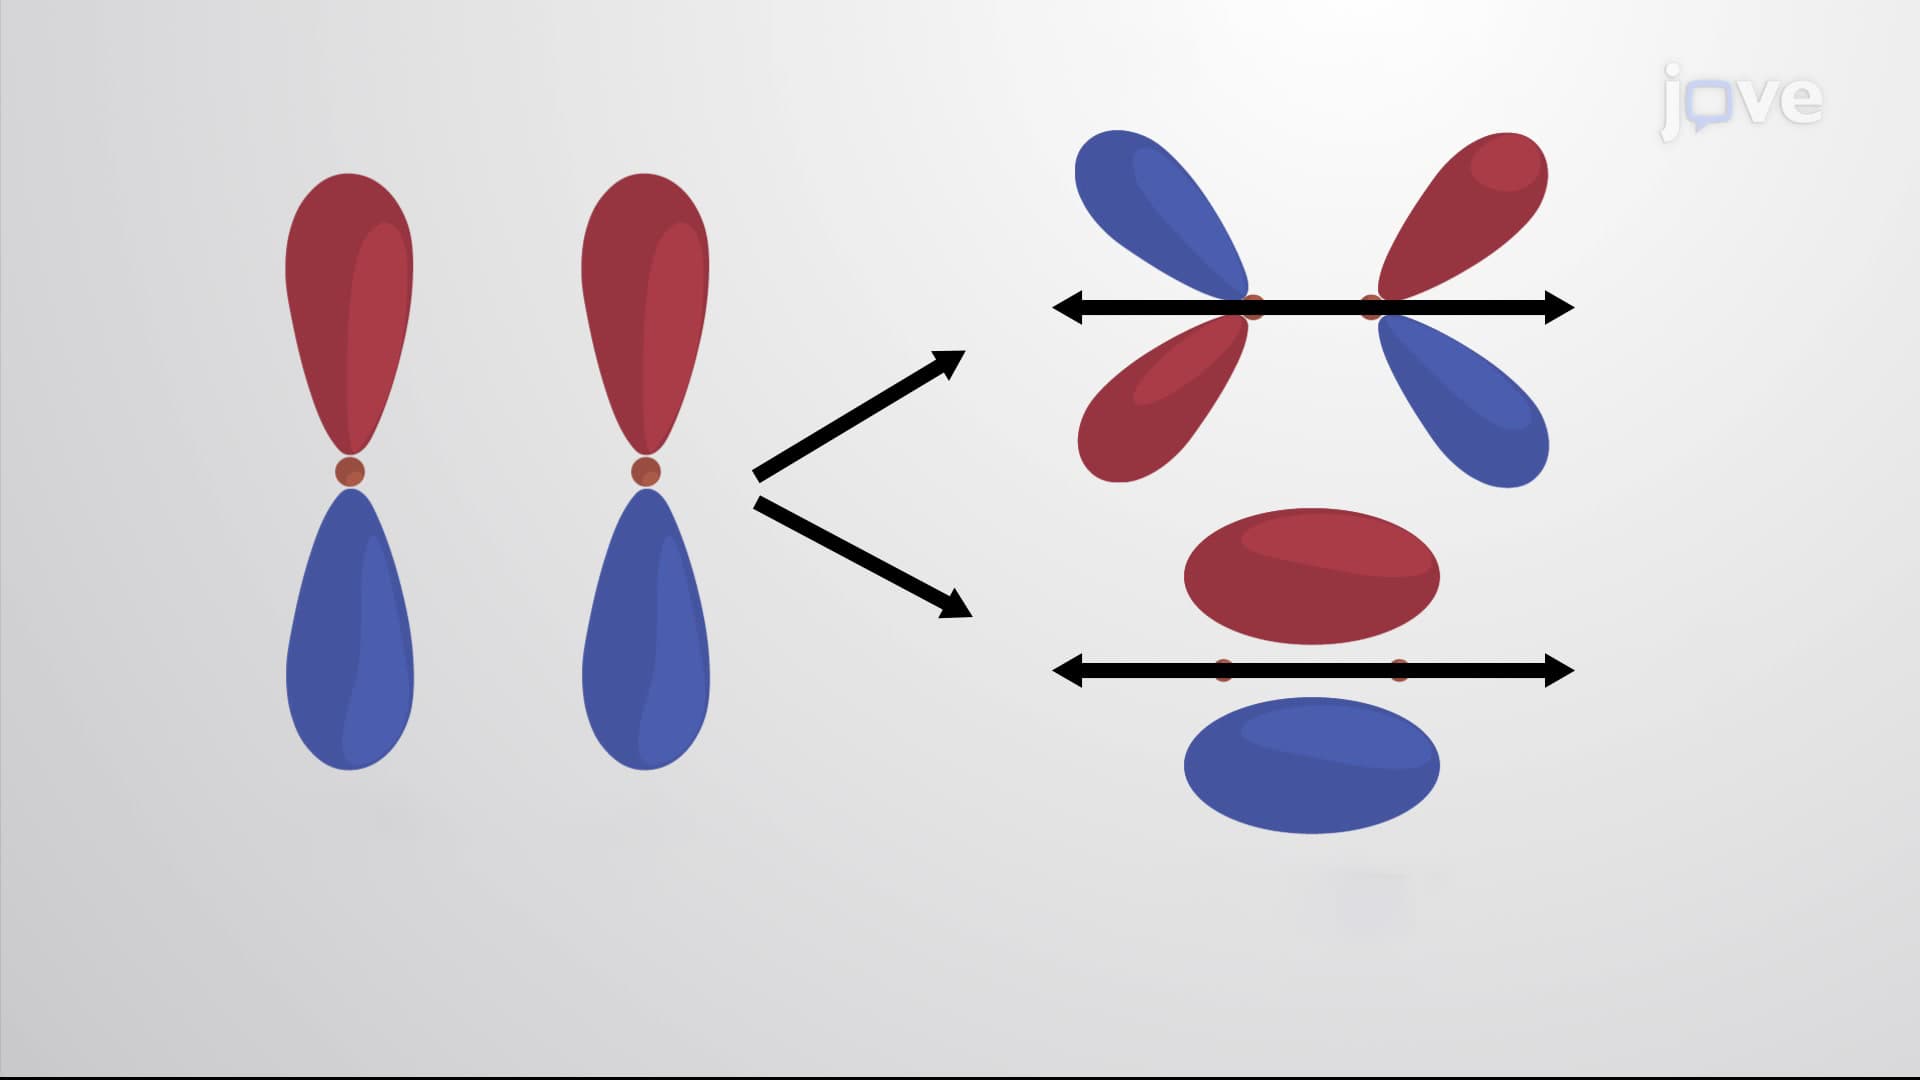 02:40
MO Theory and Covalent Bonding
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:27
The Debye–Hückel Theory of Electrolyte Solutions
 03:51
Molecular Orbital Theory II
 01:30
The Electrical Double Layer
 02:35
Molecular Orbital Theory I