Related Experiment Video
Updated: Mar 8, 2026

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Integrating Multiple Receptor Conformation Docking and Multi Dimensional QSAR for Enhancing Accuracy of Binding
Vangala Radhika1, Hassan A Jaraf1,2, Sivan S Kanth1
1Department of Chemistry, Molecular Modeling and Medicinal Chemistry Group, University College of Science, Osmania University, Hyderabad - 500007, India.

Background:
The accuracy of molecular conformation for Quantitative Structure Activity Relationship (QSAR) studies is an important criteria, and the most favourable bioactive conformer selection is a tough task. Correct ligand alignment as input for 3D-QSAR is an important step that is prone to human biases. Multiple-dimensional QSAR (mQSAR) approach provides a promising alternative to classic 3D-QSAR for drug discovery purposes.
Objective:
Obtaining ligand conformations from multiple receptor conformation docking (MRCD) will reduce the margin of error by incorporating the receptor based alignment of ligand conformations. To validate this assumption we performed 6D QSAR studies on reported HIV-1 protease inhibitors using Quasar 6.0.
Materials & Method:
The ensemble of conformation was obtained by MRCD of ligands in thirteen crystal structures of HIV-1 protease. 6D QSAR model was built using 65 cyclic urea molecules reported as HIV-1 protease inhibitors. Predictive ability of the model was validated using 35 cyclic urea molecules as test set. External predictive ability of the model was evaluated using a set of 24 HIV-1 protease inhibitors having varied structural scaffold.
Result:
6D QSAR model obtained showed a reliable cross-validated r2(q2) of 0.899, r2(classic) of 0.908 and yielded a predictive r2 (p2) of 0.527. The ratio of q2/r2 was 0.991 and p2/q2 was 0.586 for external test set.
Conclusion:
The QSAR results invariably suggest that our approach is suitable for the identification of molecules having HIV-1 protease inhibitory potency. The underlying philosophy combines flexible docking for the identification of the binding modes and 6D QSAR for their quantification.


Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
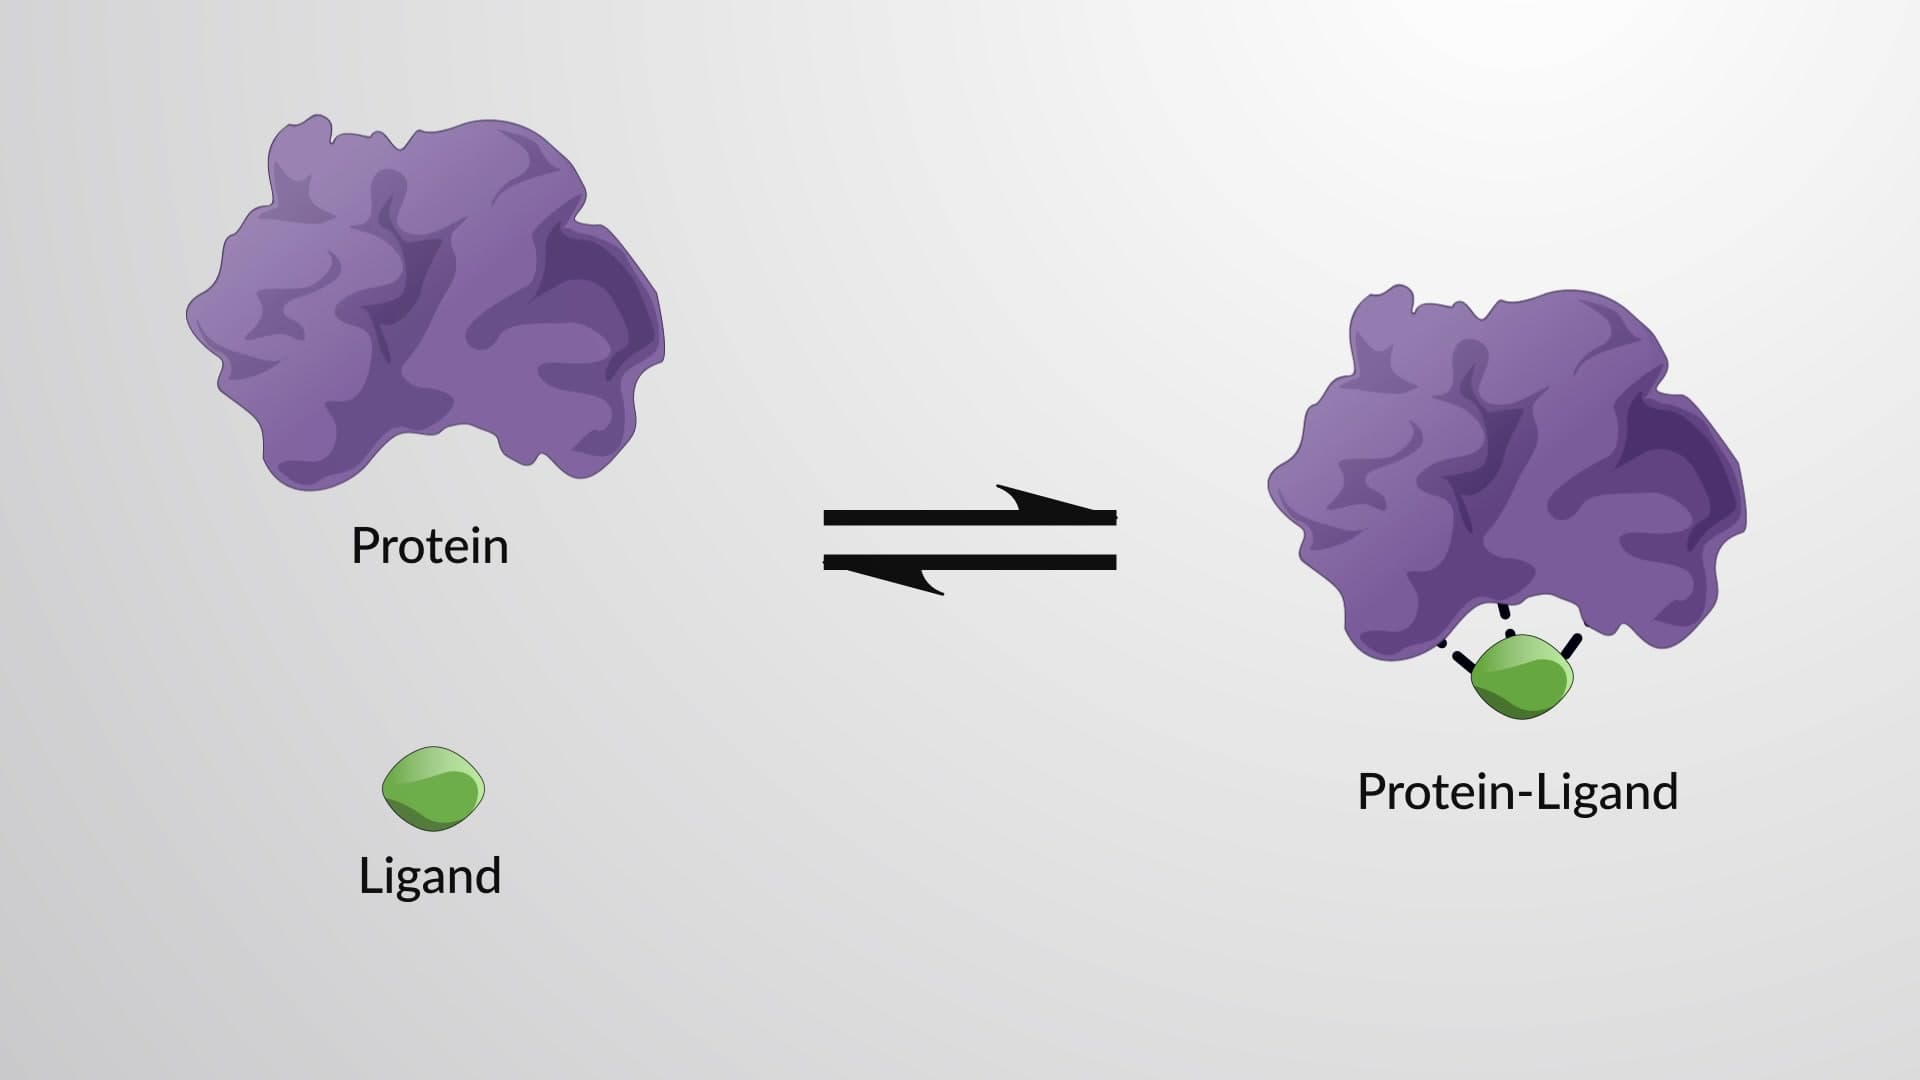
The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 00:49
Ligand Binding and Linkage