Related Experiment Video
Updated: Mar 8, 2026
Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
Rapid Design of Knowledge-Based Scoring Potentials for Enrichment of Near-Native Geometries in Protein-Protein
Alexander Sasse1, Sjoerd J de Vries1, Christina E M Schindler1
1Physik Department T38, Technische Universität München, James-Franck-Straße, Garching, Germany.

Abstract:
Protein-protein docking protocols aim to predict the structures of protein-protein complexes based on the structure of individual partners. Docking protocols usually include several steps of sampling, clustering, refinement and re-scoring. The scoring step is one of the bottlenecks in the performance of many state-of-the-art protocols. The performance of scoring functions depends on the quality of the generated structures and its coupling to the sampling algorithm. A tool kit, GRADSCOPT (GRid Accelerated Directly SCoring OPTimizing), was designed to allow rapid development and optimization of different knowledge-based scoring potentials for specific objectives in protein-protein docking. Different atomistic and coarse-grained potentials can be created by a grid-accelerated directly scoring dependent Monte-Carlo annealing or by a linear regression optimization. We demonstrate that the scoring functions generated by our approach are similar to or even outperform state-of-the-art scoring functions for predicting near-native solutions. Of additional importance, we find that potentials specifically trained to identify the native bound complex perform rather poorly on identifying acceptable or medium quality (near-native) solutions. In contrast, atomistic long-range contact potentials can increase the average fraction of near-native poses by up to a factor 2.5 in the best scored 1% decoys (compared to existing scoring), emphasizing the need of specific docking potentials for different steps in the docking protocol.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:04
Protein-protein Interfaces
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:24
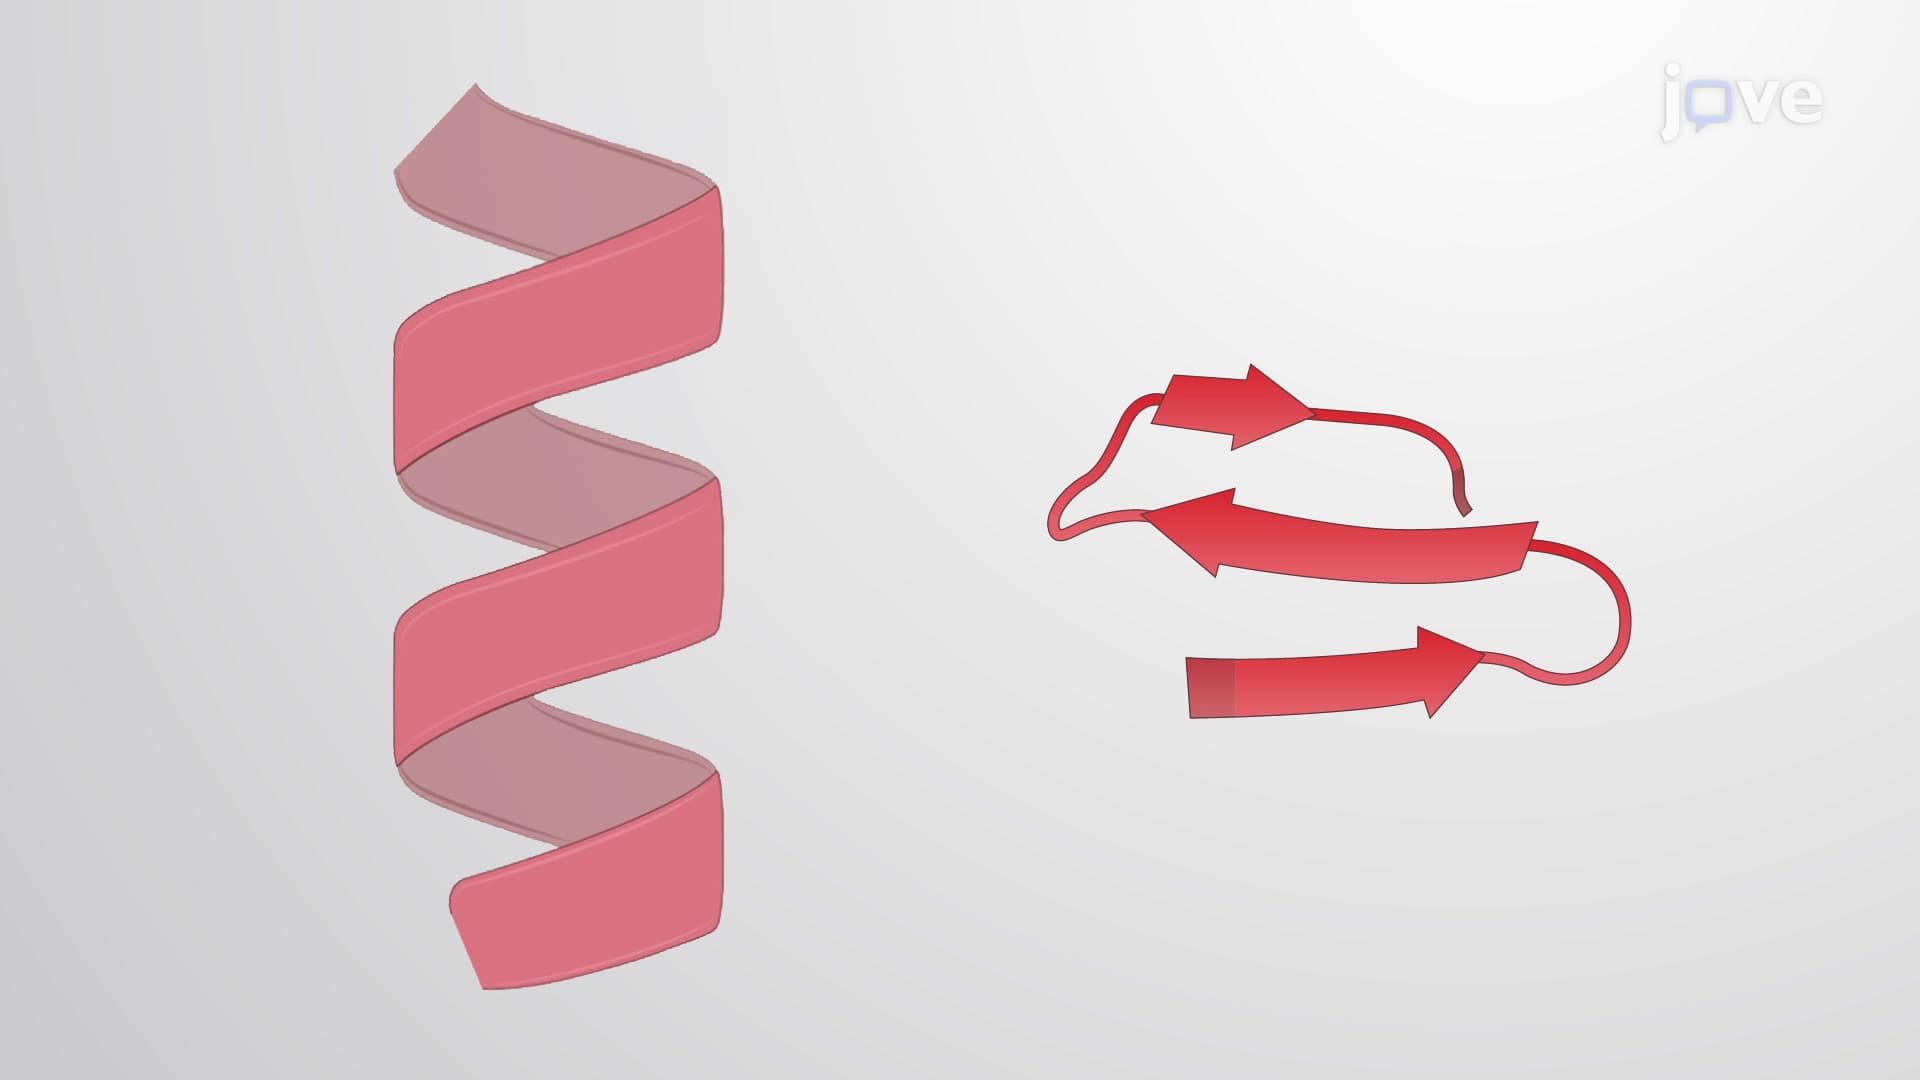
Protein Organization
The primary structure of a protein is its amino acid sequence....
 02:27
Predicting Molecular Geometry