Combined CoMFA and CoMSIA 3D-QSAR study of benzimidazole and benzothiophene derivatives with selective affinity for the CB2 cannabinoid receptor
Related Experiment Video
Updated: Mar 8, 2026
Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Javier Romero-Parra1, Hery Chung1, Ricardo A Tapia2
1Pharmacy Department, Faculty of Chemistry, Pontificia Universidad Católica de Chile, Vicuña Mackenna 4860, Santiago, Chile.

Abstract:
The preceding years have brought an exponential increase in our understanding of the endocannabinoid system (ECS), including the knowledge of CB1 and CB2 cannabinoid receptors, endocannabinoids, and the enzymes that synthesize and degrade endocannabinoids. Among these ECS components CB2 receptors have been the subject of considerable attention, primarily due to their promising therapeutic potential to treat numerous pathologies while avoiding the adverse psychotropic effects that can accompany CB1 receptor-based therapies. Recently, our research group has reported a new series of non-cytotoxic benzo[d]imidazoles and benzo[b]thiophenes displaying high CB2/CB1 selectivity index. In order to investigate the structural requirements for CB2 ligands and to derive a predictive model that can be used for the design of novel selective CB2 ligands, a three-dimensional quantitative structure-activity relationship (3D-QSAR) study was performed on the above mentioned chemical series employing comparative molecular field analysis (CoMFA) and comparative molecular similarity index analysis (CoMSIA) techniques. The CoMFA and CoMSIA models displayed high external predictability (rpred2 0.919 and 0.908) and good statistical robustness. Valuable information regarding the steric, electrostatic and hydrophobic properties of the molecules was obtained, and several modifications around both heterocycles were evaluated with the aim to generate new promising series of benzo[d]imidazoles and benzo[b]thiophenes derivatives displaying high CB2 selectivity and low toxicity.
More Related Videos
 10:17
Creating Highly Specific Chemically Induced Protein Dimerization Systems by Stepwise Phage Selection of a Combinatorial Single-Domain Antibody Library
Published on: January 14, 2020
 09:09
Preparation and Delivery of Protein Microcrystals in Lipidic Cubic Phase for Serial Femtosecond Crystallography
Published on: September 20, 2016
Related Concept Videos
 01:21
Chemotherapy-Induced Nausea and Vomiting: Cannabinoids
Two synthetic agonists of THC,...
 01:18
Directing and Steric Effects in Disubstituted Benzene Derivatives
 01:37
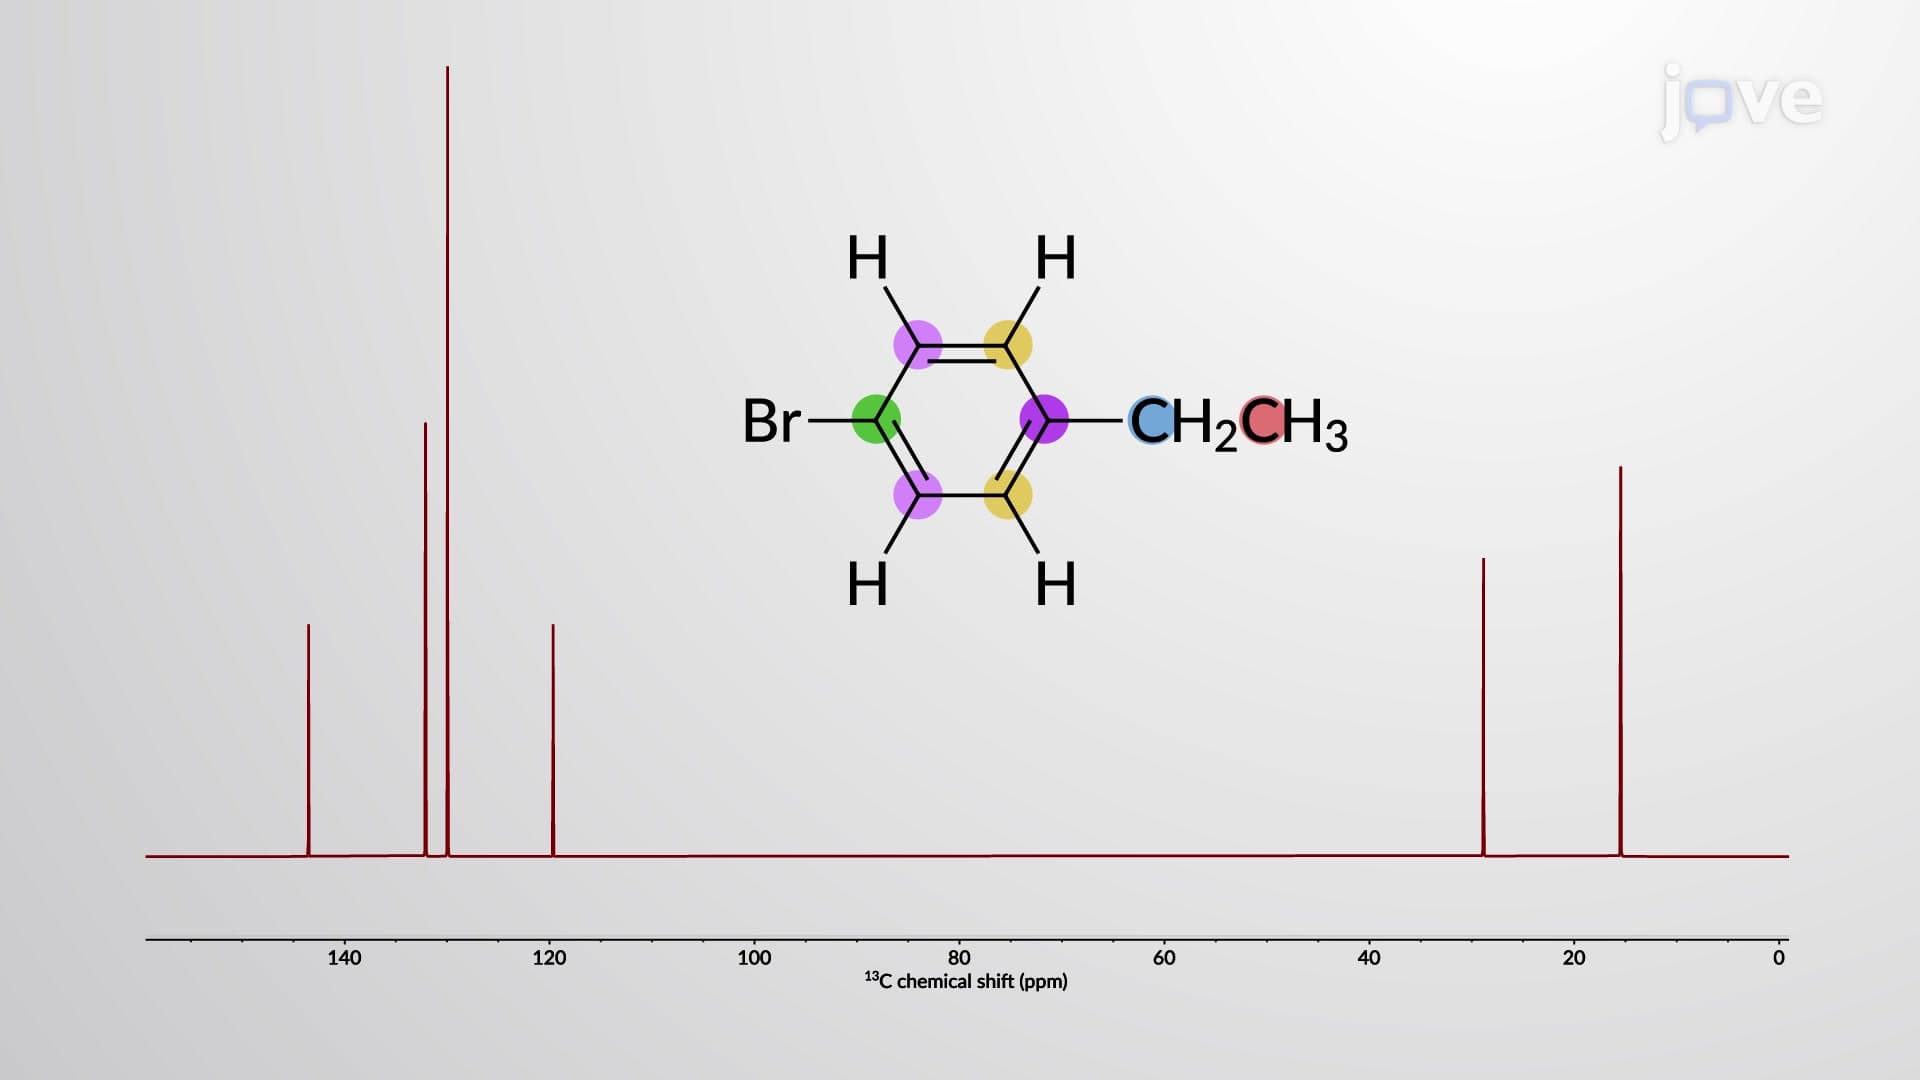
NMR Spectroscopy of Benzene Derivatives
 01:18
Structure of Benzene: Molecular Orbital Model
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:30
Quantitative Aspects of Drug-Receptor Interaction