Evaluation of DFT-D3 dispersion corrections for various structural benchmark sets
Heiner Schröder1, Jens Hühnert1, Tobias Schwabe1
1Center for Bioinformatics and Institute of Physical Chemistry, University of Hamburg, Bundesstraße 43, 20146 Hamburg, Germany.

Abstract:
We present an evaluation of our newly developed density functional theory (DFT)-D3 dispersion correction D3(CSO) in comparison to its predecessor D3(BJ) for geometry optimizations. Therefore, various benchmark sets covering bond lengths, rotational constants, and center of mass distances of supramolecular complexes have been chosen. Overall both corrections give accurate structures and show no systematic differences. Additionally, we present an optimized algorithm for the computation of the DFT-D3 gradient, which reduces the formal scaling of the gradient calculation from O(N3) to O(N2).
More Related Videos
 08:44
Measurements of Long-range Electronic Correlations During Femtosecond Diffraction Experiments Performed on Nanocrystals of Buckminsterfullerene
Published on: August 22, 2017
 09:56
Adapting Taylor Dispersion to Measure the Dispersion Coefficient of Electrolyte Solutions via an Accessible Microfluidic Setup
Published on: October 7, 2025
Related Concept Videos
 01:15
Distance Corrections
 01:20
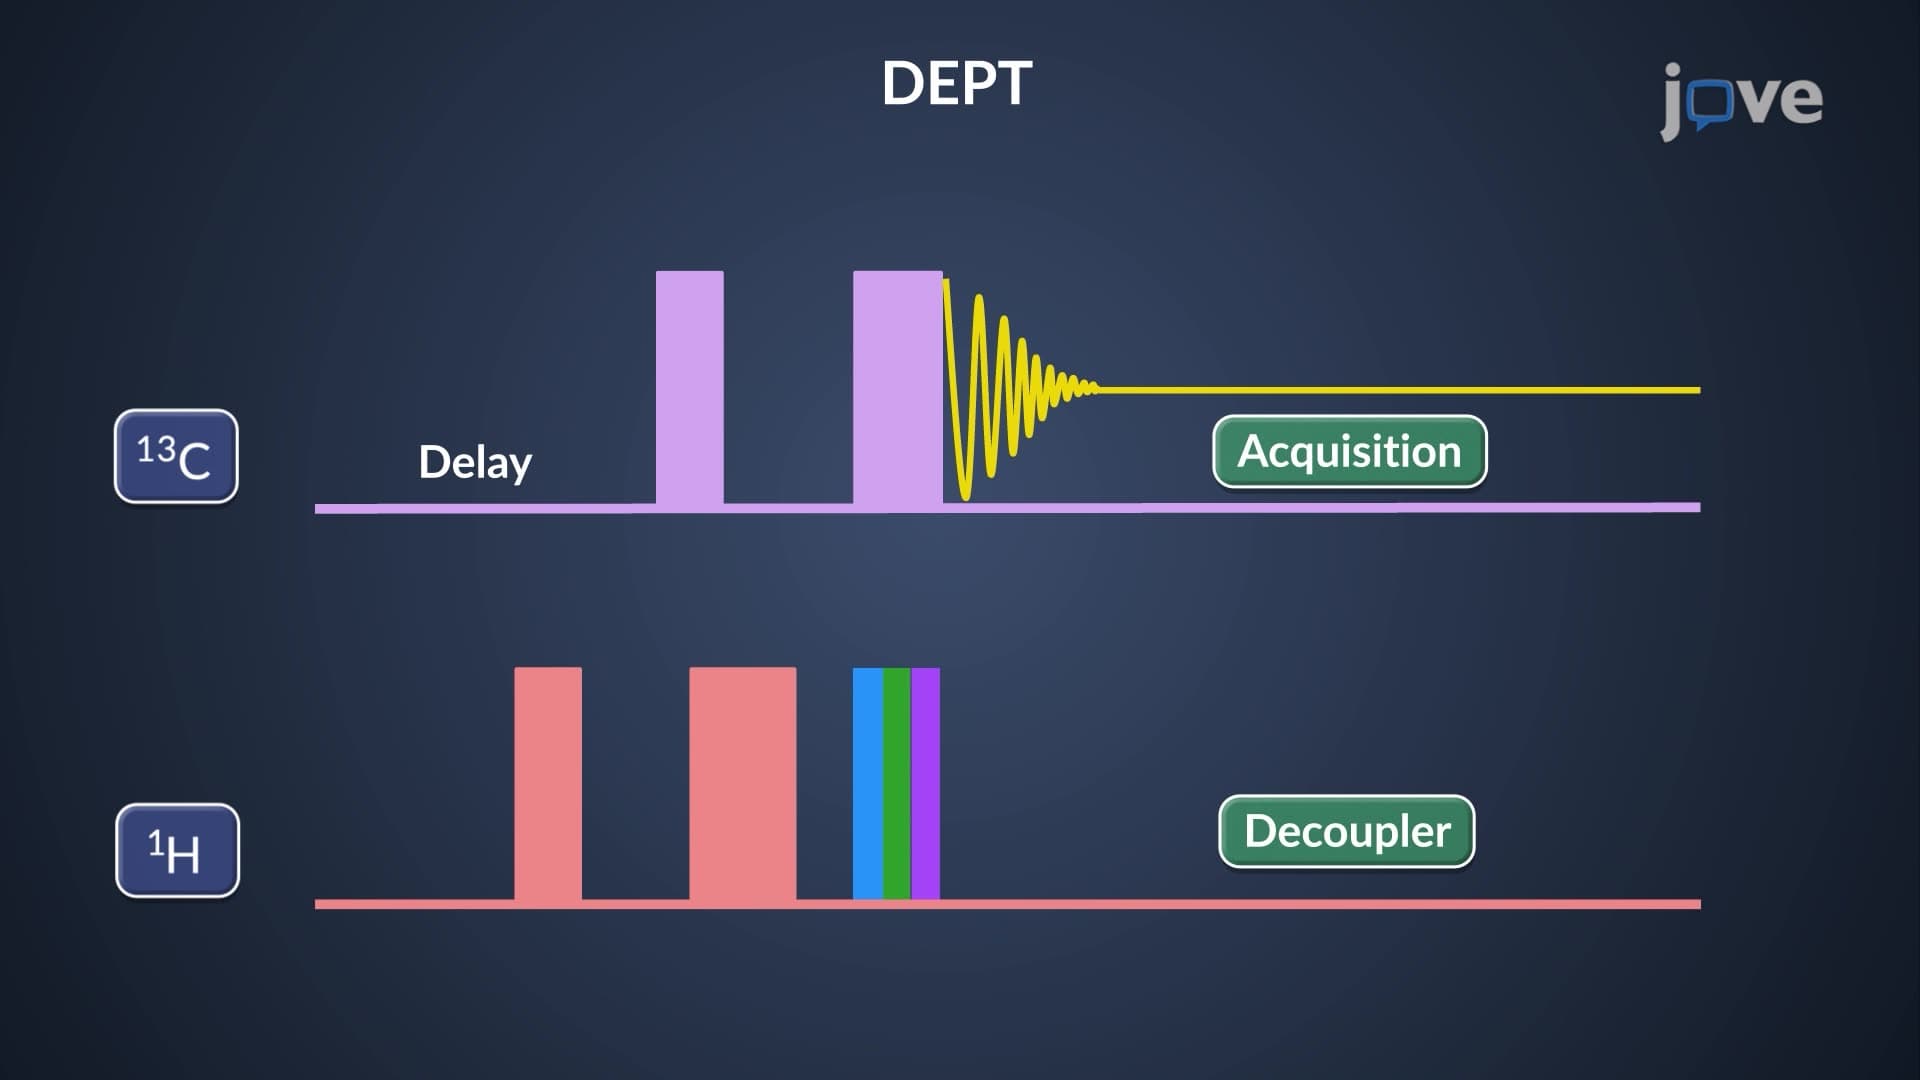
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)