Related Experiment Video
Updated: Mar 8, 2026

Analyzing Mitochondrial Transport and Morphology in Human Induced Pluripotent Stem Cell-Derived Neurons in Hereditary Spastic Paraplegia
Published on: February 9, 2020
ATPase-deficient mitochondrial inner membrane protein ATAD3A disturbs mitochondrial dynamics in dominant hereditary
Helen M Cooper1, Yang Yang2,3, Emil Ylikallio2,4
1Åbo Akademi University, Faculty of Natural Sciences and Technology, Turku, Finland.

Abstract:
De novo mutations in ATAD3A (ATPase family AAA-domain containing protein 3A) were recently found to cause a neurological syndrome with developmental delay, hypotonia, spasticity, optic atrophy, axonal neuropathy, and hypertrophic cardiomyopathy. Using whole-exome sequencing, we identified a dominantly inherited heterozygous variant c.1064G > A (p.G355D) in ATAD3A in a mother presenting with hereditary spastic paraplegia (HSP) and axonal neuropathy and her son with dyskinetic cerebral palsy, both with disease onset in childhood. HSP is a clinically and genetically heterogeneous disorder of the upper motor neurons. Symptoms beginning in early childhood may resemble spastic cerebral palsy. The function of ATAD3A, a mitochondrial inner membrane AAA ATPase, is yet undefined. AAA ATPases form hexameric rings, which are catalytically dependent on the co-operation of the subunits. The dominant-negative patient mutation affects the Walker A motif, which is responsible for ATP binding in the AAA module of ATAD3A, and we show that the recombinant mutant ATAD3A protein has a markedly reduced ATPase activity. We further show that overexpression of the mutant ATAD3A fragments the mitochondrial network and induces lysosome mass. Similarly, we observed altered dynamics of the mitochondrial network and increased lysosomes in patient fibroblasts and neurons derived through differentiation of patient-specific induced pluripotent stem cells. These alterations were verified in patient fibroblasts to associate with upregulated basal autophagy through mTOR inactivation, resembling starvation. Mutations in ATAD3A can thus be dominantly inherited and underlie variable neurological phenotypes, including HSP, with intrafamiliar variability. This finding extends the group of mitochondrial inner membrane AAA proteins associated with spasticity.
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:28
The Inner Mitochondrial Membrane
 01:42
The ADP/ATP Carrier Protein
 01:37
Energy to Drive Translocation
Generally, polypeptides are unfolded by two distinct...
 01:18
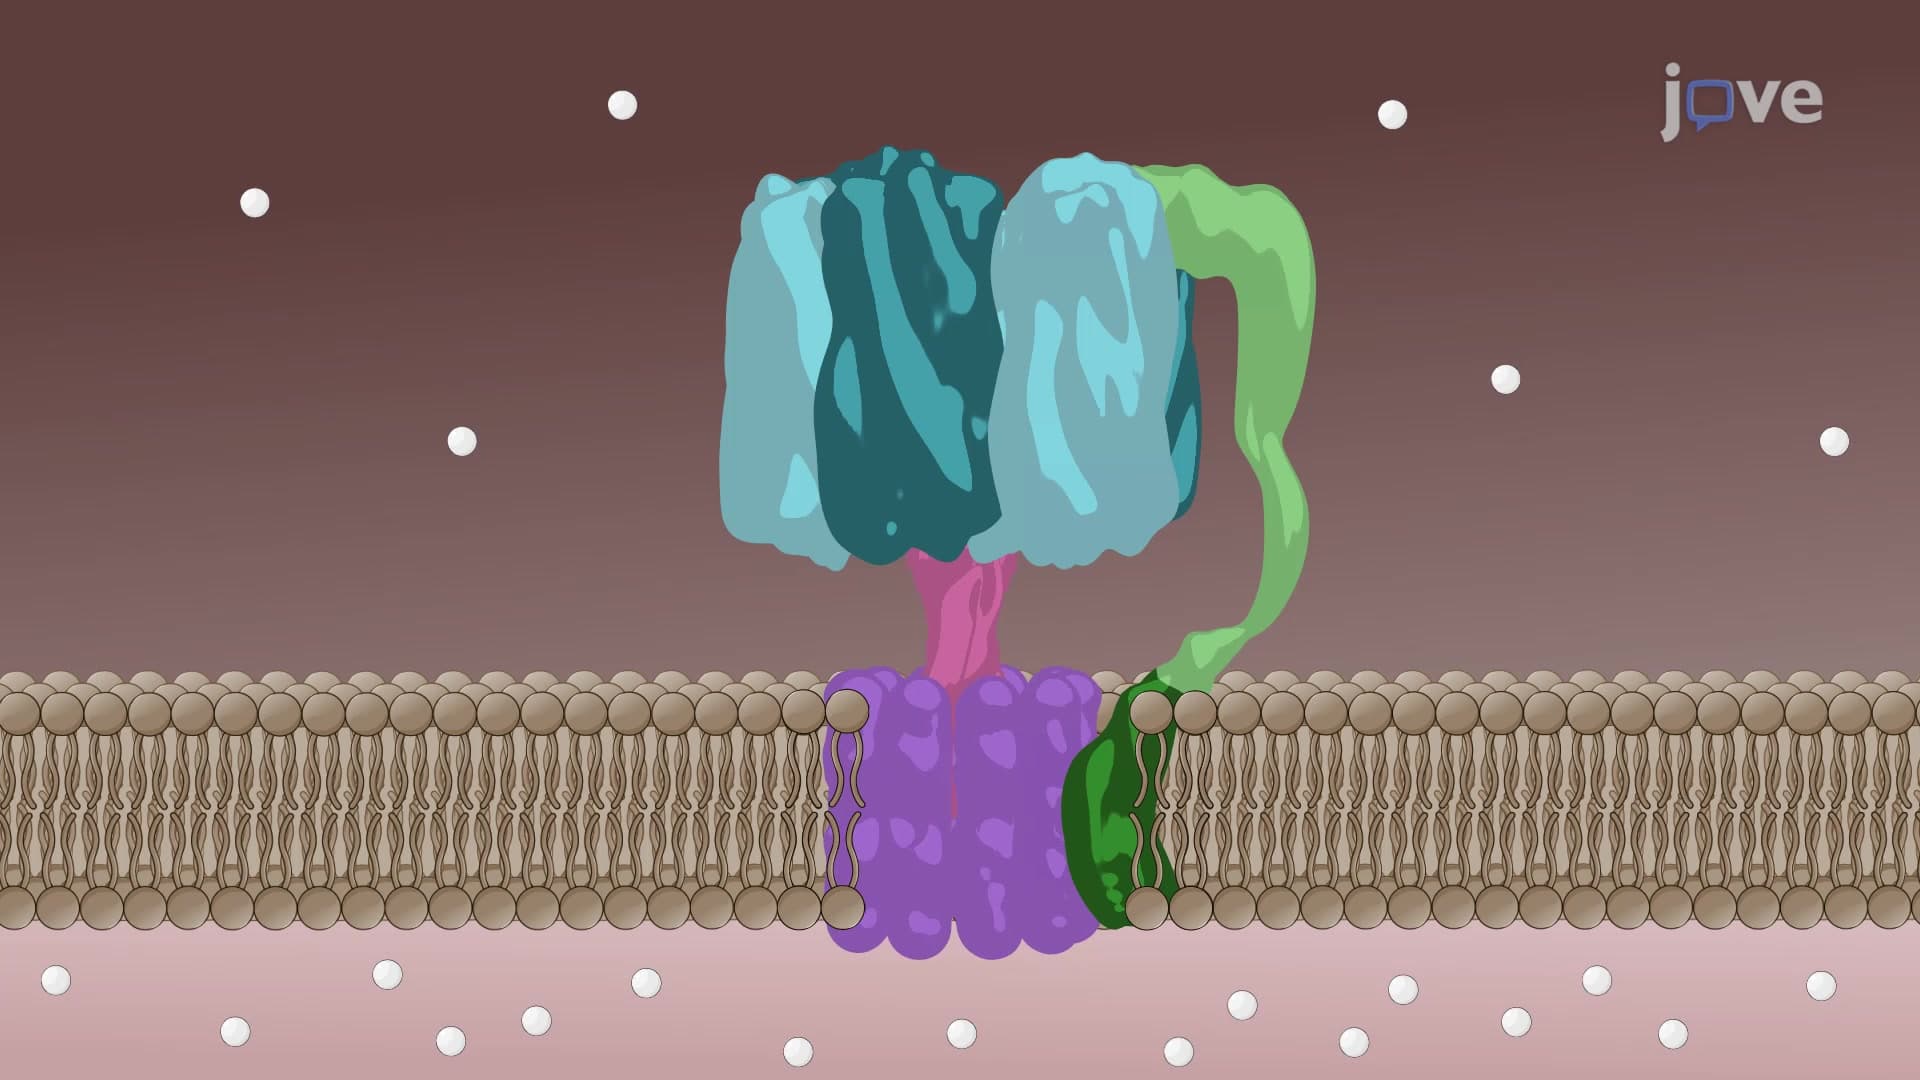
ATP Synthase: Structure
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...