Related Experiment Video
Updated: Mar 7, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Adaptively Compressed Exchange Operator for Large-Scale Hybrid Density Functional Calculations with Applications to
Wei Hu1, Lin Lin1,2, Amartya S Banerjee1
1Computational Research Division, Lawrence Berkeley National Laboratory , Berkeley, California 94720, United States.

Abstract:
Density functional theory (DFT) calculations using hybrid exchange-correlation functionals have been shown to provide an accurate description of the electronic structures of nanosystems. However, such calculations are often limited to small system sizes due to the high computational cost associated with the construction and application of the Hartree-Fock (HF) exchange operator. In this paper, we demonstrate that the recently developed adaptively compressed exchange (ACE) operator formulation [J. Chem. Theory Comput. 2016, 12, 2242-2249] can enable hybrid functional DFT calculations for nanosystems with thousands of atoms. The cost of constructing the ACE operator is the same as that of applying the exchange operator to the occupied orbitals once, while the cost of applying the Hamiltonian operator with a hybrid functional (after construction of the ACE operator) is only marginally higher than that associated with applying a Hamiltonian constructed from local and semilocal exchange-correlation functionals. Therefore, this new development significantly lowers the computational barrier for using hybrid functionals in large-scale DFT calculations. We demonstrate that a parallel planewave implementation of this method can be used to compute the ground-state electronic structure of a 1000-atom bulk silicon system in less than 30 wall clock minutes and that this method scales beyond 8000 computational cores for a bulk silicon system containing about 4000 atoms. The efficiency of the present methodology in treating large systems enables us to investigate adsorption properties of water molecules on Ag-supported two-dimensional silicene. Our computational results show that water monomer, dimer, and trimer configurations exhibit distinct adsorption behaviors on silicene. In particular, the presence of additional water molecules in the dimer and trimer configurations induces a transition from physisorption to chemisorption, followed by dissociation on Ag-supported silicene. This is caused by the enhanced effect of hydrogen bonds on charge transfer and proton transfer processes. Such a hydrogen bond autocatalytic effect is expected to have broad applications for silicene as an efficient surface catalyst for oxygen reduction reactions and water dissociation.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 01:14
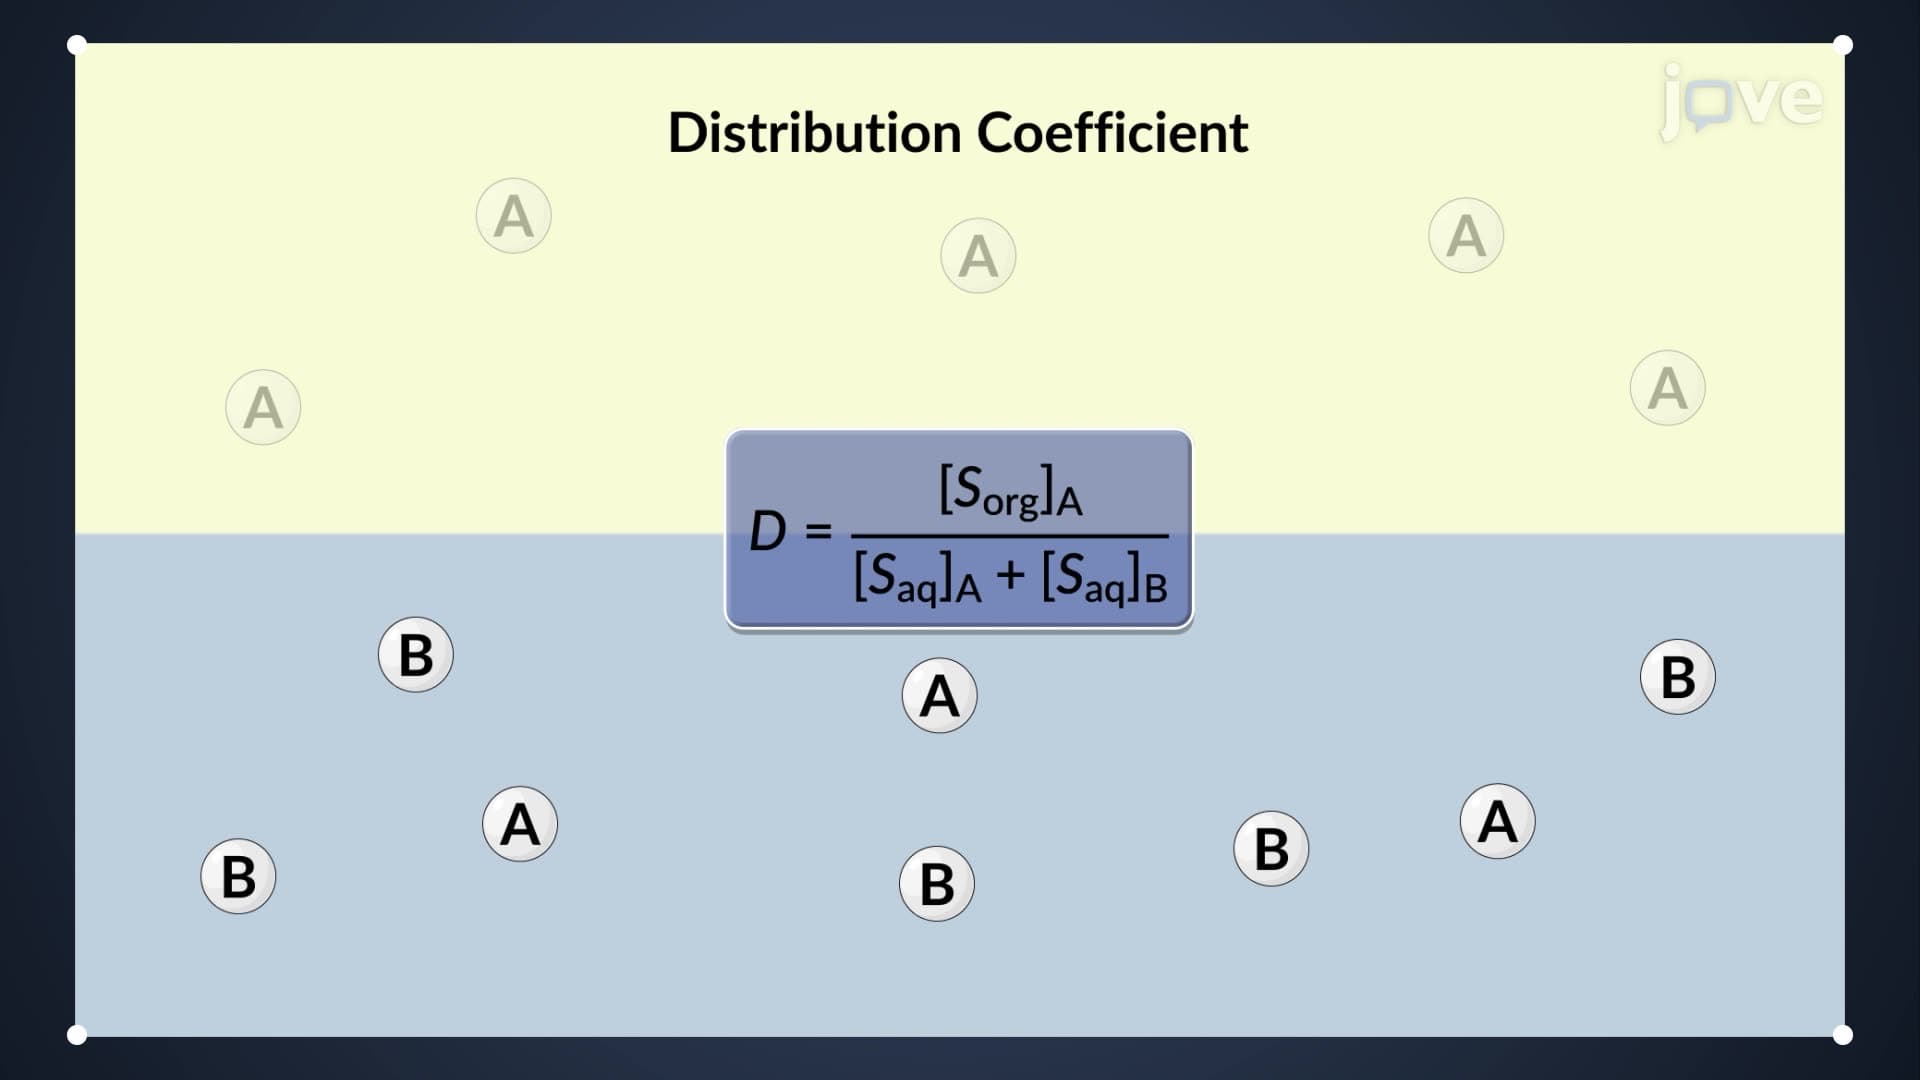
Extraction: Partition and Distribution Coefficients
For extracting a solute from an aqueous phase into an...
 01:17
Ion Exchange
 01:22
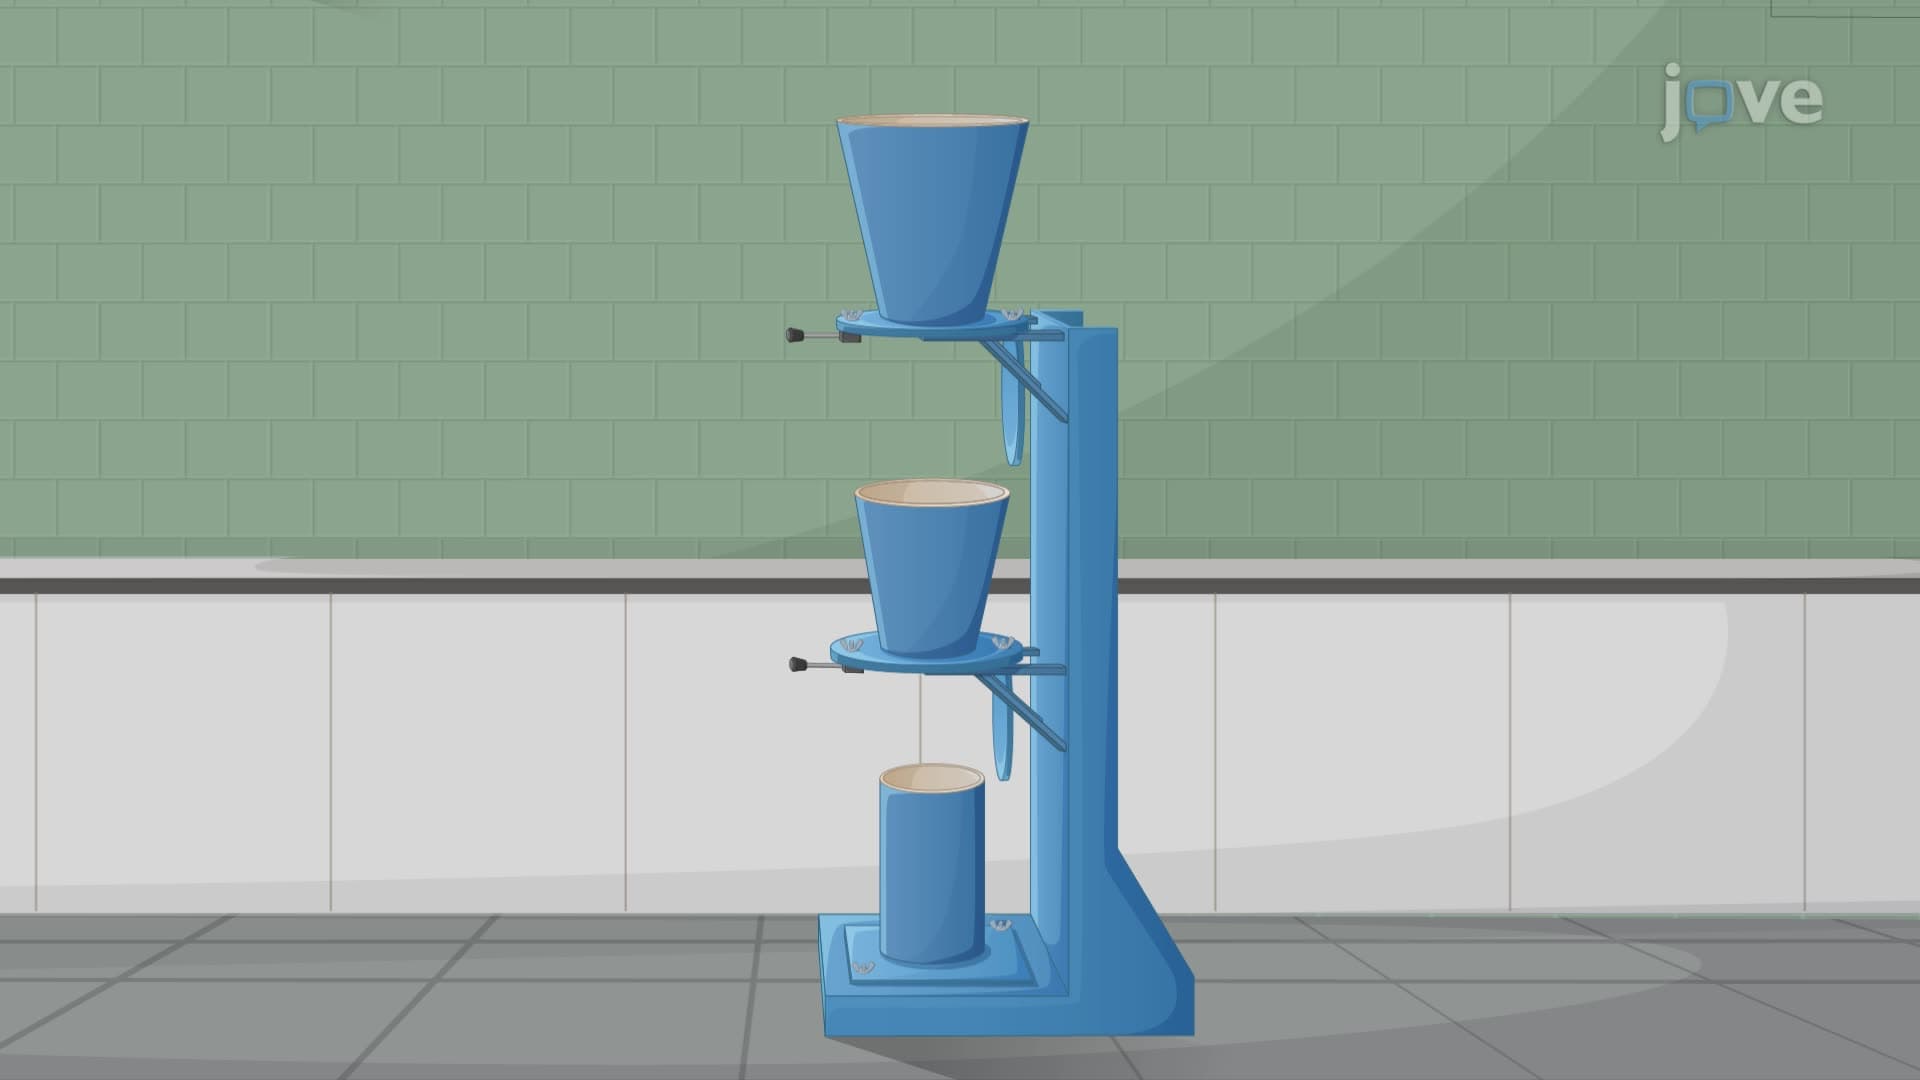
Compacting Factor test
The procedure begins by placing concrete into the upper hopper without any compaction. Once filled, the bottom door of this hopper is opened,...
 01:06
Nuclear Overhauser Enhancement (NOE)