An automatic search for similar spatial arrangements of alpha-helices and beta-strands in globular proteins
Related Experiment Video
Updated: Aug 15, 2026
A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
1Shemyakin Institute of Bioorganic Chemistry, USSR Academy of Sciences, Moscow.

Abstract:
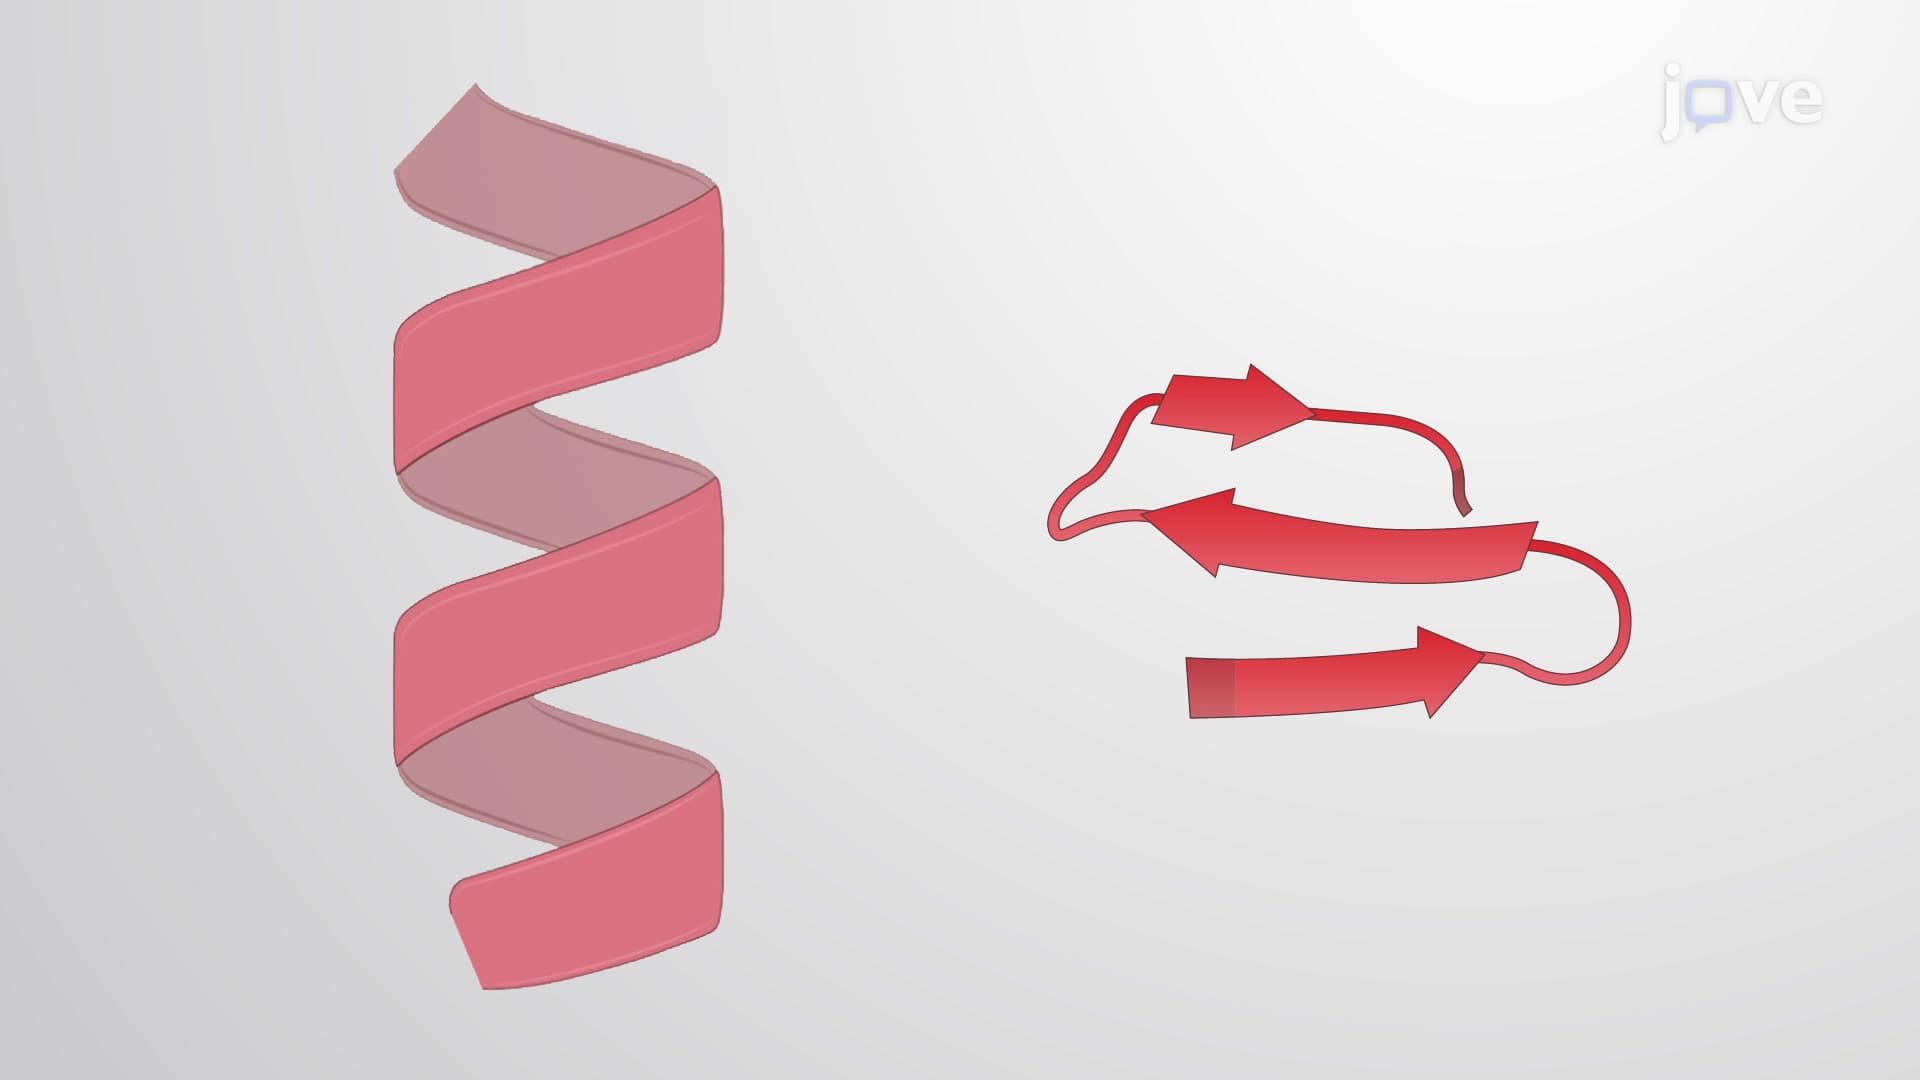
A fast search algorithm to reveal similar polypeptide backbone structural motifs in proteins is proposed. It is based on the vector representation of a polypeptide chain fold in which the elements of regular secondary structures are approximated by linear segments (Abagyan and Maiorov, J. Biomol. Struct. Dyn. 5, 1267-1279 (1988)). The algorithm permits insertions and deletions in the polypeptide chain fragments to be compared. The fast search algorithm implemented in FASEAR program is used for collecting beta alpha beta supersecondary structure units in a number of alpha/beta proteins of Brookhaven Data Bank. Variation of geometrical parameters specifying backbone chain fold is estimated. It appears that the conformation of the majority of the fragments, although almost all of them are right-handed, is quite different from that of standard beta alpha beta units. Apart from searching for specific type of secondary structure motif, the algorithm allows automatically to identify new recurrent folding patterns in proteins. It may be of particular interest for the development of tertiary template approach for prediction of protein three-dimensional structure as well for constructing artificial polypeptides with goal-oriented conformation.
Related Concept Videos
 01:13
Protein Organization
 01:22
Protein Folding
01:13Protein Organization
01:22Protein Folding
 01:24
Protein Organization
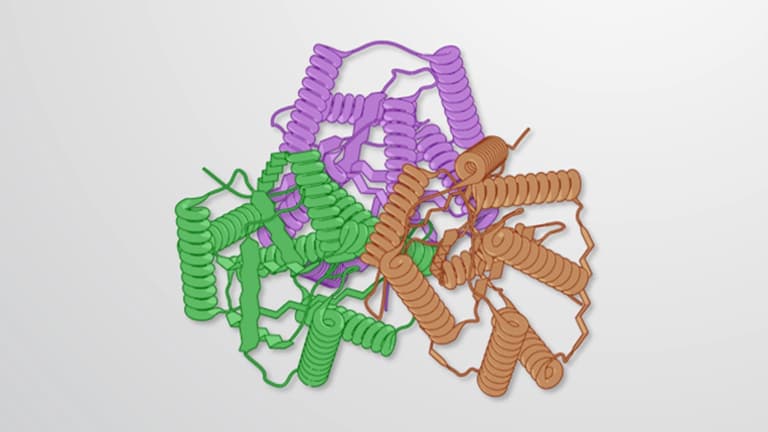
The primary structure of a protein is its amino acid sequence.
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...