A rare case of 46,XX gonadal dysgenesis and Mayer-Rokitansky-Kuster-Hauser syndrome
Related Experiment Video
Updated: Mar 7, 2026
Laparoscopic Oocyte Retrieval and Cryopreservation during Vaginoplasty for Treatment of Mayer-Rokitansky-Kuster-Hauser Syndrome
Published on: May 10, 2022
Sriharibabu Manne1, C H Veeraabhinav1, Mounica Jetti1
1Department of Medicine, G.S.L. Medical College and General Hospital, Rajahmundry, Andhra Pradesh, India.

Abstract:
46,XX gonadal dysgenesis is a rare genetically heterogeneous disorder characterized by underdeveloped ovaries with consequent, impuberism, primary amenorrhea, and hypergonadotropic hypogonadism. Mullerian agenesis or Mayer-Rokitansky-Kuster-Hauser (MRKH) syndrome is characterized by congenital aplasia of the uterus and the upper part (2/3) of the vagina in a woman with normal development of secondary sexual characteristics and a normal 46,XX karyotype. The phenotypic manifestations of MRKH syndrome may sometimes overlap with various other syndromes and require accurate delineation. The coexistence of both these disorders is extremely rare. Here, we report a case of 46,XX gonadal dysgenesis and MRKH syndrome with anatomically dispersed congenital anomalies unique among reported cases.
Related Concept Videos
 01:29
Nondisjunction
 01:21
Nondisjunction
 01:43
Sex-linked Disorders
 02:32
X and Y Chromosomes
The germline cells such as egg and sperm cells carry only half the number of chromosomes, i.e., 22 autosomes and one sex chromosome. All eggs have an X chromosome, while sperm cells can carry an X or...
 02:57
Meiosis vs. Mitosis
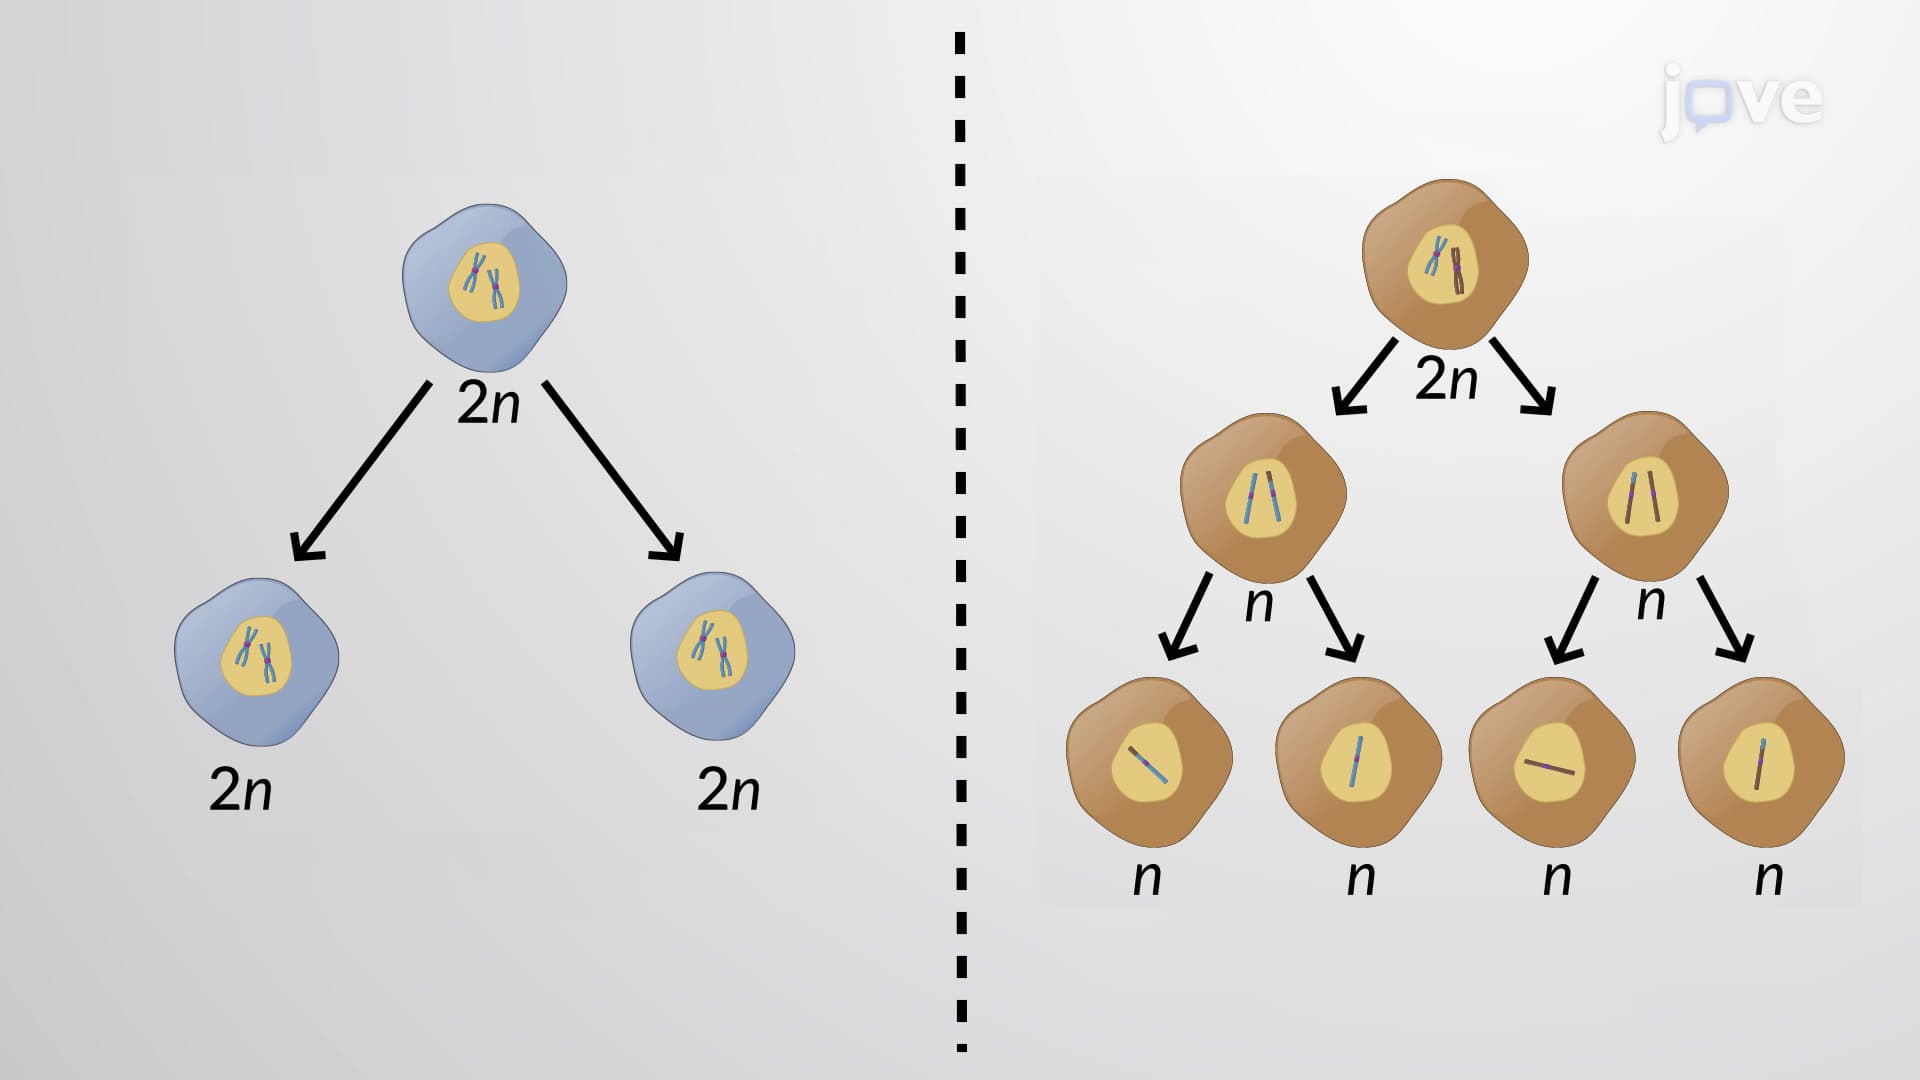
Before the start of mitosis and meiosis I, the cell synthesizes DNA, resulting in two homologous copies of each chromosome. DNA synthesis is...
 01:17
Karyotyping