Interaction Energies in Complexes of Zn and Amino Acids: A Comparison of Ab Initio and Force Field Based Calculations
Related Experiment Video
Updated: Mar 6, 2026
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Emma Ahlstrand1,2, Kersti Hermansson3, Ran Friedman1,2
1Department of Chemistry and Biomedical Sciences, Linnæus University , 391 82 Kalmar, Sweden.

Abstract:
Zinc plays important roles in structural stabilization of proteins, enzyme catalysis, and signal transduction. Many Zn binding sites are located at the interface between the protein and the cellular fluid. In aqueous solutions, Zn ions adopt an octahedral coordination, while in proteins zinc can have different coordinations, with a tetrahedral conformation found most frequently. The dynamics of Zn binding to proteins and the formation of complexes that involve Zn are dictated by interactions between Zn and its binding partners. We calculated the interaction energies between Zn and its ligands in complexes that mimic protein binding sites and in Zn complexes of water and one or two amino acid moieties, using quantum mechanics (QM) and molecular mechanics (MM). It was found that MM calculations that neglect or only approximate polarizability did not reproduce even the relative order of the QM interaction energies in these complexes. Interaction energies calculated with the CHARMM-Drude polarizable force field agreed better with the ab initio results, although the deviations between QM and MM were still rather large (40-96 kcal/mol). In order to gain further insight into Zn-ligand interactions, the free energies of interaction were estimated by QM calculations with continuum solvent representation, and we performed energy decomposition analysis calculations to examine the characteristics of the different complexes. The ligand-types were found to have high impact on the relative strength of polarization and electrostatic interactions. Interestingly, ligand-ligand interactions did not play a significant role in the binding of Zn. Finally, analysis of ligand exchange energies suggests that carboxylates could be exchanged with water molecules, which explains the flexibility in Zn binding dynamics. An exchange between carboxylate (Asp/Glu) and imidazole (His) is less likely.
More Related Videos
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:09
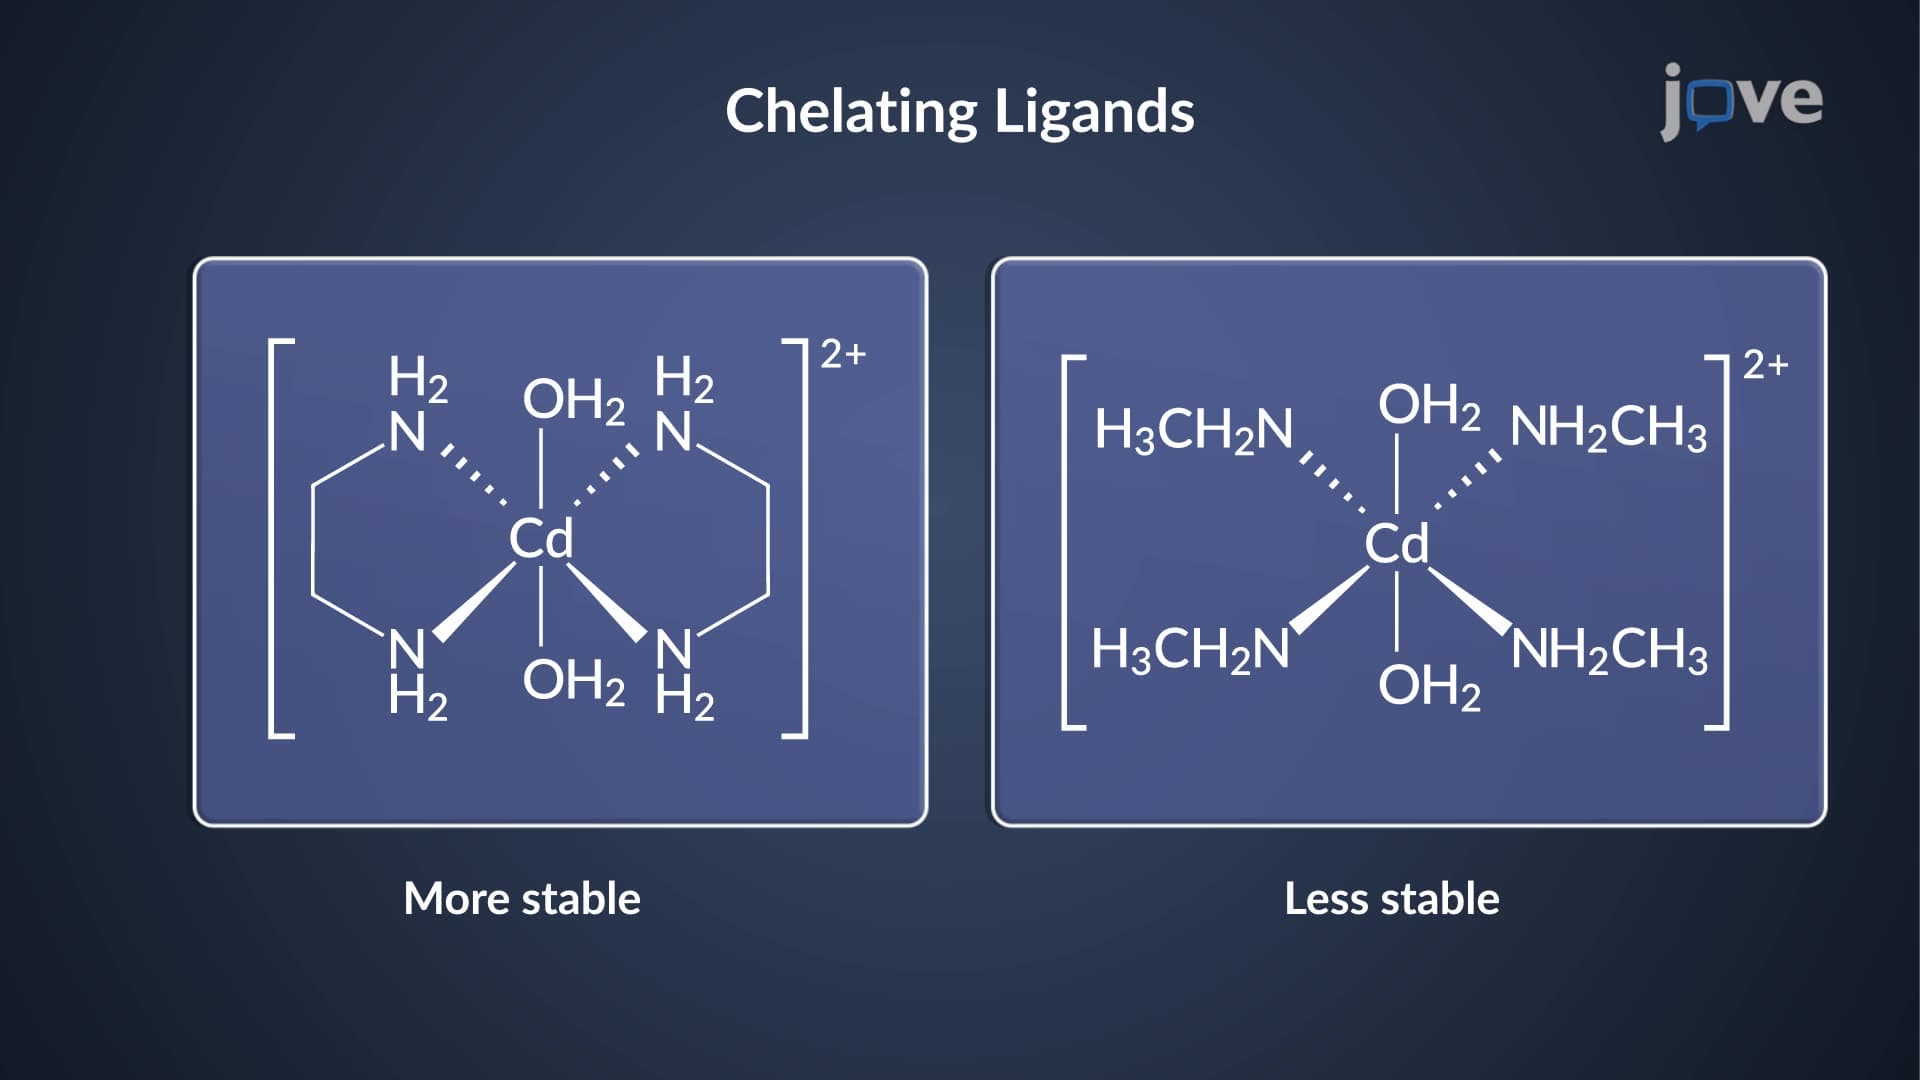
Complexation Equilibria: Factors Influencing Stability of Complexes
 01:23
Complexation Equilibria: Overview
The equilibrium constant of the complexation reaction is represented as the formation constant...
 02:42
Valence Bond Theory
 02:35
Noncovalent Attractions in Biomolecules