Related Experiment Video
Updated: Mar 6, 2026

Self-assembling Morphologies Obtained from Helical Polycarbodiimide Copolymers and Their Triazole Derivatives
Published on: February 7, 2017
Charge carrier mobility in one-dimensional aligned π-stacks of conjugated small molecules with a benzothiadiazole
1Dresden Center for Computational Materials Science (DCMS), Technische Universität Dresden (TUD), Dresden D-01062, Germany. guskova@ipfdd.de and Institute Theory of Polymers, Leibniz Institute of Polymer Research Dresden, Hohe Str. 6, Dresden D-01069, Germany.

Abstract:
A theoretical study is applied to gain insight into the microscopic electron and hole transport in benzothiadiazole-cored molecular semiconductors either with furan or thiophene flanks arranged in π-stacks. For the characterization of the energetics of the reduction and oxidation processes and their impact on the molecular geometry, the internal reorganization energy is defined for isolated molecules in the gas phase. The outer-shell reorganization energy is evaluated within the frequency-resolved cavity model and as an electrostatic contribution within the polarizable continuum model. The intermolecular electronic coupling interaction for the Marcus charge hopping is calculated using the energy splitting in dimer method, the generalized Mulliken-Hush approach and the fragment charge difference scheme. In order to probe the relation between the charge hopping rate/charge carrier mobility and the molecular organization within the π-stacks, different stacking modes are investigated: (i) dimers with a perfect registry, i.e. segregated stacking motif, when molecules are placed face-to-face, and (ii) dimers forming slipped cofacial orientations with longitudinal and transverse shifts, i.e. mixed stacking motif. Besides, the effects of molecular planarity and rigidity, influencing internal molecular relaxation upon charging, the effects of non-covalent interactions within stacks and the heteroatom replacement on the charge carrier mobility are studied. The results obtained in the simulations of one-dimensional aligned π-stacks of molecular semiconductors are compared with available experimental data for small conjugated benzothiadiazole-cored molecules with thiophene flanks and benzothiadiazole-quaterthiophene-based copolymers.
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
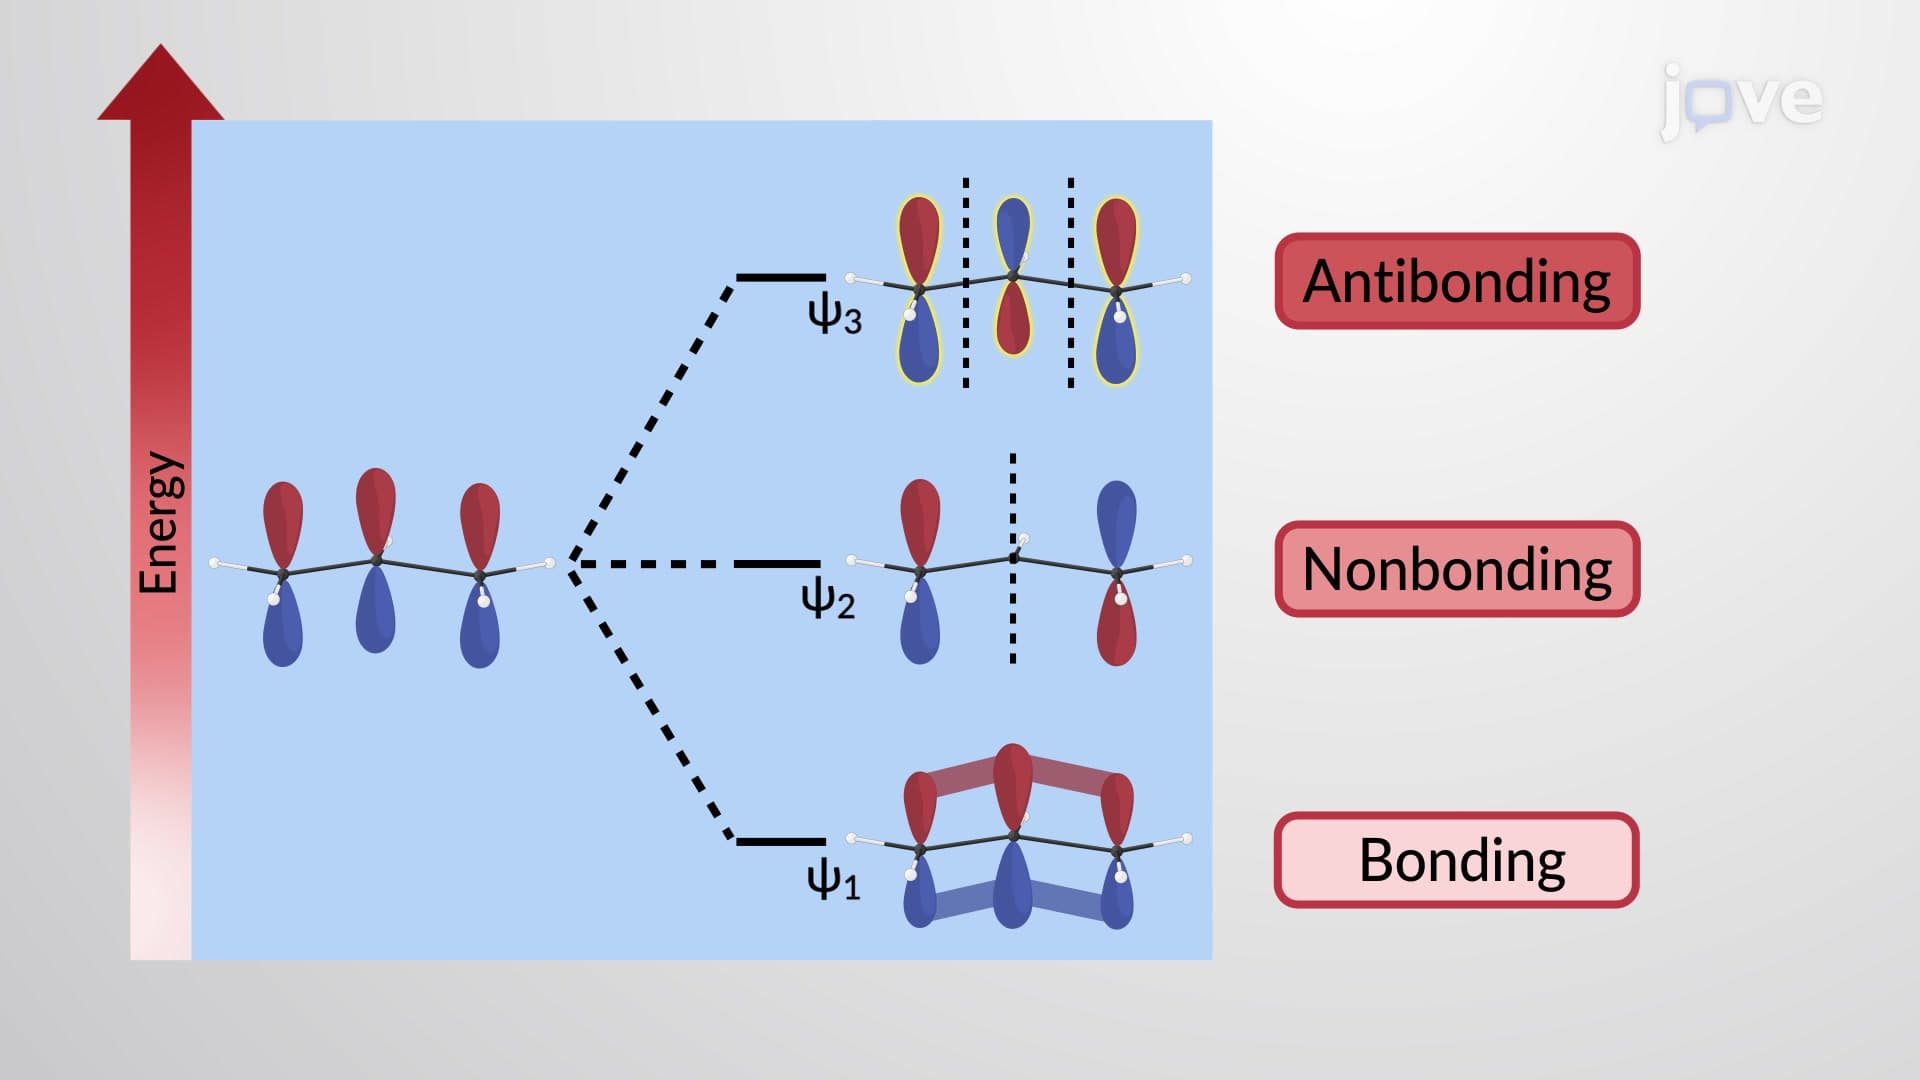
π Molecular Orbitals of the Allyl Cation and Anion
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:27
π Electron Effects on Chemical Shift: Overview
 01:21
Carrier Transport
Drift Current:
The drift of charge carriers is started by an external electric field (E). Charged particles, such as electrons and holes, experience an acceleration between collisions with lattice atoms. For electrons, this results in a drift velocity (vd) given by: