Related Experiment Video
Updated: Mar 6, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Is the Bethe-Salpeter Formalism Accurate for Excitation Energies? Comparisons with TD-DFT, CASPT2, and EOM-CCSD
Denis Jacquemin1,2, Ivan Duchemin3,4, Xavier Blase4,5
1Laboratoire CEISAM - UMR CNRS 6230, Université de Nantes , 2 Rue de la Houssinière, BP 92208, 44322 Nantes Cedex 3, France.

Abstract:
Developing ab initio approaches able to provide accurate excited-state energies at a reasonable computational cost is one of the biggest challenges in theoretical chemistry. In that framework, the Bethe-Salpeter equation approach, combined with the GW exchange-correlation self-energy, which maintains the same scaling with system size as TD-DFT, has recently been the focus of a rapidly increasing number of applications in molecular chemistry. Using a recently proposed set encompassing excitation energies of many kinds [J. Phys. Chem. Lett. 2016, 7, 586-591], we investigate here the performances of BSE/GW. We compare these results to CASPT2, EOM-CCSD, and TD-DFT data and show that BSE/GW provides an accuracy comparable to the two wave function methods. It is particularly remarkable that the BSE/GW is equally efficient for valence, Rydberg, and charge-transfer excitations. In contrast, it provides a poor description of triplet excited states, for which EOM-CCSD and CASPT2 clearly outperform BSE/GW. This contribution therefore supports the use of the Bethe-Salpeter approach for spin-conserving transitions.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:27
π Electron Effects on Chemical Shift: Overview
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:20
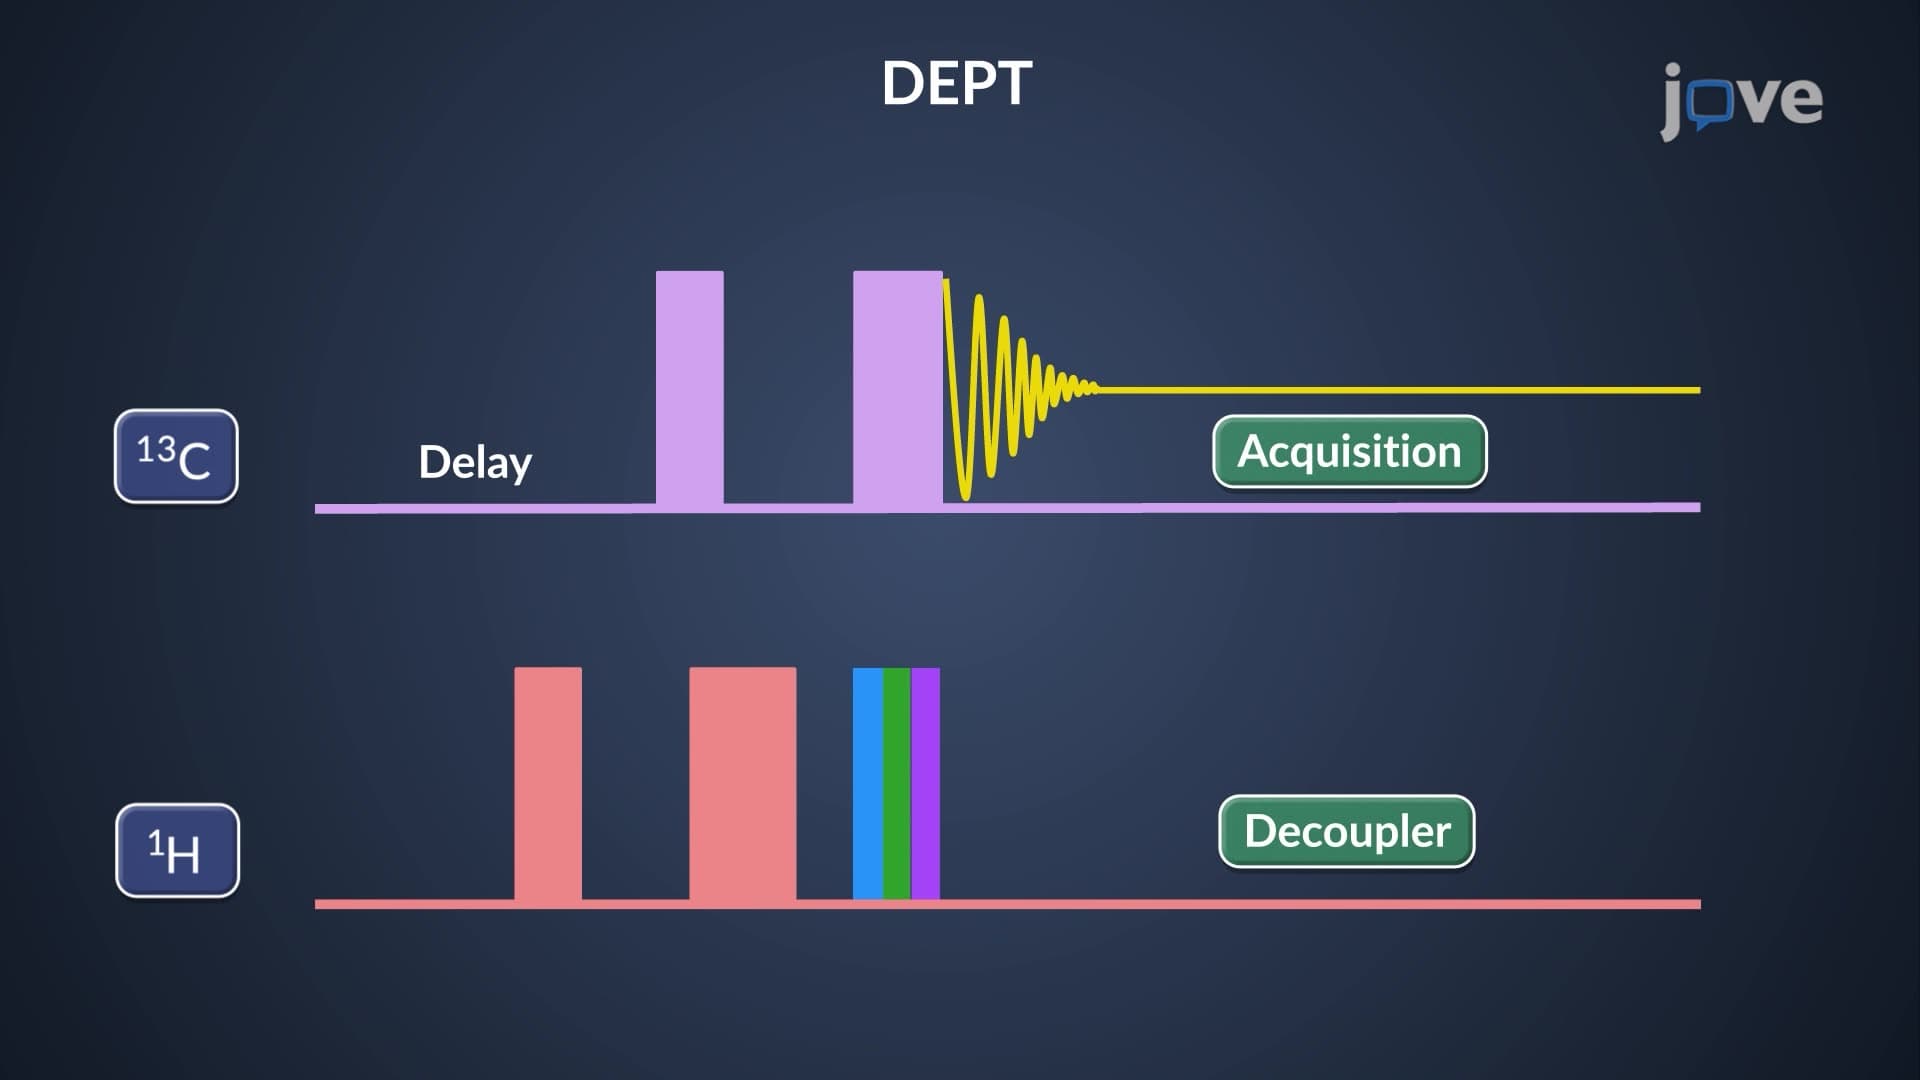
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:35
Molecular Orbital Theory I