Related Experiment Video
Updated: Mar 3, 2026

Crystal Structure of the N-terminal Domain of Ryanodine Receptor from Plutella xylostella
Published on: November 30, 2018
Mapping Ryanodine Binding Sites in the Pore Cavity of Ryanodine Receptors
Van A Ngo1, Laura L Perissinotti1, Williams Miranda1
1Center for Molecular Simulations, Department of Biological Sciences, University of Calgary, Calgary, Alberta, Canada.

Abstract:
Ryanodine (Ryd) irreversibly targets ryanodine receptors (RyRs), a family of intracellular calcium release channels essential for many cellular processes ranging from muscle contraction to learning and memory. Little is known of the atomistic details about how Ryd binds to RyRs. In this study, we used all-atom molecular dynamics simulations with both enhanced and bidirectional sampling to gain direct insights into how Ryd interacts with major residues in RyRs that were experimentally determined to be critical for its binding. We found that the pyrrolic ring of Ryd displays preference for the R4892AGGG-F4921 residues in the cavity of RyR1, which explain the effects of the corresponding mutations in RyR2 in experiments. Particularly, the mutant Q4933A (or Q4863A in RyR2) critical for both the gating and Ryd binding not only has significantly less interaction with Ryd than the wild-type, but also yields more space for Ryd and water molecules in the cavity. These results describe clear binding modes of Ryd in the RyR cavity and offer structural mechanisms explaining functional data collected on RyR blockade.
Related Concept Videos
 01:15
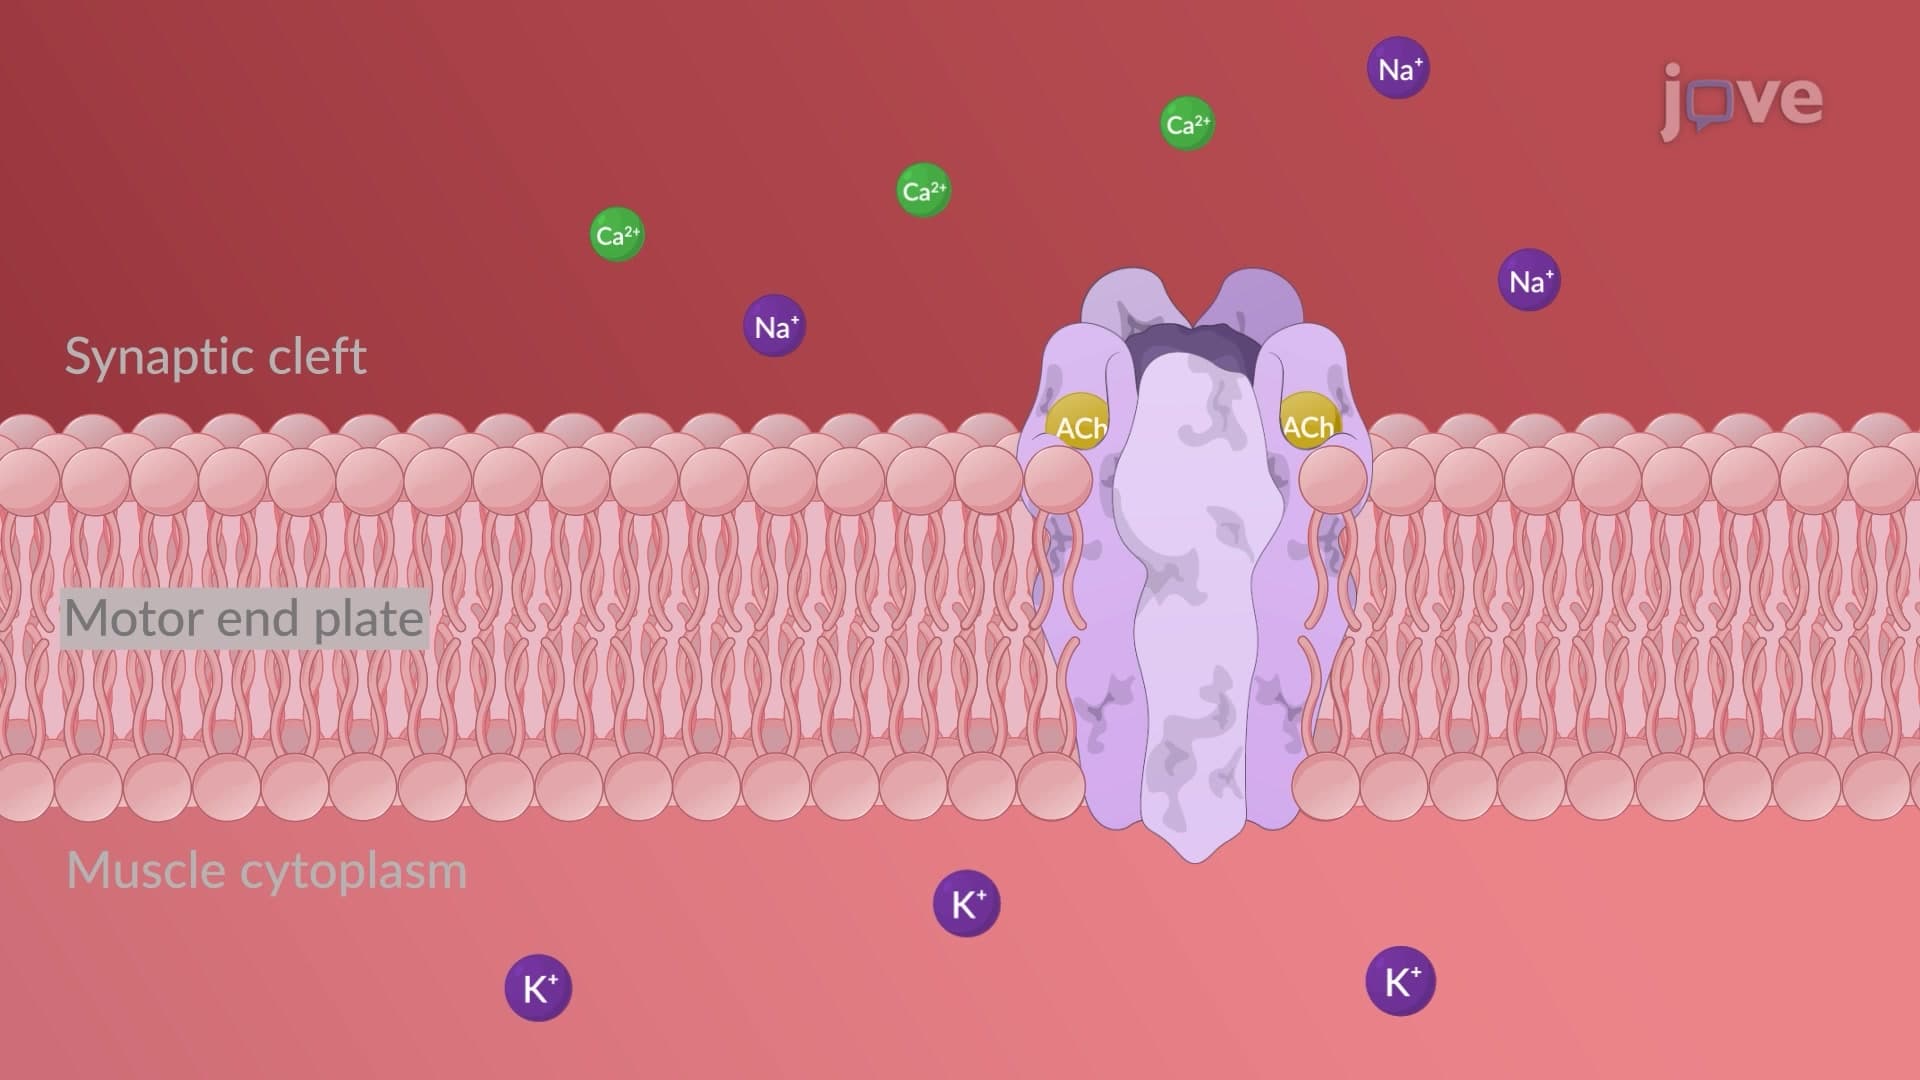
Cholinergic Receptors: Nicotinic
There are two types of nicotinic receptors: neuromuscular (NM/NM/N1) and neuronal (NN/NN/N2). The two families differ based on their location and selectivity to...
 01:18
Antihypertensive Drugs: Action of Calcium Channel Blockers
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:21
G-Protein Gated Ion Channels
Sensory...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...