Efficient Computation of Exchange Energy Density with Gaussian Basis Functions
Related Experiment Video
Updated: Mar 3, 2026
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
1Department of Chemistry, Middle Tennessee State University , Murfreesboro, Tennessee 37130, United States.

Abstract:
Density functional theory (DFT) is widely applied in chemistry and physics. Still it fails to correctly predict quantitatively or even qualitatively for systems with significant nondynamic correlation. Several DFT functionals were proposed in recent years to treat the nondynamic correlation, most of which added the exact exchange energy density as a new variable. This quantity, calculated as Hartree-Fock (HF) exchange energy density, is the computational bottleneck for calculations with these new functionals. We present an implementation of an efficient seminumerical algorithm in this paper as a solution for this computational bottleneck. The method scales quadratically with respect to the molecular size and the basis set size. The scheme, exact for the purpose of computing the HF exchange energy density, is favored for medium-sized basis sets and can be competitive even for large basis sets with efficient grids when compared with our previous approximate resolution-of-identity scheme. It can also be used as a seminumerical integration scheme to compute the HF exchange energy and matrix on a standard atom-centered grid. Calculations on a series of alanine peptides show that for large basis sets the seminumerical scheme becomes competitive to the conventional analytical method and can be about six times faster for aug-cc-pvtz basis. The practicality of the algorithm is demonstrated through a local hybrid self-consistent calculation of the acenes-20 molecule.

Related Concept Videos
 01:10
Gauss's Law: Problem-Solving
 01:07
Gauss's Law
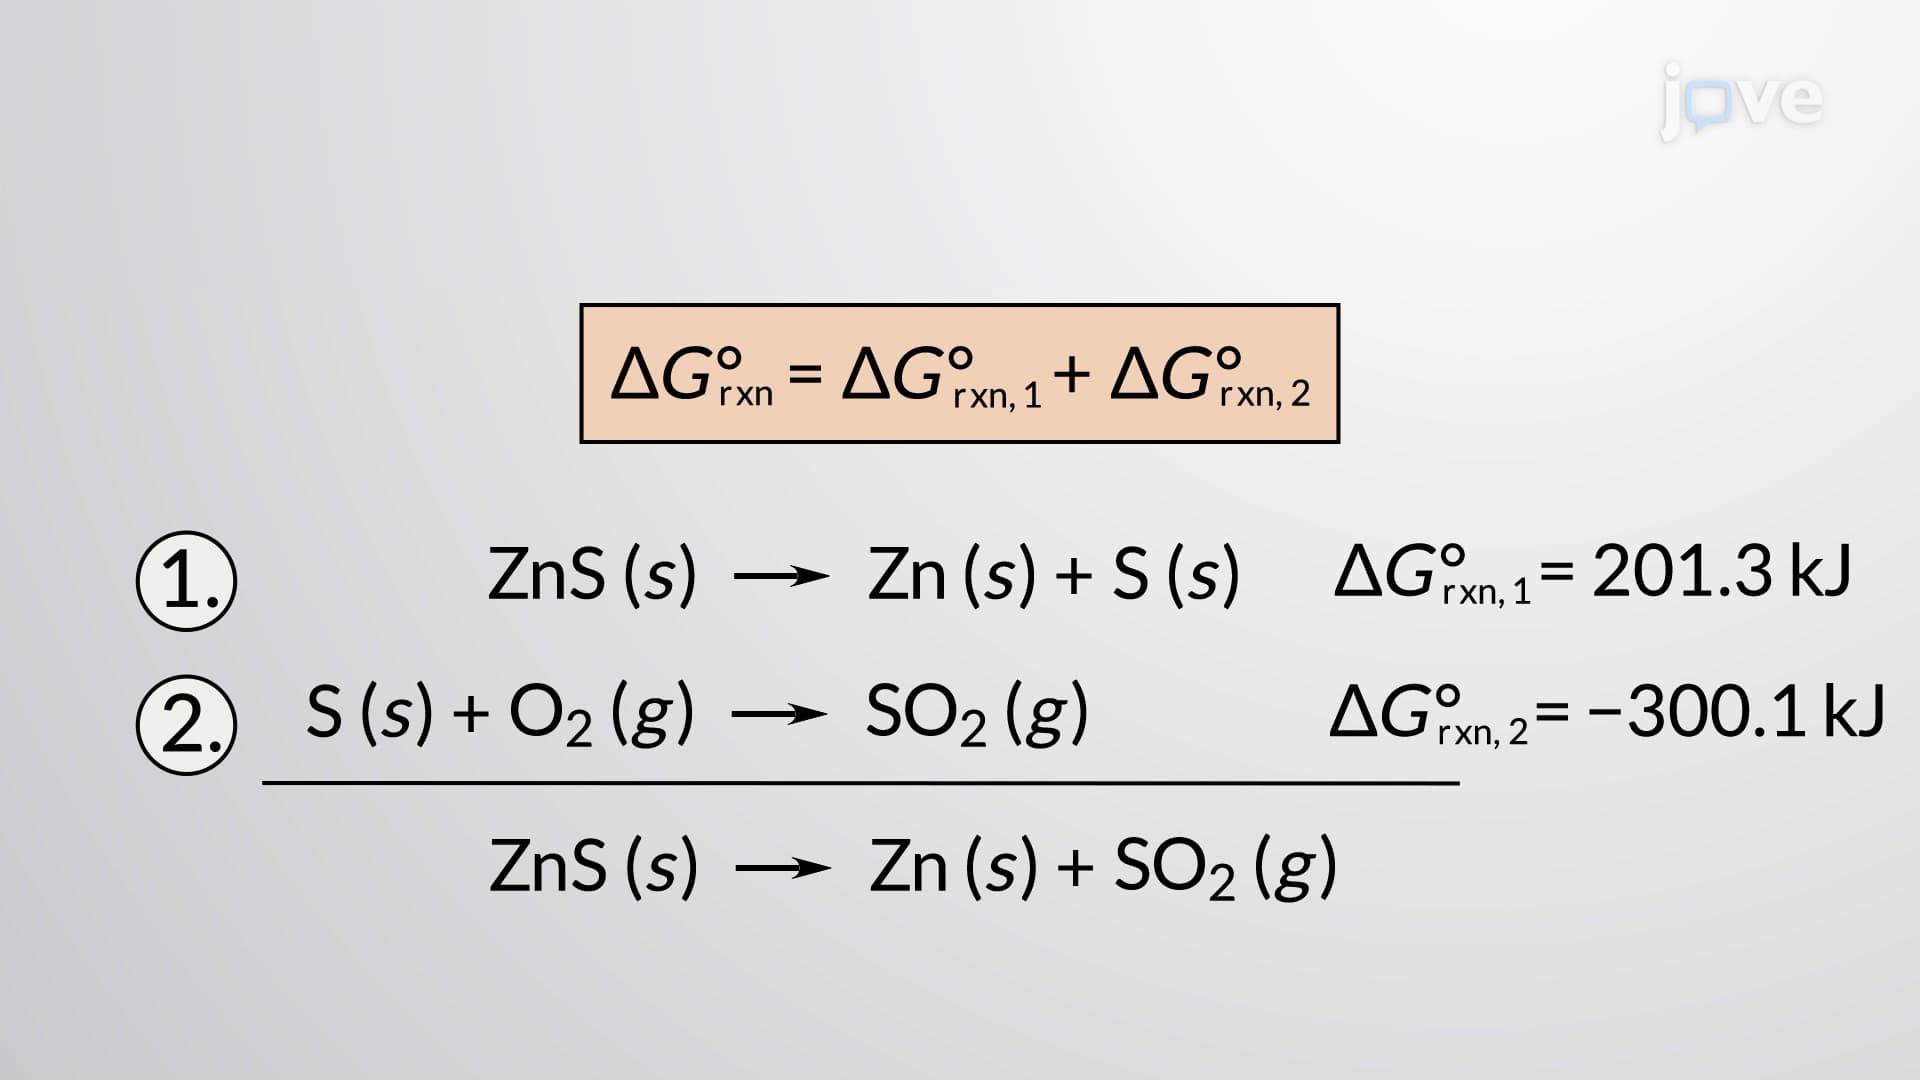 02:49
Calculating Standard Free Energy Changes
 01:28
Debye–Huckel–Onsager Conductance Equation
 03:21
The Energies of Atomic Orbitals
 01:10
Van der Waals Equation
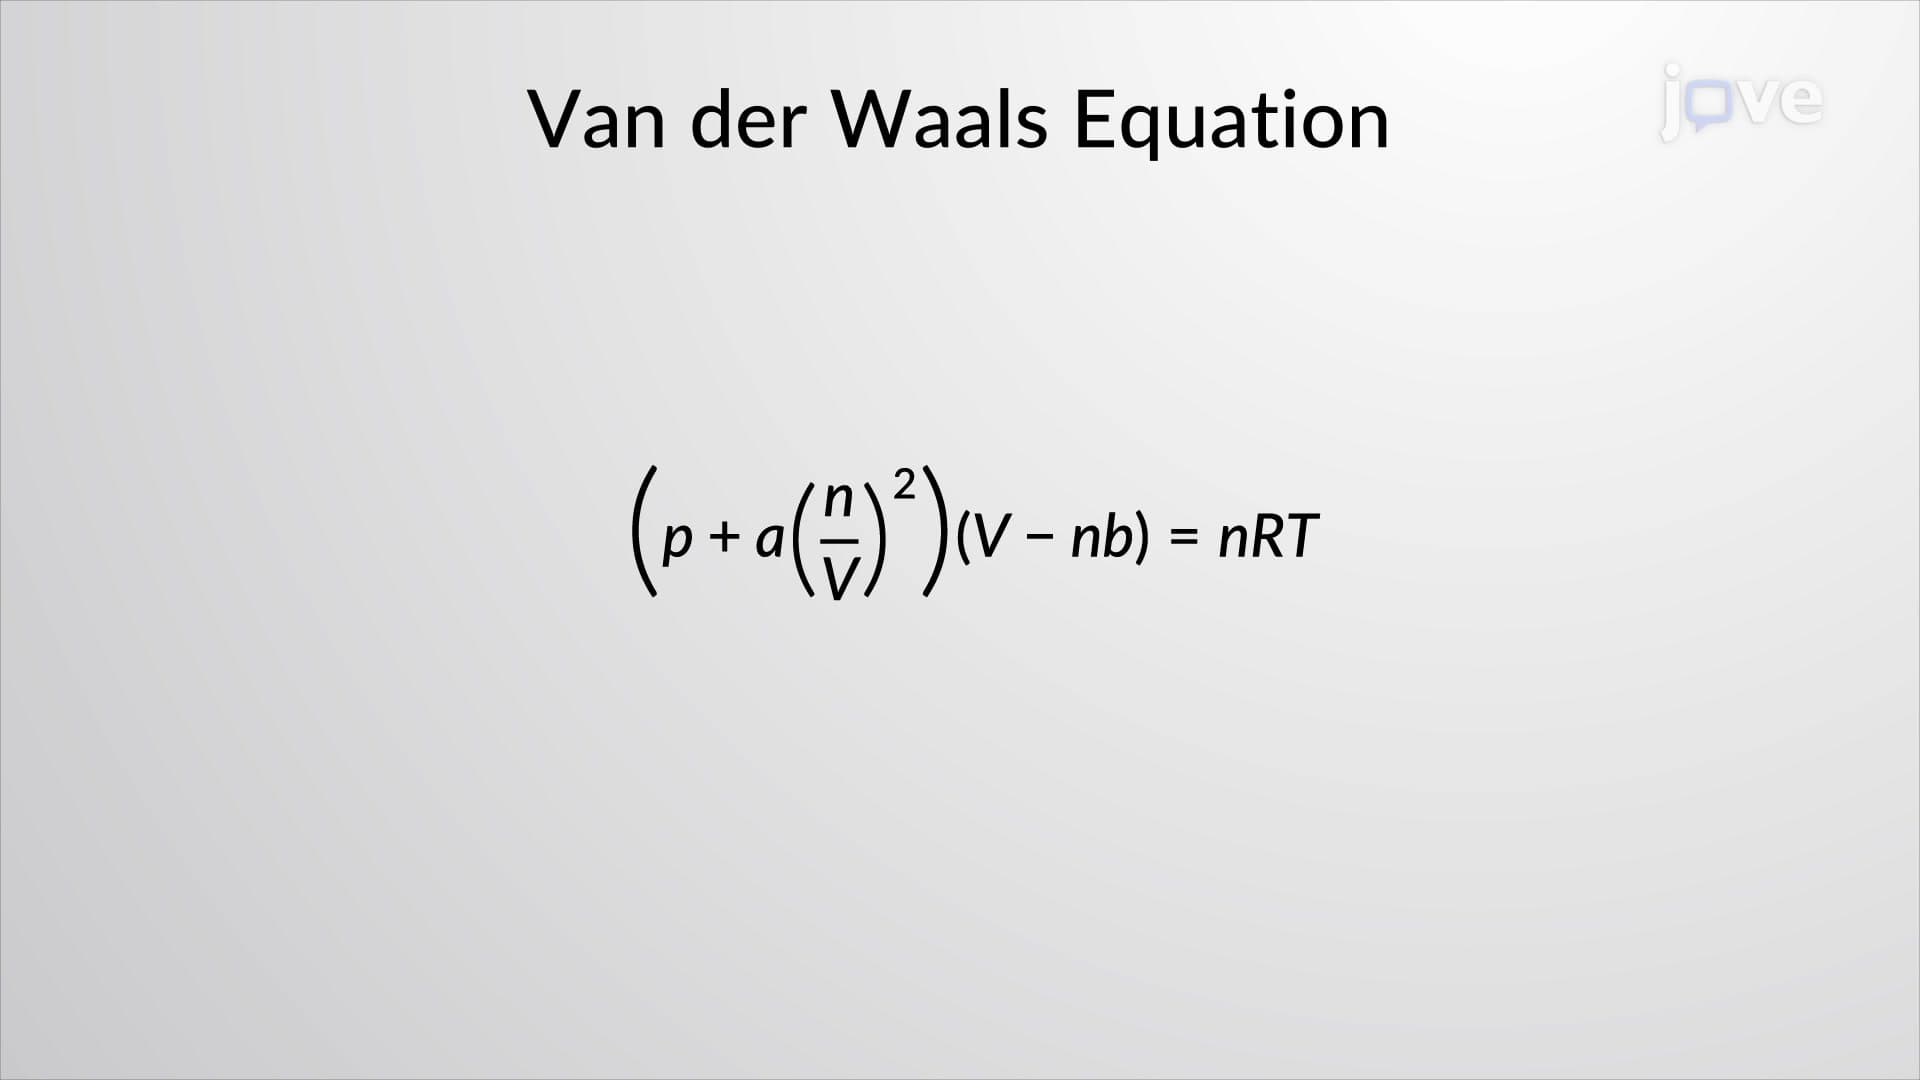
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...