
Filling in the Gaps of Mitochondrial Disease in Children
1Department of Pediatrics and Neurology, The Brooklyn Hospital, Icahn School of Medicine at Mount Sinai, Brooklyn, New York.
Pediatric Neurology
|May 4, 2017

Abstract
No abstract available in PubMed .
More Related Videos
Related Concept Videos
 01:20
Inborn Errors of Metabolism
962
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
962
 02:59
Animal Mitochondrial Genetics
9.7K
Among all the organelles in an animal cell, only mitochondria have their own independent genomes. Animal mitochondrial DNA is a double-stranded, closed-circular molecule with around 20,000 base pairs. Mitochondrial DNA is unique in that one of its two strands, the heavy, or H, -strand is guanine rich, whereas the complementary strand is cytosine rich and called the light, or L, -strand. Compared to nuclear DNA, mitochondrial DNA has a very low percentage of non-coding regions and is marked by...
9.7K
 01:45
Mitochondrial Membranes
17.5K
A single mitochondrion is a bean-shaped organelle enclosed by a double-membrane system. The outer membrane of mitochondria is smooth and contains many porins - the integral membrane transporters. Porins enable free diffusion of ions and small uncharged molecules through the outer mitochondrial membrane but limit the transport of molecules larger than 5000 Daltons. Further, the outer mitochondrial membrane forms a unique structure called membrane contact sites with other subcellular organelles,...
17.5K
 01:48
ATP Synthase: Mechanism
18.0K
In animals, the mitochondrial F1F0 ATP synthase is the key protein that synthesizes ATP molecules through a complex catalytic mechanism. While the nuclear genome encodes the majority of ATP synthase subunits, the mitochondrial genome encodes some of the enzyme's most critical components. The formation of this multi-subunit enzyme is a complex multi-step process regulated at the level of transcription, translation, and assembly. Defects in one or more of these steps can result in decreased...
18.0K
 01:27
Protein Import into the Peroxisomes
5.5K
Cells contain membrane-bound organelles called peroxisomes that oxidize organic molecules by transferring hydrogen atoms to oxygen, producing hydrogen peroxide. Peroxisomes enzymatically convert the released hydrogen peroxide into water and oxygen.
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
5.5K
 01:28
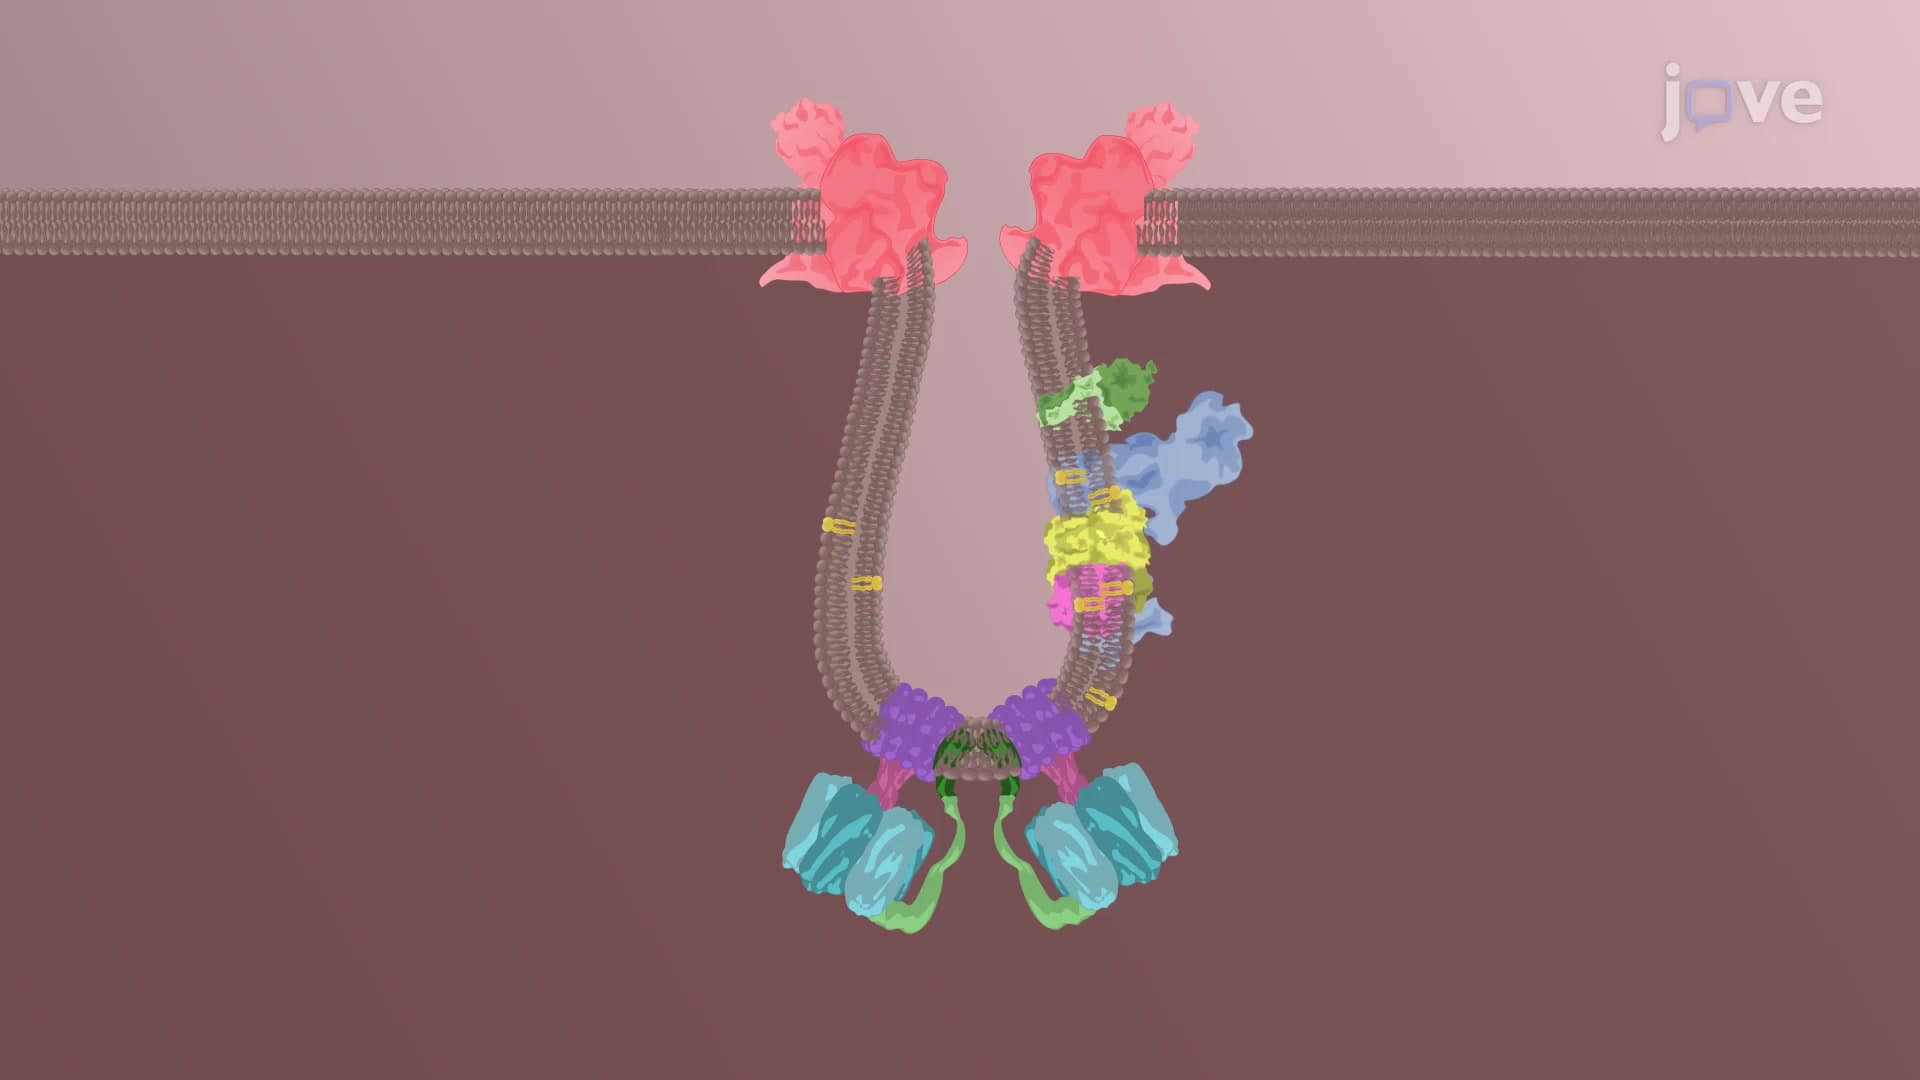
The Inner Mitochondrial Membrane
4.9K
The inner mitochondrial membrane is the primary site of ATP synthesis. The inner membrane domain that forms a smooth layer adjacent to the outer membrane is called the inner boundary membrane. This domain contains membrane transporters that drive metabolites in and out of the mitochondria. In contrast, the inner membrane network that invaginates into the matrix space is called the cristae membrane. This domain accounts for principle mitochondrial function as it accommodates the protein...
4.9K