Four faces of the interaction between ions and aromatic rings
Related Experiment Video
Updated: Mar 2, 2026
Preparation of Stable Bicyclic Aziridinium Ions and Their Ring-Opening for the Synthesis of Azaheterocycles
Published on: August 22, 2018
Dóra Papp1,2, Petra Rovó3, Imre Jákli4
1MTA-ELTE Complex Chemical Systems Research Group, H-1518 Budapest 112, P.O. Box 32, Hungary.

Abstract:
Non-covalent interactions between ions and aromatic rings play an important role in the stabilization of macromolecular complexes; of particular interest are peptides and proteins containing aromatic side chains (Phe, Trp, and Tyr) interacting with negatively (Asp and Glu) and positively (Arg and Lys) charged amino acid residues. The structures of the ion-aromatic-ring complexes are the result of an interaction between the large quadrupole moment of the ring and the charge of the ion. Four attractive interaction types are proposed to be distinguished based on the position of the ion with respect to the plane of the ring: perpendicular cation-π (CP⊥ ), co-planar cation-π (CP∥ ), perpendicular anion-π (AP⊥ ), and co-planar anion-π (AP∥ ). To understand more than the basic features of these four interaction types, a systematic, high-level quantum chemical study is performed, using the X- + C6 H6 , M+ + C6 H6 , X- + C6 F6 , and M+ + C6 F6 model systems with X- = H- , F- , Cl- , HCOO- , CH3 COO- and M+ = H+ , Li+ , Na+ , NH4+, CH3 NH3+, whereby C6 H6 and C6 F6 represent an electron-rich and an electron-deficient π system, respectively. Benchmark-quality interaction energies with small uncertainties, obtained via the so-called focal-point analysis (FPA) technique, are reported for the four interaction types. The computations reveal that the interactions lead to significant stabilization, and that the interaction energy order, given in kcal mol-1 in parentheses, is CP⊥ (23-37) > AP⊥ (14-21) > CP∥ (9-22) > AP∥ (6-16). A natural bond orbital analysis performed leads to a deeper qualitative understanding of the four interaction types. To facilitate the future quantum chemical characterization of ion-aromatic-ring interactions in large biomolecules, the performance of three density functional theory methods, B3LYP, BHandHLYP, and M06-2X, is tested against the FPA benchmarks, with the result that the M06-2X functional performs best. © 2017 Wiley Periodicals, Inc.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 09:35
Preparation of a Corannulene-functionalized Hexahelicene by CopperI-catalyzed Alkyne-azide Cycloaddition of Nonplanar Polyaromatic Units
Published on: September 18, 2016
Related Concept Videos
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:13
Electrophilic Aromatic Substitution: Fluorination and Iodination of Benzene
 01:16
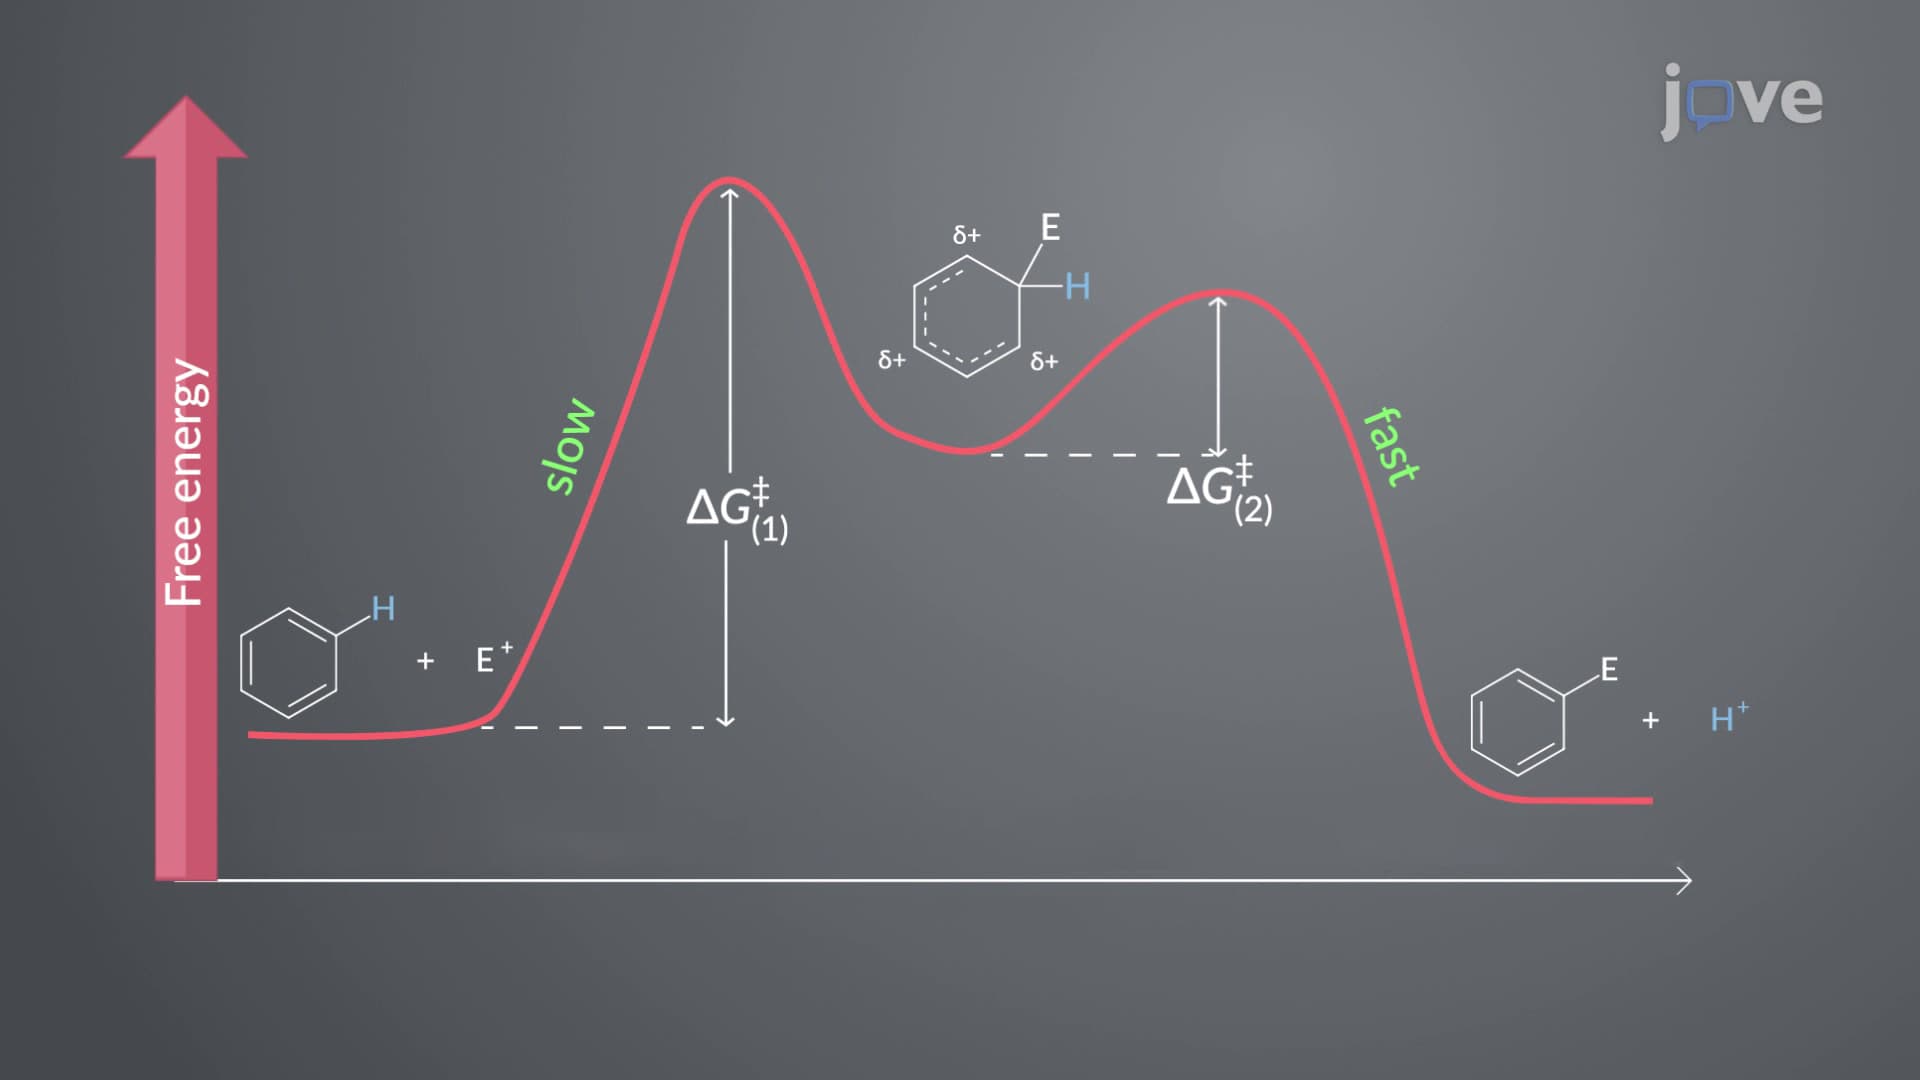
Electrophilic Aromatic Substitution: Overview