Related Experiment Video
Updated: Mar 2, 2026
An Electrochemiluminescence-Based Assay for MeCP2 Protein Variants
Published on: May 22, 2020
[MECP2 duplication syndrome: a clinical analysis of three cases and literature review]
Dan-Xia Tang1, Dong-Fang Li, Ruo-Hao Wu
1Department of Pediatrics, Sun Yat-sen Memorial Hospital, Sun Yat-sen University, Guangzhou 510210, China. luoxy33@139.com.

Insights
MECP2 duplication syndrome (MDS) is a rare pediatric disorder. Early diagnosis using array-based comparative genomic hybridization (aCGH) is crucial for children with developmental delays, recurrent infections, and seizures.
Area of Science:
- Genetics
- Pediatrics
- Neurology
Background:
- MECP2 duplication syndrome (MDS) is a rare X-linked genetic disorder.
- It primarily affects males and is characterized by severe neurodevelopmental deficits.
Observation:
- Three male patients presented with varying initial symptoms including delayed motor development, language regression, and recurrent infections.
- Hypotonia was a consistent physical finding, while epilepsy varied in presentation and management.
- Recurrent infections improved with age, but language and intellectual development showed no significant improvement.
Findings:
- Array-based comparative genomic hybridization (aCGH) confirmed MECP2 duplication on the X chromosome in all three cases, establishing the diagnosis of MDS.
- Seizure management varied, with some patients requiring antiepileptic drugs while others did not.
Implications:
- MDS should be suspected in pediatric cases with a combination of developmental delay, recurrent infections, and seizures.
- Early aCGH testing is vital for accurate and timely diagnosis of MECP2 duplication syndrome.
Abstract:
MECP2 duplication syndrome (MDS) is a rare pediatric disease and mainly manifests as delayed motor development, language loss or delay, recurrent infection, severe intellectual disability, epilepsy, autistic symptoms, and early infantile hypotonia. In this article, the three children with this disease were all boys. Cases 1 and 2 had delayed motor development, and language loss or delay as initial manifestations, and case 3 had recurrent infection as initial manifestation. Physical examination showed hypotonia and negative pathological signs in each case. Case 1 had tonic-clonic seizures and electroencephalography showed focal seizures, for which he was given oxcarbazepine, levetiracetam, and clonazepam as the antiepileptic treatment to control seizures. Case 3 experienced one absence seizure and three head-nodding seizures with normal electroencephalographic findings during these seizures, and therefore, he was not given antiepileptic treatment. In each case, recurrent infection was improved with the increase in age, but there were no significant improvements in language or intelligence. Array-based comparative genomic hybridization (aCGH) showed MECP2 duplication in X chromosome in each case, and so they were diagnosed with MDS. MDS should be considered for children with delayed development complicated by recurrent infection and epileptic seizures, and early aCGH helps with the diagnosis of this disease.
More Related Videos
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
 08:27
A Non-random Mouse Model for Pharmacological Reactivation of Mecp2 on the Inactive X Chromosome
Published on: May 22, 2019
Related Concept Videos
 01:17
Karyotyping
 03:09
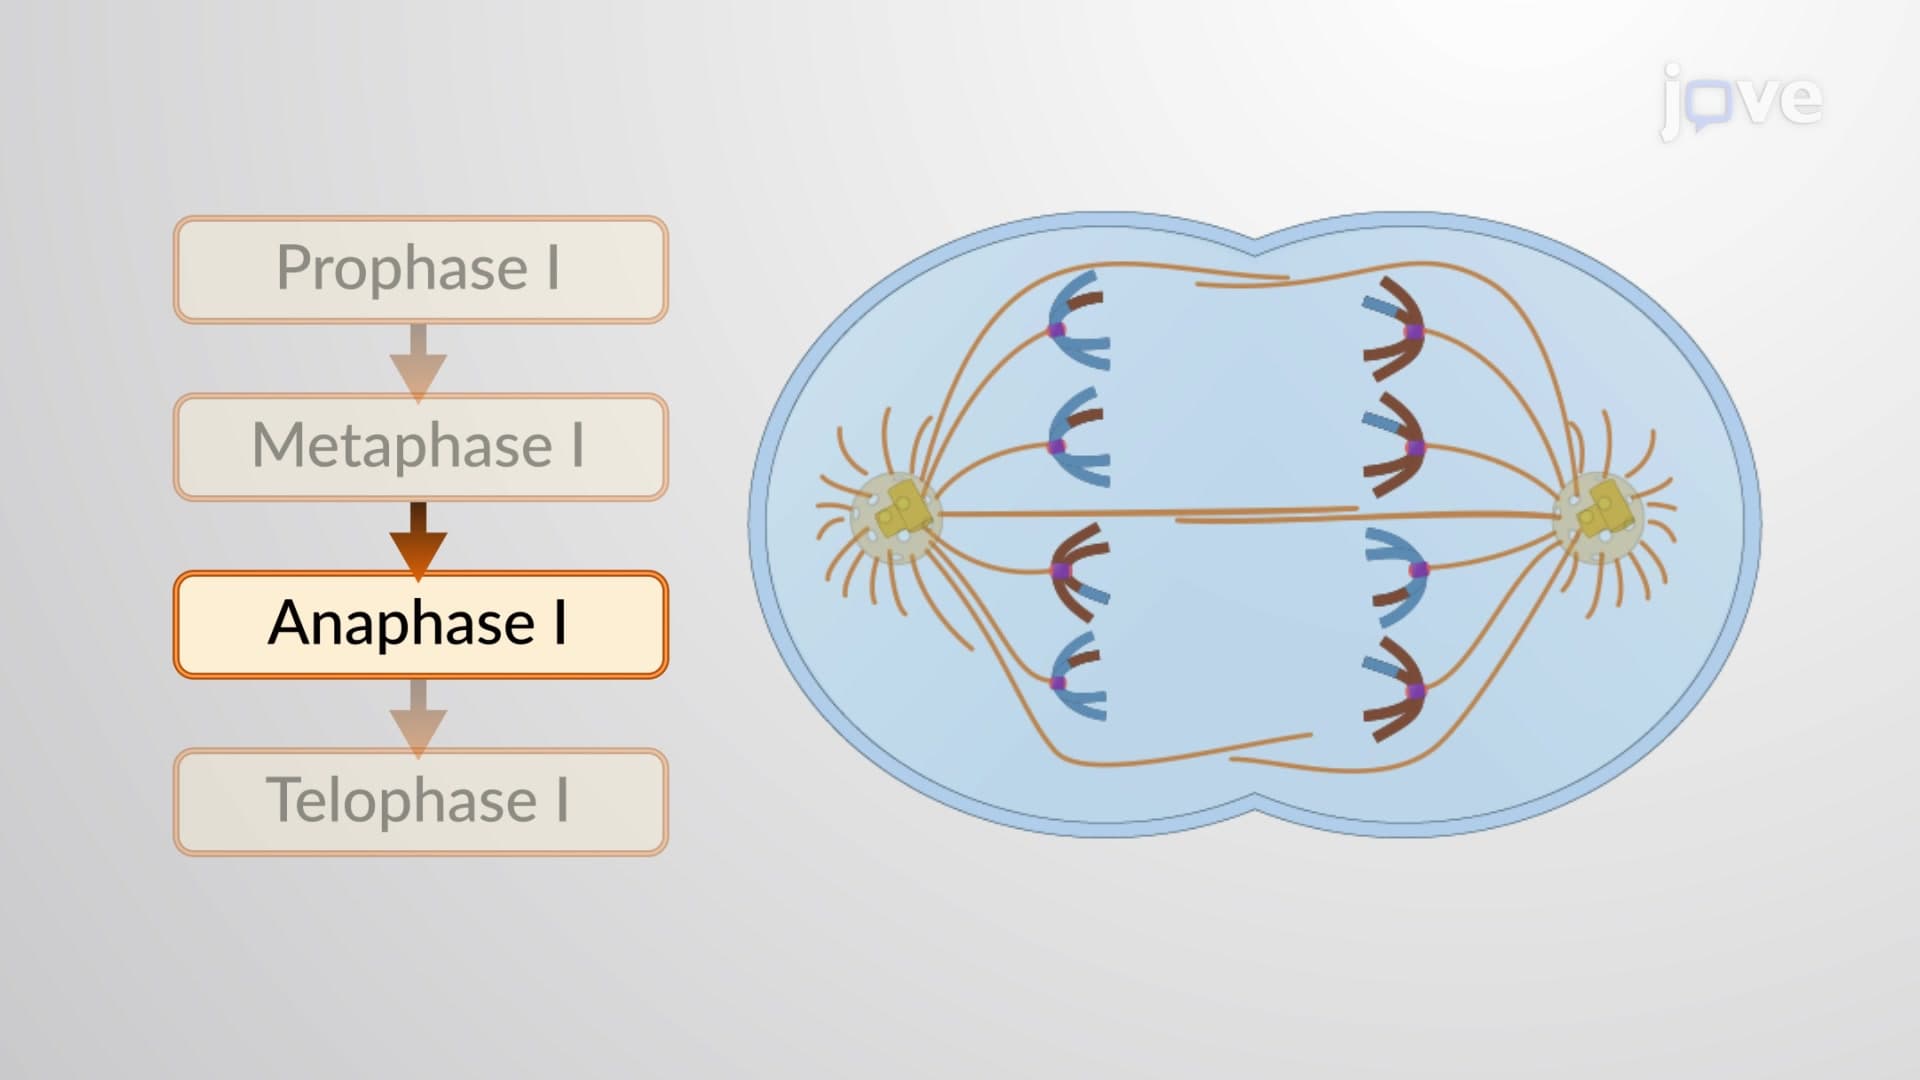
Meiosis I
Prophase I is the most extended and complex step of meiosis I characterized by synapsis, chromosome pairing, and recombination of the homologous chromosomes. This process is facilitated by a proteinaceous structure called the...
 02:23
Master Transcription Regulators