Related Experiment Video
Updated: Mar 1, 2026

Amplification of Near Full-length HIV-1 Proviruses for Next-Generation Sequencing
Published on: October 16, 2018
HIV-1 full-genome phylogenetics of generalized epidemics in sub-Saharan Africa: impact of missing nucleotide
Oliver Ratmann1, Chris Wymant2, Caroline Colijn3
1Imperial College London School of Public Health, 156430, Department of Infectious Disease Epidemiology, London, United Kingdom of Great Britain and Northern Ireland ; oliver.ratmann@imperial.ac.uk.

Abstract:
To characterize HIV-1 transmission dynamics in regions where the burden of HIV-1 is greatest, the 'Phylogenetics and Networks for Generalised HIV Epidemics in Africa' consortium (PANGEA-HIV) is sequencing full-genome viral isolates from across sub-Saharan Africa. We report the first 3,985 PANGEA-HIV consensus sequences from four cohort sites (Rakai Community Cohort Study, n=2,833; MRC/UVRI Uganda, n=701; Mochudi Prevention Project, n=359; Africa Health Research Institute Resistance Cohort, n=92). Next-generation sequencing success rates varied: more than 80% of the viral genome from the gag to the nef genes could be determined for all sequences from South Africa, 75% of sequences from Mochudi, 60% of sequences from MRC/UVRI Uganda, and 22% of sequences from Rakai. Partial sequencing failure was primarily associated with low viral load, increased for amplicons closer to the 3' end of the genome, was not associated with subtype diversity except HIV-1 subtype D, and remained significantly associated with sampling location after controlling for other factors. We assessed the impact of the missing data patterns in PANGEA-HIV sequences on phylogeny reconstruction in simulations. We found a threshold in terms of taxon sampling below which the patchy distribution of missing characters in next-generation sequences has an excess negative impact on the accuracy of HIV-1 phylogeny reconstruction, which is attributable to tree reconstruction artifacts that accumulate when branches in viral trees are long. The large number of PANGEA-HIV sequences provides unprecedented opportunities for evaluating HIV-1 transmission dynamics across sub-Saharan Africa and identifying prevention opportunities. Molecular epidemiological analyses of these data must proceed cautiously because sequence sampling remains below the identified threshold and a considerable negative impact of missing characters on phylogeny reconstruction is expected.
Related Concept Videos
 00:36
Viral Mutations
 02:54
Evolutionary Relationships through Genome Comparisons
 01:26
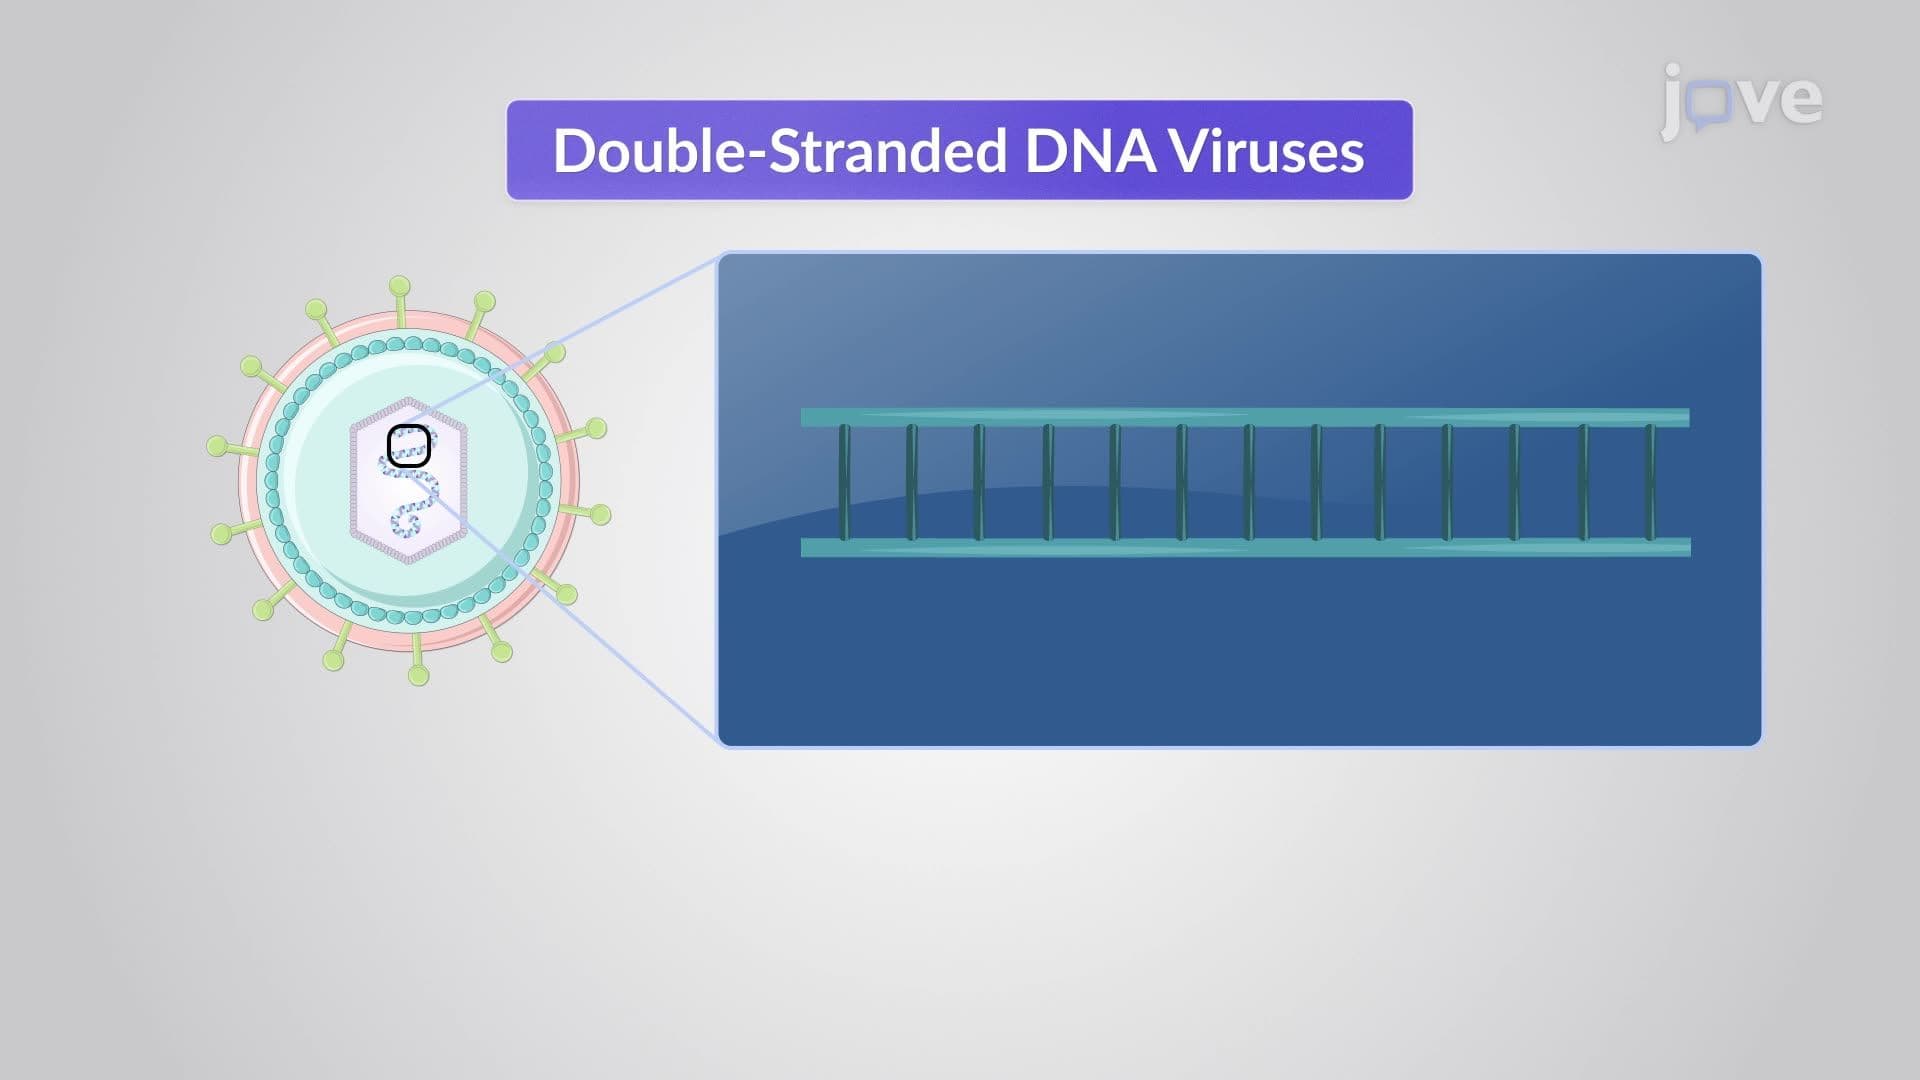
Size and Structure of Viral Genomes
 02:37
Gene Duplication and Divergence
The duplicated copies of the gene are called Paralogs. Paralogs with similar sequences and functions form a gene family. Across several species, a large number of gene families are...
 01:23
Phylogeny
 01:09
Mutation, Gene Flow, and Genetic Drift