
Determinants of the host-guest interactions between α-, β- and γ-cyclodextrins and group IA, IIA and IIIA metal
S E Angelova1, V K Nikolova, T M Dudev
1Institute of Organic Chemistry with Centre of Phytochemistry, Bulgarian Academy of Sciences, 1113 Sofia, Bulgaria.

Abstract:
The most widely used native cyclodextrins are α-, β- and γ-cyclodextrins containing six, seven or eight α-d-glucopyranoside units in the ring, respectively. Although the ligation properties of these host molecules have been extensively studied, a number of questions regarding their metal binding and selectivity remain unaddressed: to what extent do the size and flexibility of the host α-, β- and γ-cyclodextrins influence their metal affinity/selectivity? Which metal is the most preferred binding partner of α-, β- and γ-cyclodextrins? How do the charge, size and preferred coordination number of the metal cation shape its interactions with the host cyclodextrin? Can the guest metal cation inflict structural alterations in the host molecule and, if so, how do these changes correlate with the metal's properties? In the present study, by employing density functional theory (DFT) calculations combined with polarizable continuum model (PCM) computations, we try to answer these questions by evaluating the thermodynamic parameters of the IA, IIA and IIIA group metal binding to α, β- and γ-cyclodextrins. We assess how the interaction between the two binding partners depends on (1) the size, valence state and preferred coordination number of the guest metal cations, (2) the size and flexibility of the host molecule, and (3) the dielectric properties of the environment. The series of group IA (Na+ and Rb+), IIA (Mg2+ and Sr2+) and IIIA (Al3+ and In3+) metal cations have been chosen for the task as they allow us to study the effect of various metal parameters (variable charge, ionic radius and coordination number) on the strength and form of the interactions with the host cyclodextrins.
More Related Videos
 07:54
Analysis of AtHIRD11 Intrinsic Disorder and Binding Towards Metal Ions by Capillary Gel Electrophoresis and Affinity Capillary Electrophoresis
Published on: August 22, 2018
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Related Concept Videos
 01:09
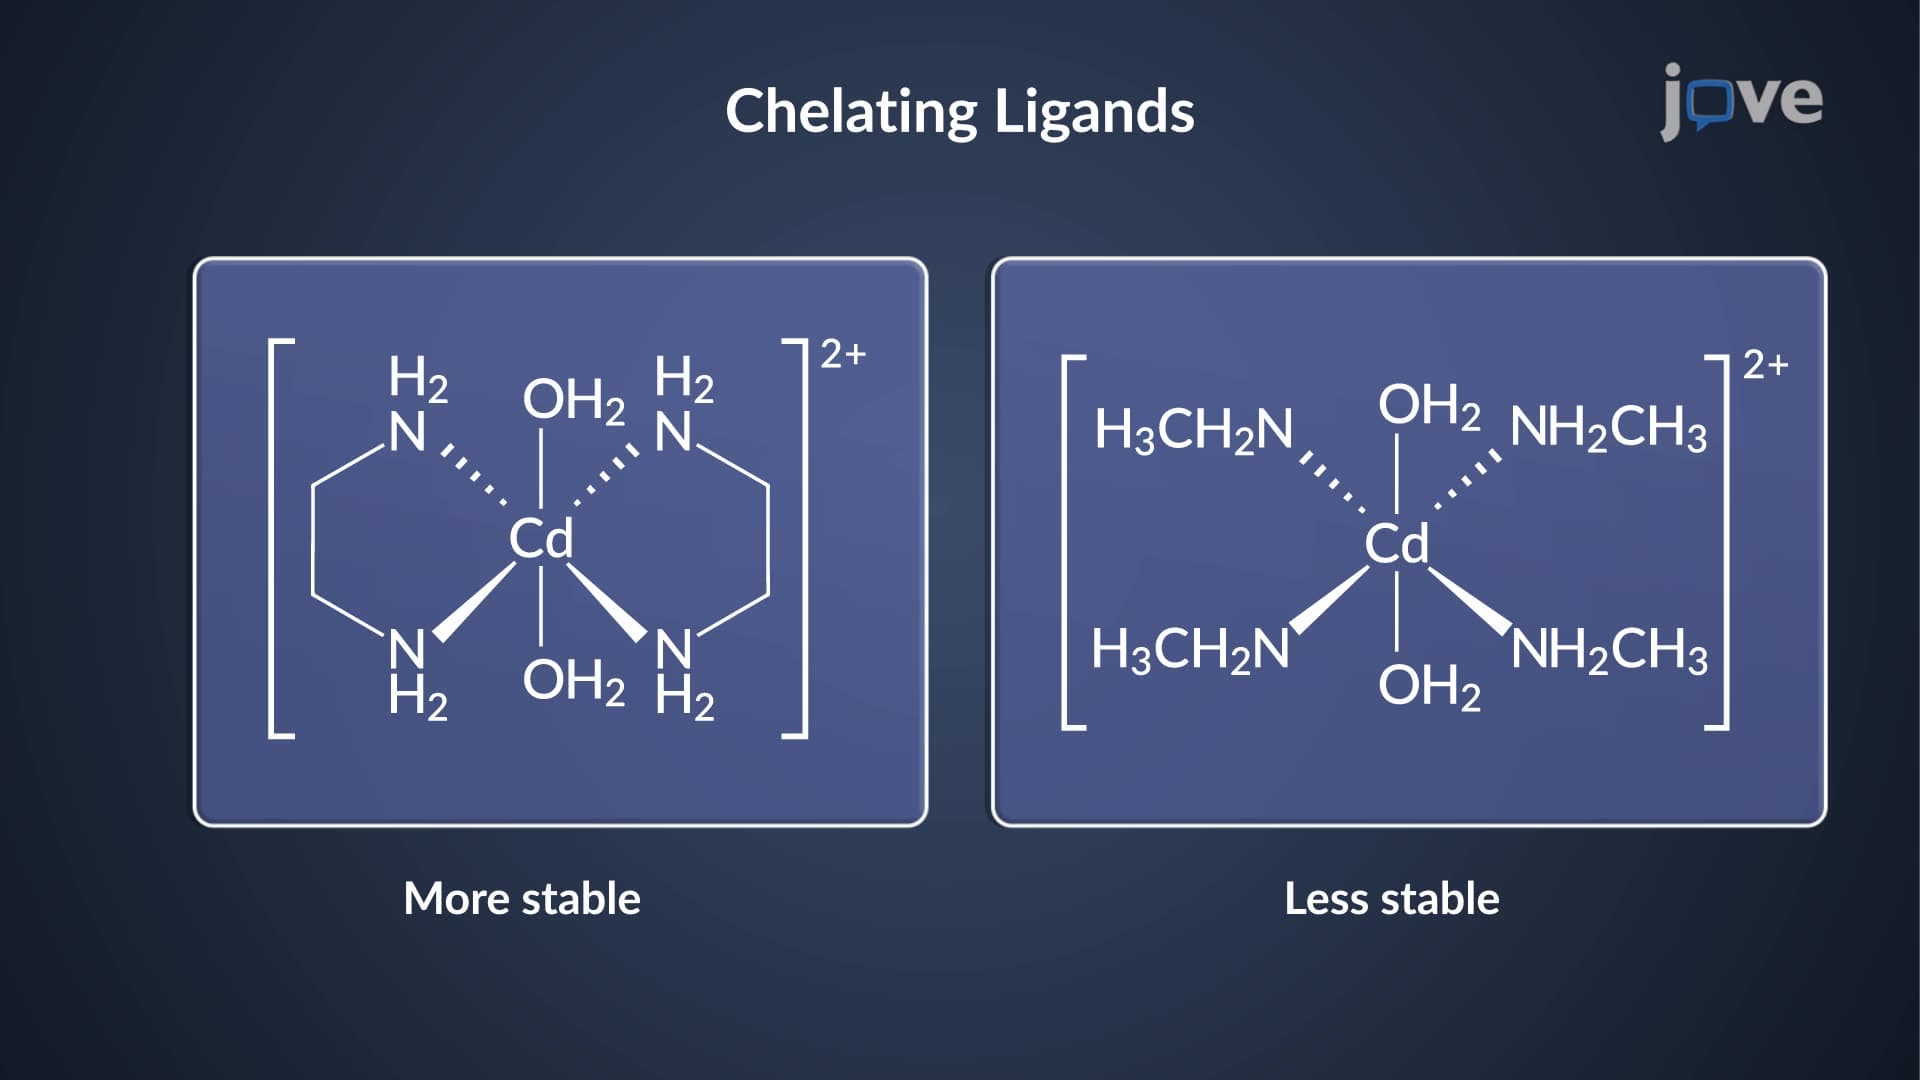
Complexation Equilibria: Factors Influencing Stability of Complexes
 01:19
Complexation Equilibria: The Chelate Effect
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 00:56
Extraction: Advanced Methods