Related Experiment Video
Updated: Aug 13, 2026

Characterization and Isolation of Mouse Primary Microglia by Density Gradient Centrifugation
Published on: February 16, 2018
Ischemia-responsive protein 94 is a key mediator of ischemic neuronal injury-induced microglial activation
Rajiv Tikamdas1, Sarthak Singhal1, Ping Zhang1
1Department of Pharmacodynamics, College of Pharmacy, University of Florida, Gainesville, Florida, USA.

Abstract:
Neuroinflammation, especially activation of microglia, the key immune cells in the brain, has been proposed to contribute to the pathogenesis of ischemic stroke. However, the dynamics and the potential mediators of microglial activation following ischemic neuronal injury are not well understood. In this study, using oxygen/glucose deprivation and reoxygenation with neuronal and microglial cell cultures as an in vitro model of ischemic neuronal injury, we set out to identify neuronal factors released from injured neurons that are capable of inducing microglial activation. Conditioned media (CM) from hippocampal and cortical neurons exposed to oxygen/glucose deprivation and reoxygenation induced significant activation of microglial cells as well as primary microglia, evidenced by up-regulation of inducible nitric oxide synthase, increased production of nitrite and reactive oxygen species, and increased expression of microglial markers. Mechanistically, neuronal ischemia-responsive protein 94 (Irp94) was a key contributor to microglial activation since significant increase in Irp94 was detected in the neuronal CM following ischemic insult and immunodepletion of Irp94 rendered ischemic neuronal CM ineffective in inducing microglial activation. Ischemic insult-augmented oxidative stress was a major facilitator of neuronal Irp94 release, and pharmacological inhibition of NADPH oxidase significantly reduced the ischemic injury-induced neuronal reactive oxygen species production and Irp94 release. Taken together, these results indicate that neuronal Irp94 may play a pivotal role in the propagation of ischemic neuronal damage. Continued studies may help identify Irp94 and/or related proteins as potential therapeutic targets and/or diagnostic/prognostic biomarkers for managing ischemia-associated brain disorders.
Insights
Injured neurons release ischemia-responsive protein 94 (Irp94), activating microglia and worsening stroke damage. Inhibiting oxidative stress reduces Irp94 release, suggesting a therapeutic target for brain disorders.
Area of Science:
- Neuroscience
- Immunology
- Cell Biology
Background:
- Neuroinflammation, particularly microglial activation, is implicated in ischemic stroke pathogenesis.
- The precise mediators and dynamics of microglial activation post-ischemic neuronal injury remain unclear.
Purpose of the Study:
- To identify specific factors released by injured neurons that trigger microglial activation.
- To elucidate the role of neuronal ischemia-responsive protein 94 (Irp94) in this process.
Main Methods:
- Utilized an in vitro model of ischemic neuronal injury (oxygen/glucose deprivation and reoxygenation) with neuronal and microglial cell cultures.
- Analyzed conditioned media from stressed neurons for factors inducing microglial activation.
- Investigated the role of Irp94 via immunodepletion and measured oxidative stress markers.
Main Results:
- Conditioned media from ischemic neurons significantly activated microglia, increasing inflammatory markers.
- Neuronal Irp94 levels were elevated in conditioned media; its depletion blocked microglial activation.
- Oxidative stress facilitated Irp94 release, which was reduced by inhibiting NADPH oxidase.
Conclusions:
- Neuronal Irp94 is a key mediator of microglial activation following ischemic injury.
- Irp94 may propagate ischemic neuronal damage, presenting a potential therapeutic target.
- Further research could establish Irp94 as a biomarker for ischemia-associated brain disorders.
Related Concept Videos
 01:31
Regulation of the Unfolded Protein Response
 01:16
Cellular Injury IV: Necrosis
 01:15
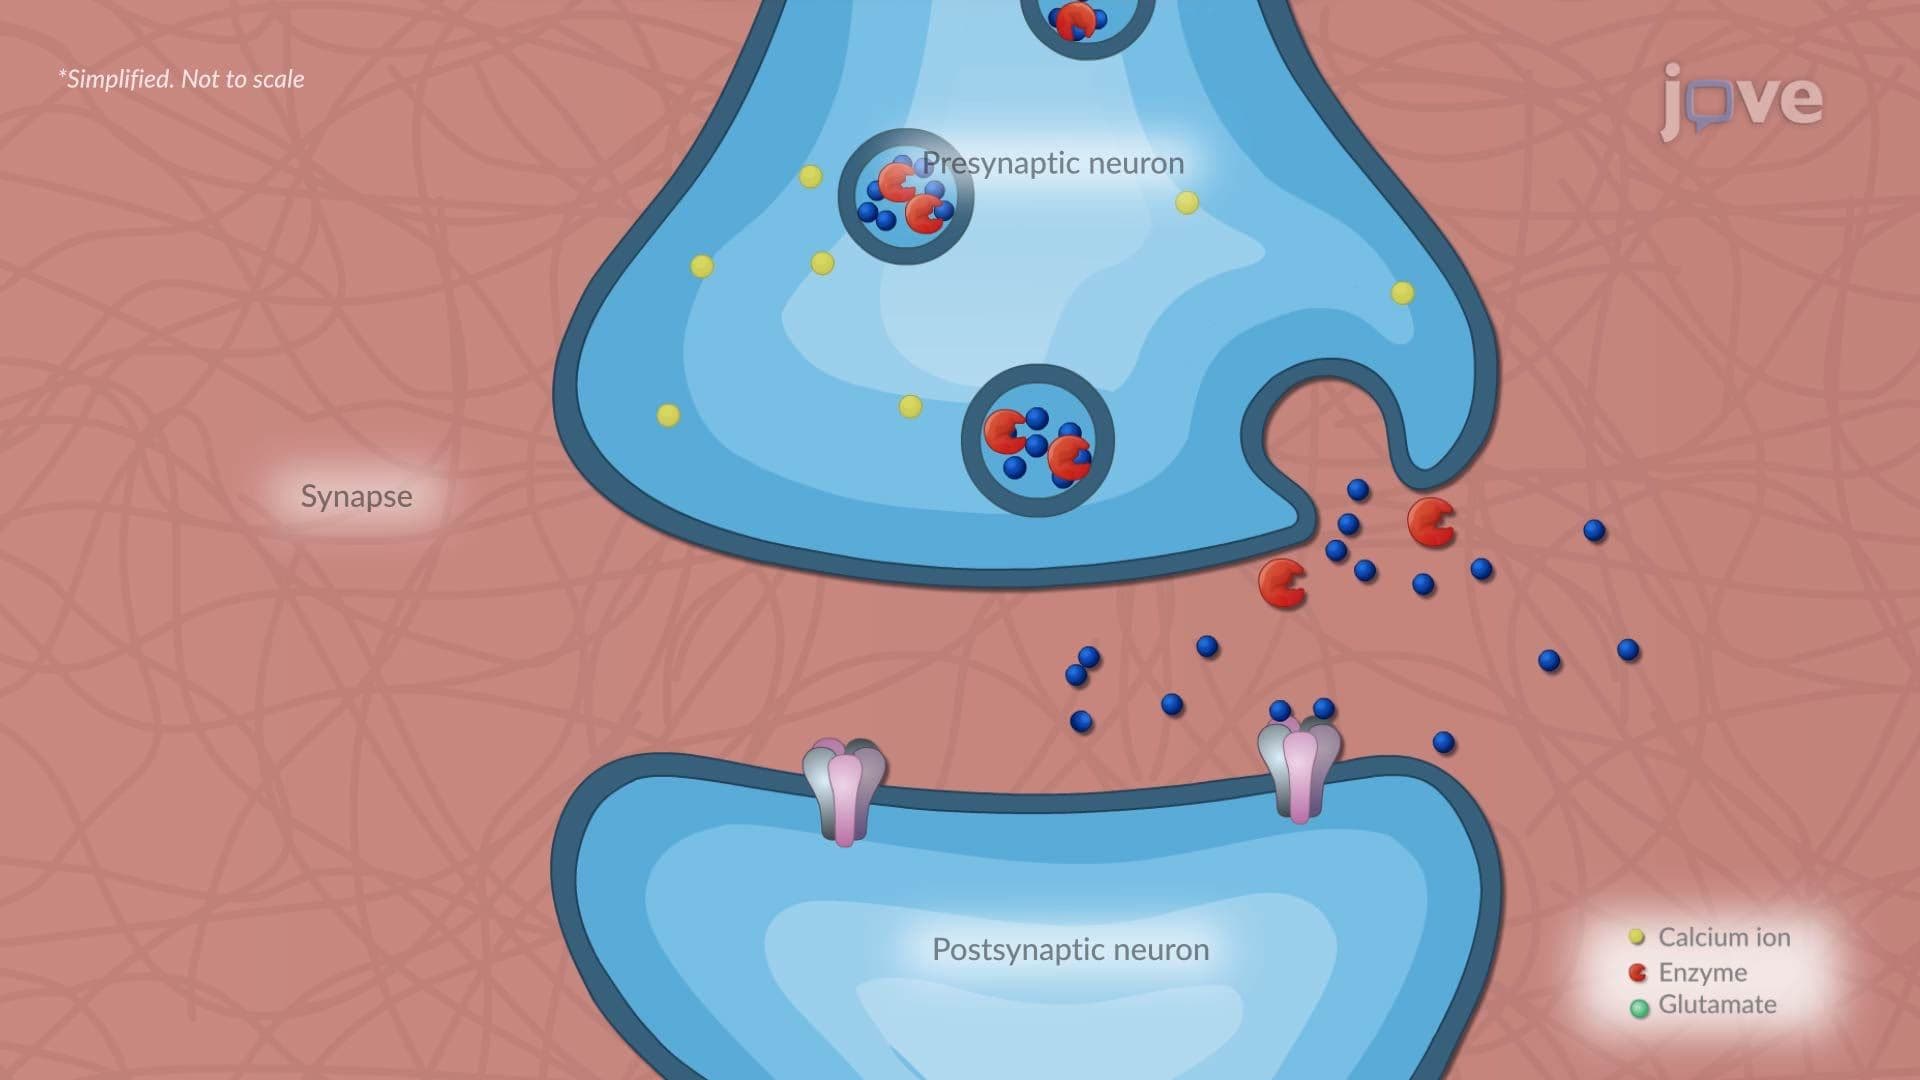
Ischemic Stroke ll: Pathophysiology
 01:25
Secondary Spinal Cord Injury llI: Pathophysiology