Related Experiment Video
Updated: Feb 27, 2026

Extraction of Non-Protein Amino Acids from Cyanobacteria for Liquid Chromatography-Tandem Mass Spectrometry Analysis
Published on: December 9, 2022
Amino acid synthesis deficiencies
1Paediatrician for Inborn Errors of Metabolism, University of Groningen, University Medical Centre Groningen, Groningen, Netherlands. t.j.de.koning@umcg.nl.

Abstract:
In recent years the number of disorders known to affect amino acid synthesis has grown rapidly. Nor is it just the number of disorders that has increased: the associated clinical phenotypes have also expanded spectacularly, primarily due to the advances of next generation sequencing diagnostics. In contrast to the "classical" inborn errors of metabolism in catabolic pathways, in which elevated levels of metabolites are easily detected in body fluids, synthesis defects present with low values of metabolites or, confusingly, even completely normal levels of amino acids. This makes the biochemical diagnosis of this relatively new group of metabolic diseases challenging. Defects in the synthesis pathways of serine metabolism, glutamine, proline and, recently, asparagine have all been reported. Although these amino acid synthesis defects are in unrelated metabolic pathways, they do share many clinical features. In children the central nervous system is primarily affected, giving rise to (congenital) microcephaly, early onset seizures and varying degrees of mental disability. The brain abnormalities are accompanied by skin disorders such as cutis laxa in defects of proline synthesis, collodion-like skin and ichthyosis in serine deficiency, and necrolytic erythema in glutamine deficiency. Hypomyelination with accompanying loss of brain volume and gyration defects can be observed on brain MRI in all synthesis disorders. In adults with defects in serine or proline synthesis, spastic paraplegia and several forms of polyneuropathy with or without intellectual disability appear to be the major symptoms in these late-presenting forms of amino acid disorders. This review provides a comprehensive overview of the disorders in amino acid synthesis.
More Related Videos
Related Concept Videos
 01:29
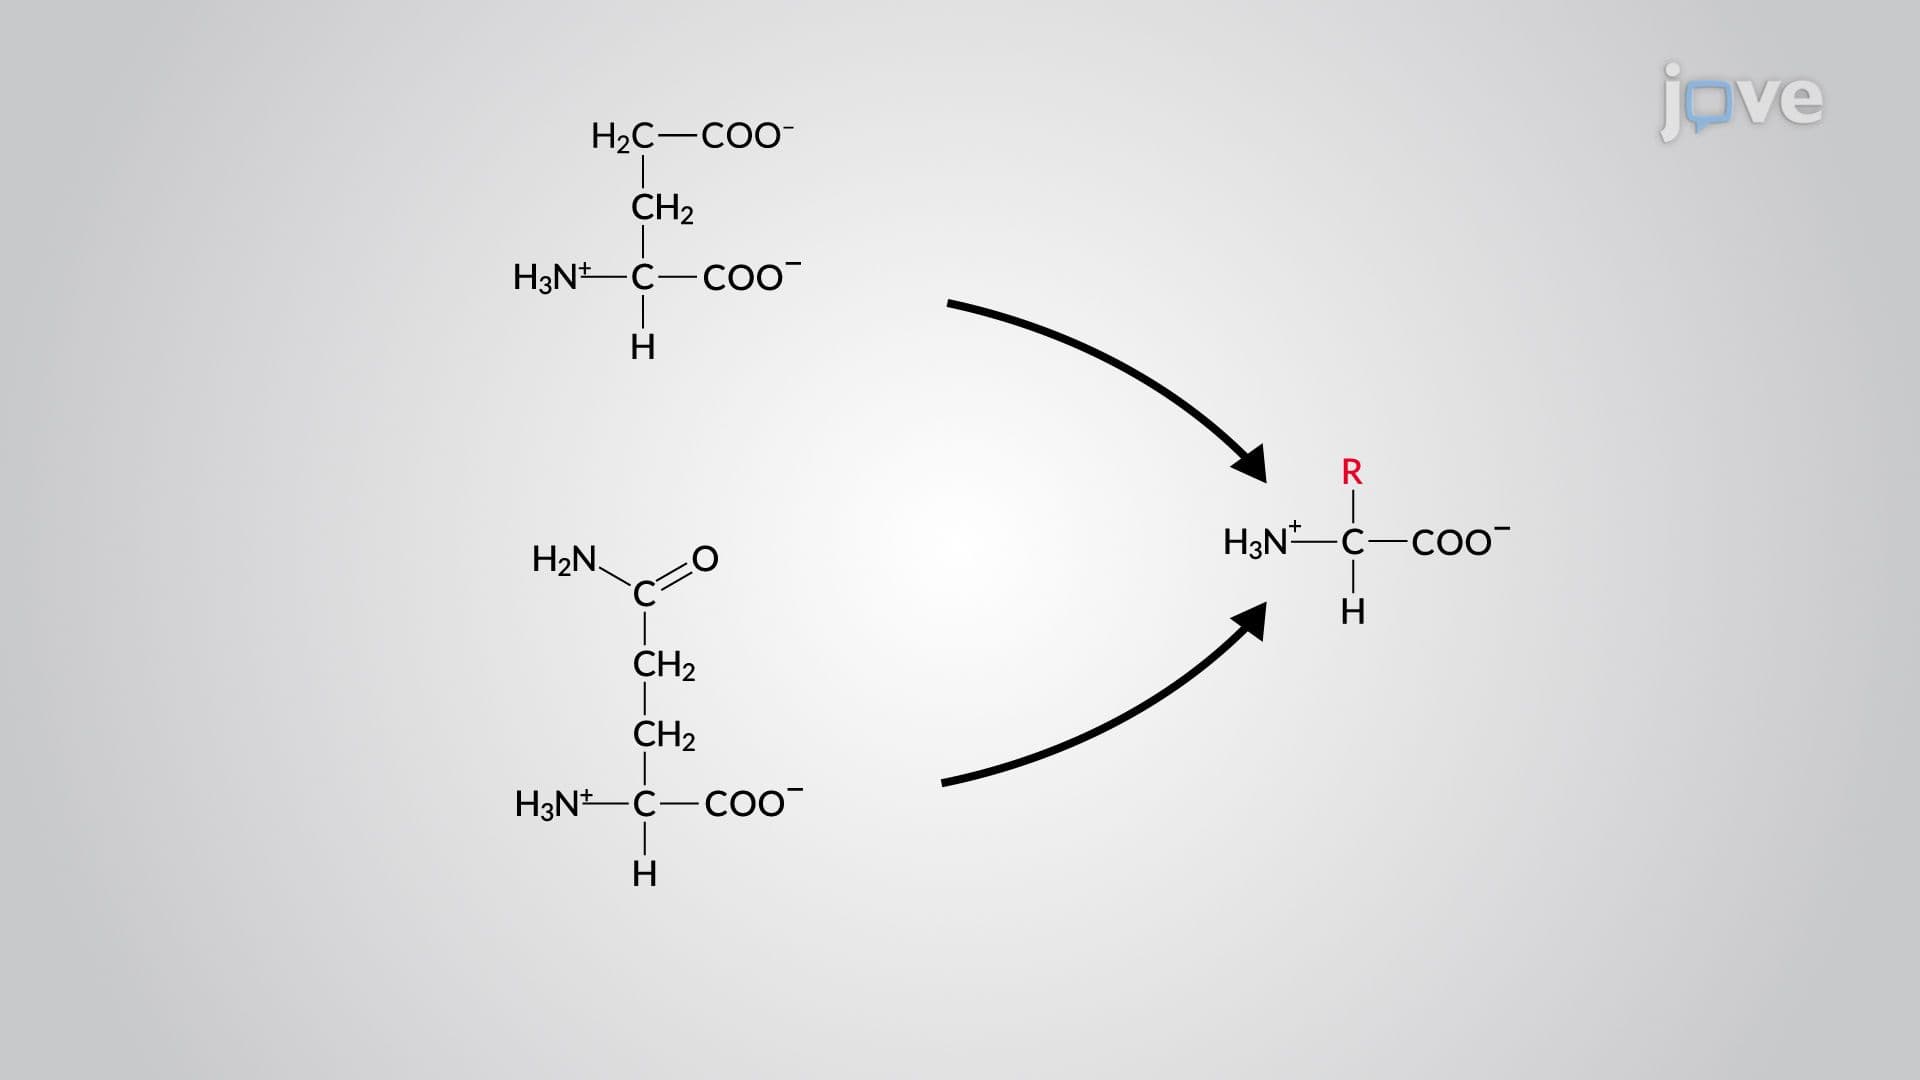
Amino Acid Biosynthetic Pathways
 01:28
Biosynthesis of Nucleic Acids
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:20
Inborn Errors of Metabolism
 01:48
ATP Synthase: Mechanism
 01:23
Urea Cycle