Related Experiment Video
Updated: Feb 27, 2026
Facile Preparation of 2Z,4E-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Combined Experimental and Computational Study on Ruthenium(II)-Catalyzed Reactions of Diynes with Aldehydes and
Yoshihiko Yamamoto1, Yuta Okude1, Shota Mori1
1Department of Basic Medicinal Sciences, Graduate School of Pharmaceutical Sciences, Nagoya University , Chikusa, Nagoya 464-8601, Japan.

Abstract:
Cycloaddition reactions of 1,6-diynes bearing methyl terminal groups with p-anisaldehyde were conducted using a cationic ruthenium catalyst with a η5-pentamethylcyclopentadienyl ligand in THF at room temperature to afford dienyl ketones via ring opening of the initially formed fused pyrans. (Z)-Stereoisomers of dienyl ketones were selectively obtained using the ruthenium catalyst, whereas previously reported rhodium catalysts produced (E)-isomers. These (E)- and (Z)-selectivities are kinetically controlled as the control experiments showed that the E/Z-isomerization of (E)-dienylketone occurs at 70 °C for 10 h to afford an E/Z-ratio of almost 1:1. The origin of this characteristic stereoselectivity for the ruthenium catalyst was attributed to the direct ring opening of the CpRu+-coordinated pyran complex intermediates on the basis of theoretical calculations [PCM (THF) M06L/SDD-6-311++G(d,p)//B3LYP/LanL2DZ-6-31G(d)] and control experiments. The (Z)-selectivity increased when the bulkiness of the diyne terminal substituents increased. Notably, the reaction of 1,6-diynes bearing tert-butyl terminal groups with various α,β-unsaturated aldehydes exclusively afforded (Z)-dienyl ketones even at 70 °C when a cationic ruthenium complex with a smaller η5-cyclopentadienyl (Cp) ligand was used as the catalyst. The same Cp complex was found to be also efficient for the hydrocarbamoylative cyclization of sterically demanding 1,6-diynes bearing tertiary or quaternary carbon tethers with N,N-dimethylformamide.
More Related Videos
Related Concept Videos
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 01:16
[4+2] Cycloaddition of Conjugated Dienes: Diels–Alder Reaction
 02:34
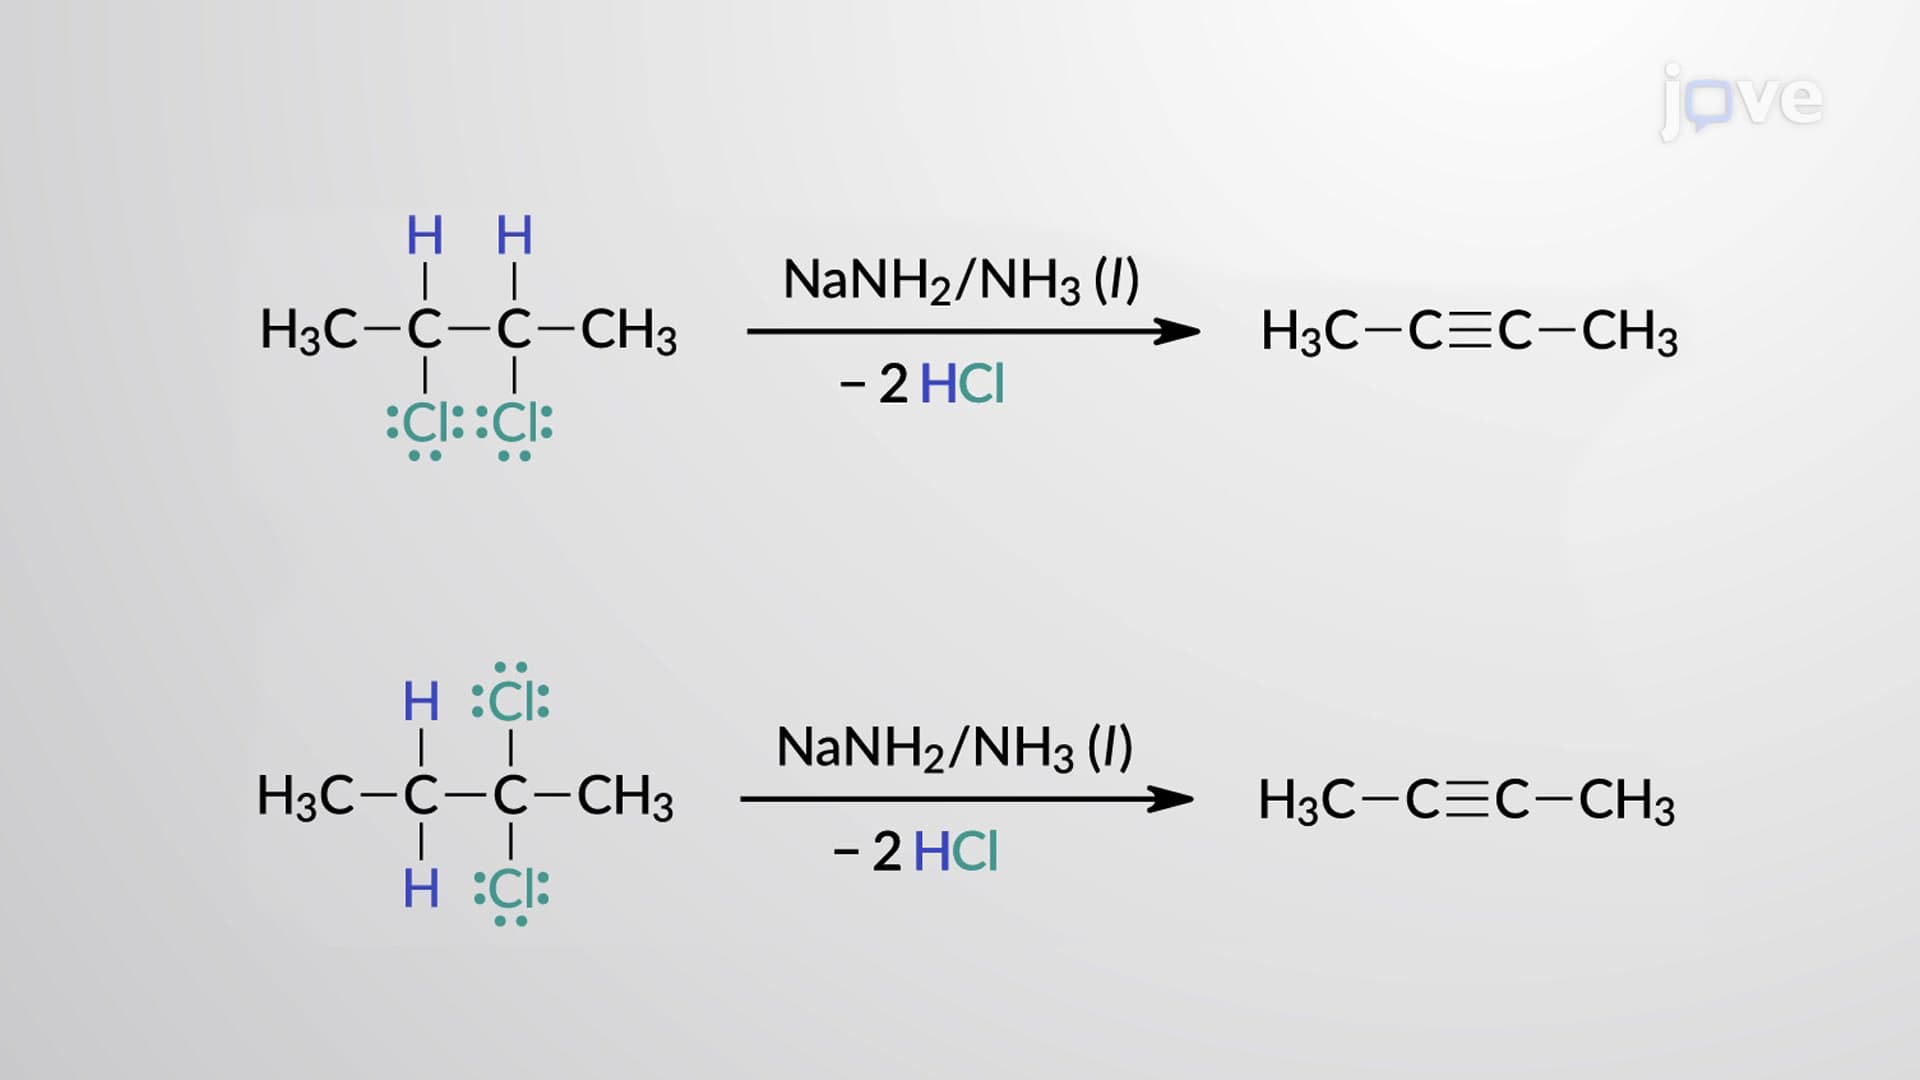
Preparation of Alkynes: Dehydrohalogenation
Alkynes can be prepared by dehydrohalogenation of vicinal or geminal dihalides in the presence of a strong base like sodium amide in liquid ammonia. The reaction proceeds with the loss of two equivalents of hydrogen halide (HX) via two successive E2 elimination reactions.
 01:33
Carboxylic Acids to Methylesters: Alkylation using Diazomethane
 01:22
Alkenes via Reductive Coupling of Aldehydes or Ketones: McMurry Reaction
 02:47
Alkynes to Aldehydes and Ketones: Hydroboration-Oxidation
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.