Related Experiment Video
Updated: Feb 26, 2026

Facile Preparation of 2Z,4E-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Two Enabling Strategies for the Stereoselective Conversion of Internal Alkynes into Trisubstituted Alkenes
Nikolas Huwyler1, Karin Radkowski1, Stephan M Rummelt1
1Max-Planck-Institut für Kohlenforschung, 45470, Mülheim/Ruhr, Germany.

Abstract:
An expedient method for the C-methylation of alkenylstannanes with formation of trisubstituted alkenes is described, which relies on the use of MeI in combination with copper thiophene-2-carboxylate (CuTC) as promotor and tetra-n-butylammonium diphenylphosphinate as an effective tin scavenger; in some cases, it proved beneficial to further supplement the mixture with catalytic amounts of Pd(PPh3 )4 . Under these conditions, the reaction is robust, high yielding, and compatible with many functional groups that might not subsist under more traditional conditions used to C-alkylate organotin derivatives. A qualitative analysis of the reaction profile suggested that the in situ formation of a reactive organocopper intermediate and its interception by MeI is only barely faster than O-methylation of the phosphinate additive by the same alkylating agent. To guarantee high yields and prevent net protodestannation from occurring, the reaction protocol had to be optimized such that these competing processes are properly decoupled. The new method is particularly well suited for the stereoselective preparation of the (E)-2-methylbut-2-en-1-ol motif that is present in numerous natural products. Alternatively, this particular target structure can be accessed starting from α-hydroxy alkenylsiloxane precursors, which get C-methylated upon exposure to CuI/LiOtBu and MeI by what is thought to be a Brook rearrangement/ alkylation sequence. The required substrates are best prepared by ruthenium-catalyzed trans-hydrosilylation or trans-hydrostannation of propargyl alcohols.
More Related Videos
 09:54
Chemoselective Preparation of 1-Iodoalkynes, 1,2-Diiodoalkenes, and 1,1,2-Triiodoalkenes Based on the Oxidative Iodination of Terminal Alkynes
Published on: September 12, 2018
 08:12
A Two-Step Protocol for Umpolung Functionalization of Ketones Via Enolonium Species
Published on: August 16, 2018
Related Concept Videos
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 02:47
Alkynes to Aldehydes and Ketones: Hydroboration-Oxidation
One of the convenient methods for the preparation of aldehydes and ketones is via hydration of alkynes. Hydroboration-oxidation of alkynes is an indirect hydration reaction in which an alkyne is treated with borane followed by oxidation with alkaline peroxide to form an enol that rapidly converts into an aldehyde or a ketone. Terminal alkynes form aldehydes, whereas internal alkynes give ketones as the final product.
 02:27
Preparation of Alkynes: Alkylation Reaction
Alkylation of terminal alkynes with primary alkyl halides in the presence of a strong base like sodium amide is one of the common methods for the synthesis of longer carbon-chain alkynes. For example, treatment of 1-propyne with sodium amide followed by reaction with ethyl bromide yields 2-pentyne.
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 02:48
Conversion of Alcohols to Alkyl Halides
 02:34
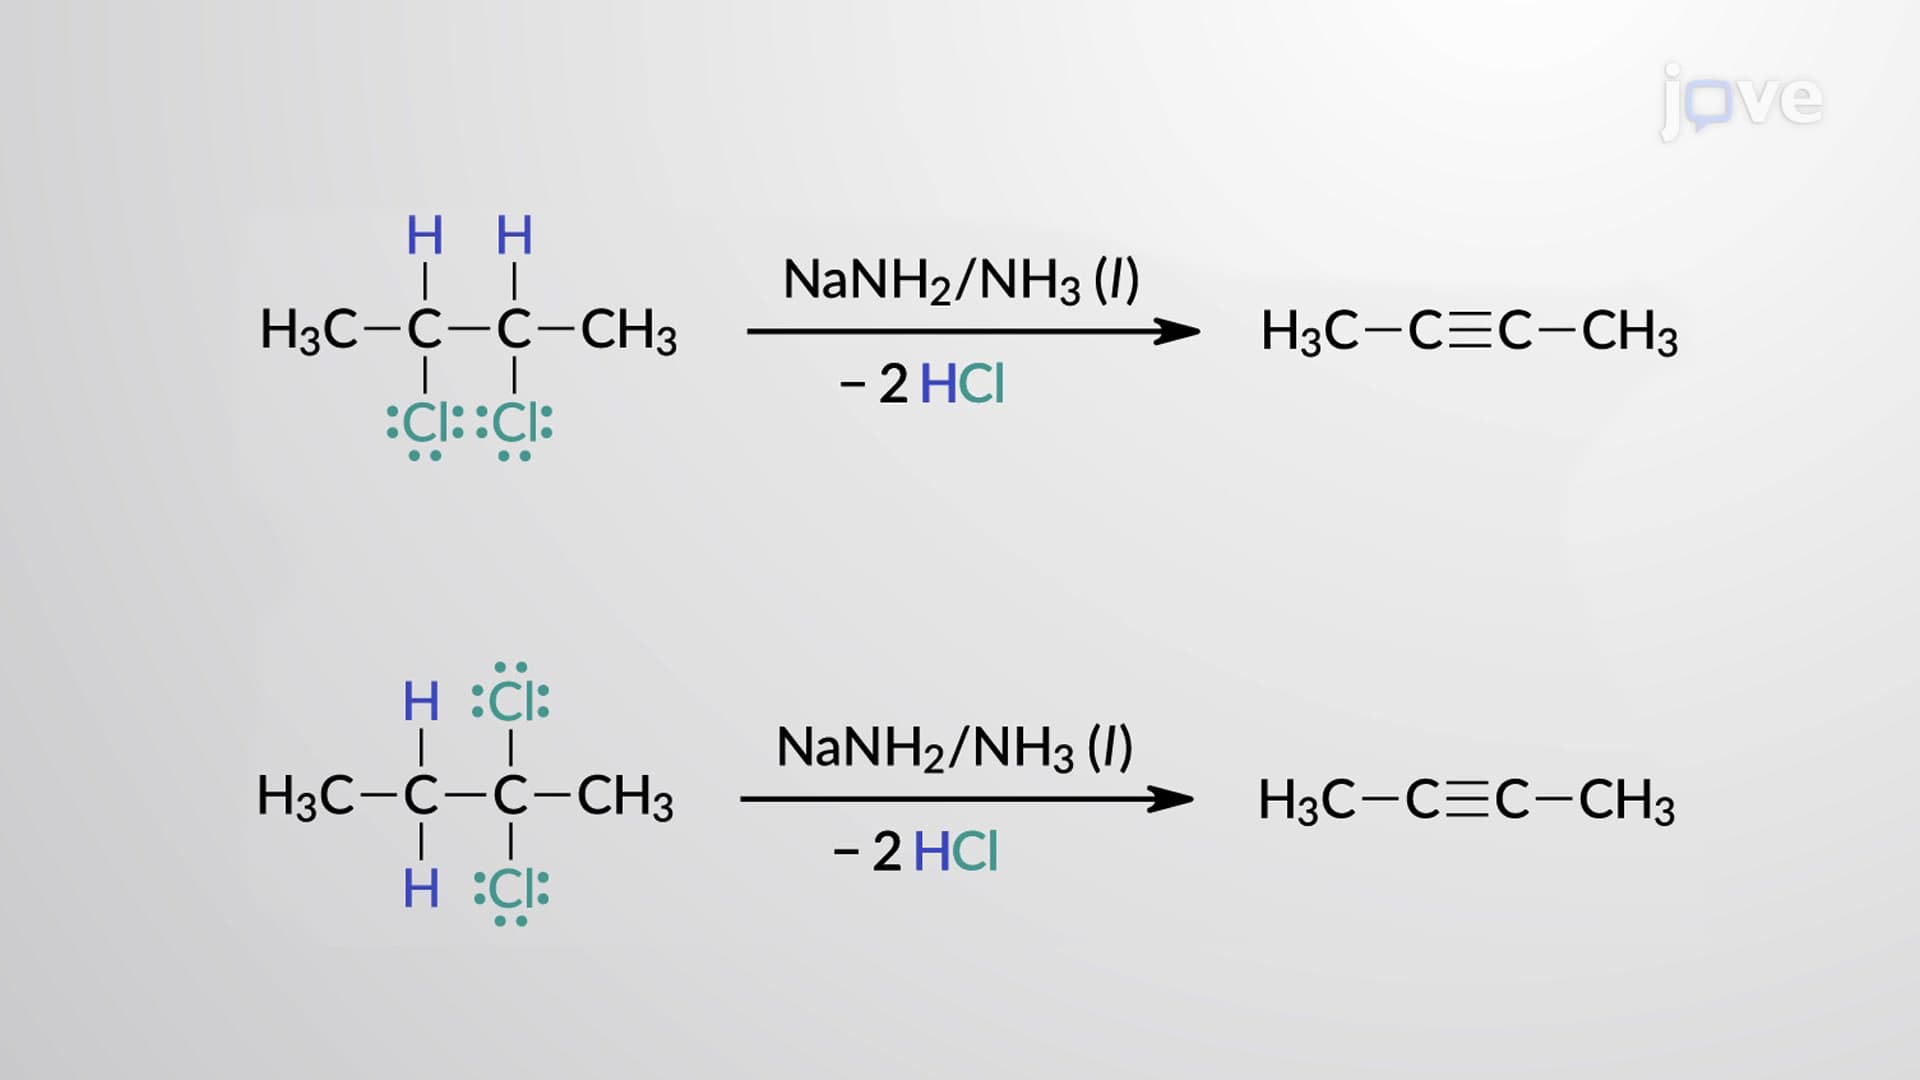
Preparation of Alkynes: Dehydrohalogenation
Alkynes can be prepared by dehydrohalogenation of vicinal or geminal dihalides in the presence of a strong base like sodium amide in liquid ammonia. The reaction proceeds with the loss of two equivalents of hydrogen halide (HX) via two successive E2 elimination reactions.