Related Experiment Video
Updated: Feb 26, 2026

Photoelectron Imaging of Anions Illustrated by 310 Nm Detachment of F−
Published on: July 27, 2018
Angular Dependence of Strong Field Ionization of CH3X (X = F, Cl, Br, or I) Using Time-Dependent Configuration
Paul Hoerner1, H Bernhard Schlegel1
1Department of Chemistry, Wayne State University , Detroit, Michigan 48202, United States.

Abstract:
Methyl halides have been used to test basis set effects on simulations of strong field ionization using time dependent configuration interaction with an absorbing potential. Standard atom centered basis sets need to be augmented by several sets of diffuse functions on each atom so that the wave function in the strong field can interact with the absorbing potential used to model ionization. An absorbing basis of 3 s functions, 2 p functions, 3 d functions, and 1 f function is sufficient for CH3F. Large absorbing basis sets with 4 s functions, 3 or 4 p functions, 4 or 5 d functions, and 2 f functions are recommended for the heavier halogens. The simulations used static fields in the 0.035-0.07 au range to explore the angular dependence of ionization of methyl halides. CH3F ionizes mainly from the methyl group; CH3Cl and CH3Br show ionization from both the methyl group and the halogen, and CH3I ionizes almost exclusively from the pπ orbitals of the iodine.

Related Concept Videos
 03:07
Electron Affinity
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:35
Hybridization of Atomic Orbitals II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:24
Hybridization of Atomic Orbitals I
 01:22
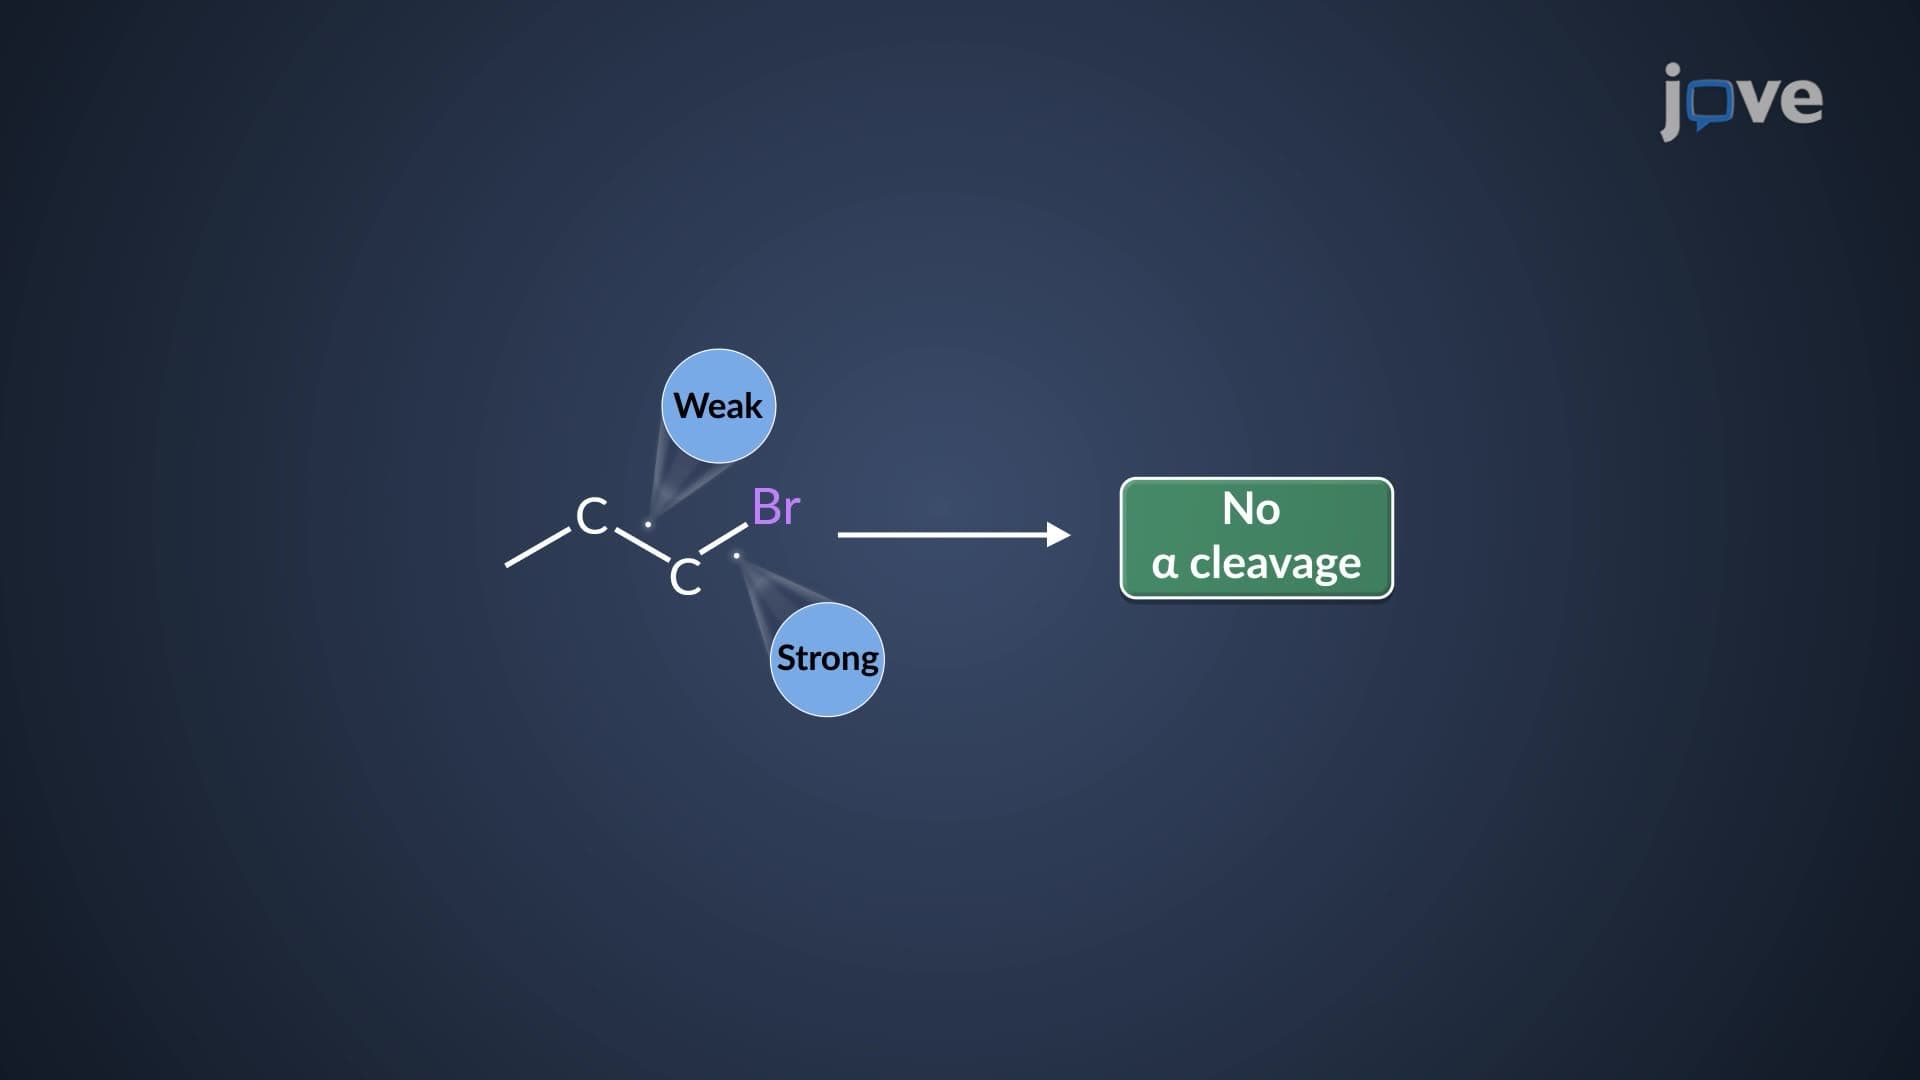
Mass Spectrometry: Alkyl Halide Fragmentation