Related Experiment Video
Updated: Feb 25, 2026
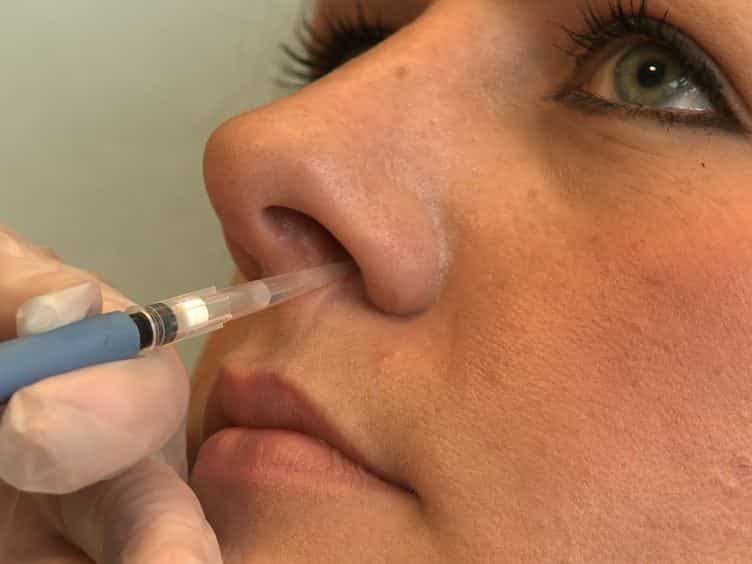

Experimental Human Pneumococcal Carriage
Published on: February 15, 2013
Genome-wide identification of lineage and locus specific variation associated with pneumococcal carriage duration
John A Lees1, Nicholas J Croucher2, David Goldblatt3
1Infection Genomics, Wellcome Trust Sanger Institute, Hinxton, United Kingdom.

Insights
Genomic variation in Streptococcus pneumoniae explains most carriage duration differences, not host factors. Prophage sequences significantly reduce carriage time, impacting bacterial competition and antibiotic resistance dynamics.
Area of Science:
- Microbiology
- Genomics
- Epidemiology
Background:
- Streptococcus pneumoniae causes invasive infant disease, particularly in low-income regions.
- Nasopharyngeal carriage is essential for disease but its duration varies.
- Current understanding of carriage duration variability is limited, primarily at the serotype level.
Purpose of the Study:
- To develop a model for calculating pneumococcal carriage episode duration.
- To investigate the contribution of genomic variation to carriage duration.
- To identify specific genomic factors influencing pneumococcal carriage duration.
Main Methods:
- Developed a model to estimate carriage duration from longitudinal swab data.
- Integrated carriage duration data with whole genome sequencing data.
- Performed a pan-genome-wide association study (GWAS) to identify genetic associations.
Main Results:
- Pneumococcal genomic variation accounted for 63% of carriage duration variation; host traits explained <5%.
- Serotype (17%), drug resistance (9%), and other loci contributed significantly to duration variation.
- Prophage sequences were associated with decreased carriage duration, independent of serotype.
Conclusions:
- Genomic factors, particularly prophages, are major drivers of pneumococcal carriage duration.
- Findings support models of pneumococcal competition and the evolution of antibiotic resistance.
- Understanding genomic influences on carriage duration can inform public health strategies.
Abstract:
Streptococcus pneumoniae is a leading cause of invasive disease in infants, especially in low-income settings. Asymptomatic carriage in the nasopharynx is a prerequisite for disease, but variability in its duration is currently only understood at the serotype level. Here we developed a model to calculate the duration of carriage episodes from longitudinal swab data, and combined these results with whole genome sequence data. We estimated that pneumococcal genomic variation accounted for 63% of the phenotype variation, whereas the host traits considered here (age and previous carriage) accounted for less than 5%. We further partitioned this heritability into both lineage and locus effects, and quantified the amount attributable to the largest sources of variation in carriage duration: serotype (17%), drug-resistance (9%) and other significant locus effects (7%). A pan-genome-wide association study identified prophage sequences as being associated with decreased carriage duration independent of serotype, potentially by disruption of the competence mechanism. These findings support theoretical models of pneumococcal competition and antibiotic resistance.

Related Concept Videos
 01:29
Modern Molecular Taxonomy
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:05
Single Nucleotide Polymorphisms-SNPs