Related Experiment Video
Updated: Feb 25, 2026

Bergmeyer Glucose Quantification for Microbiological Samples
Published on: January 17, 2025
Glycolate oxidase deficiency in a patient with congenital hyperinsulinism and unexplained hyperoxaluria
Oliver Clifford-Mobley1, Gill Rumsby2, Swati Kanodia3
1Clinical Biochemistry, Bristol Royal Infirmary, University Hospitals Bristol NHS Foundation Trust, Bristol, BS2 8HW, UK. oliver.clifford-mobley@nhs.net.

Background:
A baby girl was born at 39 weeks gestation to consanguineous Asian parents. From day 1 of life she had severe hypoglycaemia with an inappropriately elevated insulin concentration consistent with congenital hyperinsulinism (CHI), confirmed by the finding of a homozygous mutation in ABCC8 (encoding the sulfonylurea receptor 1).
Case Diagnosis/Treatment:
Urine organic acid analysis showed an incidentally elevated excretion of glycolate. Whilst this was unlikely to contribute to the hypoglycaemia, hyperglycolic aciduria is a known feature of primary hyperoxaluria type 1 (PH1); therefore oxalate was also measured in urine and found to be elevated. Sequence analysis of the genes involved in PH1 and also the two other known forms of primary hyperoxaluria revealed no pathological variants. PH1 was definitively excluded by enzyme activity analysis on a liver biopsy, which confirmed normal glyoxylate aminotransferase (AGT) activity and positive AGT immunoreactivity. Glycolate oxidase (GO) deficiency was considered, and thus gene sequencing of HAO1, which encodes GO, was performed. A homozygous change (c.493G>T p.(Gly165Cys)) was found in exon 3 of HAO1, predicted to be deleterious to protein function. Further analysis of the liver biopsy demonstrated absent GO enzyme activity, confirming GO deficiency in this case.
Conclusions:
The results lead to the conclusion that this baby has two unrelated autosomal recessive conditions, CHI and GO deficiency, and also hyperoxaluria of unknown aetiology. Deficiency of GO is a very rare disorder with only two previously published cases. It is considered to be an essentially benign inborn error of metabolism. The present case is unique in that GO deficiency is associated with persistent hyperoxaluria.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:15
Pyruvate Oxidation
First, the enzyme pyruvate dehydrogenase removes the carboxyl group from pyruvate and releases it as carbon dioxide. The stripped molecule is then oxidized and releases electrons, which are then picked up by NAD+...
 01:15
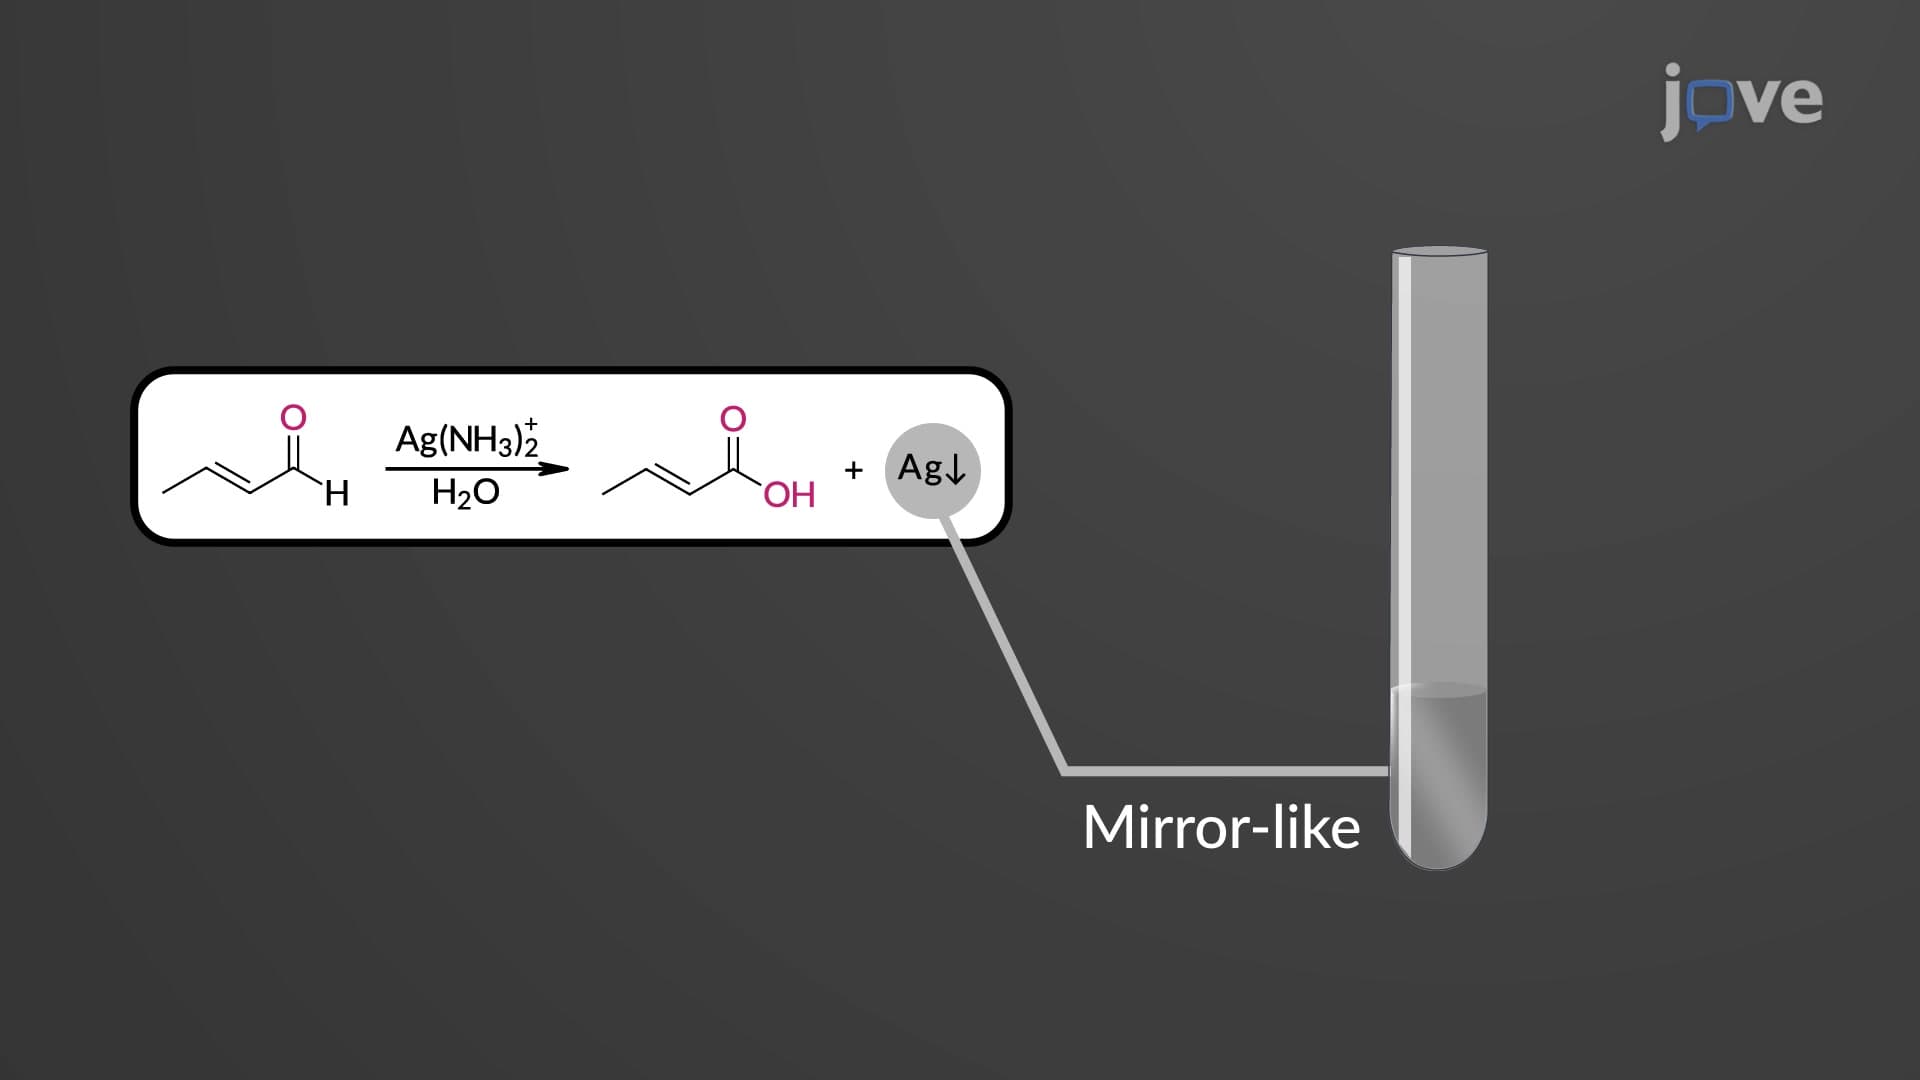
Oxidations of Aldehydes and Ketones to Carboxylic Acids
Aldehydes readily undergo oxidation in strong oxidizing agents such as potassium permanganate and chromic acid. The oxidation can also be carried out using mild oxidizing agents such as silver oxide. In fact, aldehydes can be easily oxidized...
 02:37
Oxidation of Alcohols
The process of oxidation in a chemical reaction is observed in any of the three forms:
 01:05
Proteoglycans