Related Experiment Video
Updated: Feb 25, 2026

Chemical Vapor Deposition of an Organic Magnet, Vanadium Tetracyanoethylene
Published on: July 3, 2015
Electronic structure, magnetic properties, and mixed valence character of Ce2 Ni3 Si5 from first principles
1Faculty of Science, Department of Chemistry, Helwan University, Cairo, 11795, Egypt.

Abstract:
Cerium intermetallic compounds exhibit anomalous physical properties such as heavy fermion and Kondo behaviors. Here, an ab initio study of the electronic structure, magnetic properties, and mixed valence character of Ce2 Ni3 Si5 using density functional theory (DFT) is presented. Two theoretical methods, including pure Perdew-Burke-Ernzerhof (PBE) and PBE + U, are used. In this study, Ce3+ and Ce4+ are considered as two different constituents in the unit cell. The formation energy calculations on the DFT level propose that Ce is in a stable mixed valence of 3.379 at 0 K. The calculated electronic structure shows that Ce2 Ni3 Si5 is a metallic compound with a contribution at the Fermi level from Ce 4f and Ni 3d states. With the inclusion of the effective Hubbard parameter (Ueff ), the five valence electrons of 5 Ce3+ ions are distributed only on Ce3+ 4f orbitals. Therefore, the occupied Ce3+ 4f band is located in the valence band (VB) while Ce4+ 4f orbitals are empty and Located at the Fermi level. The calculated magnetic moment in Ce2 Ni3 Si5 is only due to cerium (Ce3+ ) in good agreement with the experimental results. The Ueff value of 5.4 eV provides a reasonable magnetic moment of 0.981 μB for the unpaired electron per Ce3+ ion. These results may serve as a guide for studying present mixed valence cerium-based compounds. © 2017 Wiley Periodicals, Inc.
More Related Videos

Related Concept Videos
 03:26
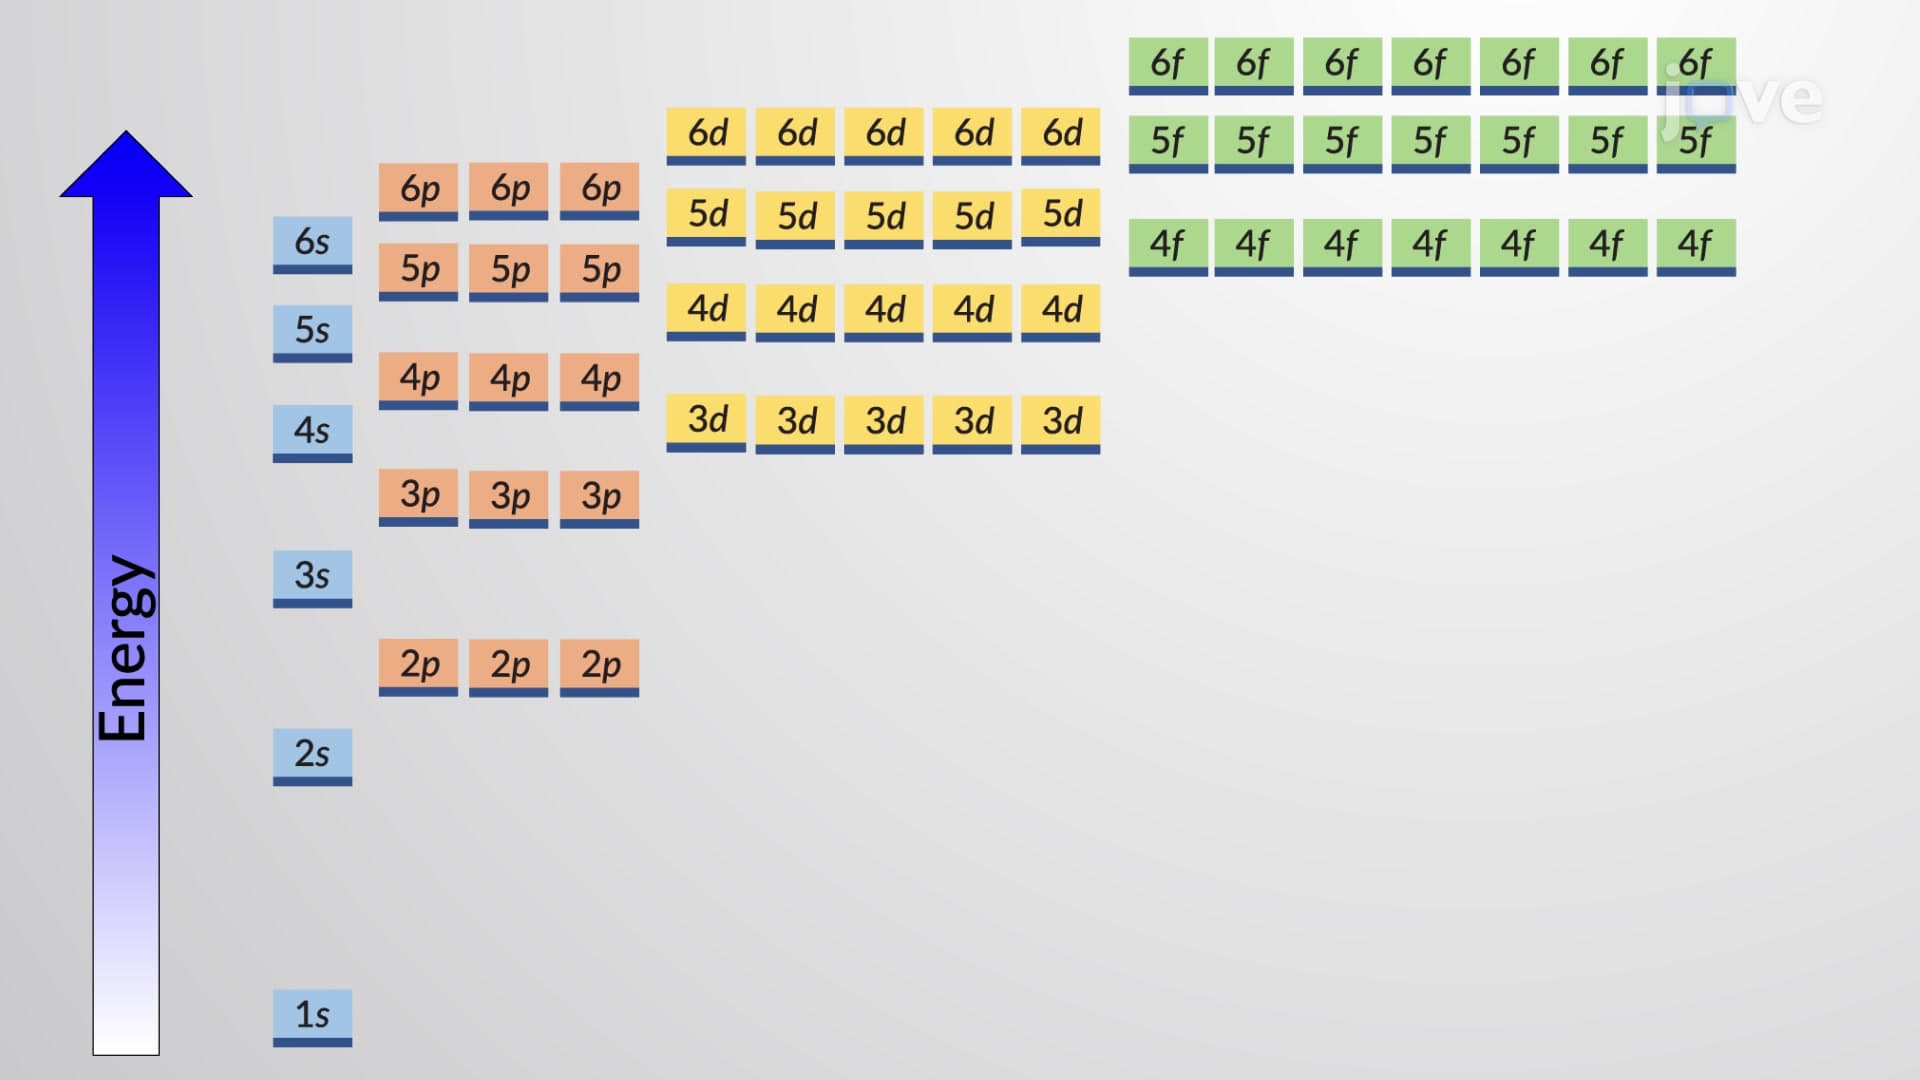
Electron Configuration of Multielectron Atoms
 02:42
Valence Bond Theory
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
 02:48
Ionic Bonding and Electron Transfer
 02:27
Predicting Molecular Geometry
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...