Related Experiment Video
Updated: Feb 25, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
An open-source framework for analyzing N-electron dynamics. II. Hybrid density functional theory/configuration
Gunter Hermann1, Vincent Pohl1, Jean Christophe Tremblay1
1Institut für Chemie und Biochemie, Freie Universität Berlin, Takustraße 3, Berlin, 14195, Germany.

Abstract:
In this contribution, we extend our framework for analyzing and visualizing correlated many-electron dynamics to non-variational, highly scalable electronic structure method. Specifically, an explicitly time-dependent electronic wave packet is written as a linear combination of N-electron wave functions at the configuration interaction singles (CIS) level, which are obtained from a reference time-dependent density functional theory (TDDFT) calculation. The procedure is implemented in the open-source Python program detCI@ORBKIT, which extends the capabilities of our recently published post-processing toolbox (Hermann et al., J. Comput. Chem. 2016, 37, 1511). From the output of standard quantum chemistry packages using atom-centered Gaussian-type basis functions, the framework exploits the multideterminental structure of the hybrid TDDFT/CIS wave packet to compute fundamental one-electron quantities such as difference electronic densities, transient electronic flux densities, and transition dipole moments. The hybrid scheme is benchmarked against wave function data for the laser-driven state selective excitation in LiH. It is shown that all features of the electron dynamics are in good quantitative agreement with the higher-level method provided a judicious choice of functional is made. Broadband excitation of a medium-sized organic chromophore further demonstrates the scalability of the method. In addition, the time-dependent flux densities unravel the mechanistic details of the simulated charge migration process at a glance. © 2017 Wiley Periodicals, Inc.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:38
Valence Bond Theory and Hybridized Orbitals
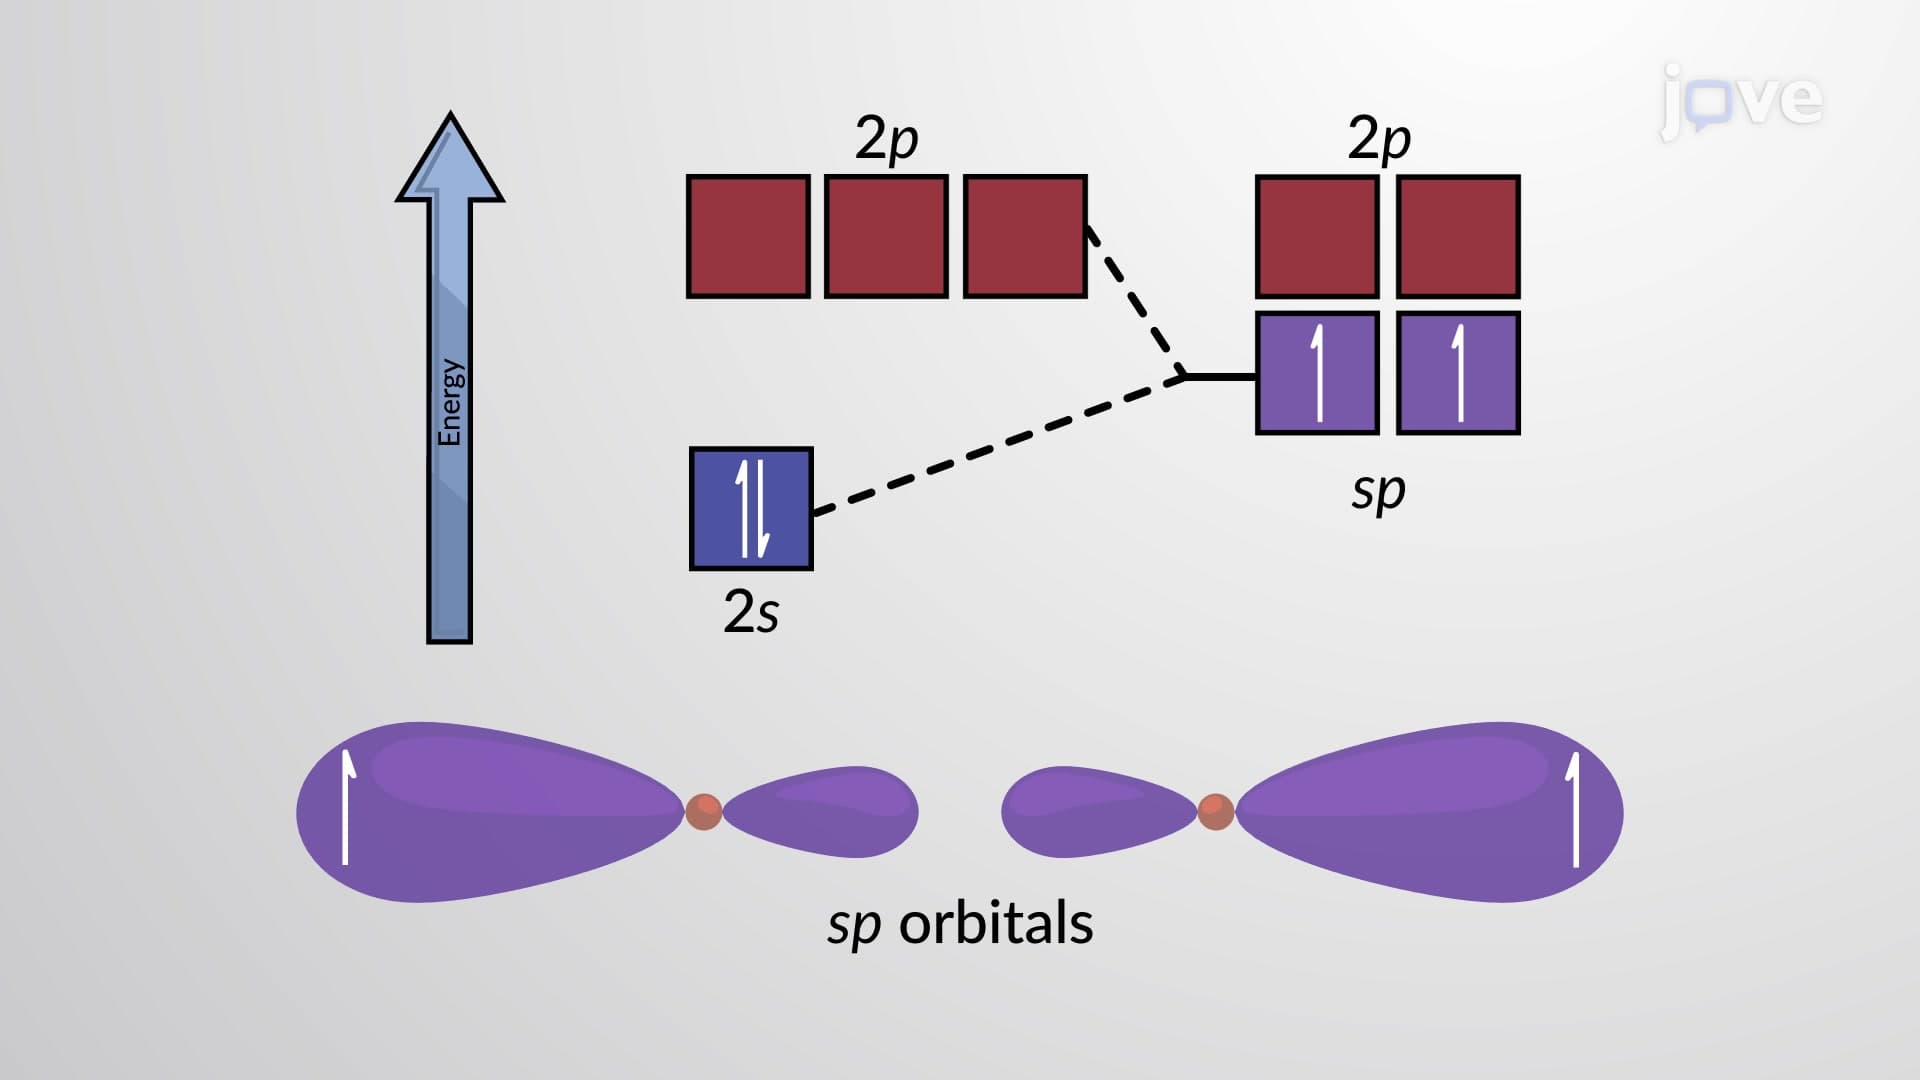
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:35
Hybridization of Atomic Orbitals II
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 03:24
Hybridization of Atomic Orbitals I
 01:27
π Electron Effects on Chemical Shift: Overview
 02:35
Molecular Orbital Theory I