Related Experiment Video
Updated: Feb 24, 2026

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Multiscale Simulation of Receptor-Drug Association Kinetics: Application to Neuraminidase Inhibitors
Fabian Zeller1, Manuel P Luitz1, Rainer Bomblies1
1Physik-Department T38, Technische Universität München , James-Franck-Str. 1, 85748 Garching, Germany.

Abstract:
A detailed understanding of the drug-receptor association process is of fundamental importance for drug design. Due to the long time scales of typical binding kinetics, the atomistic simulation of the ligand traveling from bulk solution into the binding site is still computationally challenging. In this work, we apply a multiscale approach of combined Molecular Dynamics (MD) and Brownian Dynamics (BD) simulations to investigate association pathway ensembles for the two prominent H1N1 neuraminidase inhibitors oseltamivir and zanamivir. Including knowledge of the approximate binding site location allows for the selective confinement of detailed but expensive MD simulations and application of less demanding BD simulations for the diffusion controlled part of the association pathway. We evaluate a binding criterion based on the residence time of the inhibitor in the binding pocket and compare it to geometric criteria that require prior knowledge about the binding mechanism. The method ranks the association rates of both inhibitors in qualitative agreement with experiment and yields reasonable absolute values depending, however, on the reaction criteria. The simulated association pathway ensembles reveal that, first, ligands are oriented in the electrostatic field of the receptor. Subsequently, a salt bridge is formed between the inhibitor's carboxyl group and neuraminidase residue Arg368, followed by adopting the native binding mode. Unexpectedly, despite oseltamivir's higher overall association rate, the rate into the intermediate salt-bridge state was found to be higher for zanamivir. The present methodology is intrinsically parallelizable and, although computationally demanding, allows systematic binding rate calculation on selected sets of potential drug molecules.
Related Concept Videos
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:18
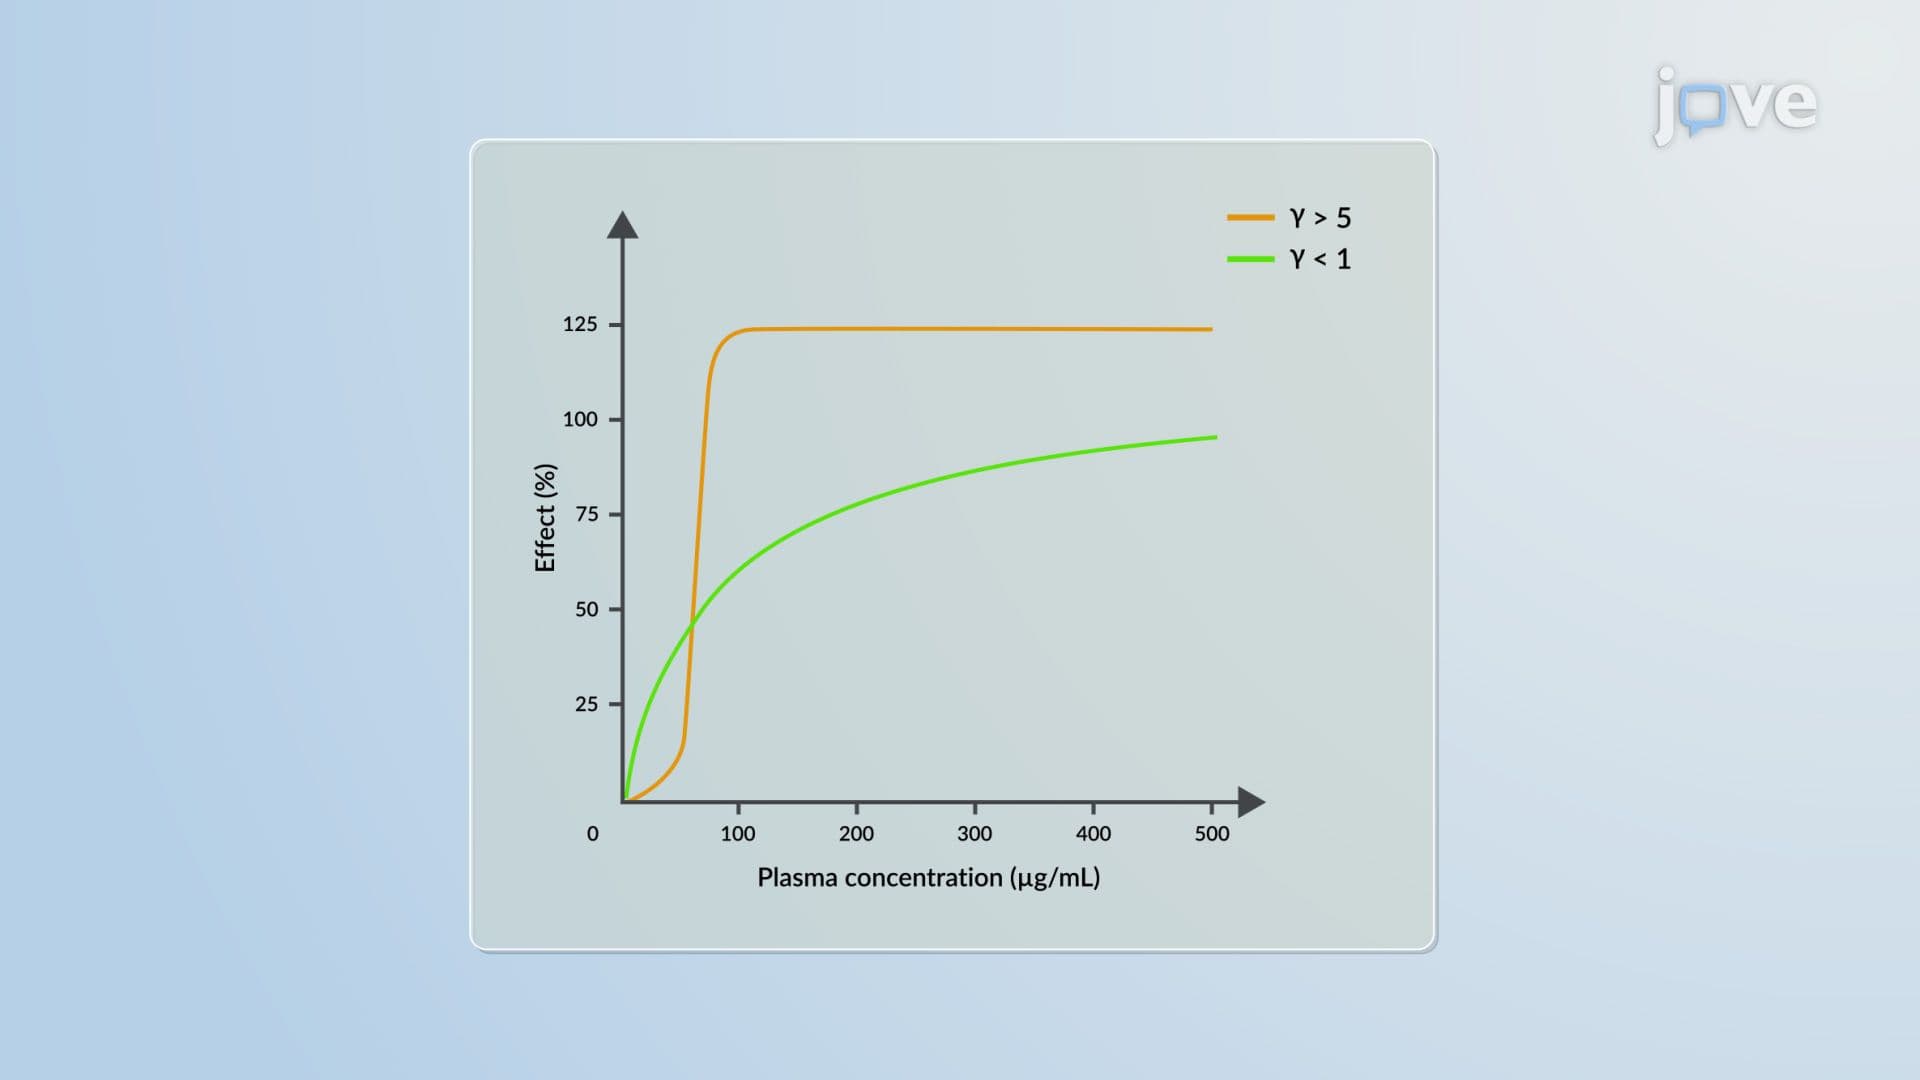
Pharmacodynamic Models: Emax Drug–Concentration Effect Model
 01:15
Impact of Pharmacokinetic–Pharmacodynamic Models: Regulatory Decisions
 01:16
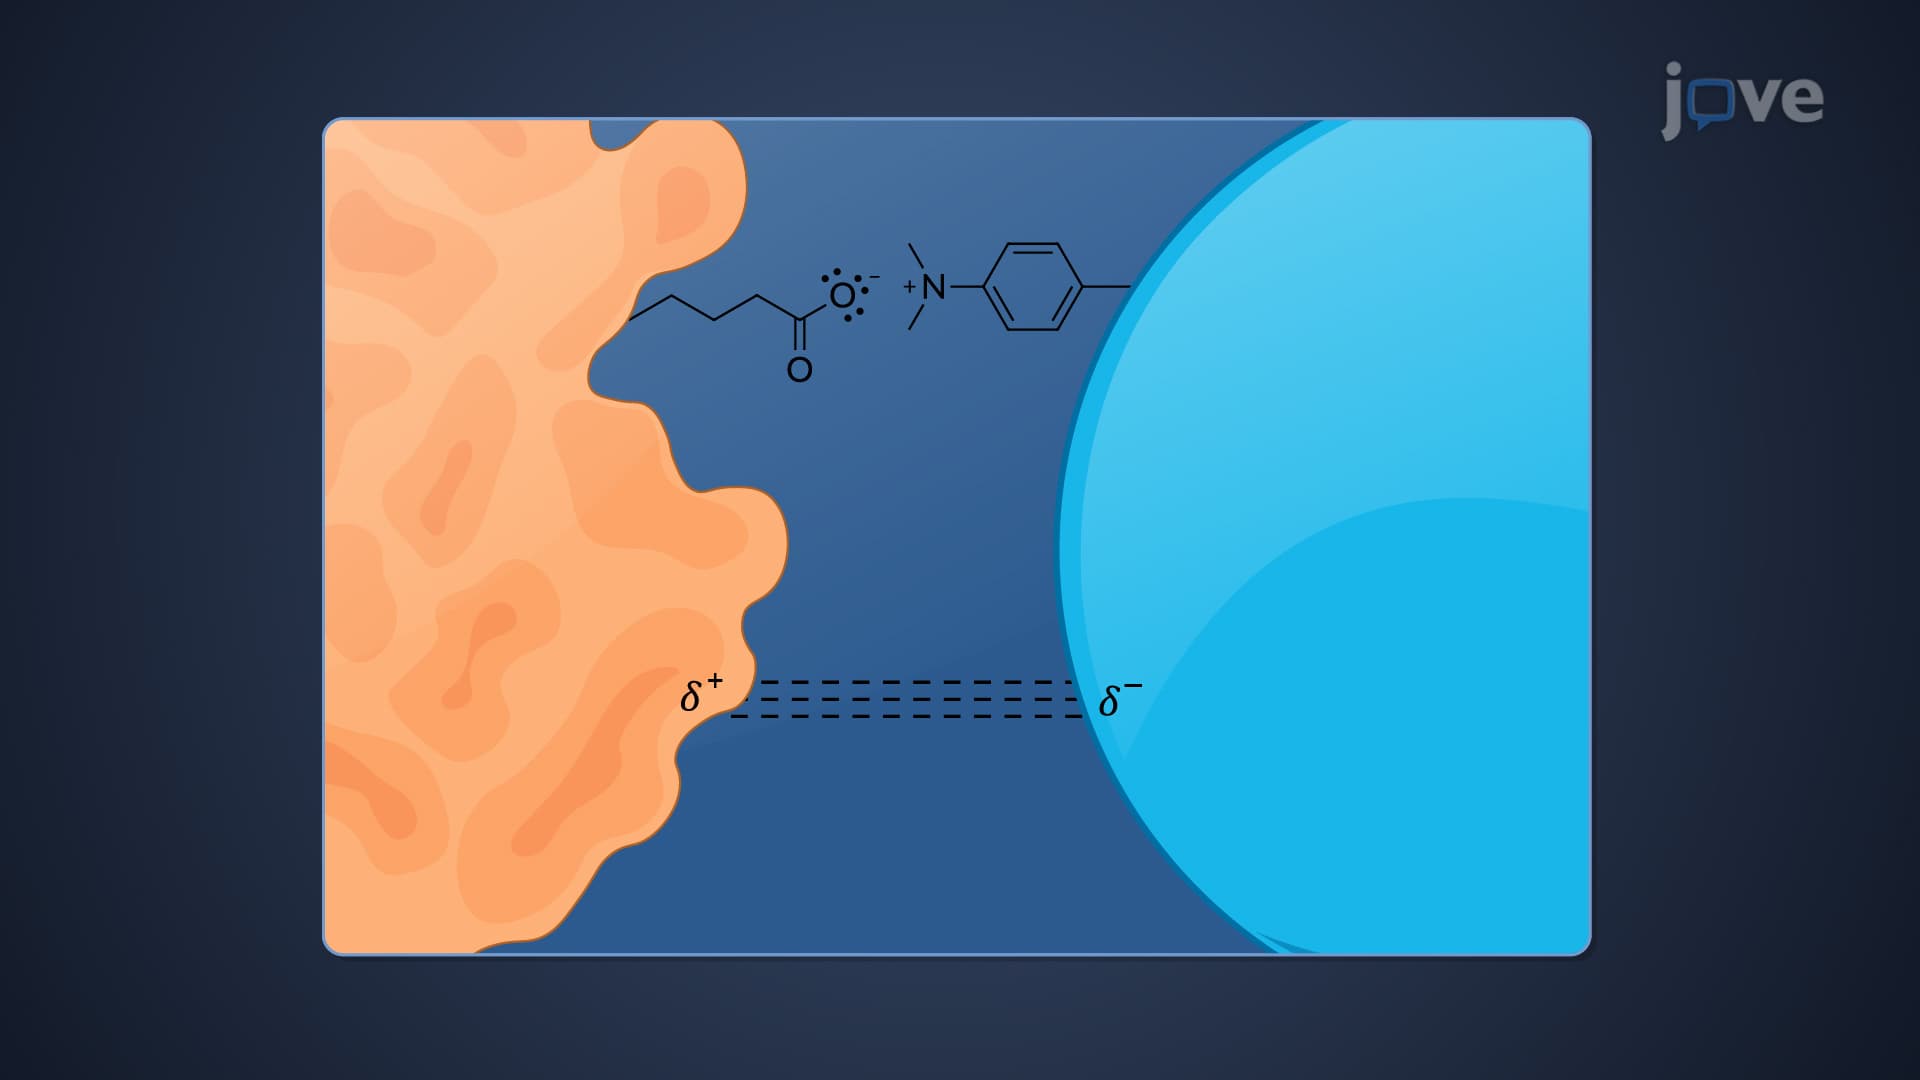
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:09
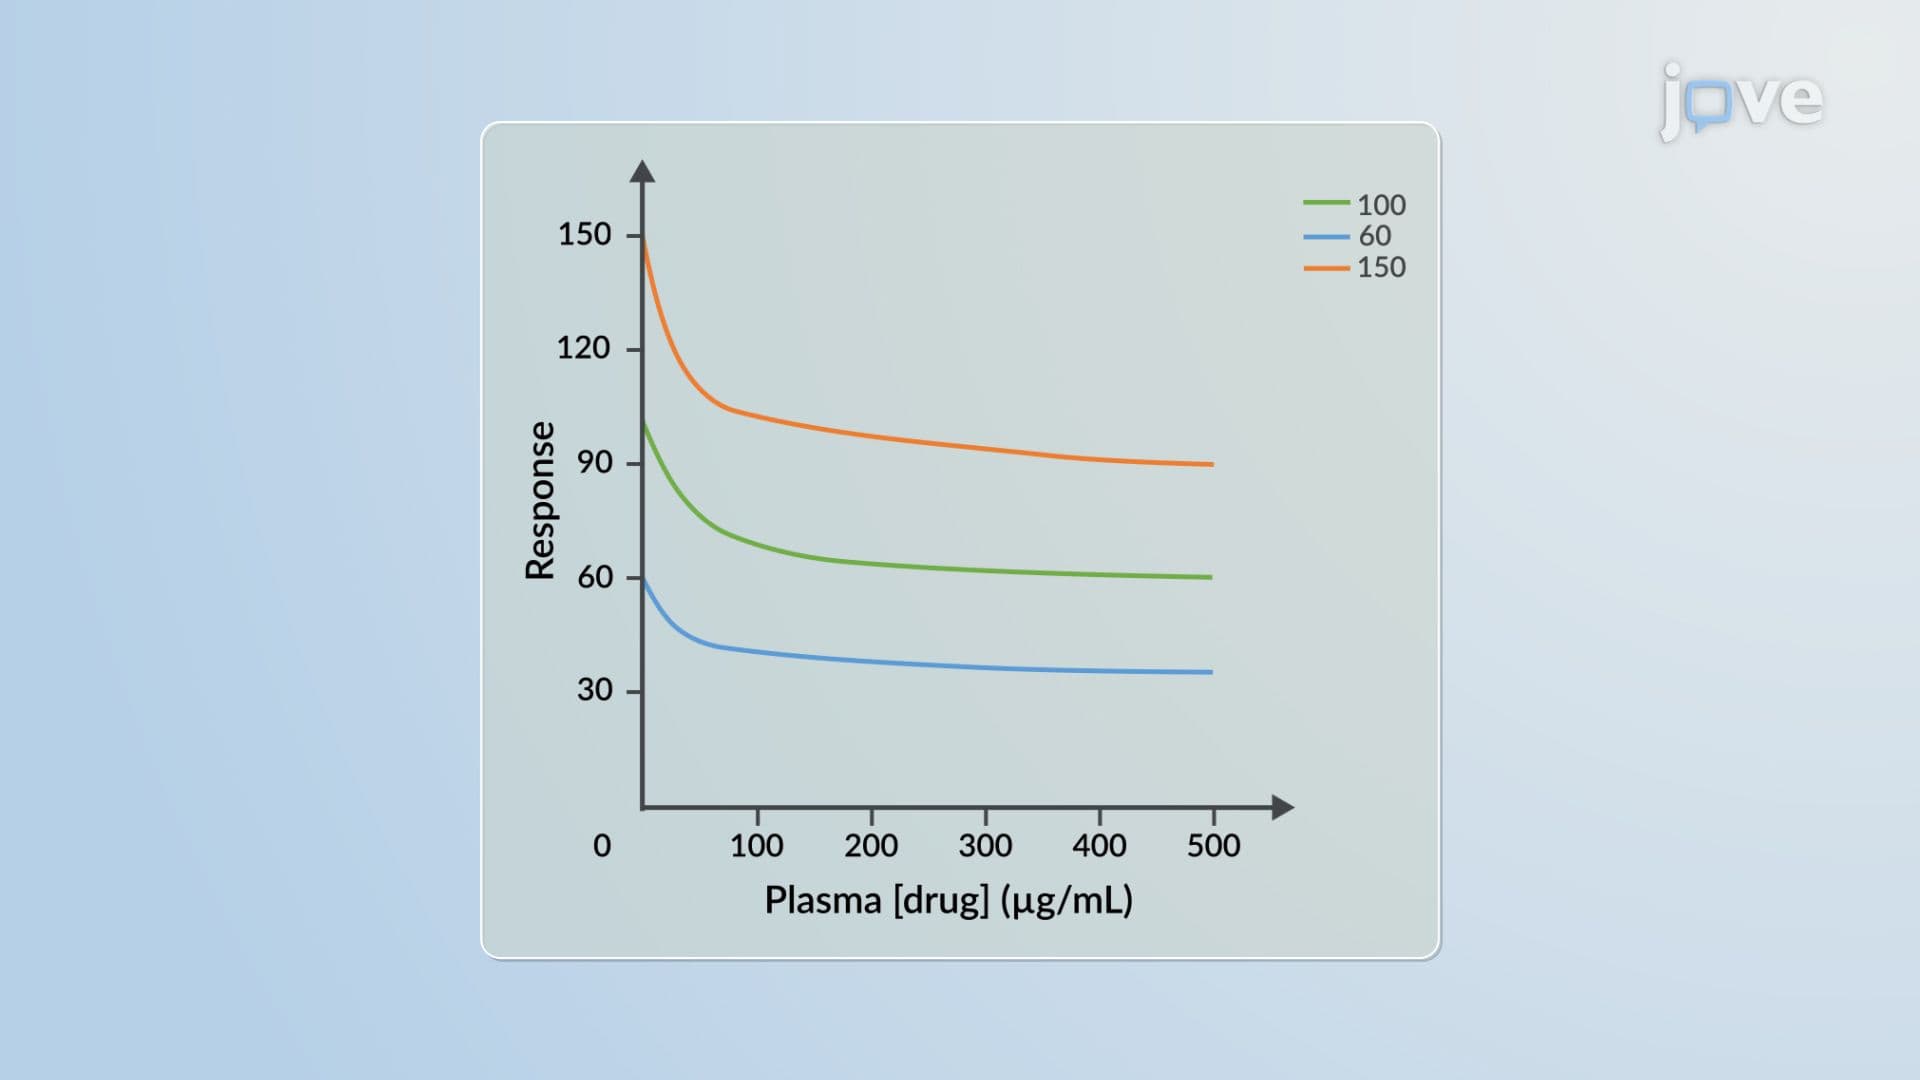
Pharmacodynamic Models: Additive and Proportional Drug Effect Model