Related Experiment Video
Updated: Feb 24, 2026

Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
Accurate gas-phase structure of para-dioxane by fs Raman rotational coherence spectroscopy and ab initio calculations
Takuya Den1, Samuel Menzi1, Hans-Martin Frey1
1Departement für Chemie und Biochemie, Universität Bern, Freiestrasse 3, CH-3000 Bern 9, Switzerland.

Abstract:
p-Dioxane is non-polar, hence its rotational constants cannot be determined by microwave rotational coherence spectroscopy (RCS). We perform high-resolution gas-phase rotational spectroscopy of para-dioxane-h8 and -d8 using femtosecond time-resolved Raman RCS in a gas cell at T = 293 K and in a pulsed supersonic jet at T∼130 K. The inertial tensor of p-dioxane-h8 is strongly asymmetric, leading to a large number of asymmetry transients in its RCS spectrum. In contrast, the d8-isotopomer is a near-oblate symmetric top that exhibits a much more regular RCS spectrum with few asymmetry transients. Fitting the fs Raman RCS transients of p-dioxane-h8 to an asymmetric-top model yields the ground-state rotational constants A0 = 5084.4(5) MHz, B0 = 4684(1) MHz, C0 = 2744.7(8) MHz, and (A0 + B0)/2 = 4884.5(7) MHz (±1σ). The analogous values for p-dioxane-d8 are A0 = 4083(2) MHz, B0 = 3925(4) MHz, C0 = 2347.1(6) MHz, and (A0 + B0)/2 = 4002.4(6) MHz. We determine the molecular structure with a semi-experimental approach involving the highly correlated coupled-cluster singles, doubles and iterated triples method and the cc-pCVXZ basis set series from double- to quadruple-zeta (X = D, T, Q). Combining the calculated vibrationally averaged rotational constants A0calc(X),B0calc(X),C0calc(X) for increasing basis-set size X with non-linear extrapolation to the experimental constants A0exp,B0exp,C0exp allows to determine the equilibrium ground state structure of p-dioxane. For instance, the equilibrium C-C and C-O bond lengths are re(CC) = 1.5135(3) Å and re(CO) = 1.4168(4) Å, and the four axial C-H bond lengths are 0.008 Å longer than the four equatorial ones. The latter is ascribed to the trans-effect (anomeric effect), i.e., the partial delocalization of the electron lone-pairs on the O atoms that are oriented trans, relative to the axial CH bonds.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 09:57
Ultrafast Time-resolved Near-IR Stimulated Raman Measurements of Functional π-conjugate Systems
Published on: February 10, 2020
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:36
Molecular Geometry and Dipole Moments
 01:20
Raman Spectroscopy: Overview
However, a small fraction of the scattered light exhibits a frequency shift due to the exchange of energy between the incident photons and...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 02:18
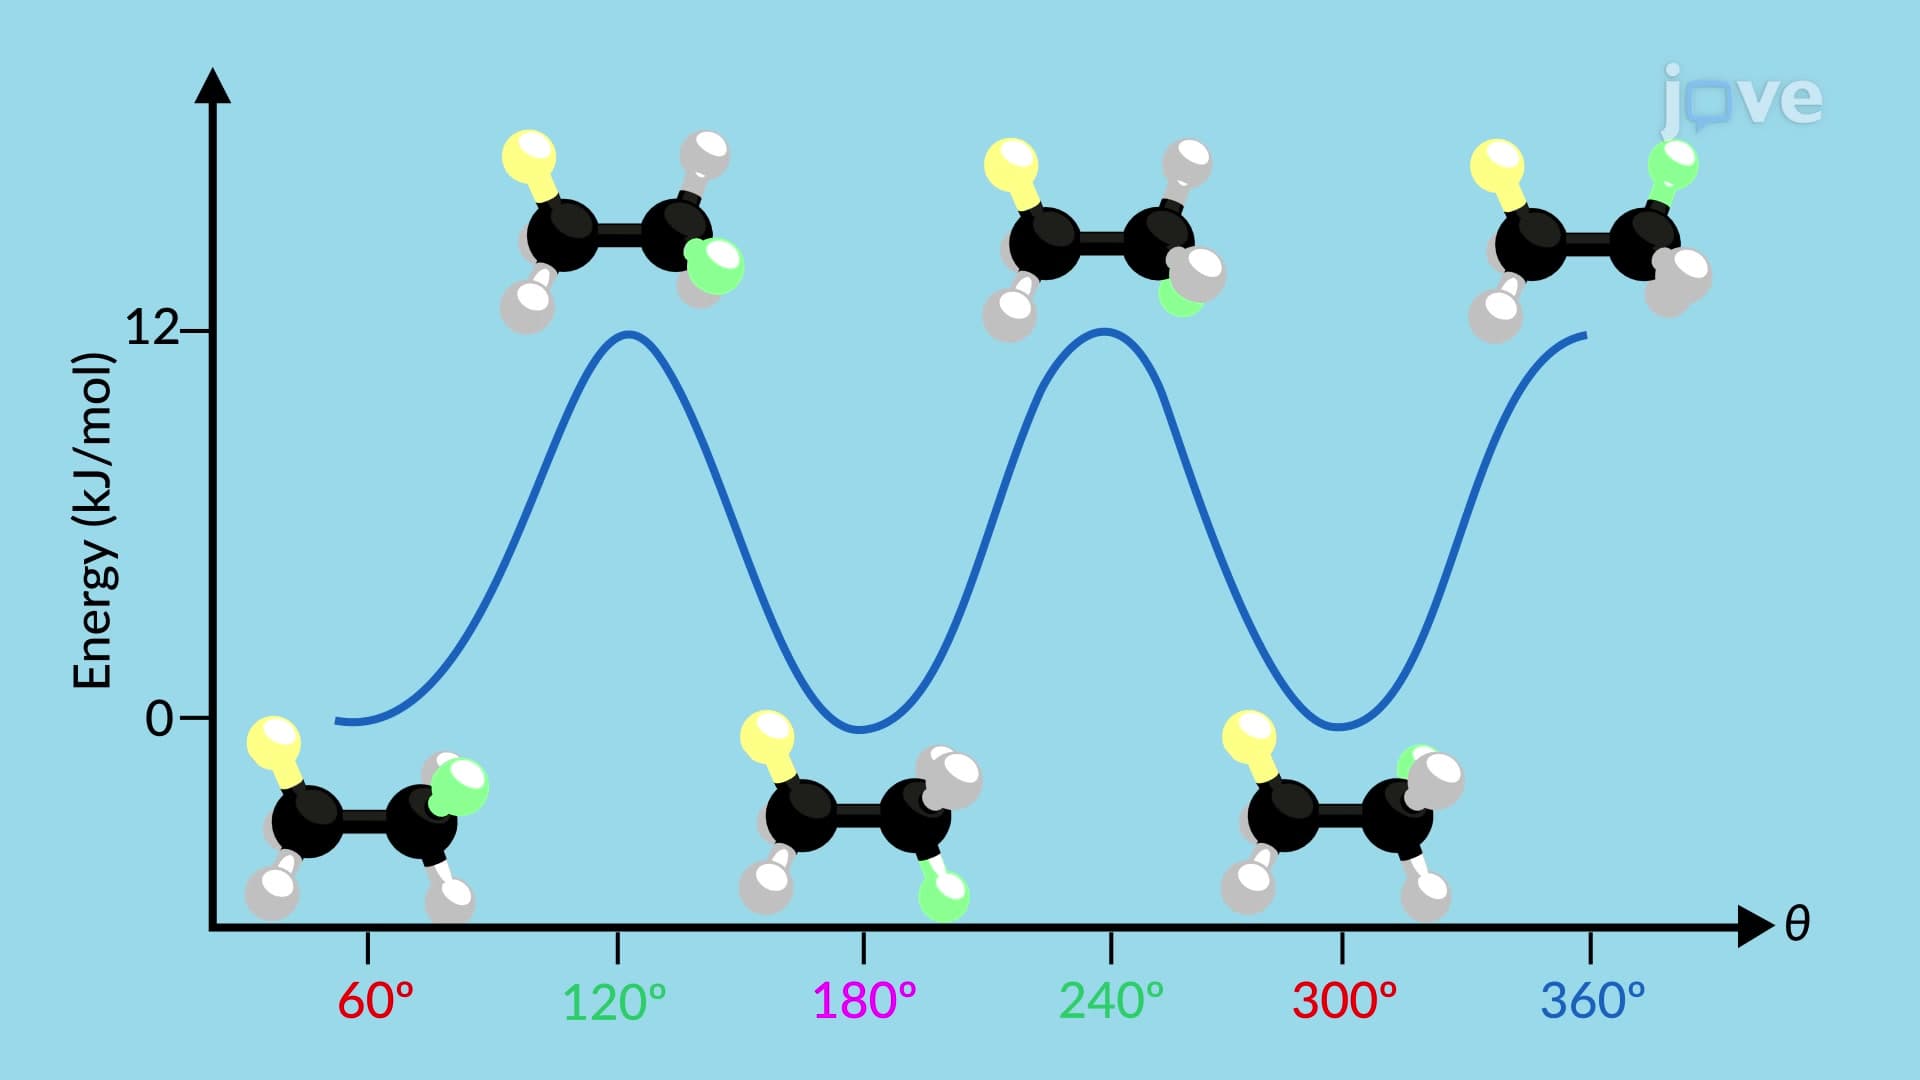
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules