

Level of Theory and Solvent Effects on DASA Absorption Properties Prediction: Comparing TD-DFT, CASPT2 and NEVPT2
Cristina García-Iriepa1, Marco Marazzi2,3
1Laboratoire Modélisation et Simulation Multi Echelle, Université Paris-Est, MSME, UMR 8208 CNRS, UPEM, 5 bd Descartes, 77454 Marne-la-Vallée, France. cristina.garciairiepa@u-pem.fr.

Abstract:
Donor-acceptor Stenhouse adducts (DASAs) are a very recent class of organic photoswitches that combine excellent properties, such as color and polarity change, a large structural modification, and excellent fatigue resistance. Despite their potential applications in different fields, very few studies have focused on rationalizing their electronic structure properties. Here, by means of different state-of-the-art theoretical methods, including solvent and vibrational effects, we show that while time dependent-density functional theory (TD-DFT) can qualitatively describe DASAs' excited states, multiconfigurational quantum chemistry methods along with dynamic electron correlation (CASPT2, NEVPT2) are required for a quantitative agreement with the experiment. This finding is reasoned based on the different charge transfer characteristics observed. Moreover, the TD-DFT computed two-photon absorption properties are reported and suggested to red-shift the absorption band, as required for biological applications.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:29
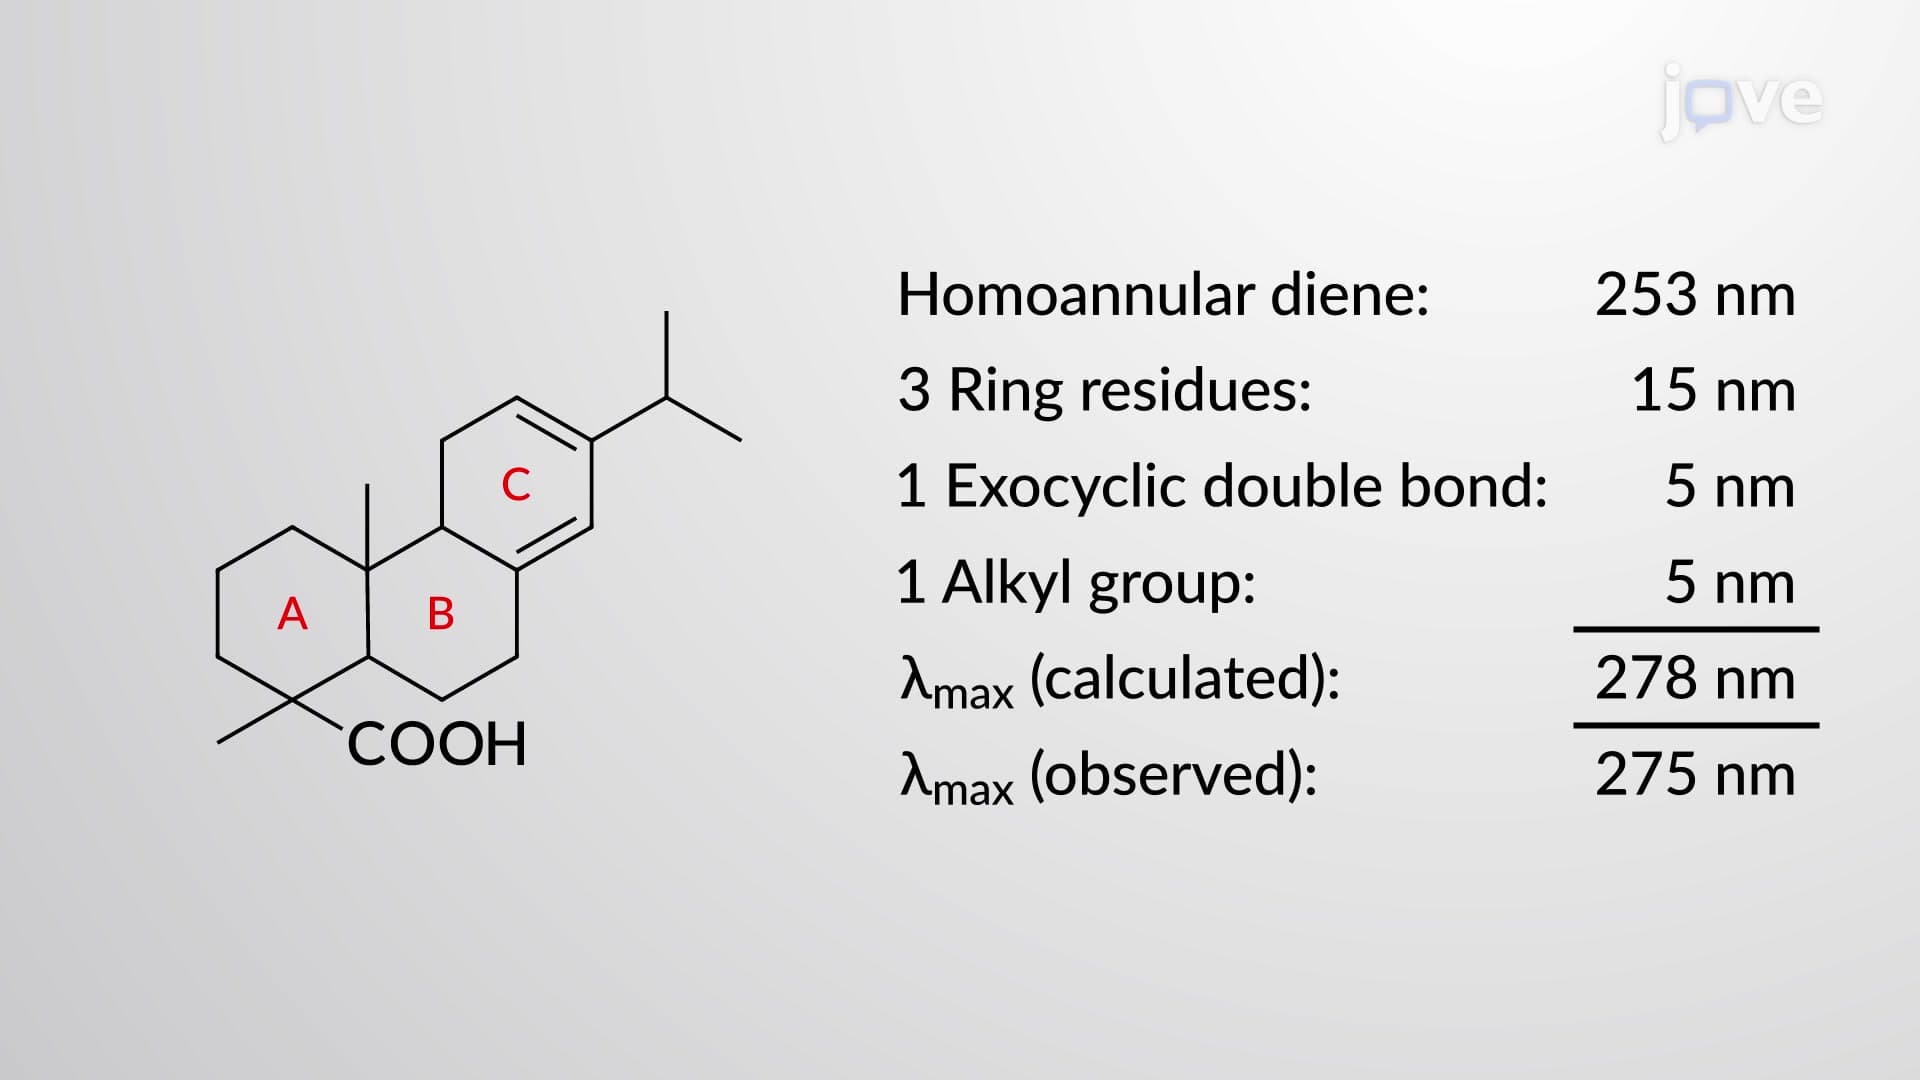
UV–Vis Spectroscopy: Woodward–Fieser Rules
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:25
Atomic Absorption Spectroscopy: Interference
Spectral interference occurs when signals from other elements or molecules overlap with the analyte signal, falsely elevating or masking the analyte's absorbance. This interference can be corrected using Zeeman,...