Related Experiment Video
Updated: Feb 23, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Stochastic GW Calculations for Molecules
Vojtěch Vlček1,2, Eran Rabani3,4, Daniel Neuhauser1
1Department of Chemistry and Biochemistry, University of California , Los Angeles, California 90095, United States.

Abstract:
Quasiparticle (QP) excitations are extremely important for understanding and predicting charge transfer and transport in molecules, nanostructures, and extended systems. Since density functional theory (DFT) within the Kohn-Sham (KS) formulation does not provide reliable QP energies, many-body perturbation techniques such as the GW approximation are essential. The main practical drawback of GW implementations is the high computational scaling with system size, prohibiting its use in extended, open boundary systems with many dozens of electrons or more. Recently, a stochastic formulation of GW (sGW) was presented (Phys. Rev. Lett. 2014, 113, 076402) with a near-linear-scaling complexity, illustrated for a series of silicon nanocrystals reaching systems of more than 3000 electrons. This advance provides a route for many-body calculations on very large systems that were impossible with previous approaches. While earlier we have shown the gentle scaling of sGW, its accuracy was not extensively demonstrated. Therefore, we show that this new sGW approach is very accurate by calculating the ionization energies of a group of sufficiently small molecules where a comparison to other GW codes is still possible. Using a set of 10 such molecules, we demonstrate that sGW provides reliable vertical ionization energies in close agreement with benchmark deterministic GW results (J. Chem. Theory Comput, 2015, 11, 5665), with mean (absolute) deviation of 0.05 and 0.09 eV. For completeness, we also provide a detailed review of the sGW theory and numerical implementation.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Related Concept Videos
 01:10
Propagation of Uncertainty from Systematic Error
 01:08
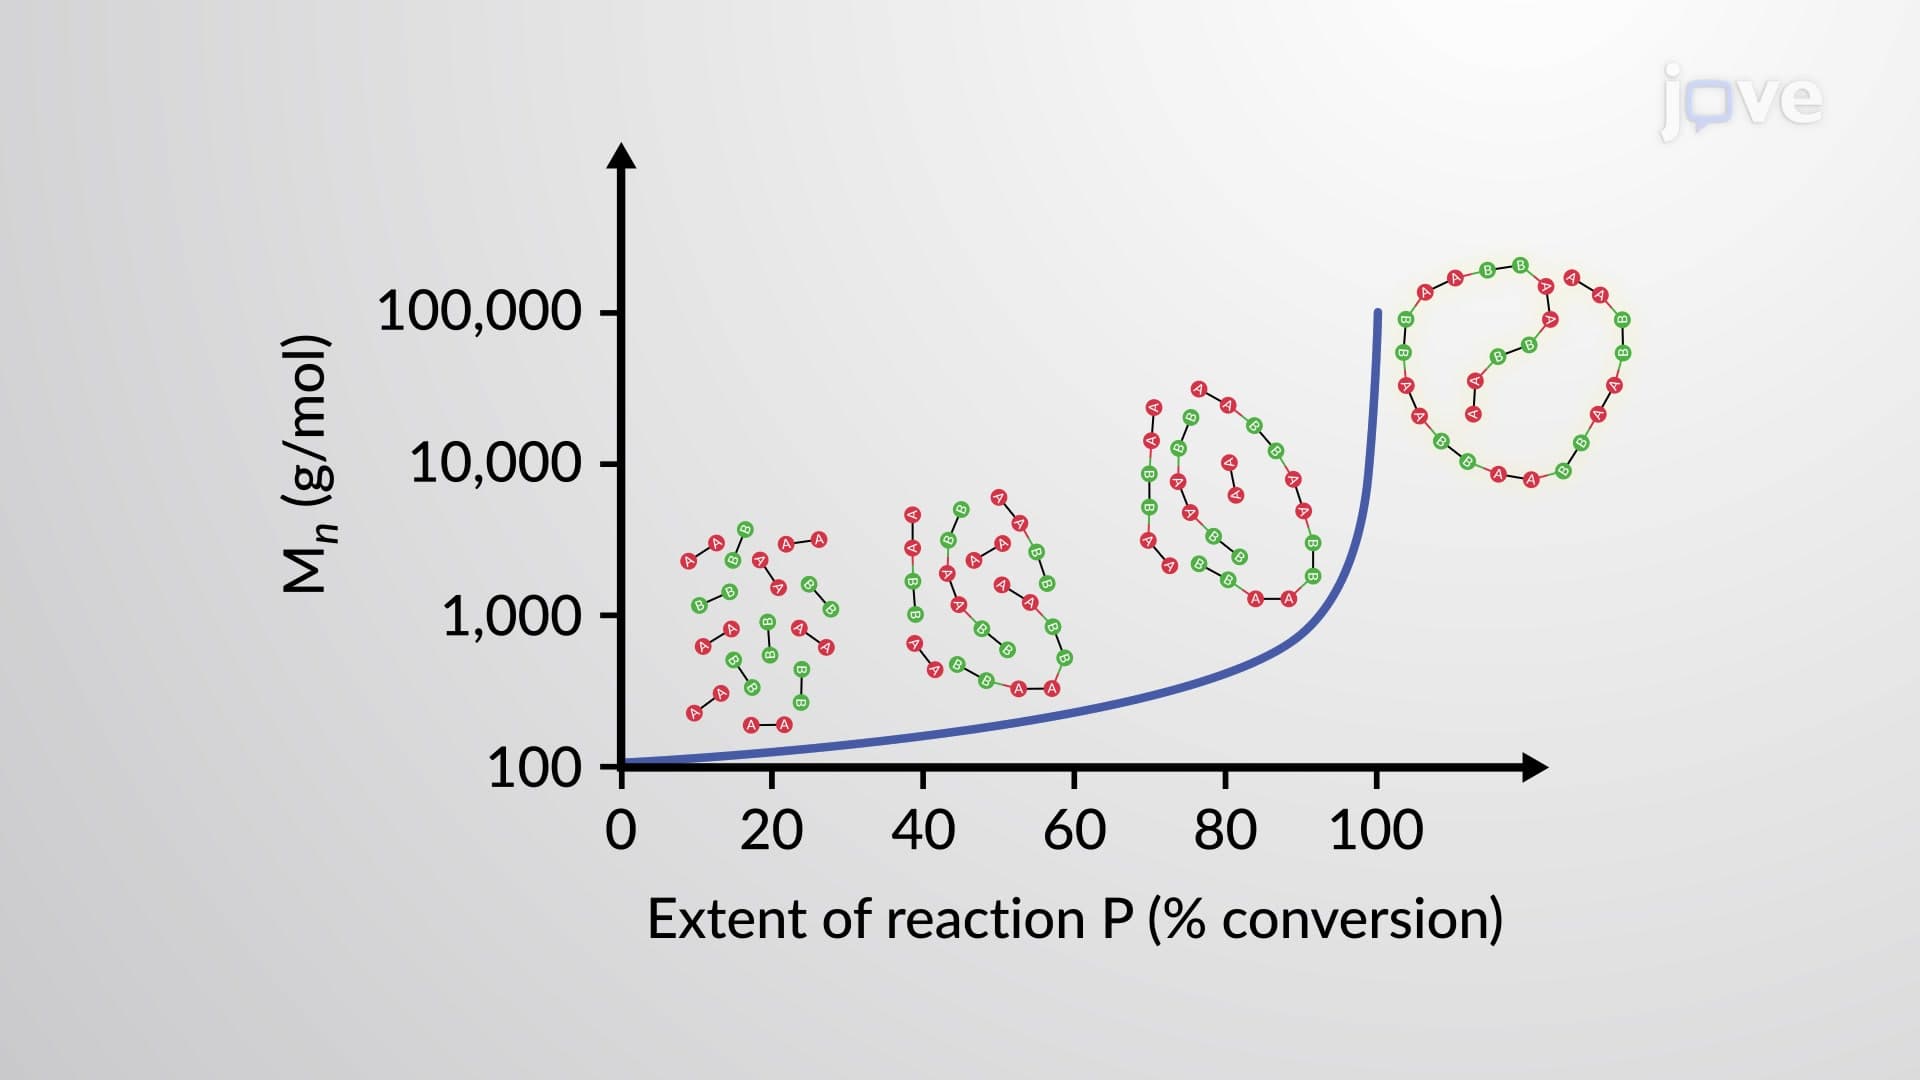
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
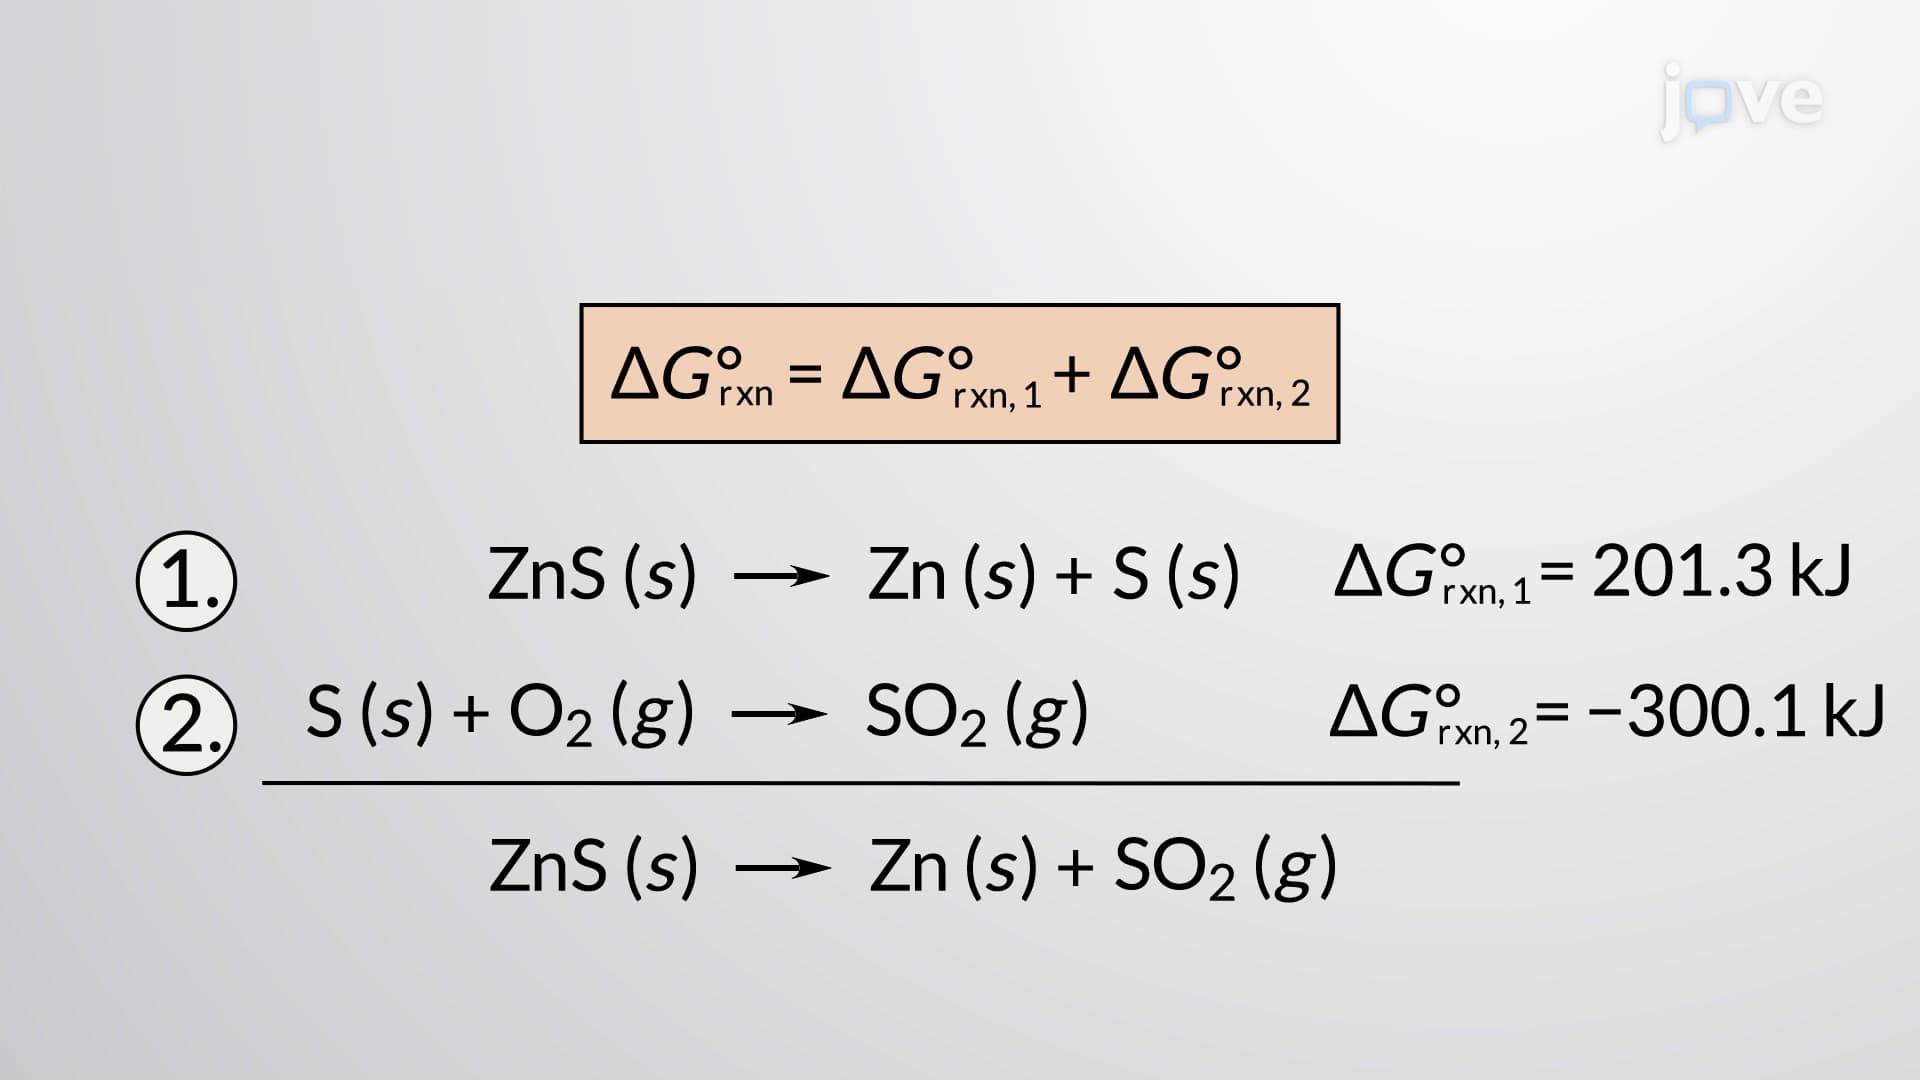 02:49
Calculating Standard Free Energy Changes
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:27
Distribution of Molecular Speeds
 01:20
Maxwell-Boltzmann Distribution: Problem Solving
This distribution function f(v) is defined by saying that the expected number N (v1,v2) of particles with speeds between v1 and v2 is given by