Related Experiment Video
Updated: Feb 23, 2026
Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
Theoretical Kinetics Analysis for Ḣ Atom Addition to 1,3-Butadiene and Related Reactions on the Ċ4H7 Potential Energy
Yang Li1, Stephen J Klippenstein2, Chong-Wen Zhou3
1Combustion Chemistry Centre, National University of Ireland , Galway, Ireland.

Abstract:
The oxidation chemistry of the simplest conjugated hydrocarbon, 1,3-butadiene, can provide a first step in understanding the role of polyunsaturated hydrocarbons in combustion and, in particular, an understanding of their contribution toward soot formation. On the basis of our previous work on propene and the butene isomers (1-, 2-, and isobutene), it was found that the reaction kinetics of Ḣ-atom addition to the C═C double bond plays a significant role in fuel consumption kinetics and influences the predictions of high-temperature ignition delay times, product species concentrations, and flame speed measurements. In this study, the rate constants and thermodynamic properties for Ḣ-atom addition to 1,3-butadiene and related reactions on the Ċ4H7 potential energy surface have been calculated using two different series of quantum chemical methods and two different kinetic codes. Excellent agreement is obtained between the two different kinetics codes. The calculated results including zero-point energies, single-point energies, rate constants, barrier heights, and thermochemistry are systematically compared among the two quantum chemical methods. 1-Methylallyl (Ċ4H71-3) and 3-buten-1-yl (Ċ4H71-4) radicals and C2H4 + Ċ2H3 are found to be the most important channels and reactivity-promoting products, respectively. We calculated that terminal addition is dominant (>80%) compared to internal Ḣ-atom addition at all temperatures in the range 298-2000 K. However, this dominance decreases with increasing temperature. The calculated rate constants for the bimolecular reaction C4H6 + Ḣ → products and C2H4 + Ċ2H3 → products are in excellent agreement with both experimental and theoretical results from the literature. For selected C4 species, the calculated thermochemical values are also in good agreement with literature data. In addition, the rate constants for H atom abstraction by Ḣ atoms have also been calculated, and it is found that abstraction from the central carbon atoms is the dominant channel (>70%) at temperatures in the range of 298-2000 K. Finally, by incorporating our calculated rate constants for both Ḣ atom addition and abstraction into our recently developed 1,3-butadiene model, we show that laminar flame speed predictions are significantly improved, emphasizing the value of this study.
More Related Videos
 07:50
Efficient Synthesis of All-Carbon Quaternary Centers via the Conjugate Addition of Functionalized Monoorganozinc Bromides
Published on: May 26, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:23
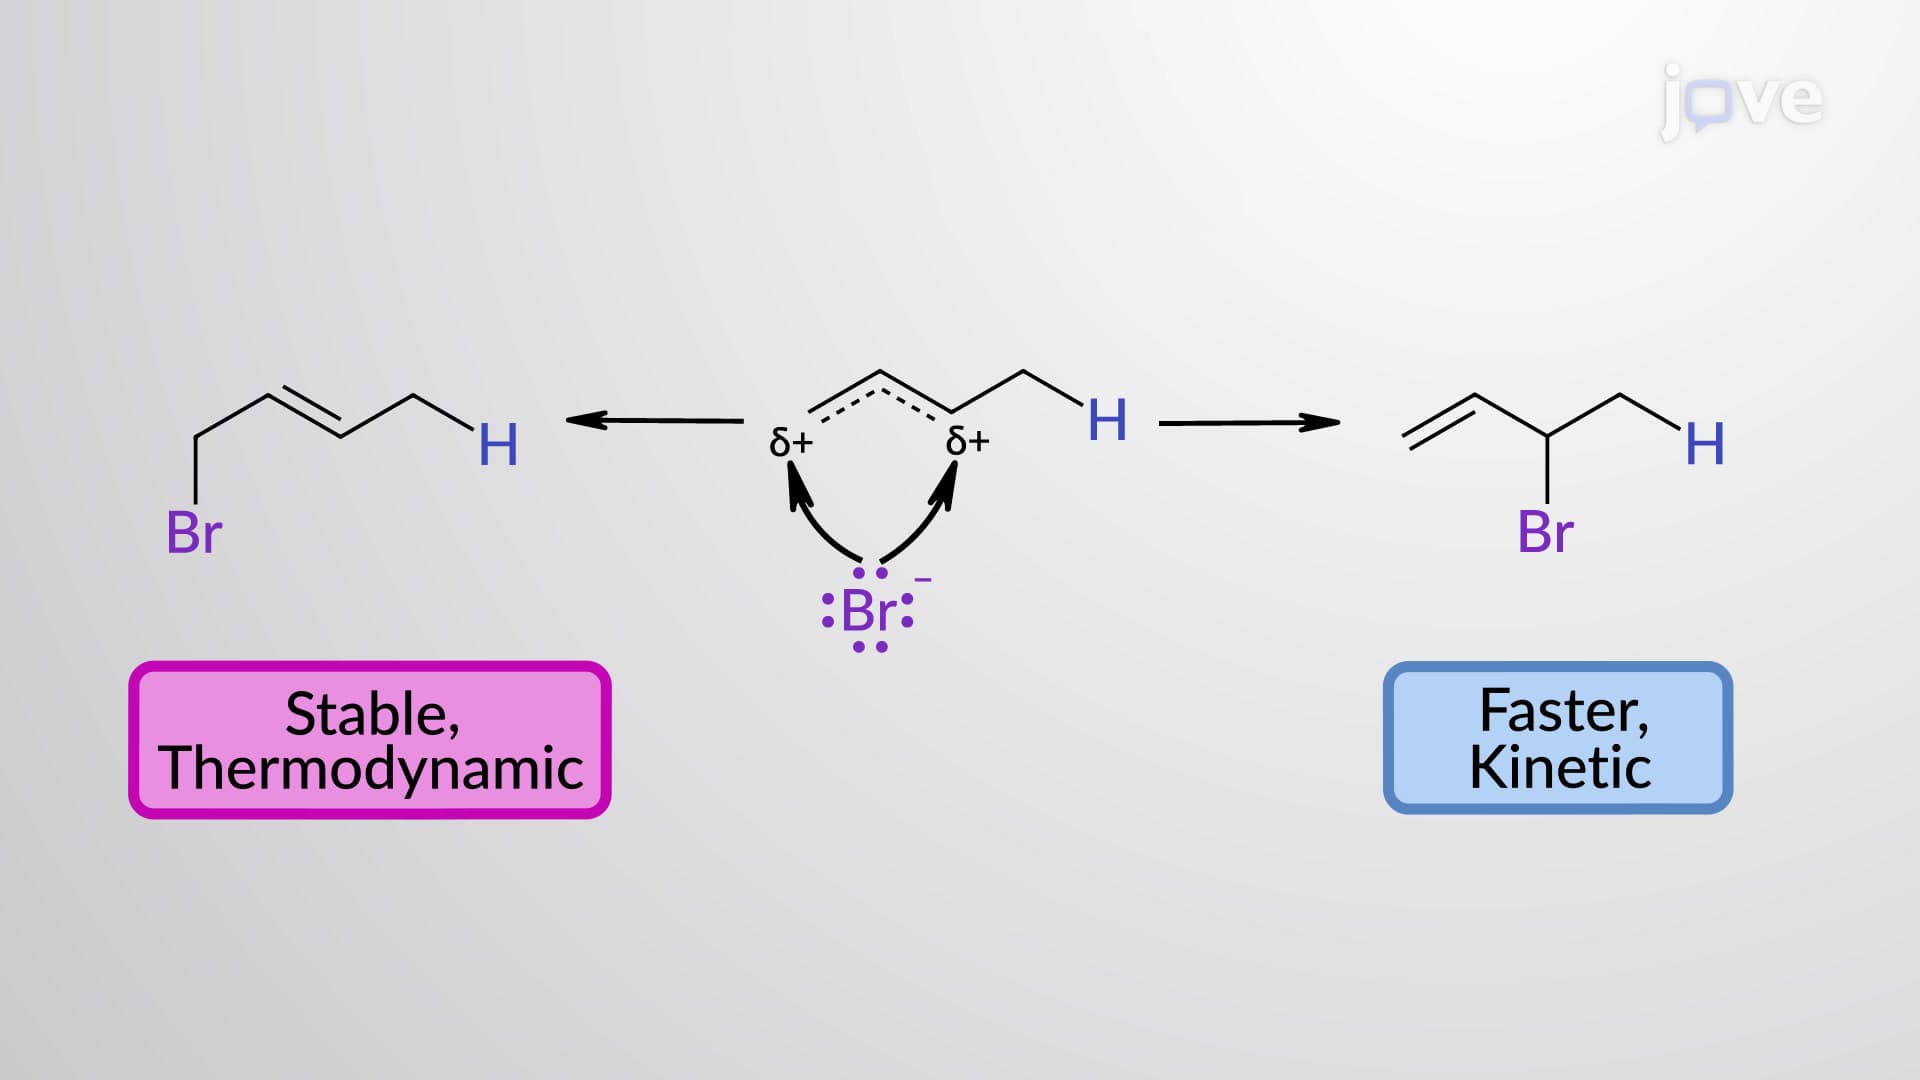
Electrophilic Addition of HX to 1,3-Butadiene: Thermodynamic vs Kinetic Control
 01:17
Electrophilic 1,2- and 1,4-Addition of HX to 1,3-Butadiene
 01:14
Electrophilic 1,2- and 1,4-Addition of X2 to 1,3-Butadiene
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 01:32
Radical Anti-Markovnikov Addition to Alkenes: Thermodynamics