In silico screening of drug-membrane thermodynamics reveals linear relations between bulk partitioning and the
Roberto Menichetti1, Kiran H Kanekal1, Kurt Kremer1
1Max Planck Institute for Polymer Research, Ackermannweg 10, 55128 Mainz, Germany.

Abstract:
The partitioning of small molecules in cell membranes-a key parameter for pharmaceutical applications-typically relies on experimentally available bulk partitioning coefficients. Computer simulations provide a structural resolution of the insertion thermodynamics via the potential of mean force but require significant sampling at the atomistic level. Here, we introduce high-throughput coarse-grained molecular dynamics simulations to screen thermodynamic properties. This application of physics-based models in a large-scale study of small molecules establishes linear relationships between partitioning coefficients and key features of the potential of mean force. This allows us to predict the structure of the insertion from bulk experimental measurements for more than 400 000 compounds. The potential of mean force hereby becomes an easily accessible quantity-already recognized for its high predictability of certain properties, e.g., passive permeation. Further, we demonstrate how coarse graining helps reduce the size of chemical space, enabling a hierarchical approach to screening small molecules.
Related Concept Videos
 01:23
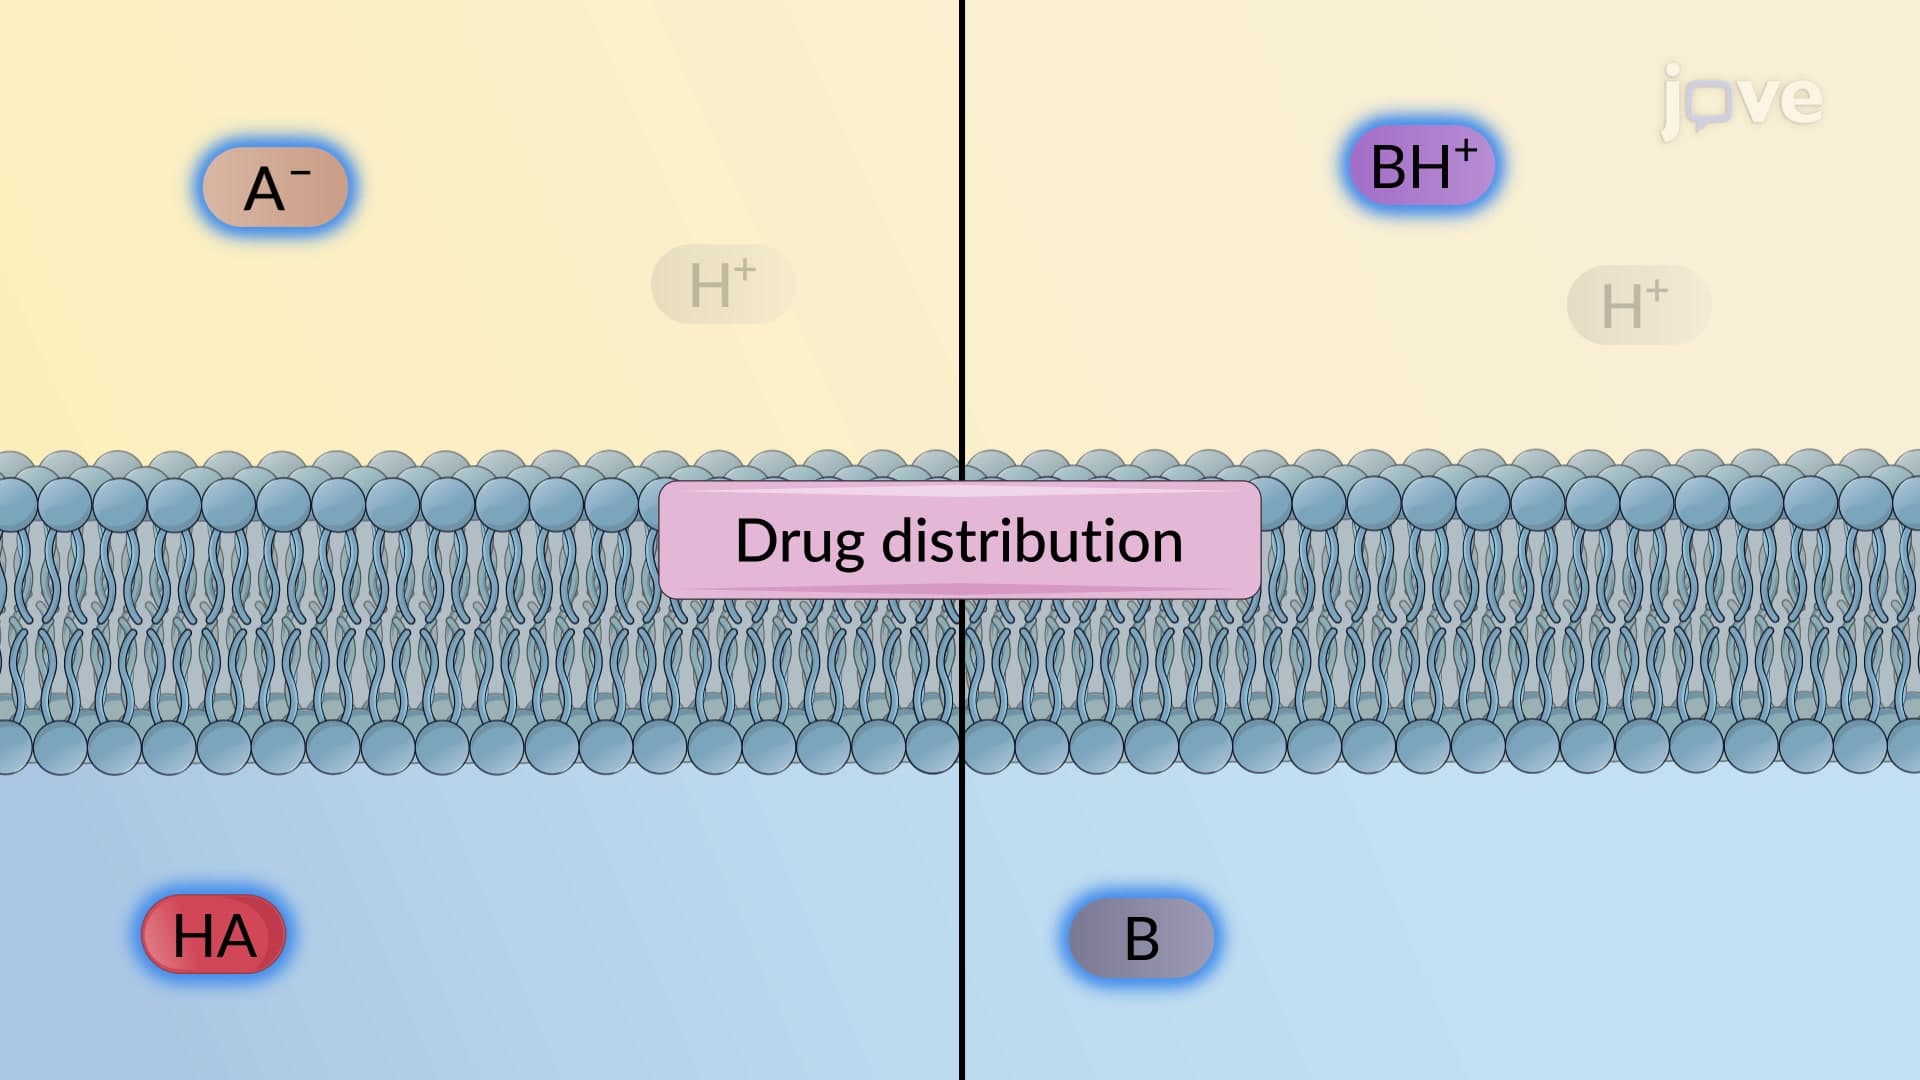
Drug Absorption Mechanism: Passive Membrane Transport
 01:21
Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH
A drug's pKa and the pH of the gastrointestinal (GI) tract play crucial roles...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:14
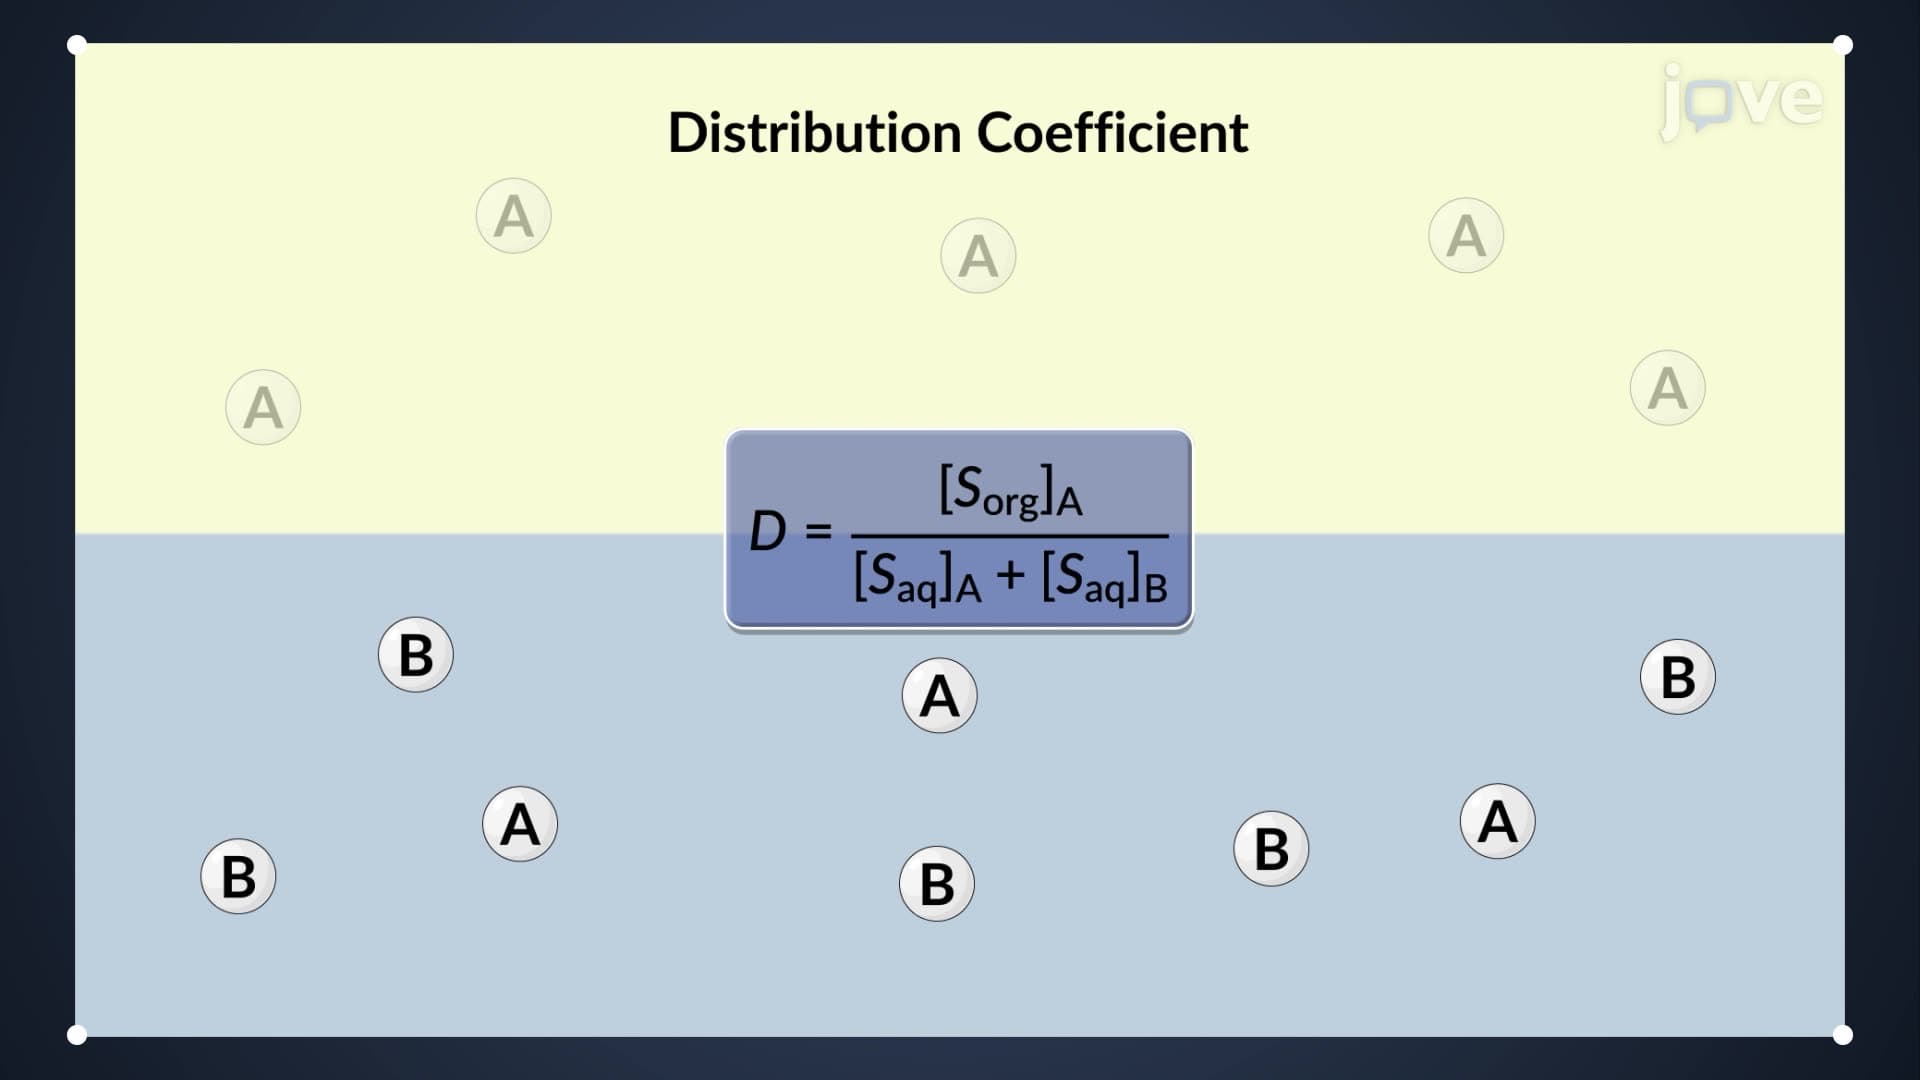
Extraction: Partition and Distribution Coefficients
For extracting a solute from an aqueous phase into an...
 02:34
Comparing Intermolecular Forces: Melting Point, Boiling Point, and Miscibility
Temporary attractive forces like dispersion are present in all molecules, whether they are polar or nonpolar. They...