Related Experiment Video
Updated: Feb 21, 2026

Methods for Evaluating the Role of c-Fos and Dusp1 in Oncogene Dependence
Published on: January 7, 2019
Inhibition of USP10 induces degradation of oncogenic FLT3
Ellen L Weisberg1, Nathan J Schauer2, Jing Yang2
1Department of Medical Oncology, Dana-Farber Cancer Institute, Harvard Medical School, Boston, Massachusetts, USA.

Abstract:
Oncogenic forms of the kinase FLT3 are important therapeutic targets in acute myeloid leukemia (AML); however, clinical responses to small-molecule kinase inhibitors are short-lived as a result of the rapid emergence of resistance due to point mutations or compensatory increases in FLT3 expression. We sought to develop a complementary pharmacological approach whereby proteasome-mediated FLT3 degradation could be promoted by inhibitors of the deubiquitinating enzymes (DUBs) responsible for cleaving ubiquitin from FLT3. Because the relevant DUBs for FLT3 are not known, we assembled a focused library of most reported small-molecule DUB inhibitors and carried out a cellular phenotypic screen to identify compounds that could induce the degradation of oncogenic FLT3. Subsequent target deconvolution efforts allowed us to identify USP10 as the critical DUB required to stabilize FLT3. Targeting of USP10 showed efficacy in preclinical models of mutant-FLT3 AML, including cell lines, primary patient specimens and mouse models of oncogenic-FLT3-driven leukemia.
Insights
Targeting deubiquitinating enzymes (DUBs) with small molecules can promote proteasome-mediated degradation of FLT3. Inhibiting USP10 effectively degraded FLT3 and showed efficacy in preclinical models of acute myeloid leukemia (AML).
Area of Science:
- Biochemistry
- Molecular Biology
- Oncology
Background:
- Oncogenic FLT3 mutations drive acute myeloid leukemia (AML) but are subject to rapid drug resistance.
- Resistance mechanisms include FLT3 point mutations and increased FLT3 expression, limiting kinase inhibitor efficacy.
- A complementary approach is needed to overcome resistance and improve therapeutic outcomes in FLT3-mutant AML.
Purpose of the Study:
- To identify deubiquitinating enzymes (DUBs) that stabilize FLT3.
- To develop a pharmacological strategy to induce proteasome-mediated degradation of oncogenic FLT3.
- To evaluate the therapeutic potential of targeting FLT3-stabilizing DUBs in preclinical AML models.
Main Methods:
- Assembled a library of small-molecule DUB inhibitors.
- Performed a cellular phenotypic screen to identify compounds inducing FLT3 degradation.
- Conducted target deconvolution to identify the specific DUB responsible for FLT3 stabilization.
- Tested the efficacy of USP10 inhibition in various preclinical models of FLT3-mutant AML.
Main Results:
- Identified USP10 as the critical DUB required for FLT3 stabilization.
- Demonstrated that USP10 inhibition leads to proteasome-mediated degradation of FLT3.
- Showed that targeting USP10 has significant efficacy in cell lines, primary patient specimens, and mouse models of FLT3-mutant AML.
Conclusions:
- USP10 is a key regulator of FLT3 stability.
- Inhibiting USP10 represents a promising therapeutic strategy for overcoming resistance in FLT3-mutant AML.
- Targeting DUBs offers a complementary approach to kinase inhibitors for treating AML.
More Related Videos
Related Concept Videos
 02:23
Abnormal Proliferation
 01:31
The Intrinsic Apoptotic Pathway
 01:12
Loss of Tumor Suppressor Gene Functions
When the tumor suppressor genes develop mutations or are lost, cells start growing out of control, leading to cancer. However, a single functional copy of the tumor suppressor gene is enough for the cells to maintain their normal functions and cell...
 01:20
The JAK-STAT Signaling Pathway
 01:22
PI3K/mTOR/AKT Signaling Pathway
 02:58
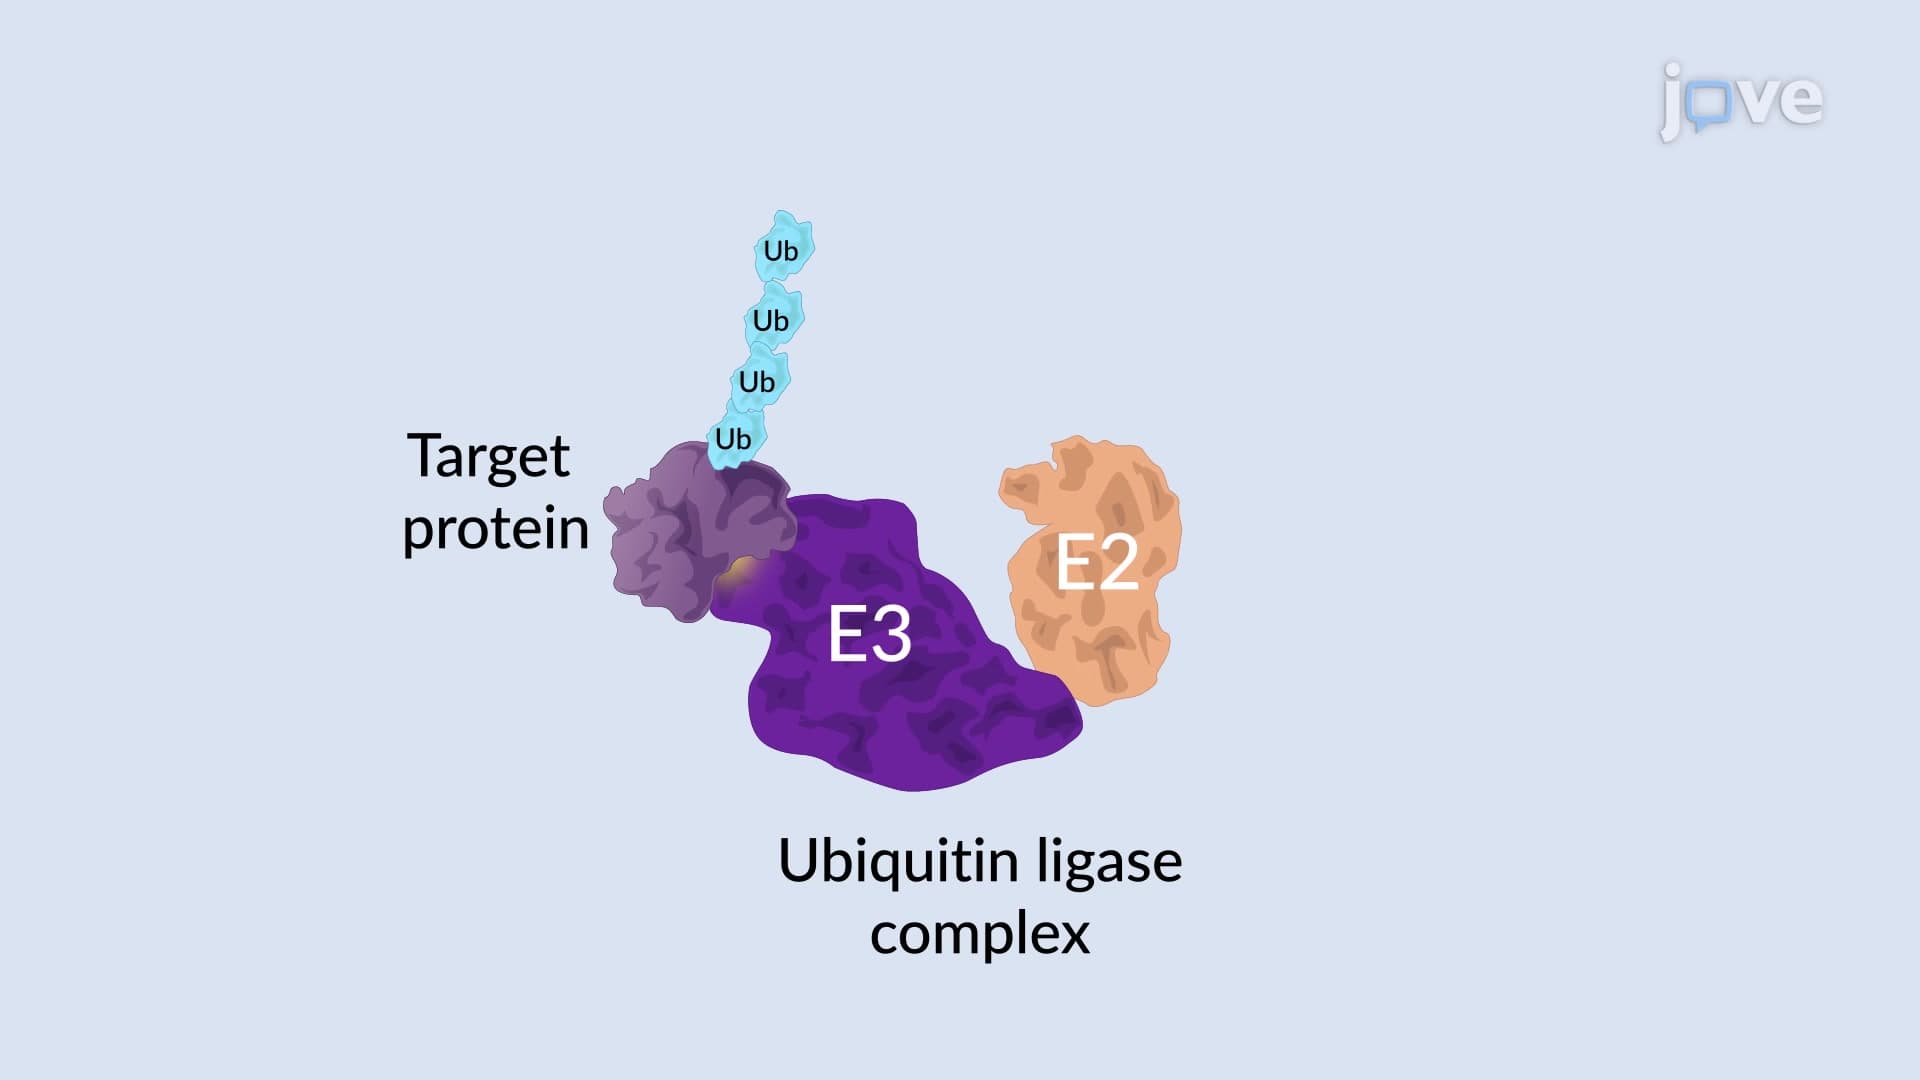
Regulated Protein Degradation
Protein degradation plays two important roles in the cells. It helps to protect cells from misfolded or damaged proteins before they lead to a...