Assessing Density Functionals Using Many Body Theory for Hybrid Perovskites
Related Experiment Video
Updated: Feb 20, 2026
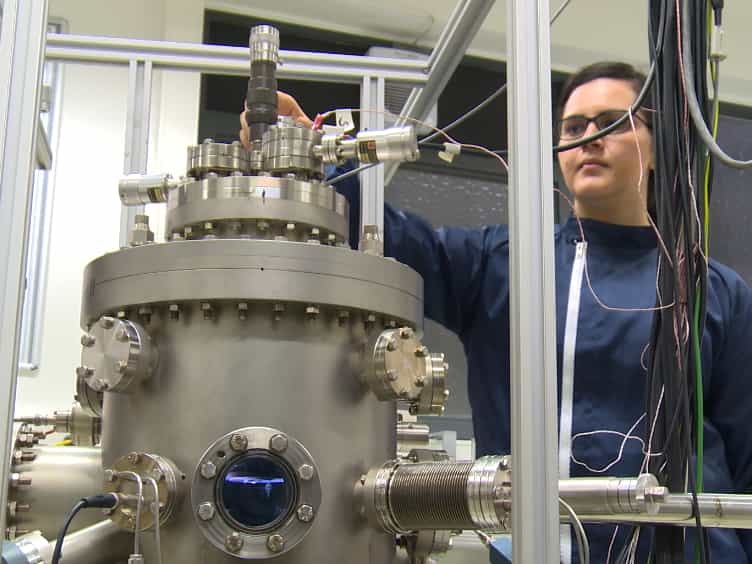
Influence of Hybrid Perovskite Fabrication Methods on Film Formation, Electronic Structure, and Solar Cell Performance
Published on: February 27, 2017
Menno Bokdam1, Jonathan Lahnsteiner1, Benjamin Ramberger1
1University of Vienna, Faculty of Physics and Center for Computational Materials Sciences, Sensengasse 8/12, A-1090 Vienna, Austria.

Abstract:
Which density functional is the "best" for structure simulations of a particular material? A concise, first principles, approach to answer this question is presented. The random phase approximation (RPA)-an accurate many body theory-is used to evaluate various density functionals. To demonstrate and verify the method, we apply it to the hybrid perovskite MAPbI_{3}, a promising new solar cell material. The evaluation is done by first creating finite temperature ensembles for small supercells using RPA molecular dynamics, and then evaluating the variance between the RPA and various approximate density functionals for these ensembles. We find that, contrary to recent suggestions, van der Waals functionals do not improve the description of the material, whereas hybrid functionals and the strongly constrained appropriately normed (SCAN) density functional yield very good agreement with the RPA. Finally, our study shows that in the room temperature tetragonal phase of MAPbI_{3}, the molecules are preferentially parallel to the shorter lattice vectors but reorientation on ps time scales is still possible.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:24
Hybridization of Atomic Orbitals I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:35
Hybridization of Atomic Orbitals II