Backbone flexibility of extended metal atom chains. Ab initio molecular dynamics for Cr3(dpa)4X2 (X = NCS, CN, NO3) in gas and crystalline phases
M Spivak1, V Arcisauskaite, X López
1Departament de Quimica Fisica i Inorgànica, Universitat Rovira i Virgili, Marcel·lí Domingo 1, 43007 Tarragona, Spain. coen.degraaf@urv.cat.

Abstract:
Recently published static DFT and CASSCF/CASPT2 calculations depicted extremely flat Potential Energy Surfaces (PESs) for the Cr-Cr flexibility of Cr3(dpa)4X2 (X = NCS-, CN-, NO3-) extended metal atom chains (EMACs) (M. Spivak, et al., Dalton Trans., 2017, 46, 6202). We herein explore the thermal and crystal packing effects on the structure of EMACs using ab initio molecular dynamics (MD). Car-Parrinello DFT-based simulations of the isolated molecules show that thermal energy favors asymmetric arrangements of the Cr3 chain due, in part, to the bending of the axial ligands (X) and the increased X-Cr distance, both of which weaken X → Cr σ-donation. This effect is even more prominent in the crystalline phase due to the interaction between the axial ligands of neighboring molecules in the unit cell. This could explain the typical discrepancies between the experimental and theoretical characterization of Cr3 EMACs observed in the literature.
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
 02:18
X-ray Crystallography
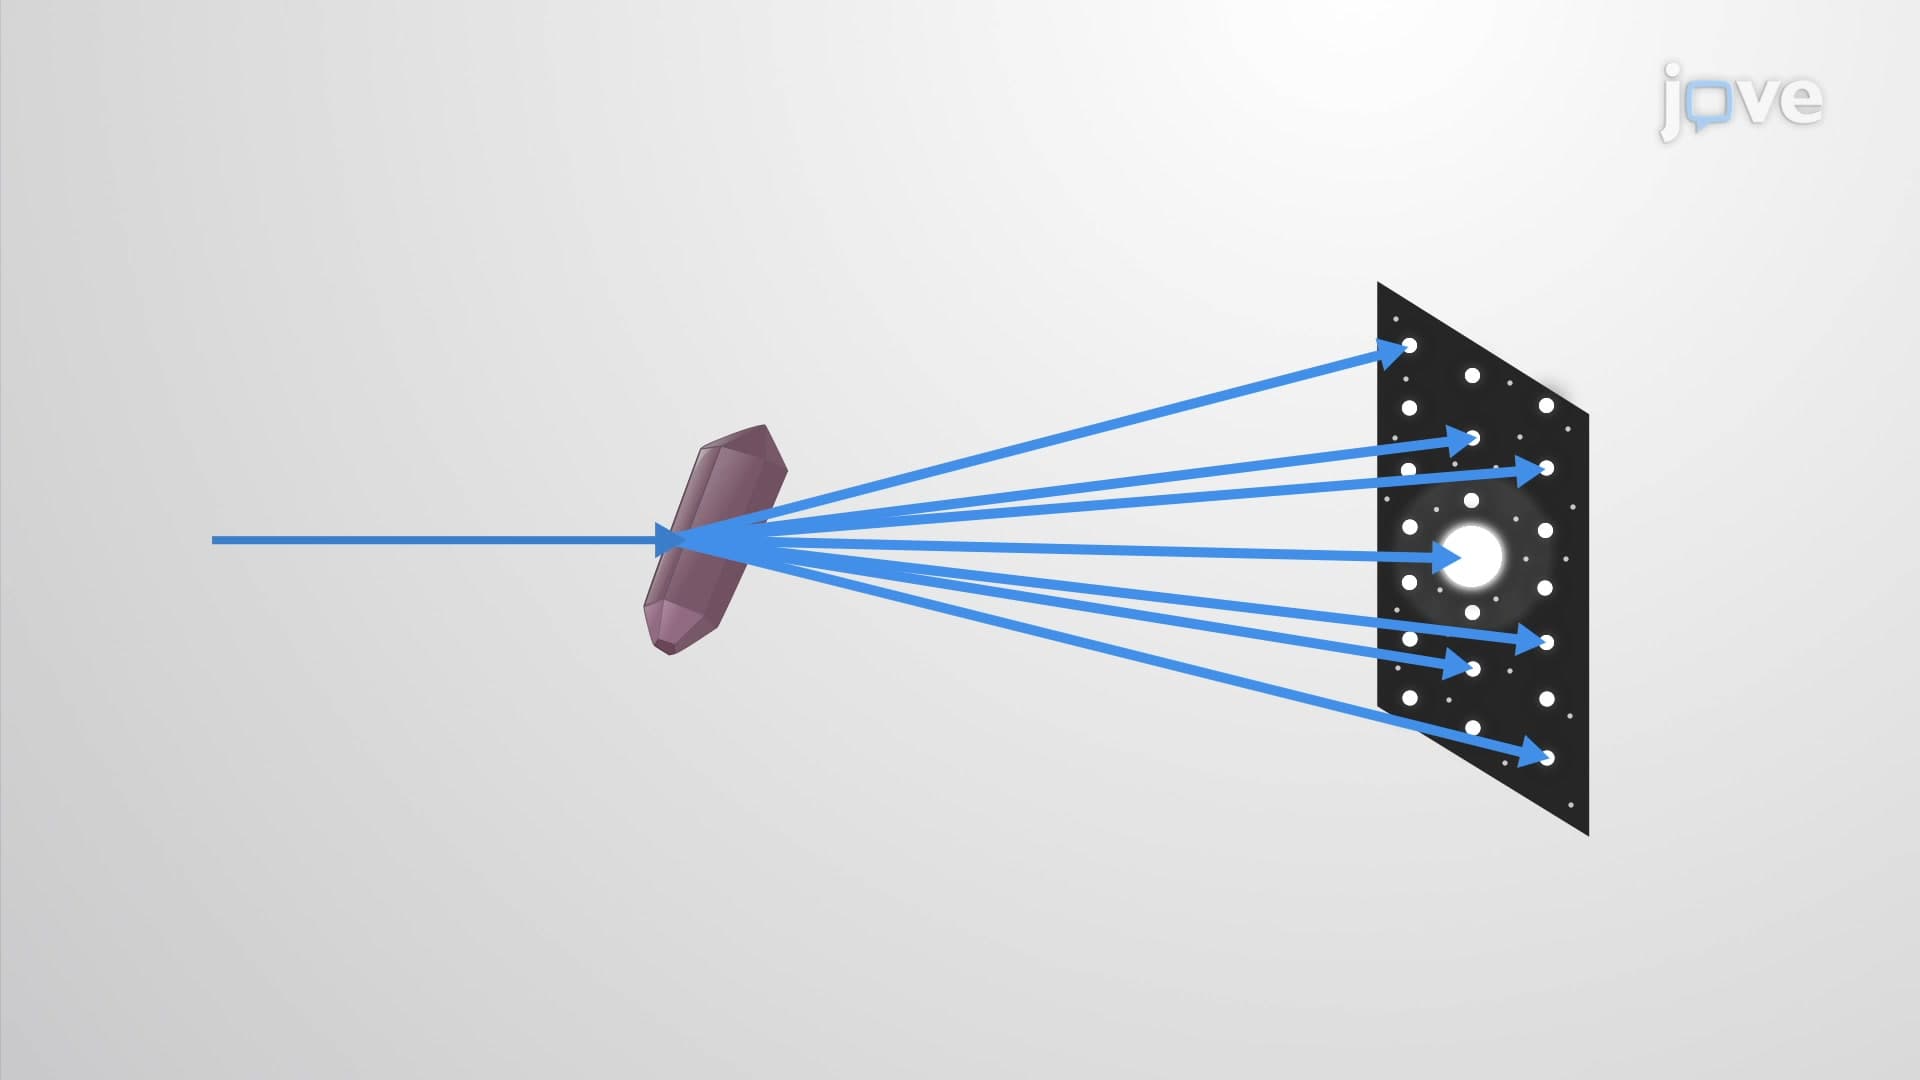
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...