Internal force corrections with machine learning for quantum mechanics/molecular mechanics simulations
Jingheng Wu1, Lin Shen1, Weitao Yang1
1Department of Chemistry, Duke University, Durham, North Carolina 27708, USA.

Abstract:
Ab initio quantum mechanics/molecular mechanics (QM/MM) molecular dynamics simulation is a useful tool to calculate thermodynamic properties such as potential of mean force for chemical reactions but intensely time consuming. In this paper, we developed a new method using the internal force correction for low-level semiempirical QM/MM molecular dynamics samplings with a predefined reaction coordinate. As a correction term, the internal force was predicted with a machine learning scheme, which provides a sophisticated force field, and added to the atomic forces on the reaction coordinate related atoms at each integration step. We applied this method to two reactions in aqueous solution and reproduced potentials of mean force at the ab initio QM/MM level. The saving in computational cost is about 2 orders of magnitude. The present work reveals great potentials for machine learning in QM/MM simulations to study complex chemical processes.


Related Concept Videos
 01:30
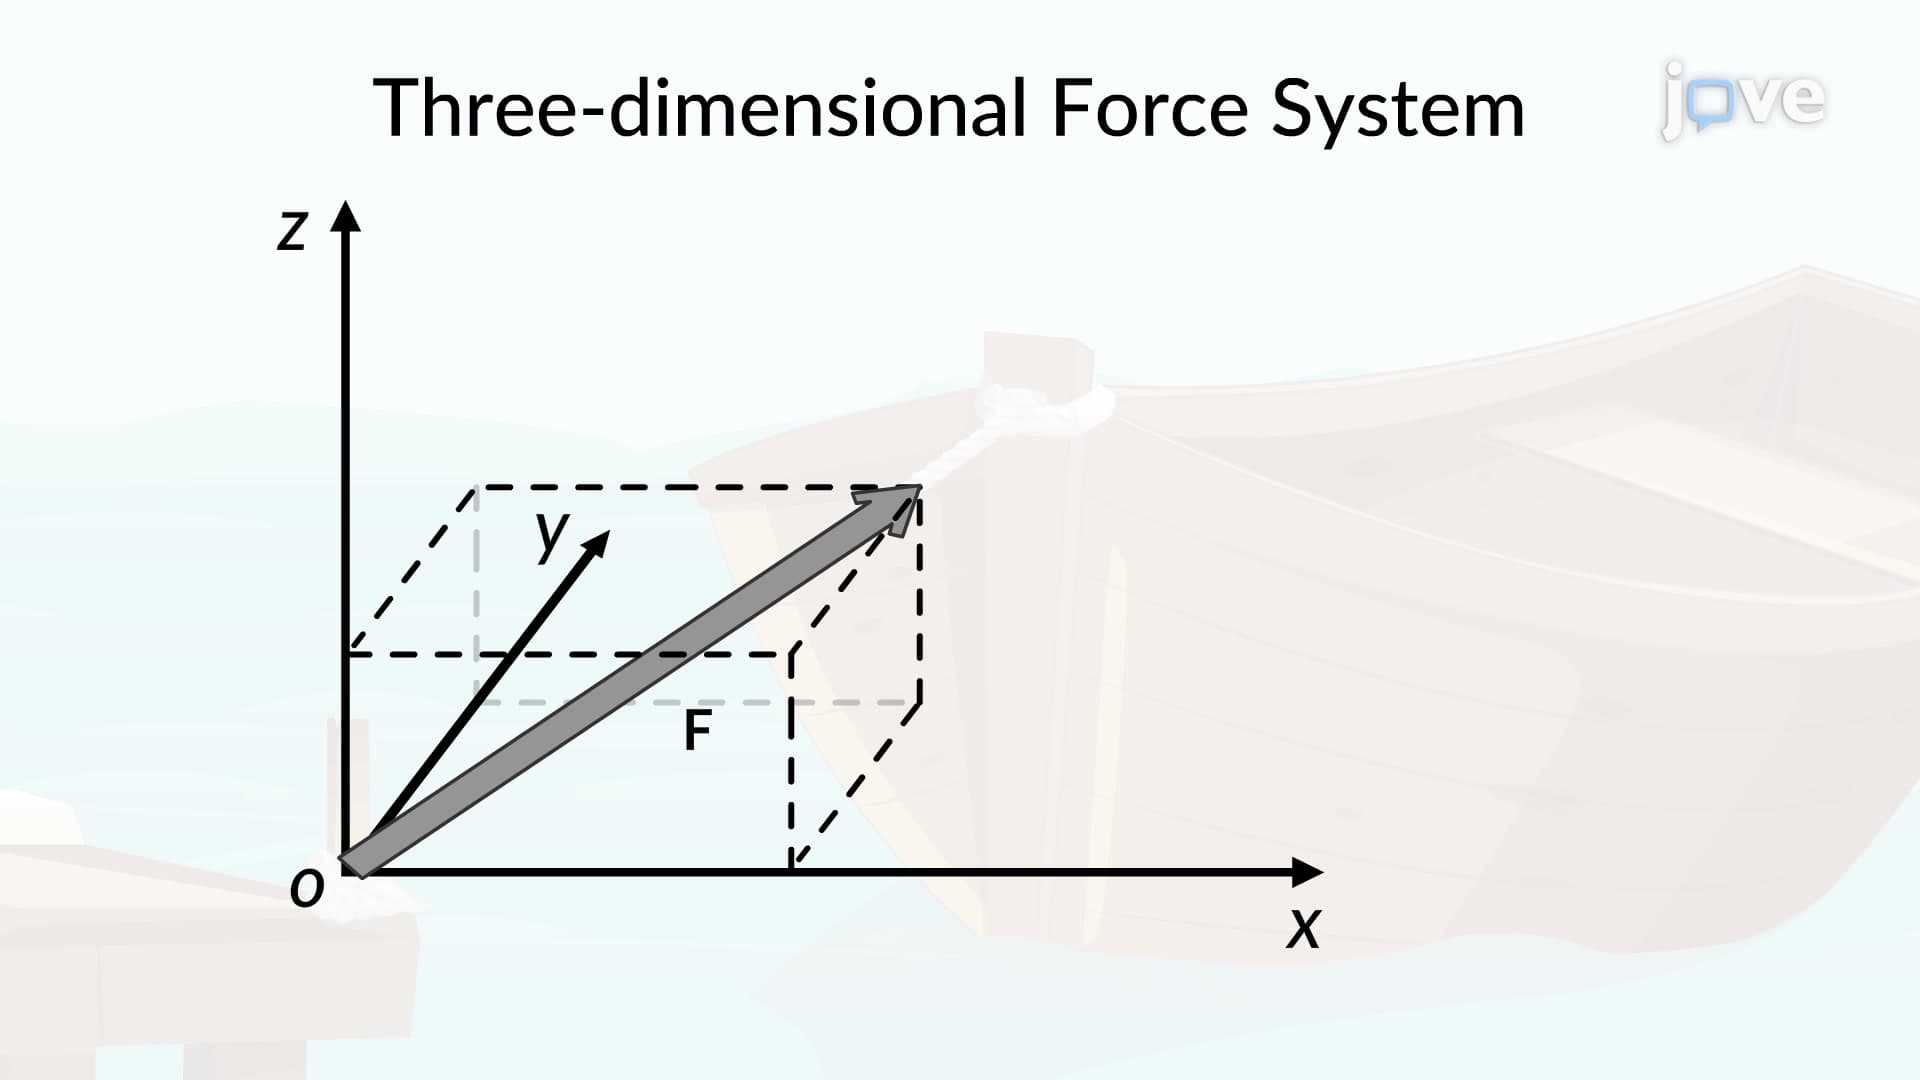
Three-Dimensional Force System
 01:30
Three-Dimensional Force System:Problem Solving
To solve a three-dimensional force system, first resolve each force into its respective scalar components. Do this using...
 02:45
The Quantum-Mechanical Model of an Atom
 02:27
Predicting Molecular Geometry
 01:17
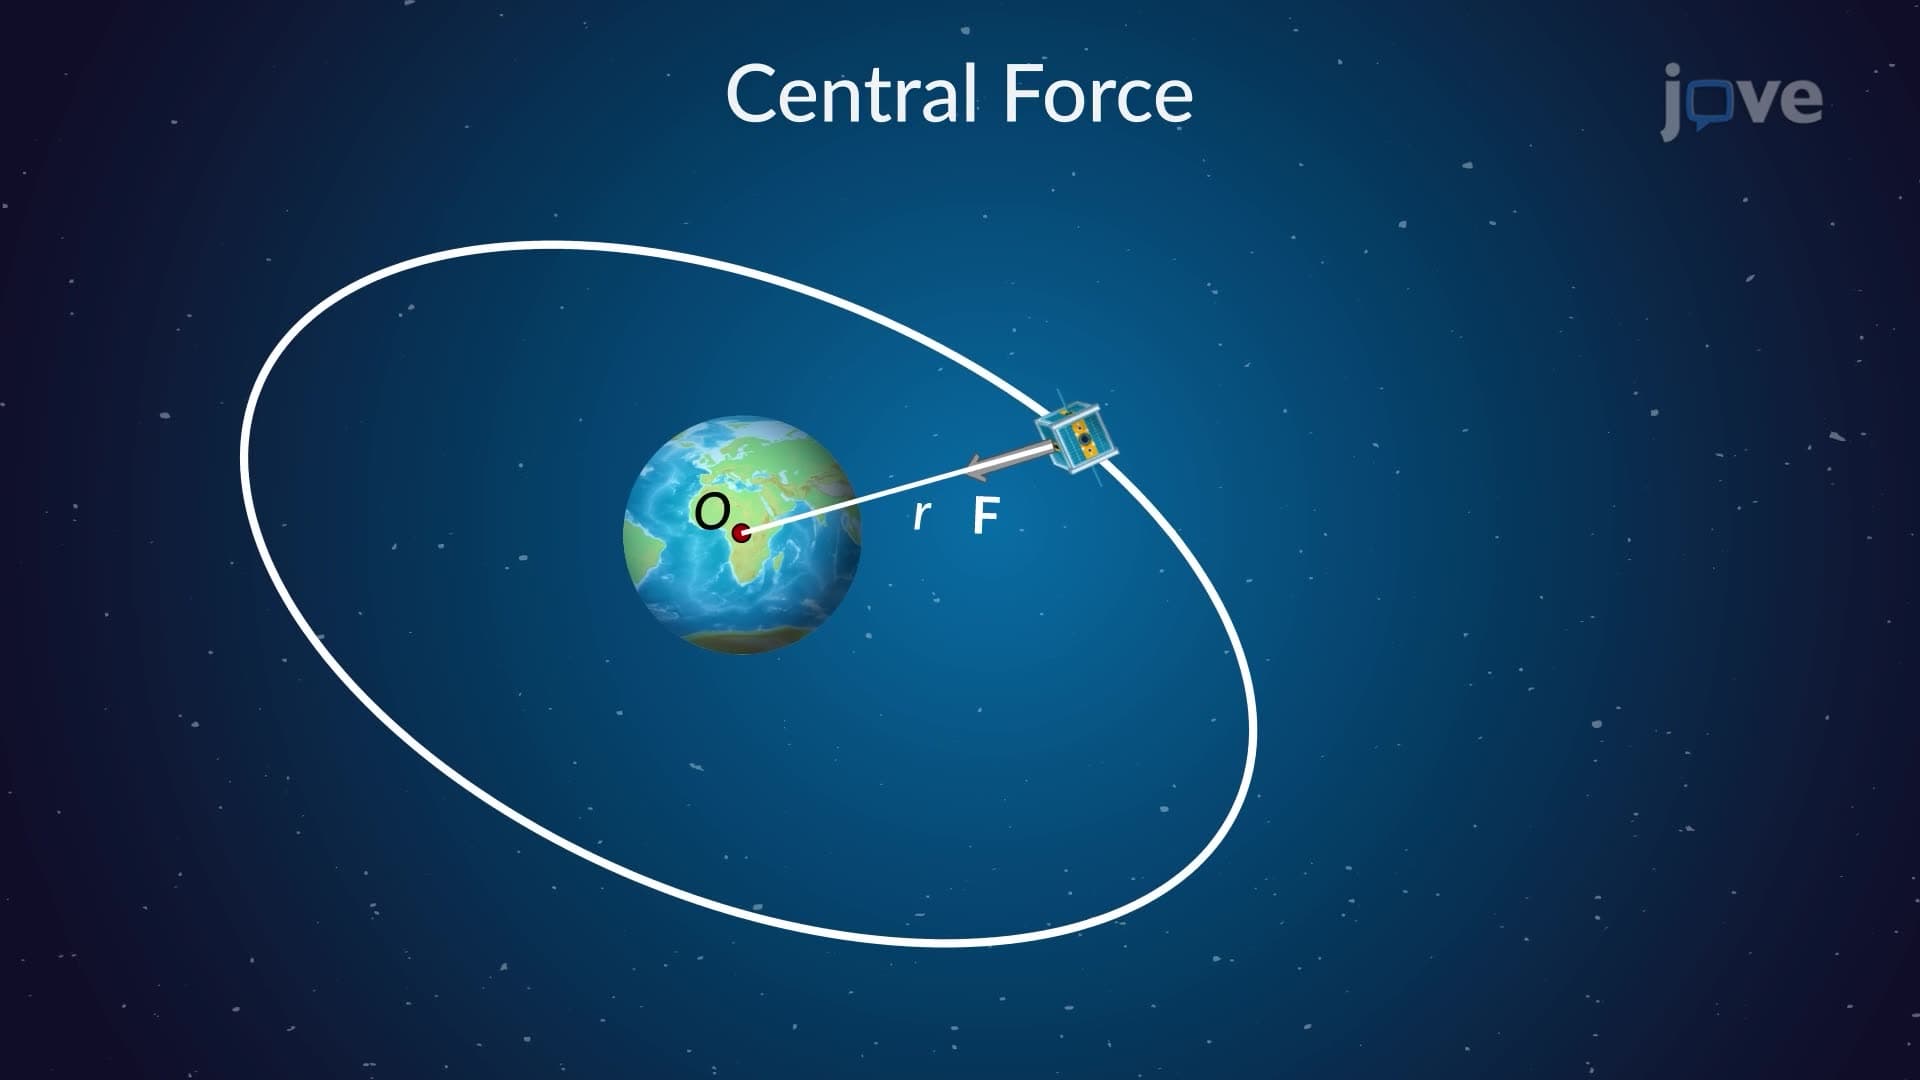
Central-Force Motion
 01:17
Non-conservative Forces
Also unlike their conservative counterparts, they are path-dependent; where the object starts and stops does matter. For example, a grinding wheel applies a...