Related Experiment Video
Updated: Feb 19, 2026
Standardized Measurement of Nasal Membrane Transepithelial Potential Difference NPD
Published on: September 13, 2018
Cystic fibrosis newborn screening: outcome of infants with normal sweat tests
Claire Edmondson1, Christopher Grime1, Ammani Prasad2
1Paediatric Respiratory Medicine, Royal Brompton Hospital, London, UK.

Abstract:
Newborn babies positively screened for cystic fibrosis (CF) (high serum immunoreactive trypsin (IRT) with DNA analysis) are referred for a diagnostic sweat test, which may be normal (sweat chloride <30 mmol/L). Unless two gene mutations are identified during Newborn screening (NBS), the babies are discharged from follow-up. We wished to check that none had subsequently developed symptoms suggestive of CF. We retrospectively reviewed patient notes and contacted general practitioners of all babies with a negative sweat test, conducted in one of the four paediatric specialist CF centres in London, over the first 6 years of screening in South East England.Of 511 babies referred, 95 (19%) had a normal sweat test. Five (5%) had CF diagnosed genetically, two of them on extended genome sequencing after clinical suspicion. Eleven (12%) were designated as CF screen positive inconclusive diagnosis (CFSPID); one of the five CF children was originally designated as CFSPID. Seventy-nine (83%) were assumed to be false-positive cases and discharged; follow-up data were available for 51/79 (65%); 32/51 (63%) had no health issues, 19/51 (37%) had other significant non-CF pathology.These results are reassuring in that within the limitations of those lost to follow-up, CF symptoms have not emerged in the discharged children. The high non-CF morbidity in these children may relate to known causes of high IRT at birth. Clinicians need to be aware that a child can have CF despite a normal sweat test following NBS, and if symptoms suggest the diagnosis, further testing, including extended genome sequencing, is required.
More Related Videos
 08:00
Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
 05:56
Implementation of Non-invasive Point of Care Transient Elastography for Evaluation of Liver Disease in Pediatric Populations with Cystic Fibrosis
Published on: August 29, 2025
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:20
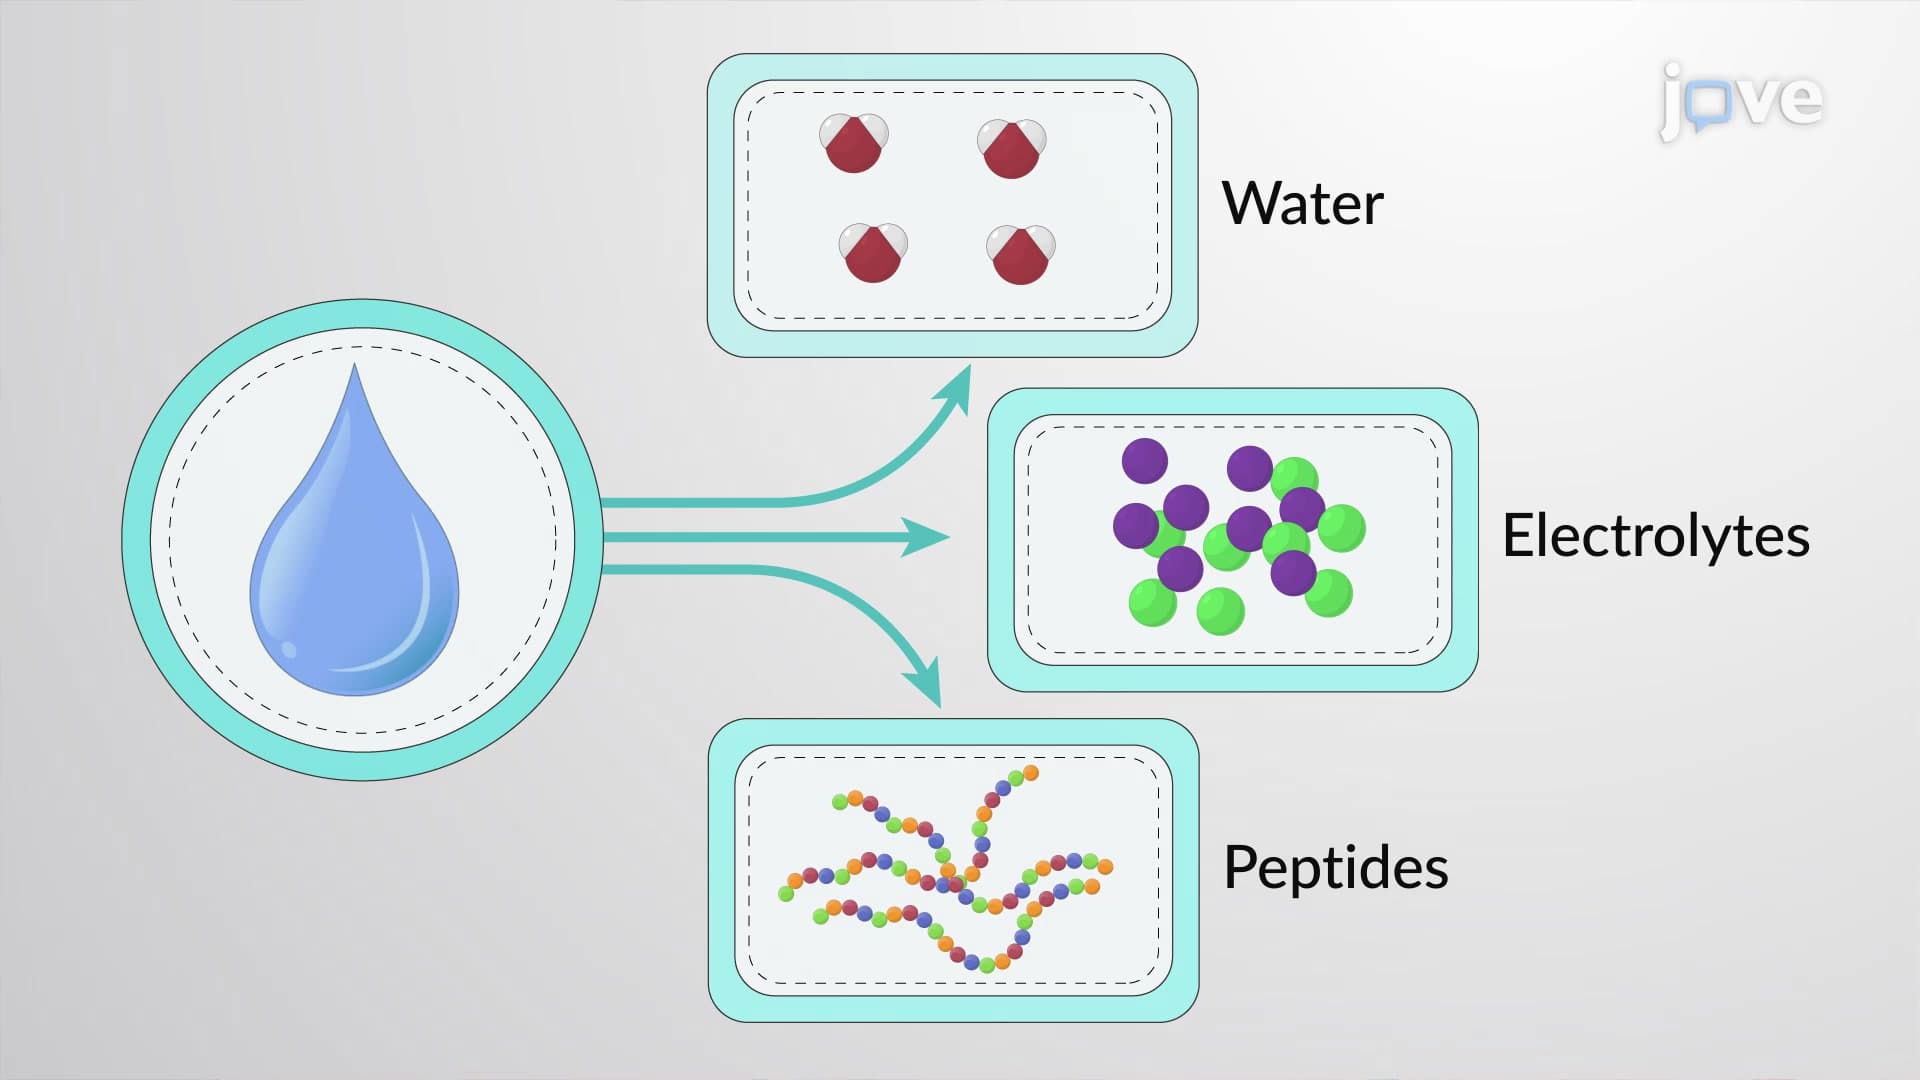
Accessory Structures of the Skin: Sweat Glands
Sweat glands are classified as merocrine glands; that is, the secretions are excreted by exocytosis through a duct without affecting the cells of the gland. There...
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes: