Related Experiment Video
Updated: Feb 19, 2026

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Using physics-based pose predictions and free energy perturbation calculations to predict binding poses and relative
Christina Athanasiou1, Sofia Vasilakaki1, Dimitris Dellis2
1Biomedical Research Foundation, Academy of Athens, 4 Soranou Ephessiou, 115 27, Athens, Greece.

Abstract:
Computer-aided drug design has become an integral part of drug discovery and development in the pharmaceutical and biotechnology industry, and is nowadays extensively used in the lead identification and lead optimization phases. The drug design data resource (D3R) organizes challenges against blinded experimental data to prospectively test computational methodologies as an opportunity for improved methods and algorithms to emerge. We participated in Grand Challenge 2 to predict the crystallographic poses of 36 Farnesoid X Receptor (FXR)-bound ligands and the relative binding affinities for two designated subsets of 18 and 15 FXR-bound ligands. Here, we present our methodology for pose and affinity predictions and its evaluation after the release of the experimental data. For predicting the crystallographic poses, we used docking and physics-based pose prediction methods guided by the binding poses of native ligands. For FXR ligands with known chemotypes in the PDB, we accurately predicted their binding modes, while for those with unknown chemotypes the predictions were more challenging. Our group ranked #1st (based on the median RMSD) out of 46 groups, which submitted complete entries for the binding pose prediction challenge. For the relative binding affinity prediction challenge, we performed free energy perturbation (FEP) calculations coupled with molecular dynamics (MD) simulations. FEP/MD calculations displayed a high success rate in identifying compounds with better or worse binding affinity than the reference (parent) compound. Our studies suggest that when ligands with chemical precedent are available in the literature, binding pose predictions using docking and physics-based methods are reliable; however, predictions are challenging for ligands with completely unknown chemotypes. We also show that FEP/MD calculations hold predictive value and can nowadays be used in a high throughput mode in a lead optimization project provided that crystal structures of sufficiently high quality are available.


Related Concept Videos
 02:18
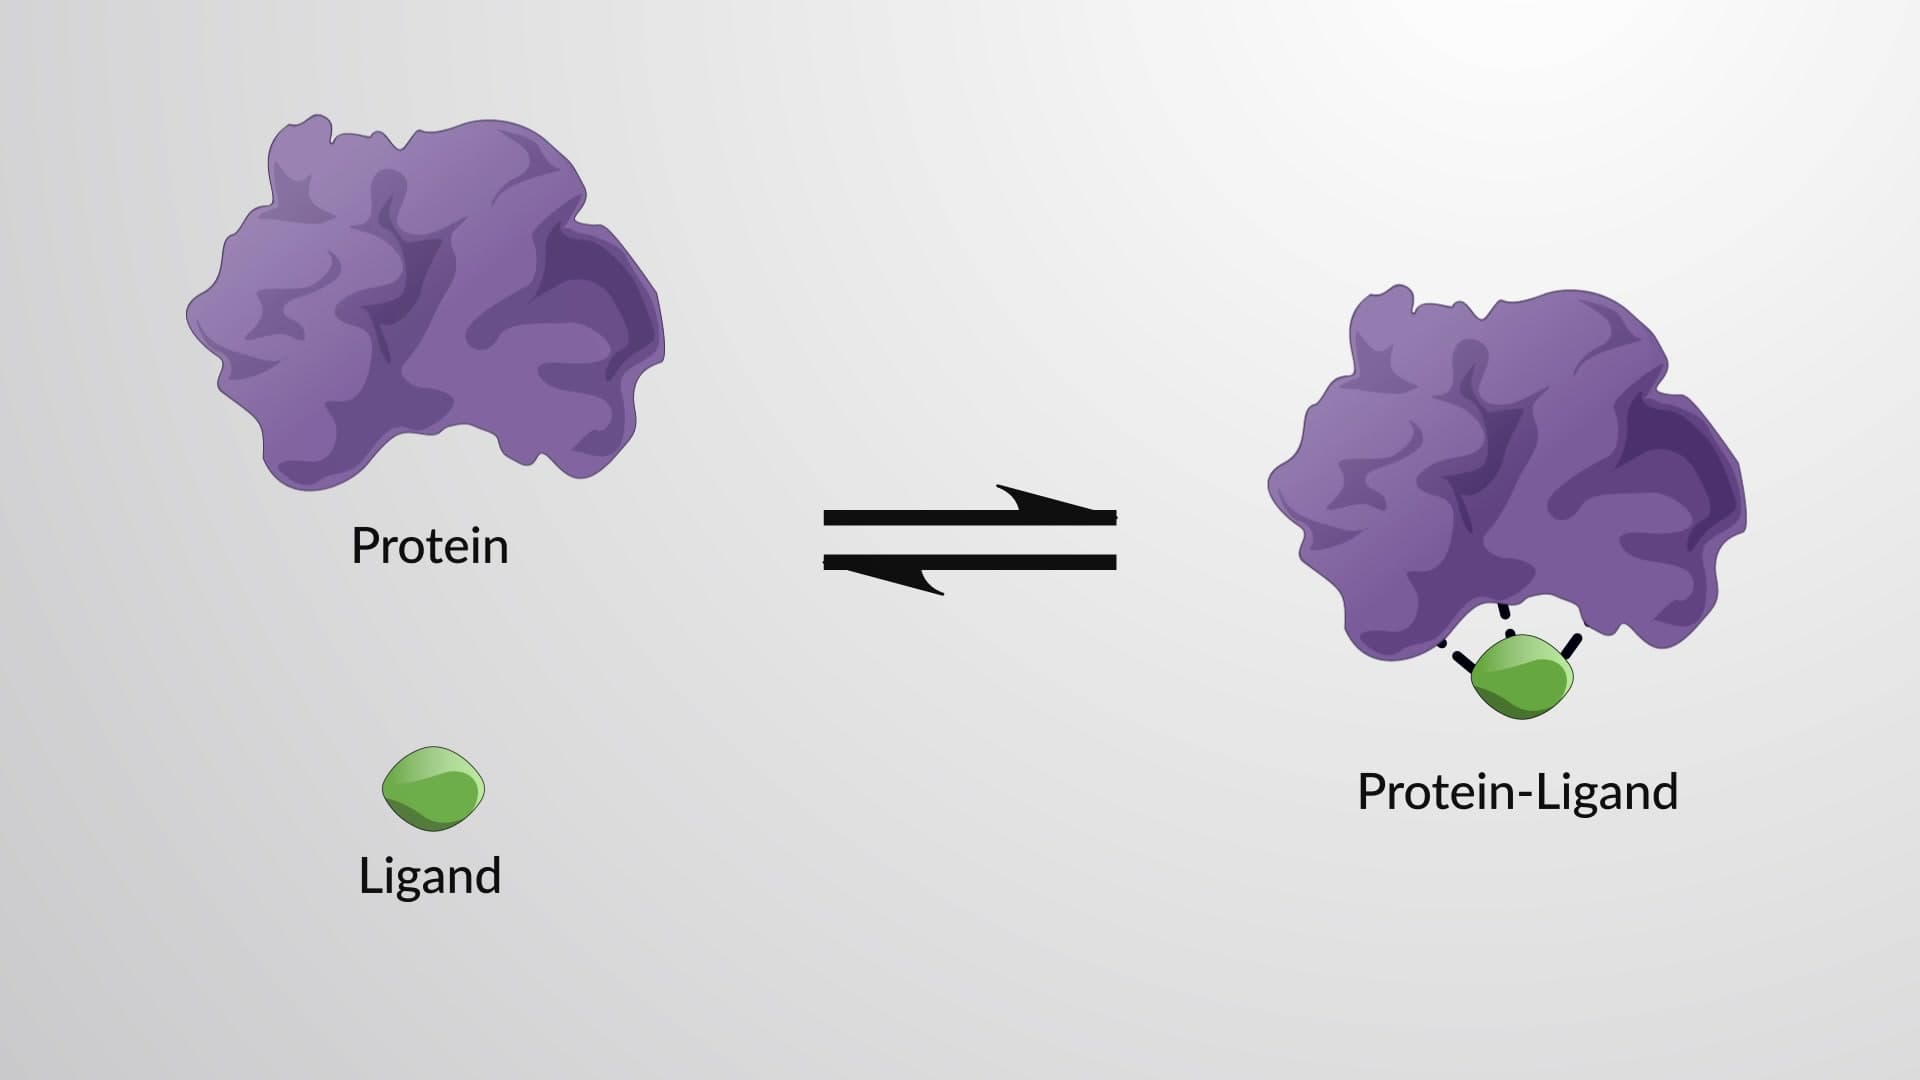
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:04
Protein-protein Interfaces
 01:13
Force and Potential Energy in One Dimension