Related Experiment Video
Updated: Feb 18, 2026


Solid-phase Synthesis of [4.4] Spirocyclic Oximes
Published on: February 6, 2019
Structure-based design of hERG-neutral antihypertensive oxazalone and imidazolone derivatives
Busecan Aksoydan1, Isik Kantarcioglu1, Ismail Erol2
1Computational Biology and Molecular Simulations Laboratory, Department of Biophysics, School of Medicine, Bahcesehir University (BAU), Istanbul, Turkey.

Abstract:
Angiotensin II receptor type 1 (AT1) antagonists are the most recent drug class against hypertension. Recently first crystal structure of AT1 receptor is deposited to the protein data bank (PDB ID: 4YAY). In this work, several molecular screening methods such as molecular docking and de novo design studies were performed and it is found that oxazolone and imidazolone derivatives reveal similar/better interaction energy profiles compared to the FDA approved sartan molecules at the binding site of the AT1 receptor. A database consisting of 3500-fragments were used to enumerate de novo designed imidazolone and oxazolone derivatives and hereby more than 50000 novel small molecules were generated. These derivatives were then used in high throughput virtual screening simulations (Glide/HTVS) to find potent hit molecules. In addition, virtual screening of around 18 million small drug-like compounds from ZINC database were screened at the binding pocket of the AT1 receptor via Glide/HTVS method. Filtered structures were then used in more sophisticated molecular docking simulations protocols (i.e., Glide/SP; Glide/XP; Glide/IFD; Glide/QPLD, and GOLD). However, the K+ ion channel/drug interactions should also be considered in studies implemented in molecular level against their cardiovascular risks. Thus, selected compounds with high docking scores via all diverse docking algorithms are also screened at the pore domain regions of human ether-a-go-go-related gene (hERG1) K+ channel to remove the high affinity hERG1 blocking compounds. High docking scored compounds at the AT1 with low hERG1 affinity is considered for long molecular dynamics (MD) simulations. Post-processing analysis of MD simulations assisted for better understanding of molecular mechanism of studied compounds at the binding cavity of AT1 receptor. Results of this study can be useful for designing of novel and safe AT1 inhibitors.
More Related Videos
Related Concept Videos
 01:15
Antihypertensive Drugs: Thiazide-Class Diuretics
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:23
Antihypertensive Drugs: Vasodilators
 01:28
Antihypertensive Drugs: Types of β-Blockers
 01:26
Heart Failure Drugs: Inhibitors of Renin-Angiotensin System
 01:17
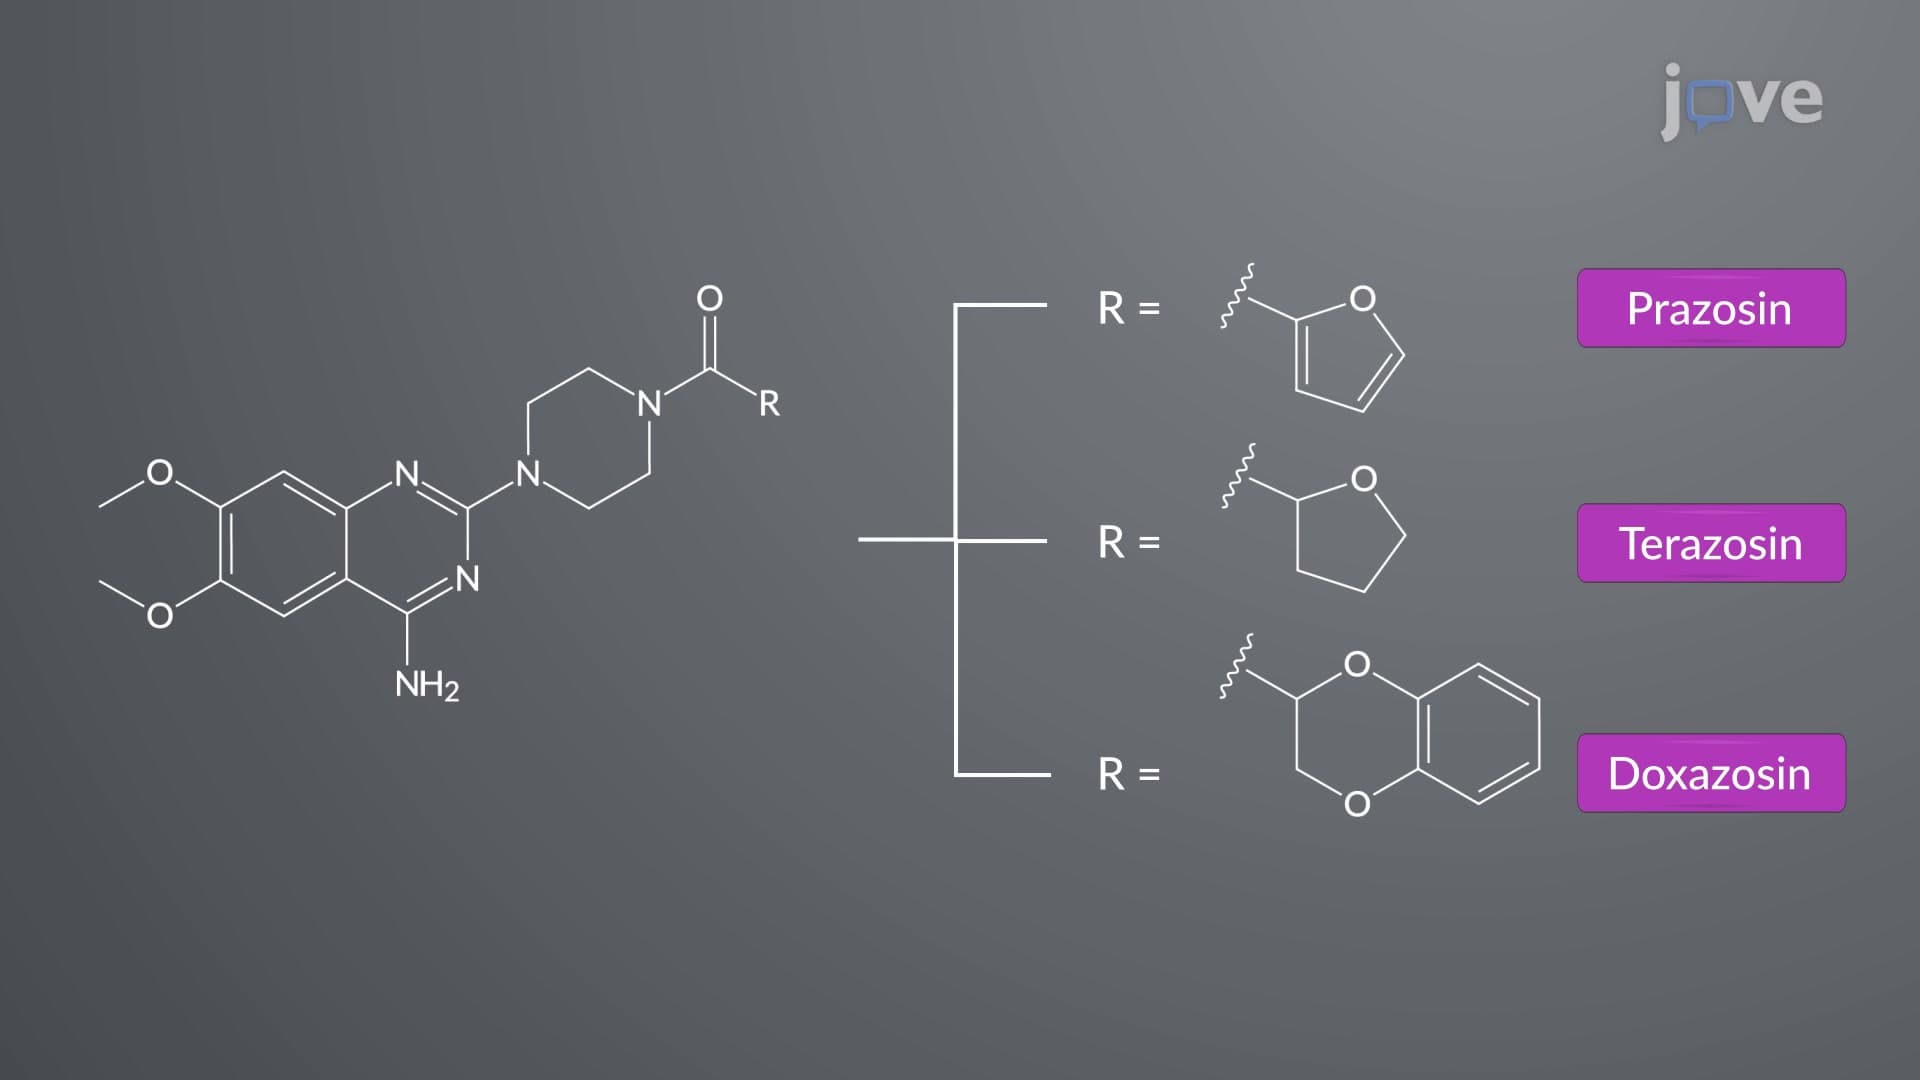
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...