Related Experiment Video
Updated: Feb 18, 2026

Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
The electronic and optical properties of the sulvanite compounds: a many-body perturbation and time-dependent density
W F Espinosa-García1,2, S Pérez-Walton3, J M Osorio-Guillén2
1Facultad de Ingenierías, Grupo de Investigación en Modelamiento y Simulación Computacional, Universidad de San Buenaventura-Medellín, Carrera 56C No 51-110, Medellín, Colombia.

Abstract:
We have studied, by means of first-principles calculations, the electronic and optical properties of the sulvanite family: Cu3MX4 (M = V, Nb, Ta and X = S, Se), which, due to its broad range of gaps and chemical stability, have emerged as promising materials for technological applications such as photovoltaics and transparent conductivity. To address the reliability of those properties we have used semi-local and hybrid functionals (PBEsol, HSE06), many-body perturbation theory (G0W0 approximation and Bethe-Salpeter equation), and time-dependent density functional theory (revised bootstrap kernel) to calculate the quasi-particle dispersion relation, band gaps, the imaginary part of the macroscopic dielectric function and the absorption coefficient. The calculated valence band maximum and the conduction band minimum are located at the R and X-points, respectively. The calculated gaps using PBEsol are between 0.81 and 1.88 eV, with HSE06 into 1.73 and 2.94 eV, whereas the G0W0 values fall into the 1.91-3.19 eV range. The calculated dielectric functions and absorption coefficients show that all these compounds present continuous excitonic features when the Bethe-Salpeter equation is used. Contrarily, the revised bootstrap kernel is incapable of describing the excitonic spectra. The calculated optical spectra show that Cu3VS4 and Cu3MSe4 have good absorption in the visible, whereas Cu3NbS4 and Cu3TaS4 have it on the near ultraviolet.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:58
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Related Concept Videos
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 01:10
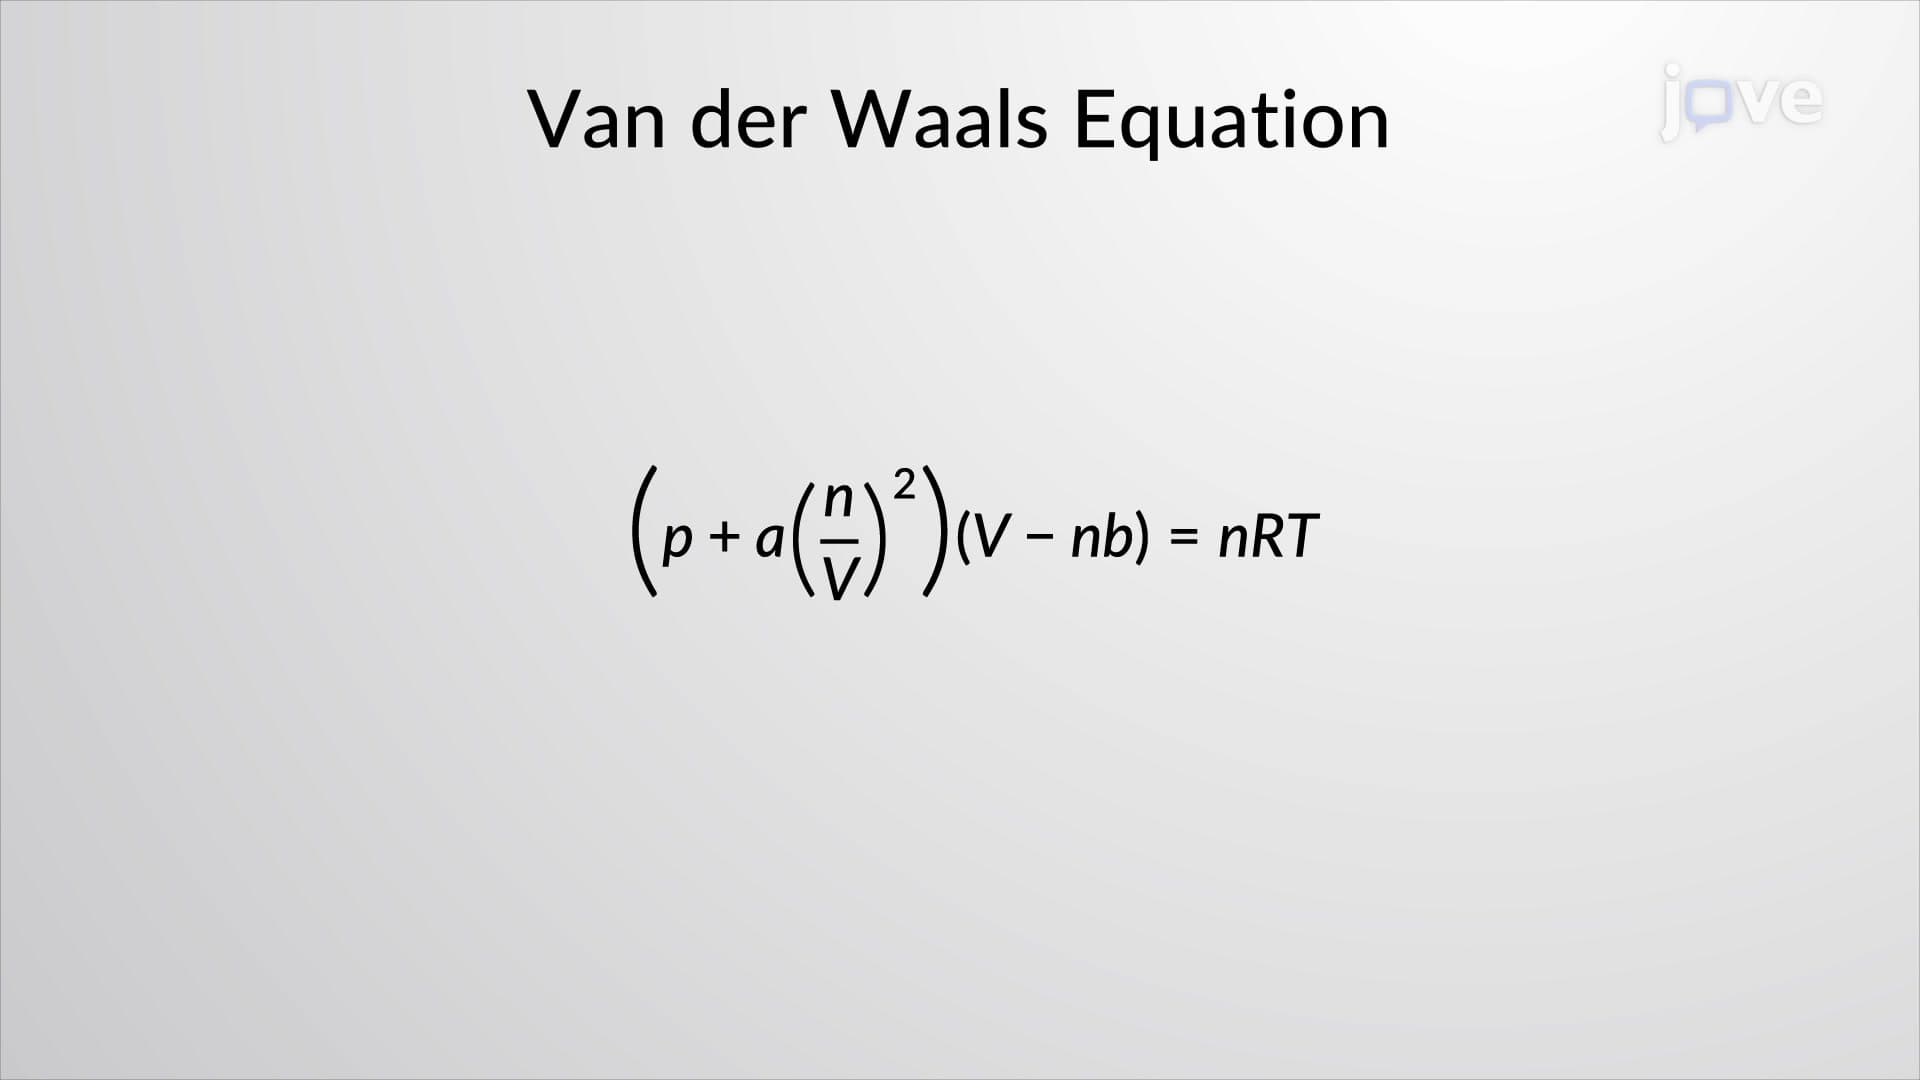
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
Van der Waals Interactions