Related Experiment Video
Updated: Feb 17, 2026
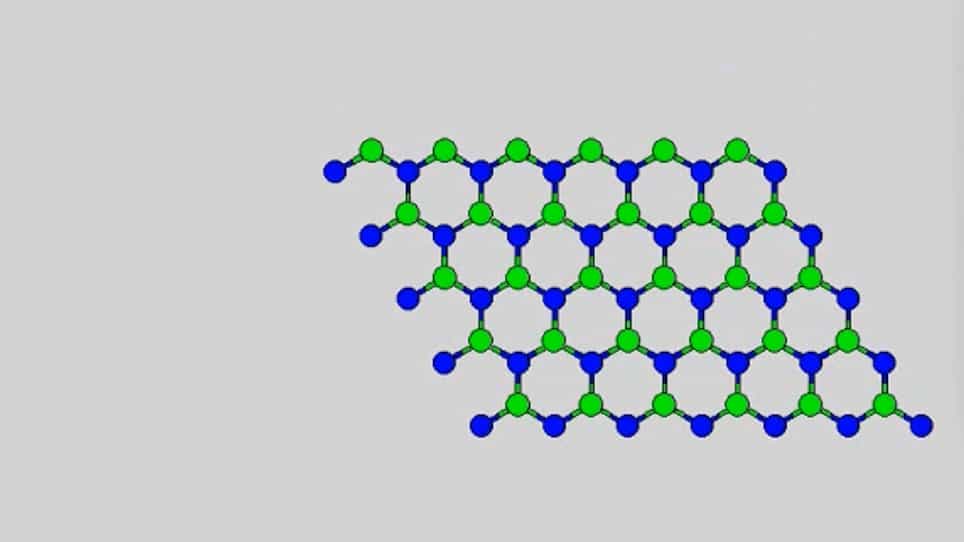
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Dual-Functional Tamm-Dancoff Approximation with Self-Interaction-Free Orbitals: Vertical Excitation Energies and
Yinan Shu1, Kelsey A Parker1, Donald G Truhlar1
1Department of Chemistry, Chemical Theory Center, and Minnesota Supercomputing Institute, University of Minnesota , Minneapolis, Minnesota 55455-0431, United States.

Abstract:
Recently we have developed the dual-functional Tamm-Dancoff approximation (DF-TDA) method. DF-TDA is an alternative to linear-response time-dependent density functional theory (LR-TDDFT) with the advantage of providing a correct double-cone topology of S1/S0 conical intersections. In the DF-TDA method, we employ different functionals, which are denoted G and F, for orbital optimization and Hamiltonian construction. We use the notation DF-TDA/G:F. In the current work, we propose that G be the same as F except for having 100% Hartree-Fock exchange. We use the notation F100 to denote functional F with this modification. A motivation for this is that functionals with 100% Hartree-Fock exchange are one-electron self-interaction-free. Here we validate the use of F100/M06 to compute vertical excitation energies and the global potential energy surface of ammonia near a conical intersection to further validate the F100 method for photochemical problems.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 03:35
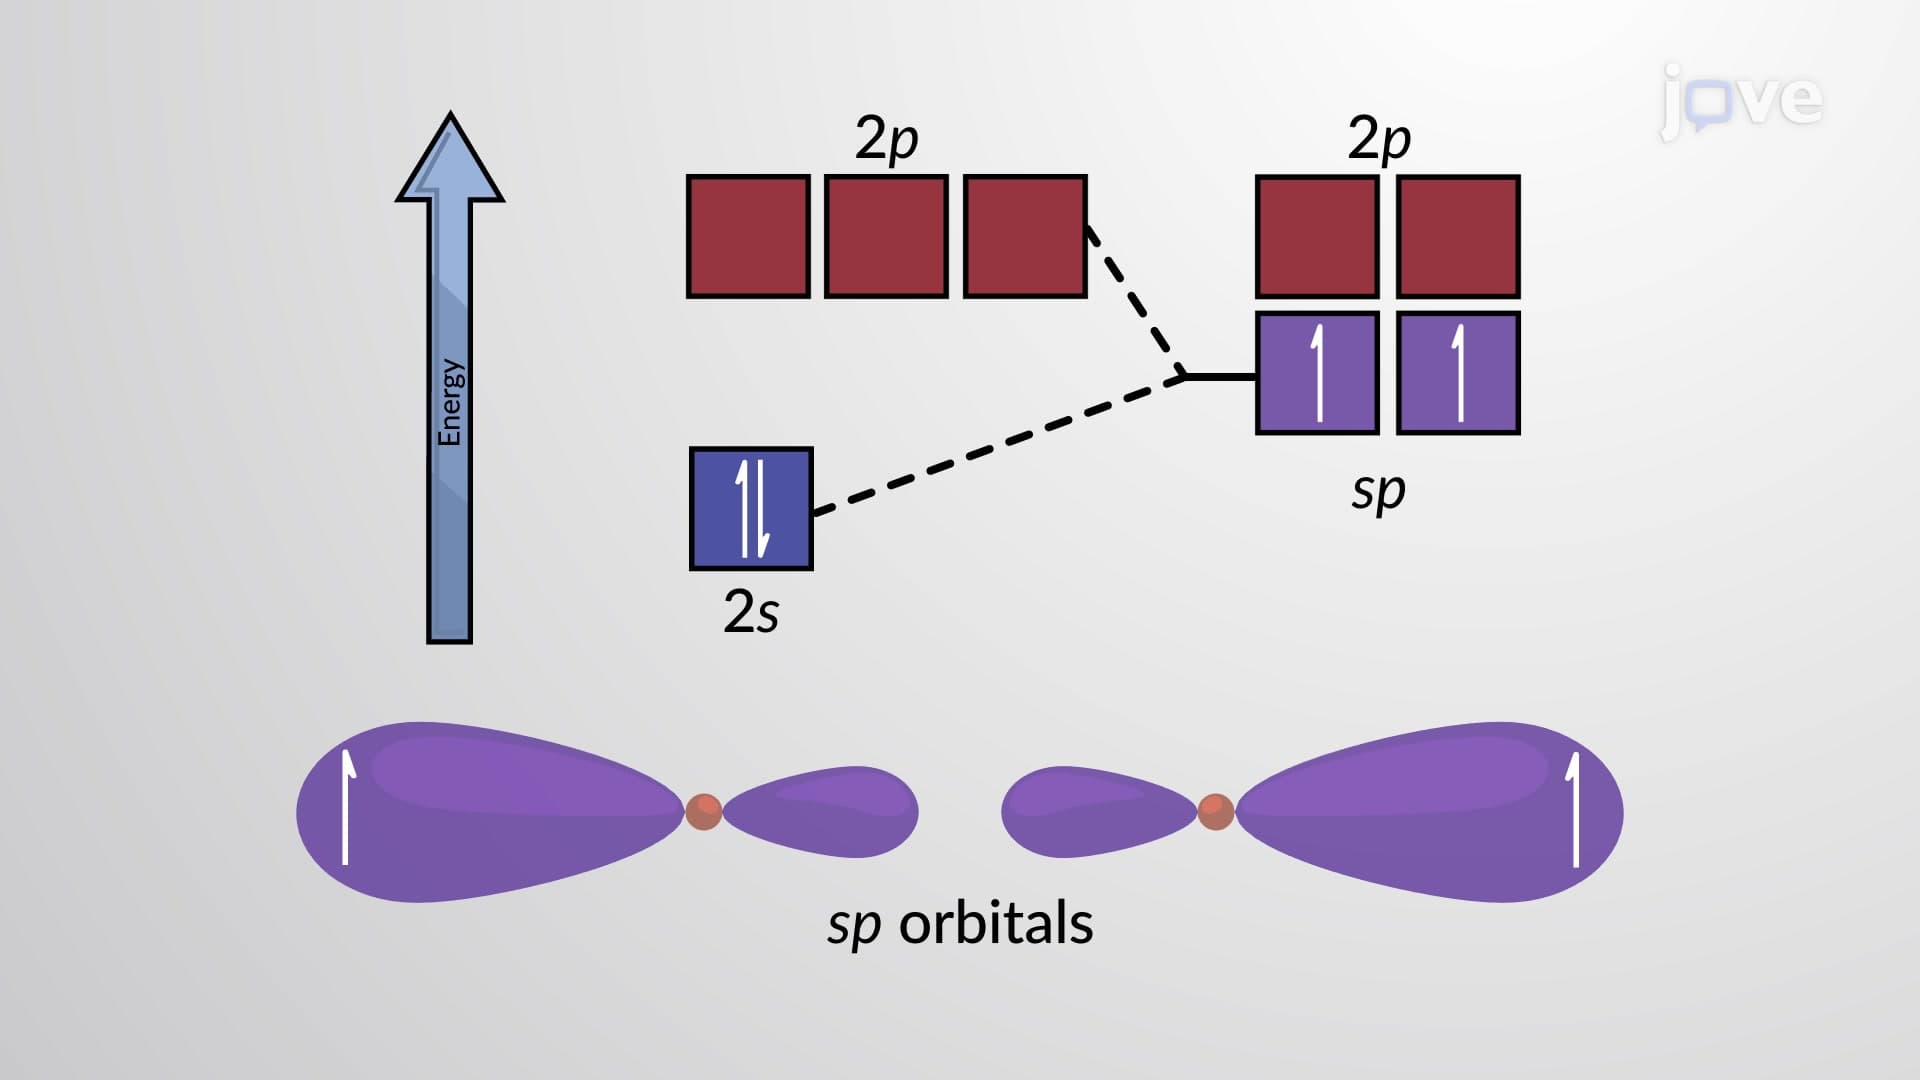
Hybridization of Atomic Orbitals II
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:24
Hybridization of Atomic Orbitals I
 02:44
Atomic Orbitals
 01:10
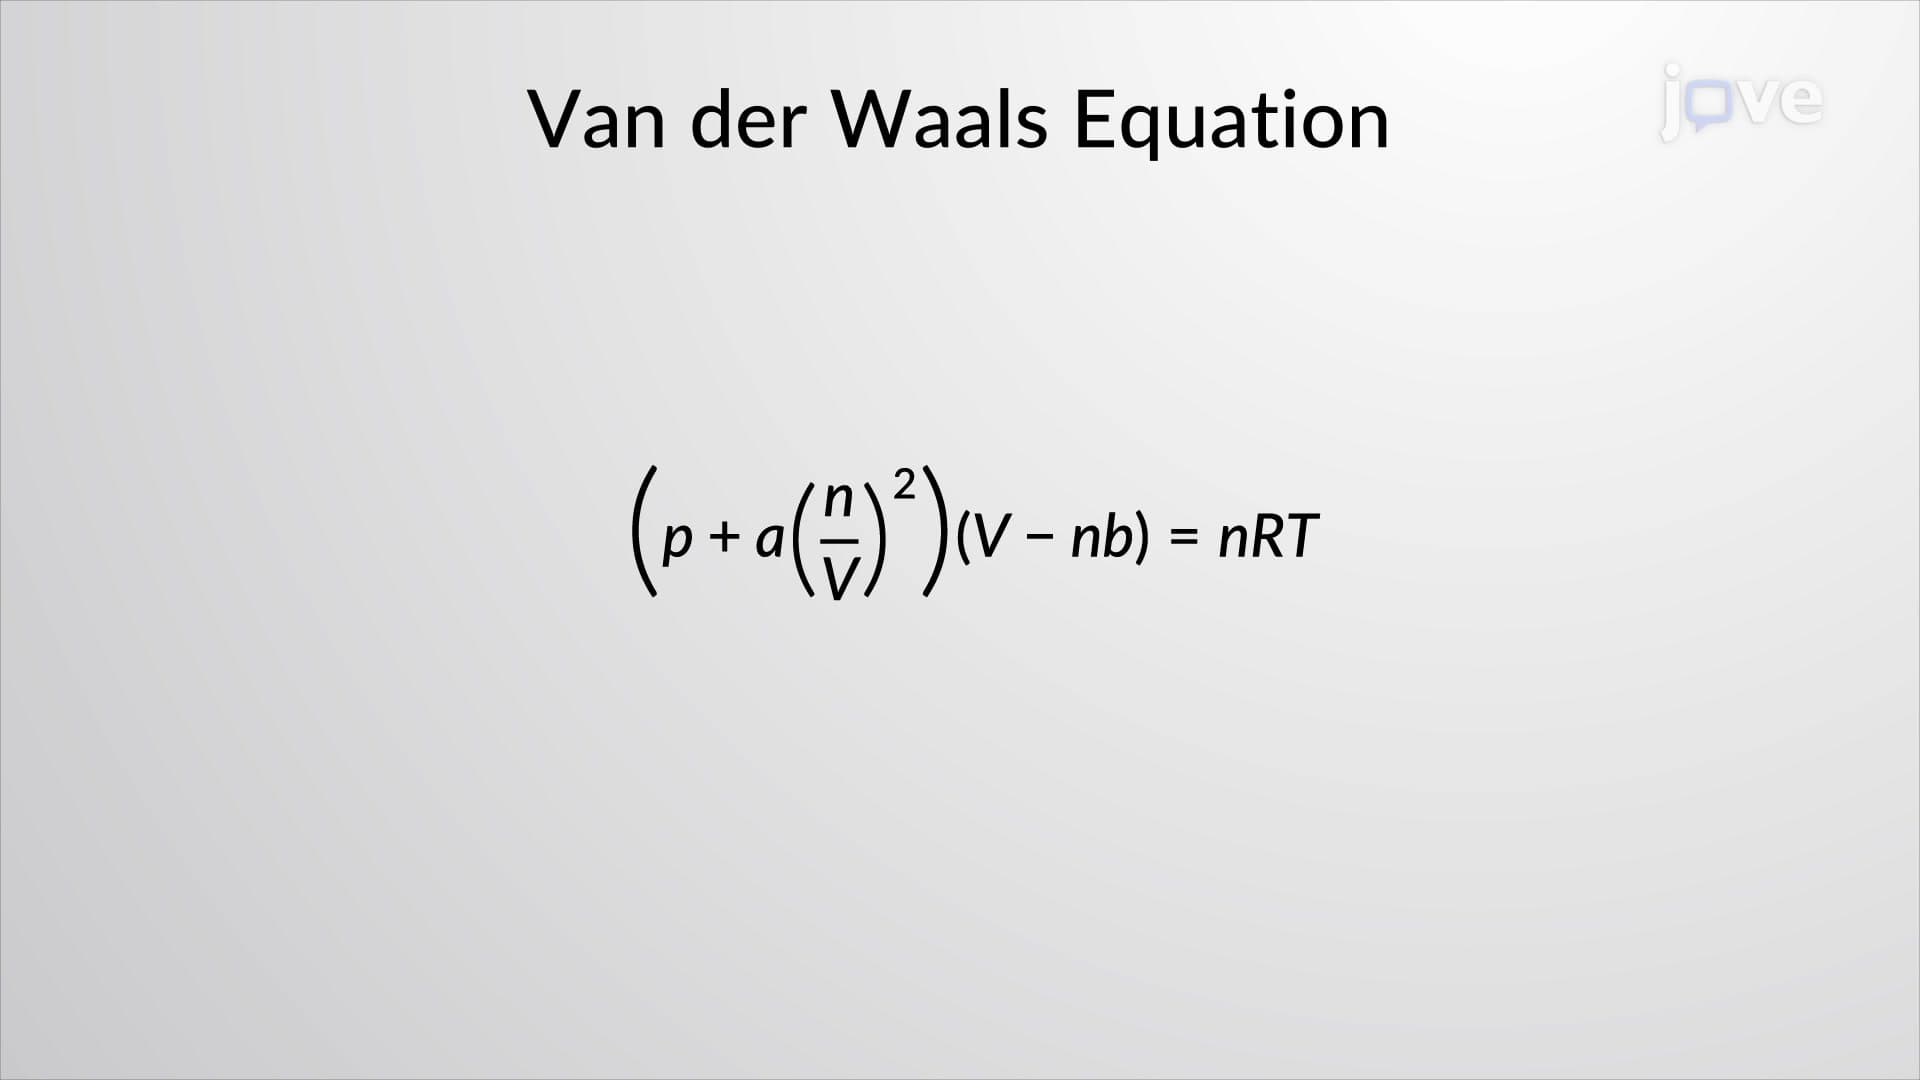
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:21
The Energies of Atomic Orbitals