Related Experiment Video
Updated: Feb 17, 2026
Synthesis and Exfoliation of Discotic Zirconium Phosphates to Obtain Colloidal Liquid Crystals
Published on: May 25, 2016
Geometry optimization of zirconium sulfophenylphosphonate layers by molecular simulation methods
Jakub Škoda1, Miroslav Pospíšil2, Petr Kovář1
1Charles University, Faculty of Mathematics and Physics, Ke Karlovu 3, 121 16, Prague 2, Czech Republic.

Abstract:
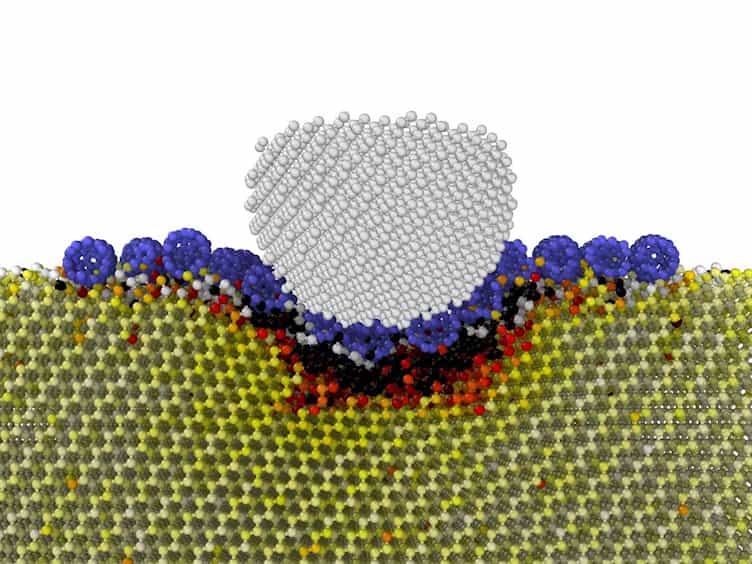
Classical molecular simulation methods were used for a detailed structural description of zirconium 4-sulfophenylphosphonate and zirconium phenylphosphonate 4-sulfophenylphosphonates with general formula Zr(HO3SC6H4PO3) x (C6H5PO3)2-x ·yH2O (x = 0.7-2; y = 0 or 2). First, models describing the structure of zirconium 4-sulfophenylphosphonate (x = 2) were calculated for the hydrated (y = 2) and dehydrated (y = 0) compounds. Subsequently, models for two mixed zirconium phenylphosphonate 4-sulfophenylphosphonates (x = 1.3 and 0.7) were calculated. Optimized models suggest that the presence of water molecules between sulfo groups creates a water-sulfonate layer with a system of hydrogen bonds. We suppose that this arrangement is the reason for a higher proton conductivity of the hydrated samples compared to dehydrated samples. When the water molecules are removed, a small decrease in the basal spacing (around 0.06 Å) is observed. This behavior is confirmed by the simulated models, where no significant changes in the structure on dehydration were observed except the absence of the water molecules and a lower number of hydrogen bonds between two adjacent sulfonate sheets. Due to the good crystallinity of the samples and the presence of sharp non-basal peaks in their X-ray diffraction patterns, Miller indices of the non-basal peaks in the diffraction patterns calculated from the models can be compared with those found in the experimental data. This allowed us to precisely describe for example (15 5-2) planes, from which mutual distances of the phenyl rings were determined to be 2.62 Å. Graphical Abstract Detailed ball and stick view into the interlayer structure of ZrSPhP1.3.
Related Concept Videos
 02:52
VSEPR Theory and the Basic Shapes
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:27
Predicting Molecular Geometry