Related Experiment Video
Updated: Feb 17, 2026
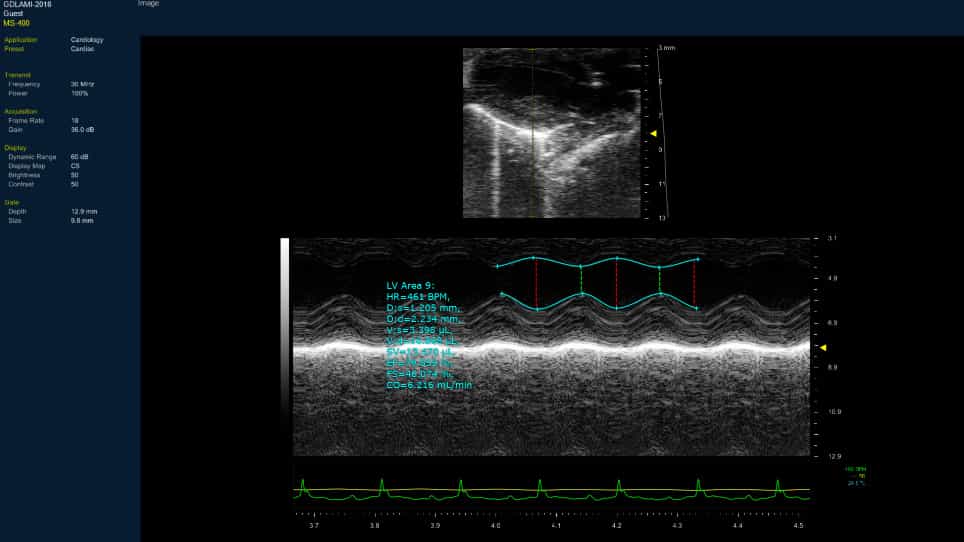
Investigating the Pathogenesis of MYH7 Mutation Gly823Glu in Familial Hypertrophic Cardiomyopathy using a Mouse Model
Published on: August 8, 2022
Role of titin in cardiomyopathy: from DNA variants to patient stratification
James S Ware1,2, Stuart A Cook1,2,3
1National Heart and Lung Institute, Imperial College London, Royal Brompton Hospital, Sydney Street, London SW3 6NP, UK.

Abstract:
Dilated cardiomyopathy (DCM) affects approximately 1 in 250 individuals and is the leading indication for heart transplantation. DCM is often familial, and the most common genetic predisposition is a truncating variation in the giant sarcomeric protein, titin, which occurs in up to 15% of ambulant patients with DCM and 25% of end-stage or familial cases. In this article, we review the evidence for the role of titin truncation in the pathogenesis of DCM and our understanding of the molecular mechanisms and pathophysiological consequences of variation in the gene encoding titin (TTN). Such variation is common in the general population (up to 1% of individuals), and we consider key features that discriminate variants with disease-causing potential from those that are benign. We summarize strategies for clinical interpretation of genetic variants for use in the diagnosis of patients and the evaluation of their relatives. Finally, we consider the contemporary and potential future role for genetic stratification in cardiomyopathy and in the general population, evaluating titin variation as a predictor of outcome and treatment response for precision medicine.
Related Concept Videos
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:25
Cardiomyopathy I: Introduction and Classification
 01:30
Cardiomyopathy II: Dilated Cardiomyopathy
 01:29
Cardiomyopathy IV: Restrictive Cardiomyopathy
 02:30
Histone Variants at the Centromere
 01:29
Pharmacogenomics: Identification of New Drug Targets