Mechanistic Insights into Solvent and Ligand Dependency in Cu(I)-Catalyzed Allylic Alkylation with gem-Diborylalkanes
Related Experiment Video
Updated: Feb 16, 2026
Facile Preparation of 2Z,4E-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Qi Zhang1,2, Bing Wang3, Jia-Qin Liu1,2
1Institute of Industry & Equipment Technology, Hefei University of Technology , Hefei 230009, China.

Abstract:
The recent Cu-catalyzed allylic substitution reaction between gem-diboryalkane and allyl electrophiles shows intriguing solvent and ligand-controlled regioselectivity. The α-alkylation product was obtained in DMF solvent, while γ-alkylation product was obtained in dioxane solvent and the dioxane and NHC ligand situation. In the present study, density functional theory calculations have been used to investigate the reaction mechanism and origin of the regioselectivity. For both dioxane and DMF, γ-alkylation undergoes successive oxidative addition (CH2Bpin trans to leaving group) and direct Cγ-C reductive elimination. The α-alkylation is found to undergo oxidative addition (CH2Bpin trans to leaving group), isomerization, and Cα-C reductive elimination rather than the previously proposed oxidative addition (-CH2Bpin cis to the leaving group) and Cα-C reductive elimination. The γ-alkylation and α-alkylation is, respectively, favorable for dioxane and DMF solvent, which is consistent with the γ- and α-selectivity in experiment. The solvent interferes the isomerization step, thereby affects the relative facility of the α- and γ-alkylation. Further investigation shows that η1-intermediate formation promoted by solvent is the rate-determining step of the isomerization. The stronger electron-donating ability of DMF than dioxane facilitates the η1-intermediate formation and finally results in the easier isomerization in DMF. For dioxane and NHC situation, in the presence of neutral NHC ligand, the -PO4Et2 group tightly coordinates with the Cu center after the oxidative addition, preventing the isomerization process. The regioselectivity is determined by the relative facility of the oxidative addition step. Therefore, the favorable oxidative addition (in which -CH2Bpin trans to the leaving group) results in the facility of γ-alkylation.
More Related Videos
 12:08
Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
 09:45
Accessing Valuable Ligand Supports for Transition Metals: A Modified, Intermediate Scale Preparation of 1,2,3,4,5-Pentamethylcyclopentadiene
Published on: March 20, 2017
Related Concept Videos
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 01:20
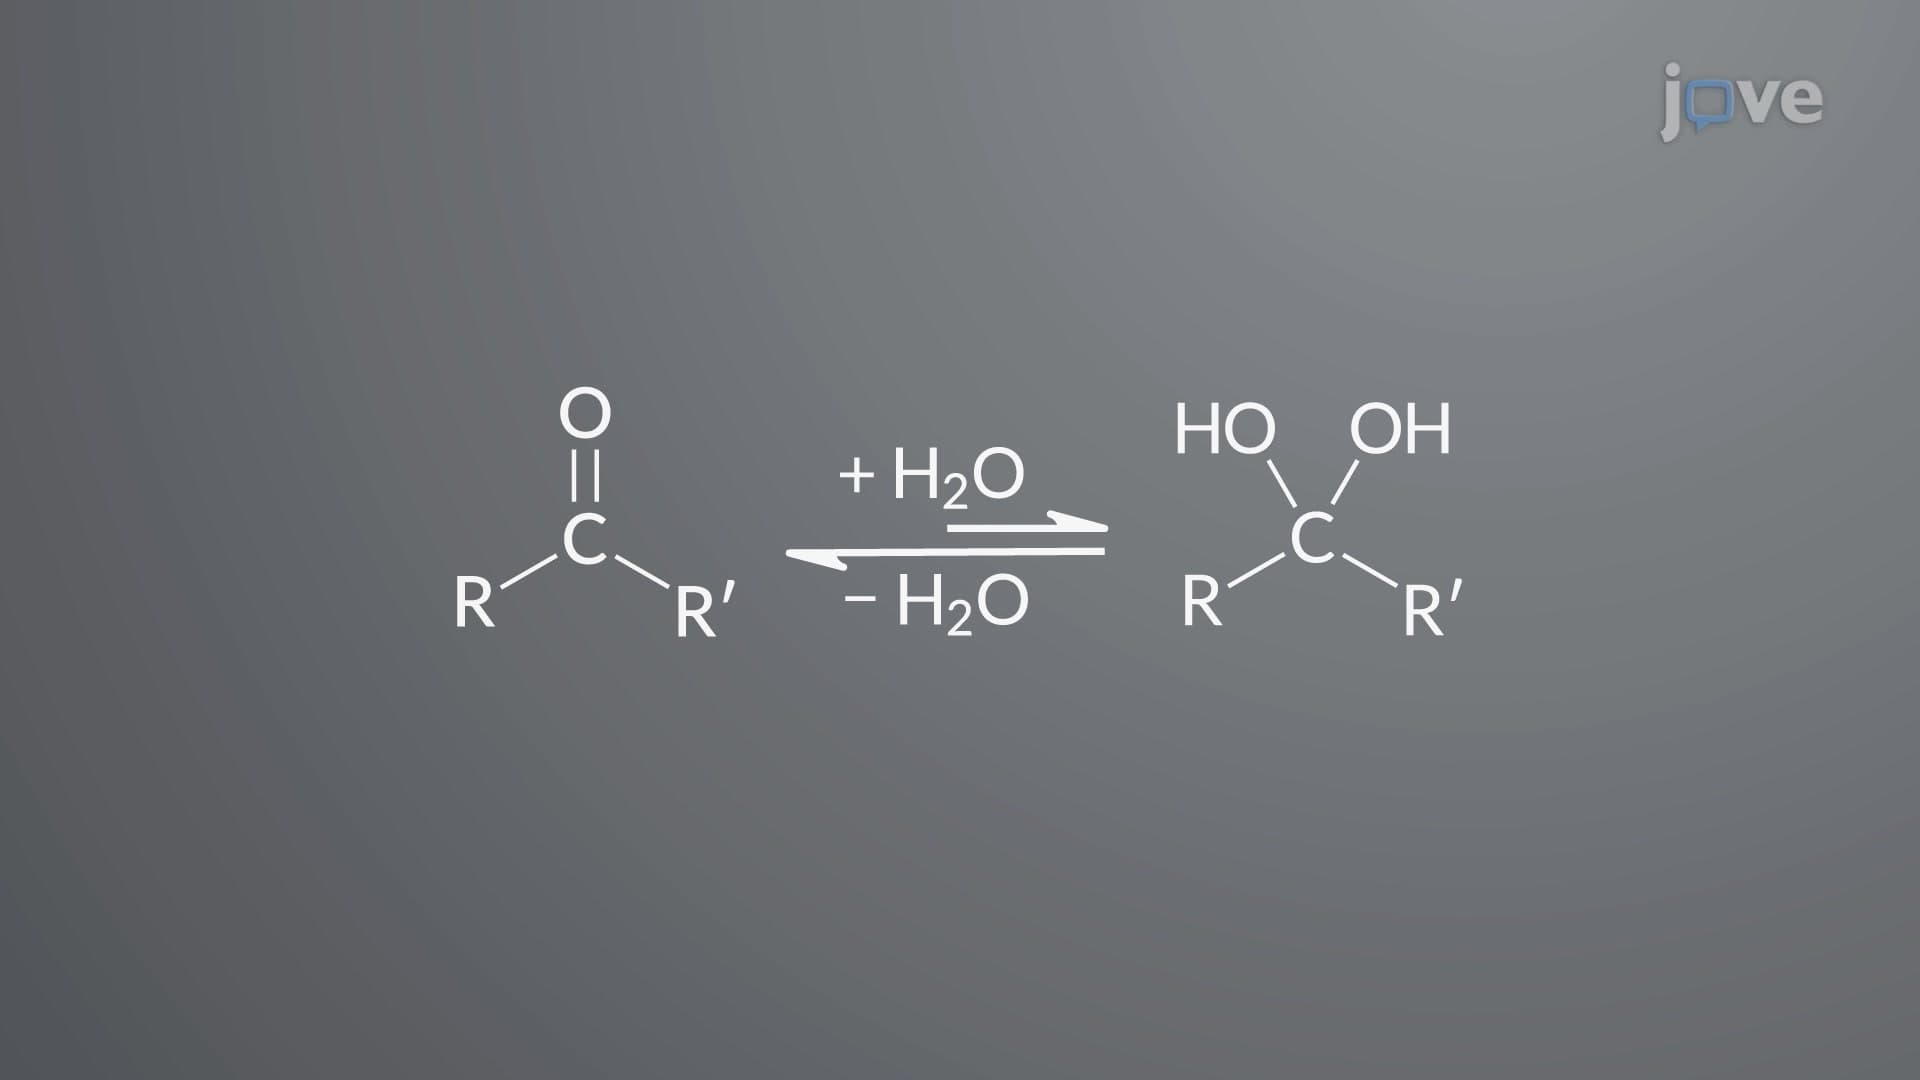
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 01:27
Radical Substitution: Allylic Bromination
 01:18
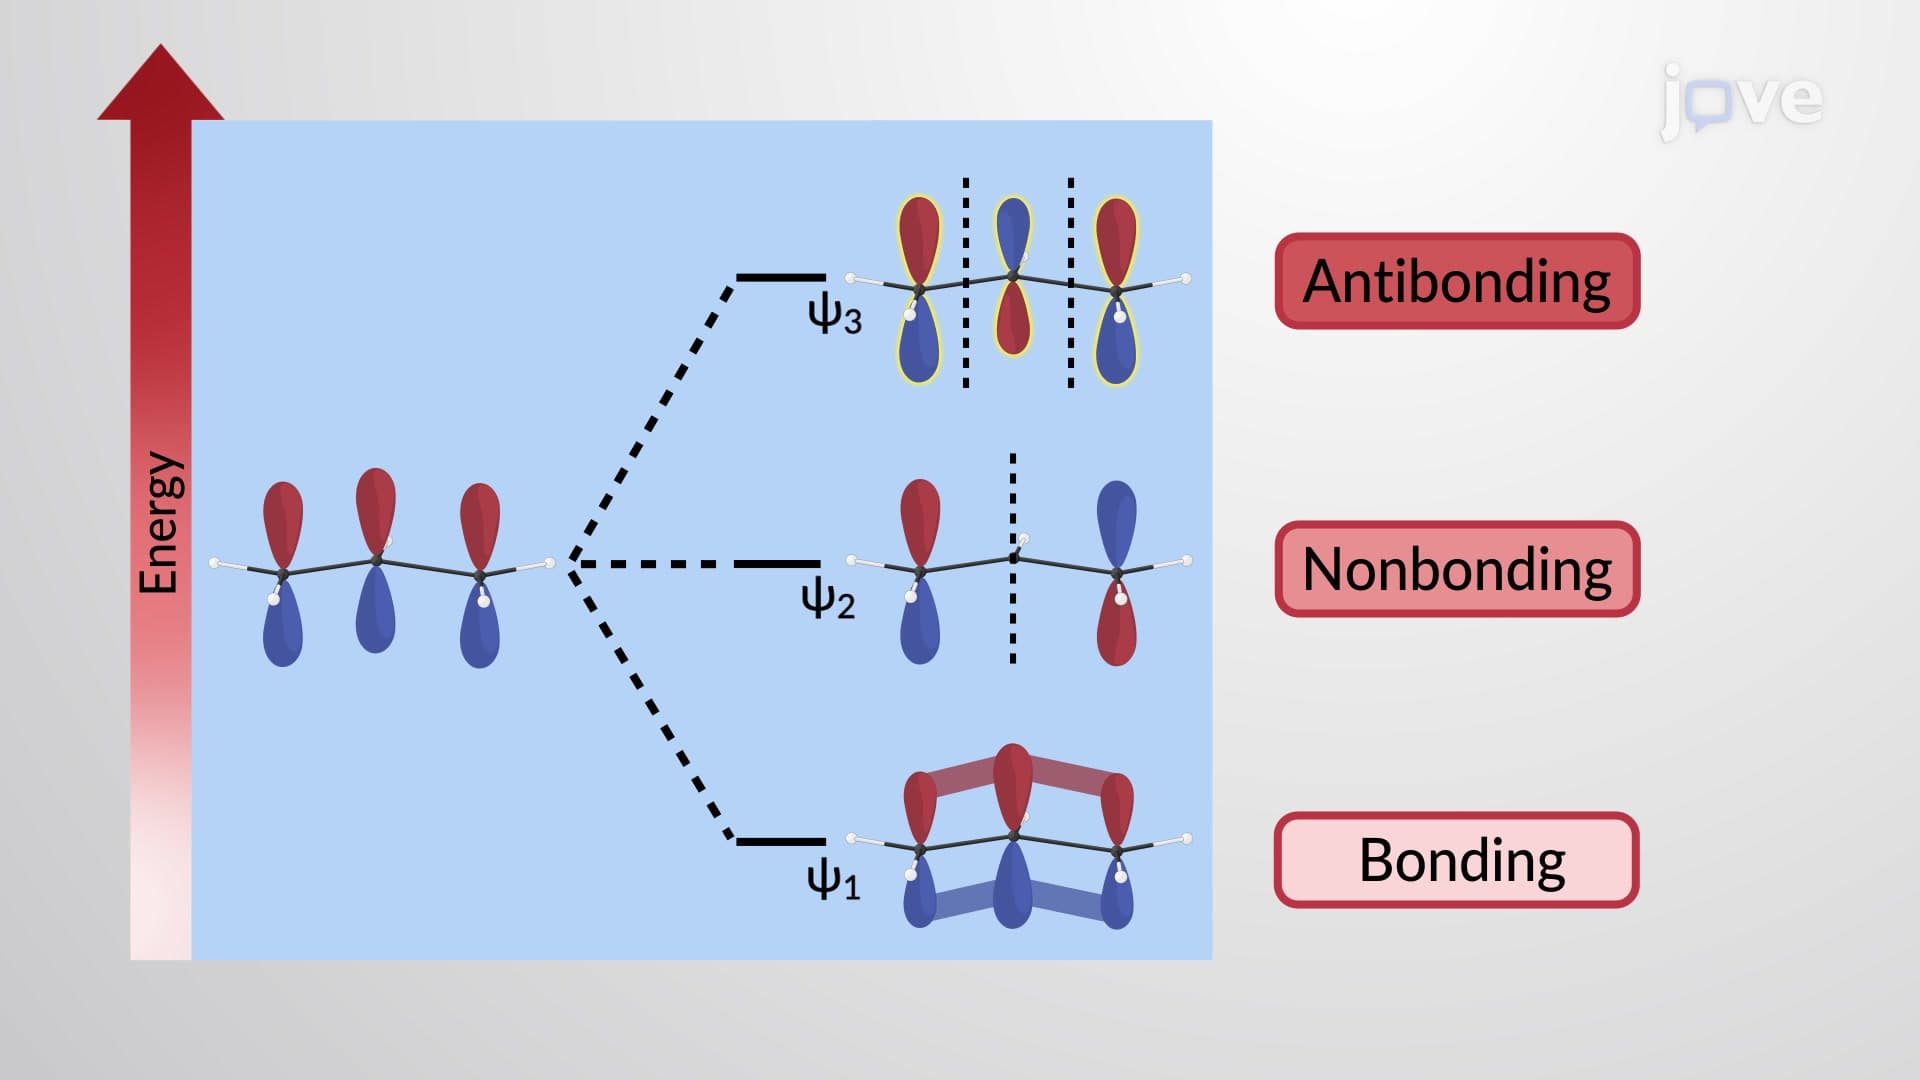
π Molecular Orbitals of the Allyl Cation and Anion
 01:16
[4+2] Cycloaddition of Conjugated Dienes: Diels–Alder Reaction