Related Experiment Video
Updated: Jan 19, 2026
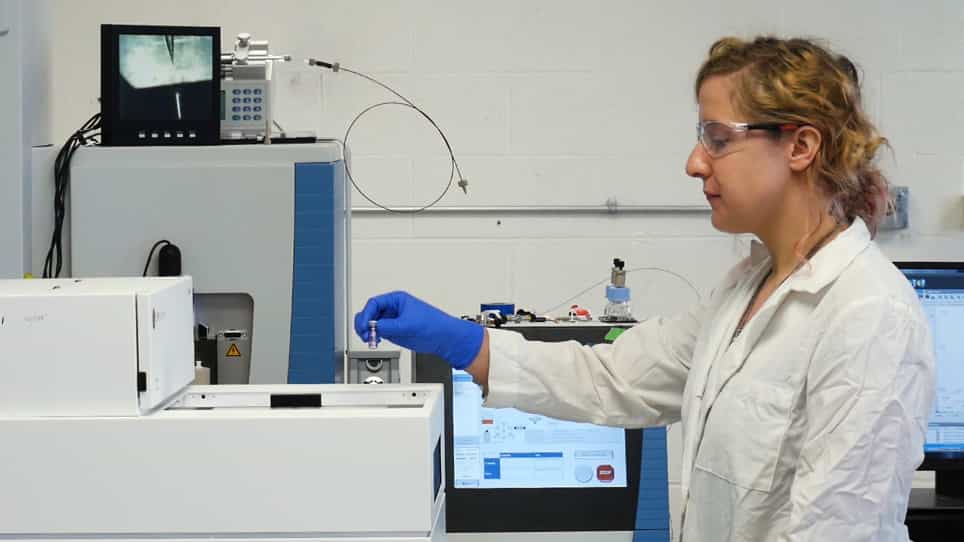

Large-scale Top-down Proteomics Using Capillary Zone Electrophoresis Tandem Mass Spectrometry
Published on: October 24, 2018
Optimization of mass spectrometric parameters improve the identification performance of capillary zone
Zhenbin Zhang1, Norman J Dovichi1
1Department of Chemistry and Biochemistry, University of Notre Dame, Notre Dame, IN 46556, USA.

Abstract:
The effects of MS1 injection time, MS2 injection time, dynamic exclusion time, intensity threshold, and isolation width were investigated on the numbers of peptide and protein identifications for single-shot bottom-up proteomics analysis using CZE-MS/MS analysis of a Xenopus laevis tryptic digest. An electrokinetically pumped nanospray interface was used to couple a linear-polyacrylamide coated capillary to a Q Exactive HF mass spectrometer. A sensitive method that used a 1.4 Th isolation width, 60,000 MS2 resolution, 110 ms MS2 injection time, and a top 7 fragmentation produced the largest number of identifications when the CZE loading amount was less than 100 ng. A programmable autogain control method (pAGC) that used a 1.4 Th isolation width, 15,000 MS2 resolution, 110 ms MS2 injection time, and top 10 fragmentation produced the largest number of identifications for CZE loading amounts greater than 100 ng; 7218 unique peptides and 1653 protein groups were identified from 200 ng by using the pAGC method. The effect of mass spectrometer conditions on the performance of UPLC-MS/MS was also investigated. A fast method that used a 1.4 Th isolation width, 30,000 MS2 resolution, 45 ms MS2 injection time, and top 12 fragmentation produced the largest number of identifications for 200 ng UPLC loading amount (6025 unique peptides and 1501 protein groups). This is the first report where the identification number for CZE surpasses that of the UPLC at the 200 ng loading level. However, more peptides (11476) and protein groups (2378) were identified by using UPLC-MS/MS when the sample loading amount was increased to 2 μg with the fast method. To exploit the fast scan speed of the Q-Exactive HF mass spectrometer, higher sample loading amounts are required for single-shot bottom-up proteomics analysis using CZE-MS/MS.
More Related Videos
Related Concept Videos
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:21
Mass Spectrometry: Complex Analysis
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...