Related Experiment Video
Updated: Feb 15, 2026

Protocol for the Synthesis of Ortho-trifluoromethoxylated Aniline Derivatives
Published on: January 19, 2016
The ortho-benzyne cation is not planar
D Kaiser1, E Reusch, P Hemberger
1Institute of Physical and Theoretical Chemistry, University of Würzburg, Am Hubland, D-97074 Würzburg, Germany. ingo.fischer@uni-wuerzburg.de bernd.engels@uni-wuerzburg.de.

Abstract:
A recent review on the photoionisation of the C6H4 isomer ortho-benzyne suggests that bands reported in earlier photoelectron spectra might be due to side products or contaminations, while computations raise doubts, whether the cation has a planar geometry. We therefore reinvestigate the photoionisation of ortho-benzyne, generated by pyrolysis from benzocyclobutenedione, by photoion mass-selected threshold photoelectron (ms-TPE) spectroscopy using synchrotron radiation. The experiments are accompanied by a theoretical study that investigates the structure of the ortho-benzyne cation systematically as a function of the computational method, up to CASPT2(11,14) ab initio computations. Our study leads to a re-evaluation of the ionisation energy of ortho-benzyne. It reveals that the ortho-benzyne cation has indeed a twisted C2 geometry rather than a C2v structure. A vertical ionisation energy IEvert of 9.77 eV and an adiabatic ionisation energy of IEad = 9.56 eV are computed for ortho-benzyne. A Franck-Condon simulation of the photoelectron spectrum based on the CASPT2 results and including three electronic states of the cation is in agreement with the experiment and yields IEad = 9.51 eV (+50 meV/-100 meV). Since this value is in contrast with previous work, the ionisation energy has to be revised based on our study. Computational methods based on density functional theory give a reasonable description of the cationic ground state, but fail for the corresponding excited electronic states that are indispensible for a proper assignment of the photoelectron spectrum.
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:14
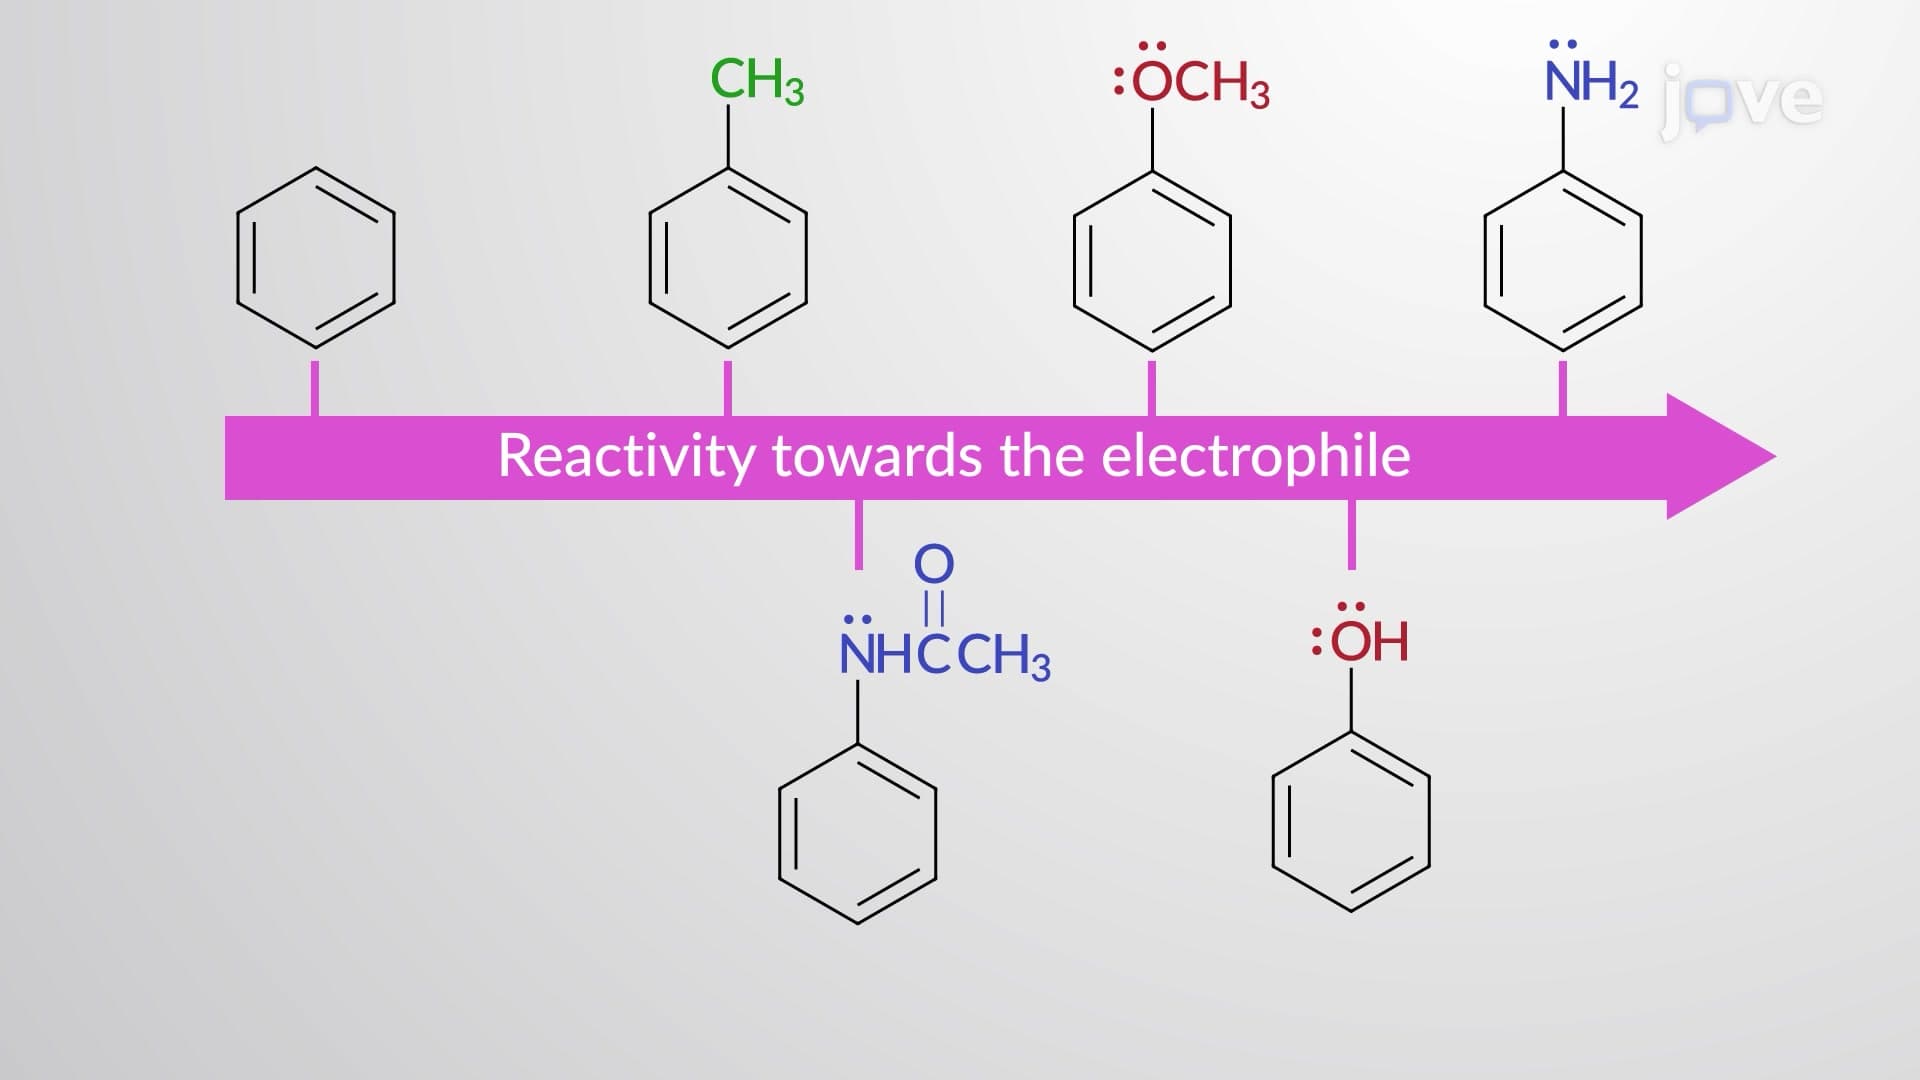
Directing Effect of Substituents: ortho–para-Directing Groups
 01:24
ortho–para-Directing Deactivators: Halogens
 01:22
Planar Rigid-Body Motion
Planar motion is typically divided into three distinct categories. The first is rectilinear translation, demonstrated by a subway train that moves along...
 01:27
Gauss's Law: Planar Symmetry
 00:57
Cationic Chain-Growth Polymerization: Mechanism