Related Experiment Video
Updated: Feb 13, 2026

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Composite and Low-Cost Approaches for Molecular Crystal Structure Prediction
Luc M LeBlanc1, Alberto Otero-de-la-Roza2, Erin R Johnson1
1Department of Chemistry , Dalhousie University , 6274 Coburg Road , P.O. Box 15000, Halifax , Nova Scotia , Canada B3H 4R2.

Abstract:
Molecular crystal structure prediction (CSP) requires evaluating differences in lattice energy between candidate crystal structures accurately and efficiently. In this work, we explore and compare several low-cost alternatives to dispersion-corrected density-functional theory (DFT) in the plane-waves/pseudopotential approximation, the most accurate and general approach used for CSP at present. Three types of low-cost methods are considered: DFT with a small basis set of finite-support numerical orbitals (the SIESTA method), dispersion-corrected Gaussian small or minimal-basis-set Hartree-Fock and DFT with additional empirical corrections (HF-3c and PBEh-3c), and self-consistent-charge dispersion-corrected density-functional tight binding (SCC-DFTB3-D3). In addition, we study the performance of composite methods that comprise a geometry optimization using a low-cost approach followed by a single-point calculation using the accurate but comparatively expensive B86bPBE-XDM functional. All methods were tested for their abilities to produce absolute lattice energies, relative lattice energies, and crystal geometries. We show that assessing various methods by their ability to produce absolute lattice energies can be misleading and that relative lattice energies are a much better indicator of performance in CSP. The EE14 set of relative solubilities of homochiral and heterochiral chiral crystals is proposed for relative lattice-energy benchmarking. Our results show that PBE-D2 plus a DZP basis set of numerical orbitals coupled with a final B86bPBE-XDM single-point calculation gives excellent performance at a fraction of the cost of a full B86bPBE-XDM calculation, although the results are sensitive to the particular details of the computational protocol. The B86bPBE-XDM//PBE-D2/DZP method was subsequently tested in a practical CSP application from our recent work on the crystal structure of the enantiopure and racemate forms of 1-aza[6]helicene, a chiral organic semiconductor. Our results show that this multilevel method is able to correctly reproduce the energy ranking of both crystal forms.

Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 01:22
Sperm Structure and Semen Composition
 02:34
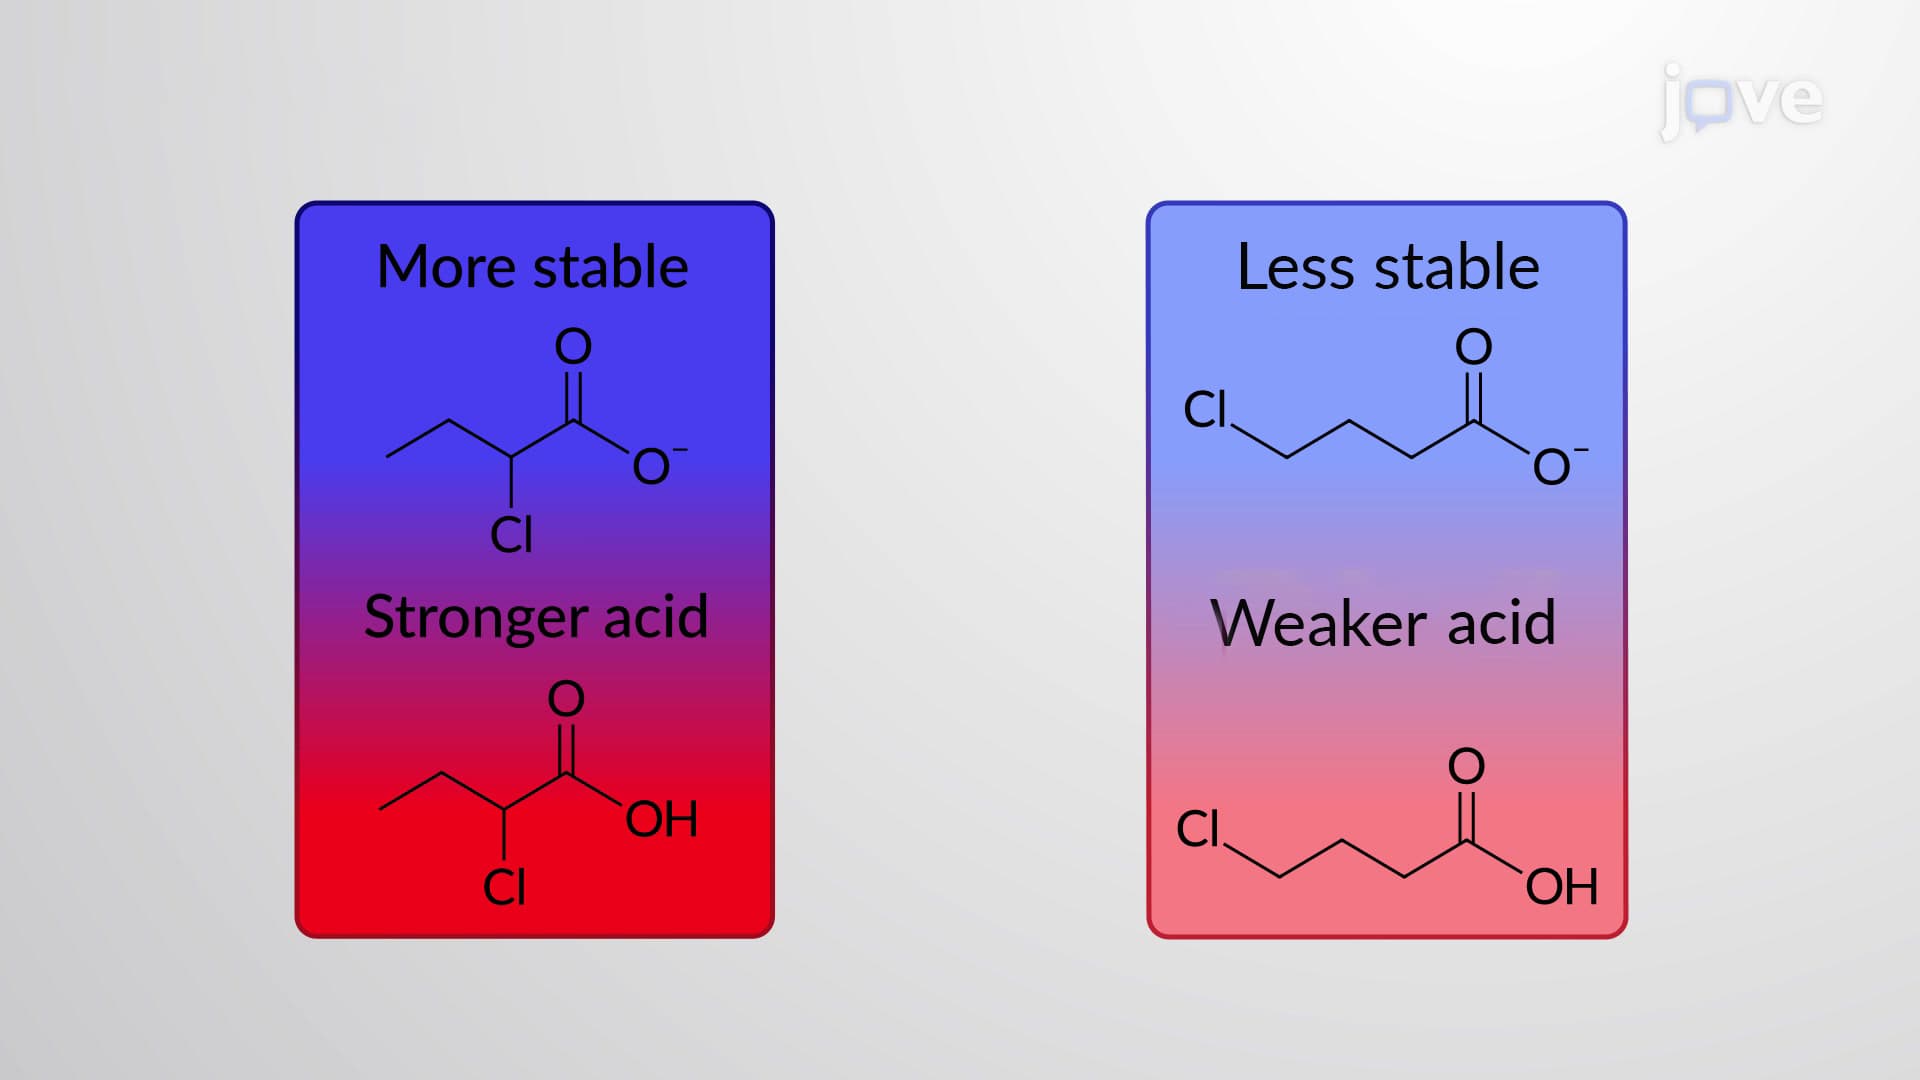
Molecular Structure and Acidity
The size effect explains the change in atomic size on acidity. When comparing the acids formed from elements that belong to the same column in the periodic table, their atomic sizes...
 02:54
Lewis Structures of Molecular Compounds and Polyatomic Ions
 03:05
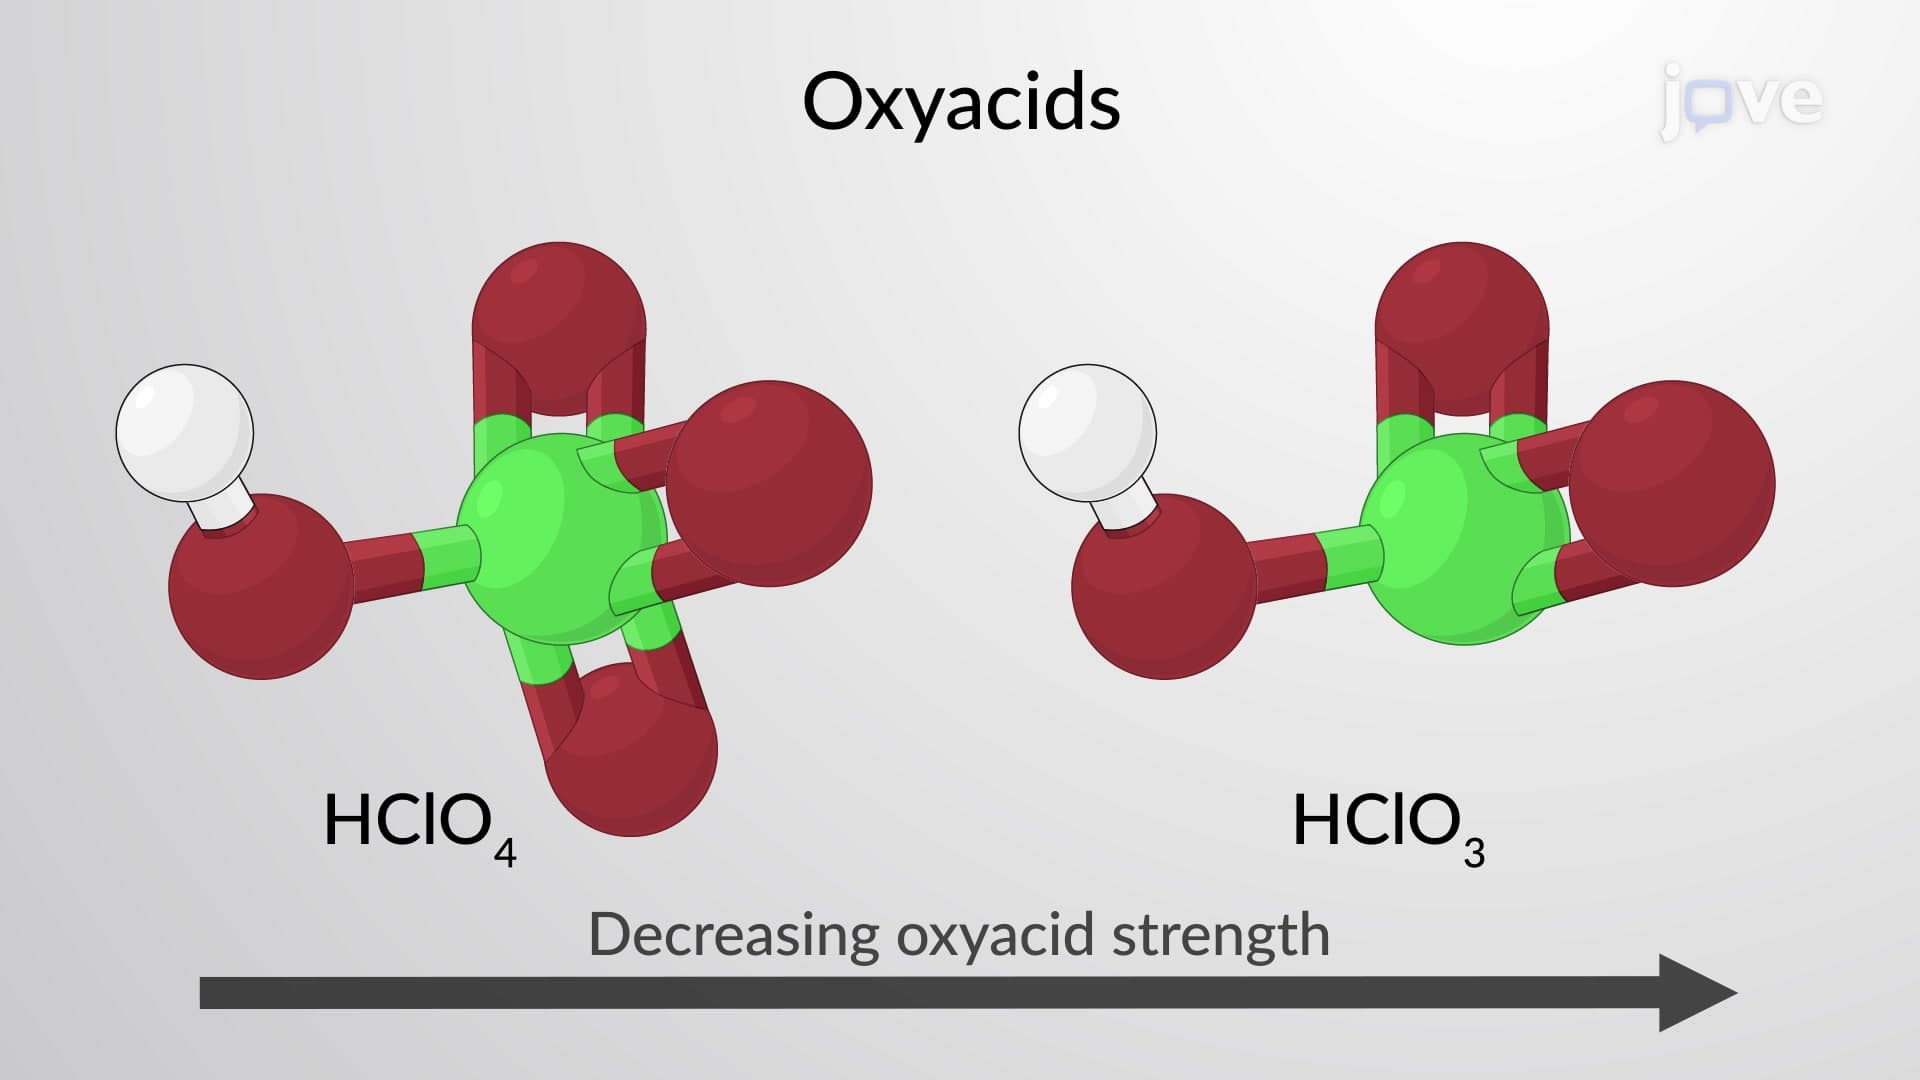
Acid Strength and Molecular Structure
In the absence of any leveling effect, the acid strength of binary compounds of hydrogen with nonmetals (A) increases as the H-A bond strength decreases down a group in the periodic table. For group 17, the order of increasing acidity is HF < HCl < HBr < HI. Likewise, for group 16, the order of increasing acid strength is H2O < H2S < H2Se < H2Te. Across a row in the periodic table, the acid strength of binary hydrogen compounds increases with increasing...