Suberanilohydroxamic acid prevents TGF-β1-induced COX-2 repression in human lung fibroblasts post-transcriptionally by TIA-1 downregulation
Related Experiment Video
Updated: Feb 13, 2026
Reprogramming Mouse Embryonic Fibroblasts with Transcription Factors to Induce a Hemogenic Program
Published on: December 16, 2016
Alice Pasini1, Oliver J Brand2, Gisli Jenkins2
1Division of Respiratory Medicine, University of Nottingham School of Medicine, City Hospital, Nottingham NG5 1PB, United Kingdom; Department of Electrical, Electronic and Information Engineering "Guglielmo Marconi" (DEI), University of Bologna, Via Venezia 52, 47521 Cesena, FC, Italy.

Abstract:
Cyclooxygenase-2 (COX-2), with its main antifibrotic metabolite PGE2, is regarded as an antifibrotic gene. Repressed COX-2 expression and deficient PGE2 have been shown to contribute to the activation of lung fibroblasts and excessive deposition of collagen in pulmonary fibrosis. We have previously demonstrated that COX-2 expression in lung fibroblasts from patients with idiopathic pulmonary fibrosis (IPF) is epigenetically silenced and can be restored by epigenetic inhibitors. This study aimed to investigate whether COX-2 downregulation induced by the profibrotic cytokine transforming growth factor-β1 (TGF-β1) in normal lung fibroblasts could be prevented by epigenetic inhibitors. We found that COX-2 protein expression and PGE2 production were markedly reduced by TGF-β1 and this was prevented by the pan-histone deacetylase inhibitor suberanilohydroxamic acid (SAHA) and to a lesser extent by the DNA demethylating agent Decitabine (DAC), but not by the G9a histone methyltransferase (HMT) inhibitor BIX01294 or the EZH2 HMT inhibitor 3-deazaneplanocin A (DZNep). However, chromatin immunoprecipitation assay revealed that the effect of SAHA was unlikely mediated by histone modifications. Instead 3'-untranslated region (3'-UTR) luciferase reporter assay indicated the involvement of post-transcriptional mechanisms. This was supported by the downregulation by SAHA of the 3'-UTR mRNA binding protein TIA-1 (T-cell intracellular antigen-1), a negative regulator of COX-2 translation. Furthermore, TIA-1 knockdown by siRNA mimicked the effect of SAHA on COX-2 expression. These findings suggest SAHA can prevent TGF-β1-induced COX-2 repression in lung fibroblasts post-transcriptionally through a novel TIA-1-dependent mechanism and provide new insights into the mechanisms underlying its potential antifibrotic activity.
Insights
The antifibrotic gene cyclooxygenase-2 (COX-2) is downregulated by TGF-β1. Epigenetic inhibitor SAHA prevents this repression via a novel post-transcriptional mechanism involving TIA-1, offering potential antifibrotic therapy.
Area of Science:
- Molecular Biology
- Cell Biology
- Pulmonary Medicine
Background:
- Cyclooxygenase-2 (COX-2) and its metabolite PGE2 are antifibrotic.
- Reduced COX-2/PGE2 contributes to pulmonary fibrosis by activating lung fibroblasts.
- Epigenetic silencing of COX-2 in idiopathic pulmonary fibrosis (IPF) can be reversed by inhibitors.
Purpose of the Study:
- To investigate if epigenetic inhibitors can prevent transforming growth factor-β1 (TGF-β1)-induced COX-2 downregulation in normal lung fibroblasts.
- To elucidate the underlying molecular mechanisms of COX-2 regulation by TGF-β1 and potential epigenetic intervention.
Main Methods:
- Treatment of normal lung fibroblasts with TGF-β1 and various epigenetic inhibitors (SAHA, Decitabine, BIX01294, DZNep).
- Assessment of COX-2 protein expression and PGE2 production.
- Chromatin immunoprecipitation assays, 3'-untranslated region (3'-UTR) luciferase reporter assays, and small interfering RNA (siRNA) knockdown of TIA-1.
Main Results:
- TGF-β1 significantly reduced COX-2 protein and PGE2 production.
- SAHA and Decitabine, but not BIX01294 or DZNep, prevented TGF-β1-induced COX-2 downregulation.
- SAHA's effect was mediated by post-transcriptional mechanisms, specifically the downregulation of TIA-1, a negative regulator of COX-2 translation, independent of histone modifications.
Conclusions:
- SAHA prevents TGF-β1-induced COX-2 repression in lung fibroblasts through a novel TIA-1-dependent post-transcriptional mechanism.
- These findings offer new insights into the antifibrotic potential of SAHA and its mechanism of action.
- The study highlights a new therapeutic target and strategy for managing pulmonary fibrosis.
Related Concept Videos
 01:52
Eukaryotic Transcription Inhibitors
Eukaryotic transcription inhibitors usually contain two distinct domains, a...
 02:42
Acidity of 1-Alkynes
The acidic strength of hydrocarbons follows the order: Alkynes > Alkenes > Alkanes. The strength of an acid is commonly expressed in units of pKa — the lower the pKa, the stronger the acid. Among the hydrocarbons, terminal alkynes have lower pKa values and are, therefore, more acidic. For example, the pKa values for ethane, ethene, and acetylene are 51, 44, and 25, respectively, as shown here.
 02:27
Predicting Products: SN1 vs. SN2
With increased substitution on the alkyl halide,...
 02:16
Transcription Factors
 02:23
Master Transcription Regulators
 02:42
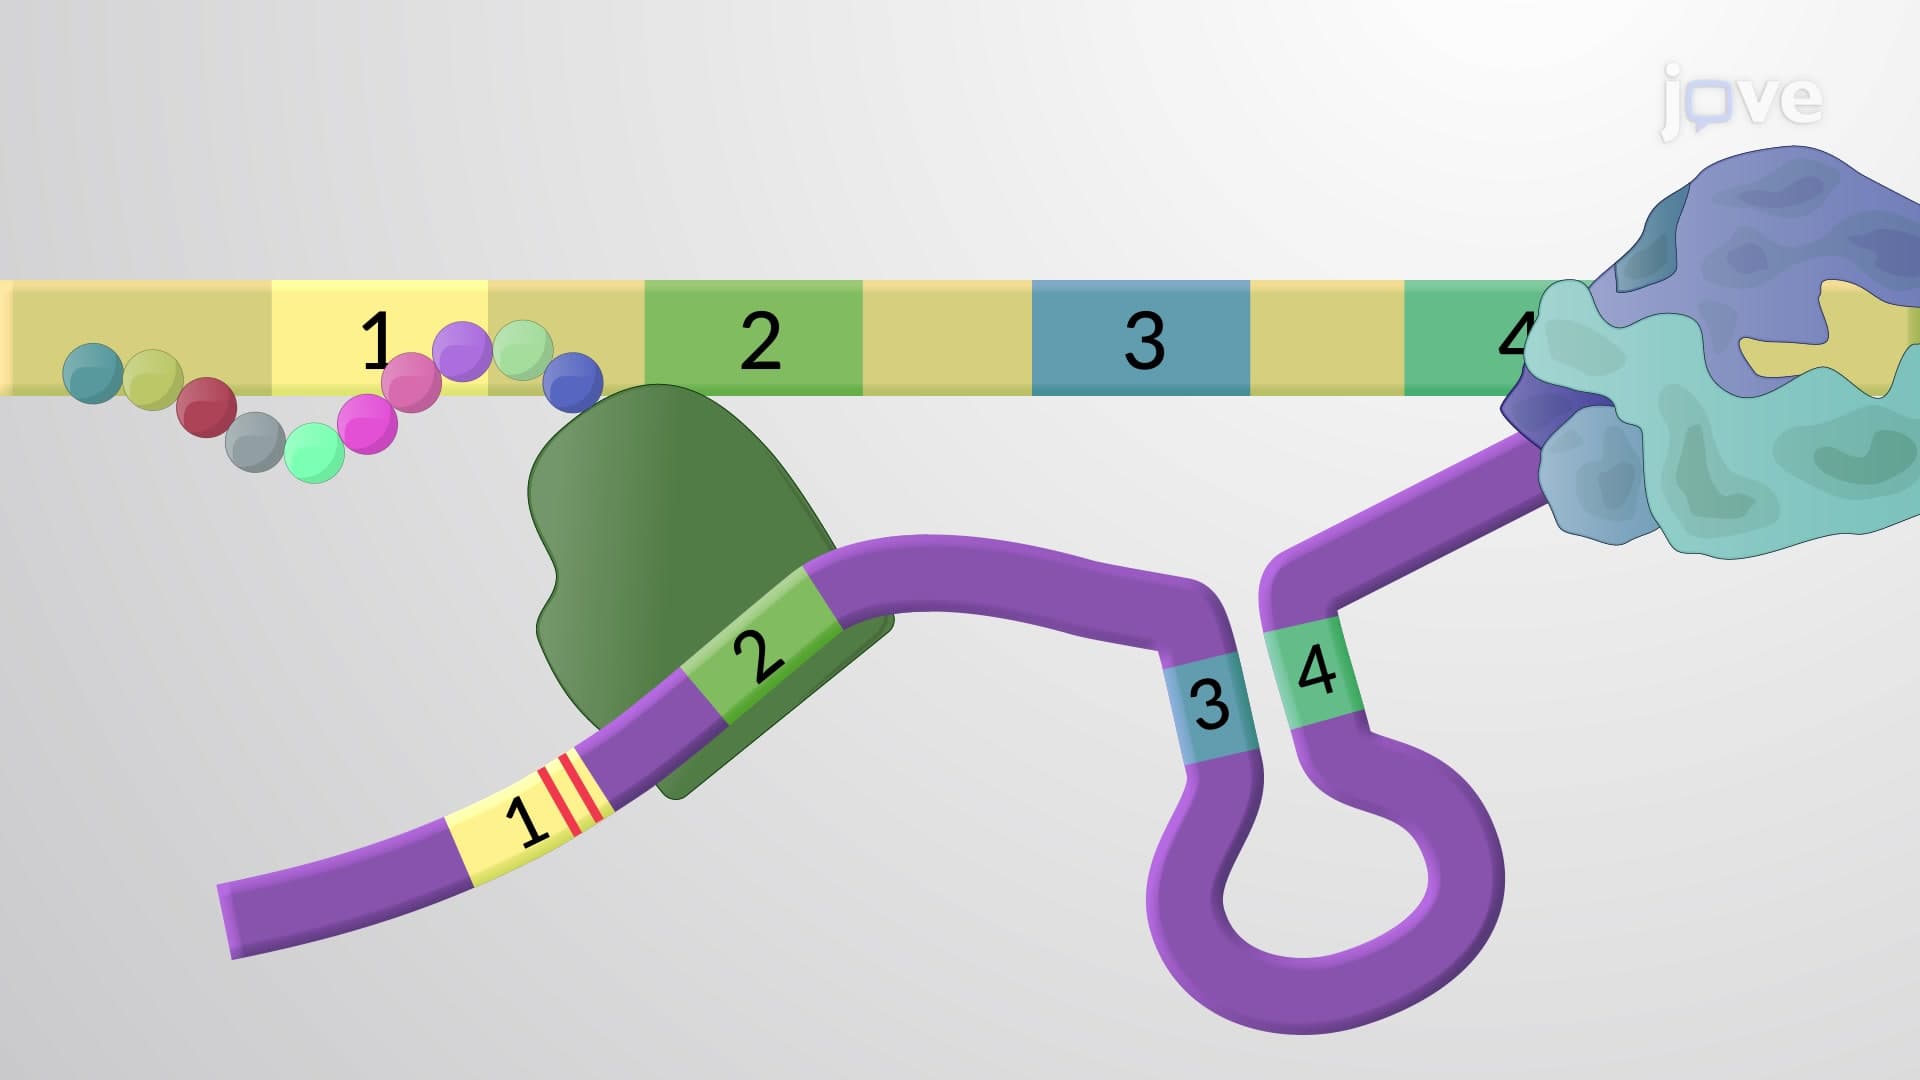
Transcription Attenuation in Prokaryotes
There are several different mechanisms used to attenuate transcription. In ribosome mediated...